Related Experiment Video
Updated: Jun 26, 2025
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Exploring the effects of mono-bromination on hole-electron transport and distribution in dibenzofuran and
1Computational & Theoretical Physics Laboratory, PG & Research Department of Physics, Thiruvalluvar Govt. Arts College, Rasipuram, 637408, Tamil Nadu, India. deepakvijaysalem@gmail.com.

Context:
This study delves into hole-electron transport and distribution properties inherent in mono-brominated dibenzofuran (DBF) and dibenzothiophene (DBT) isomers. As determined by frontier molecular orbitals, all brominated structures have narrower bandgaps than their primary structures. The TD-DFT calculation showed that 2BDBT had the highest absorption wavelength of all molecules at 315.35 nm. Notably, the study unveils remarkably low electron and hole reorganization energies due to bromine substitution in DBF and DBT molecules. Specifically, the 4BDBF has the lowest hole reorganization energy of all DBF configurations, 0.229 eV. In addition, 3BDBF has 0.226 eV less electron reorganization energy than all other molecules. Compared to DBT, 3BDBT has the lowest electron reorganization energy of 0.254 eV. Overall, this research sheds significant light on the fundamental electronic and hole transport characteristics of bromine-substituted DBF and DBT isomers, highlighting their promising role in polymer design as donors/acceptors for advanced organic electronic applications.
Methods:
Molecular structures were optimized using Density Functional Theory (DFT) B3LYP/6-311 + + G (d, p) level of theory, and the study further elucidates these molecules' energy levels and absorption spectra through Time-Dependent Density Functional Theory TD-DFT; these calculations were performed using Gaussian 09W software package. The key parameters such as reorganization energies, Electron Localization Function map, Laplacian Bond Order, and NCI-RDG were meticulously examined for the molecules with the results of DFT calculations were analyzed and displayed by utilizing the software packages VMD 1.9.4 and Multiwfn 3.8, aiming to comprehend their charge transport and distribution properties.
Related Concept Videos
 01:13
Electrophilic Aromatic Substitution: Fluorination and Iodination of Benzene
 02:46
Halogenation of Alkenes
Consider the bromination of cyclopentene. Molecular bromine is polarized in the proximity of the π electrons of cyclopentene. An electrophilic bromine atom adds across the double bond, forming a cyclic bromonium ion intermediate.
 01:18
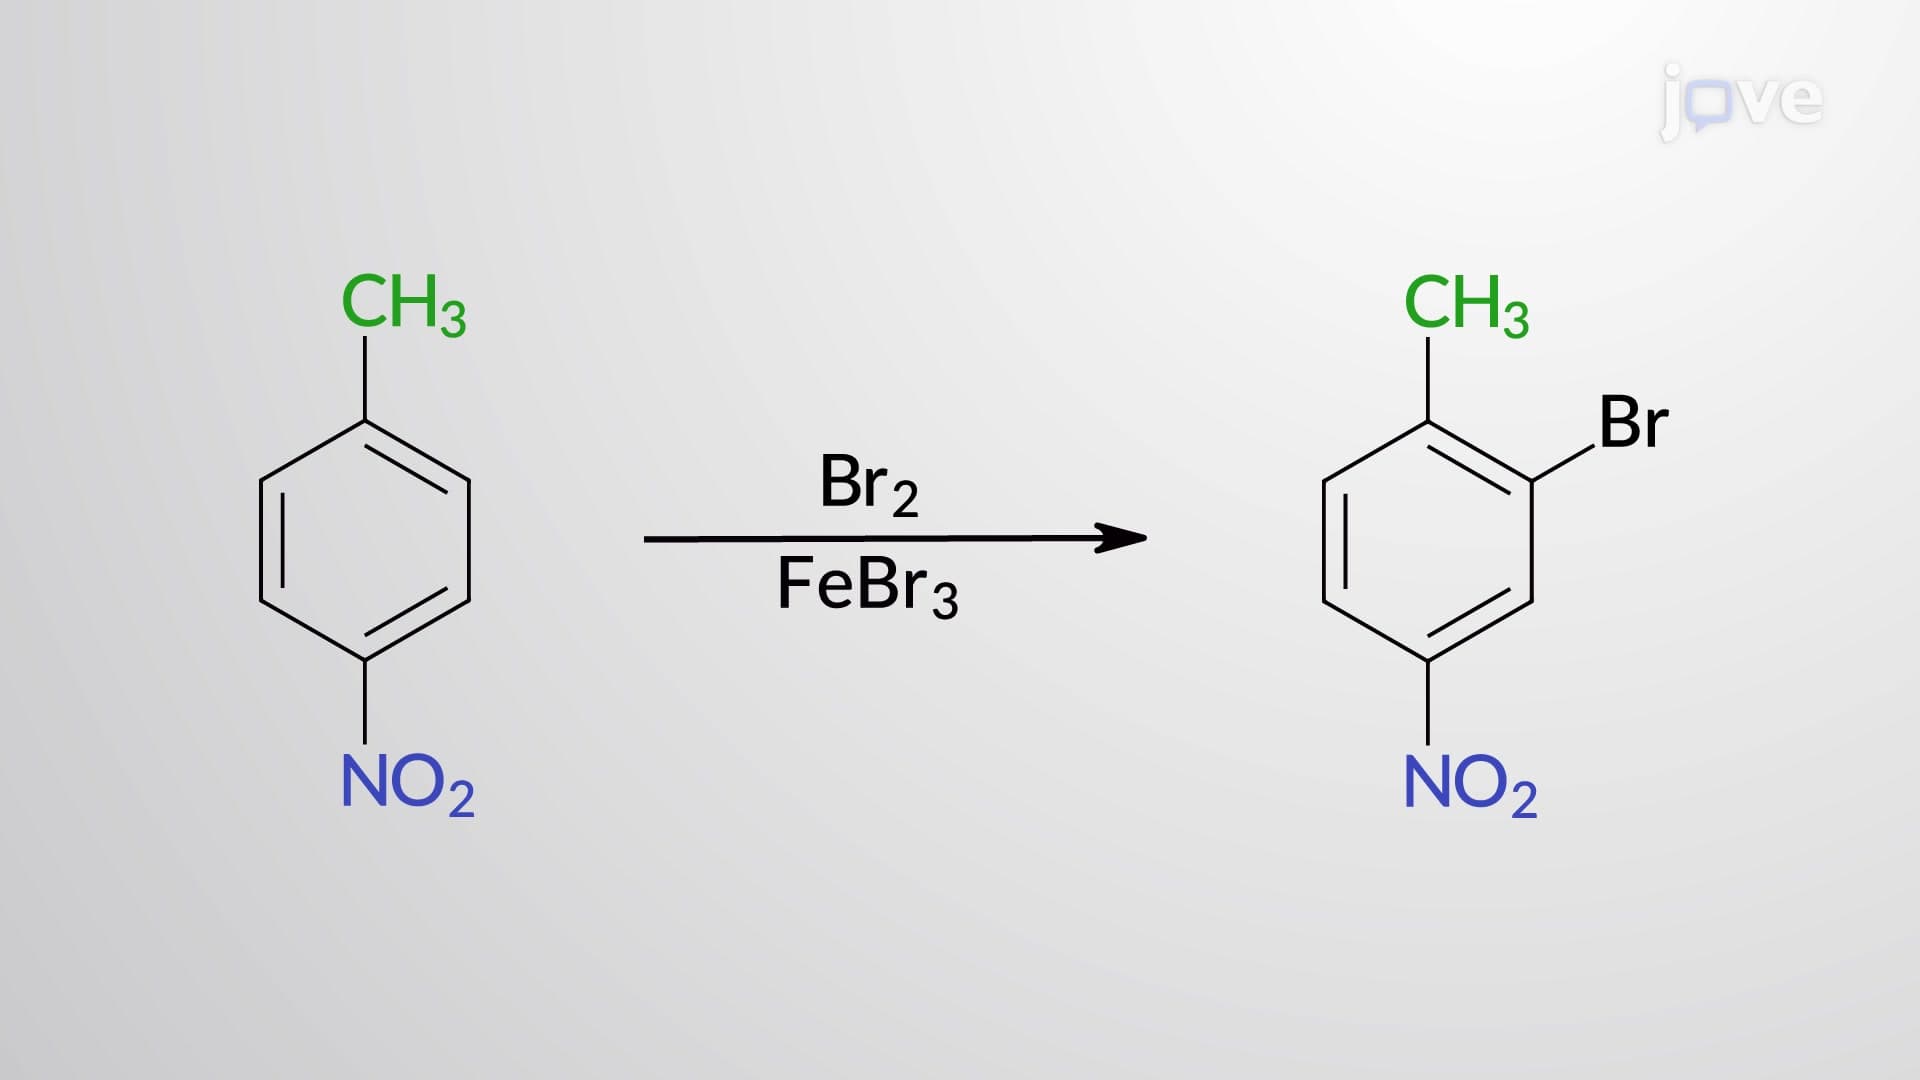
Directing and Steric Effects in Disubstituted Benzene Derivatives
 04:01
VSEPR Theory and the Effect of Lone Pairs
 01:34
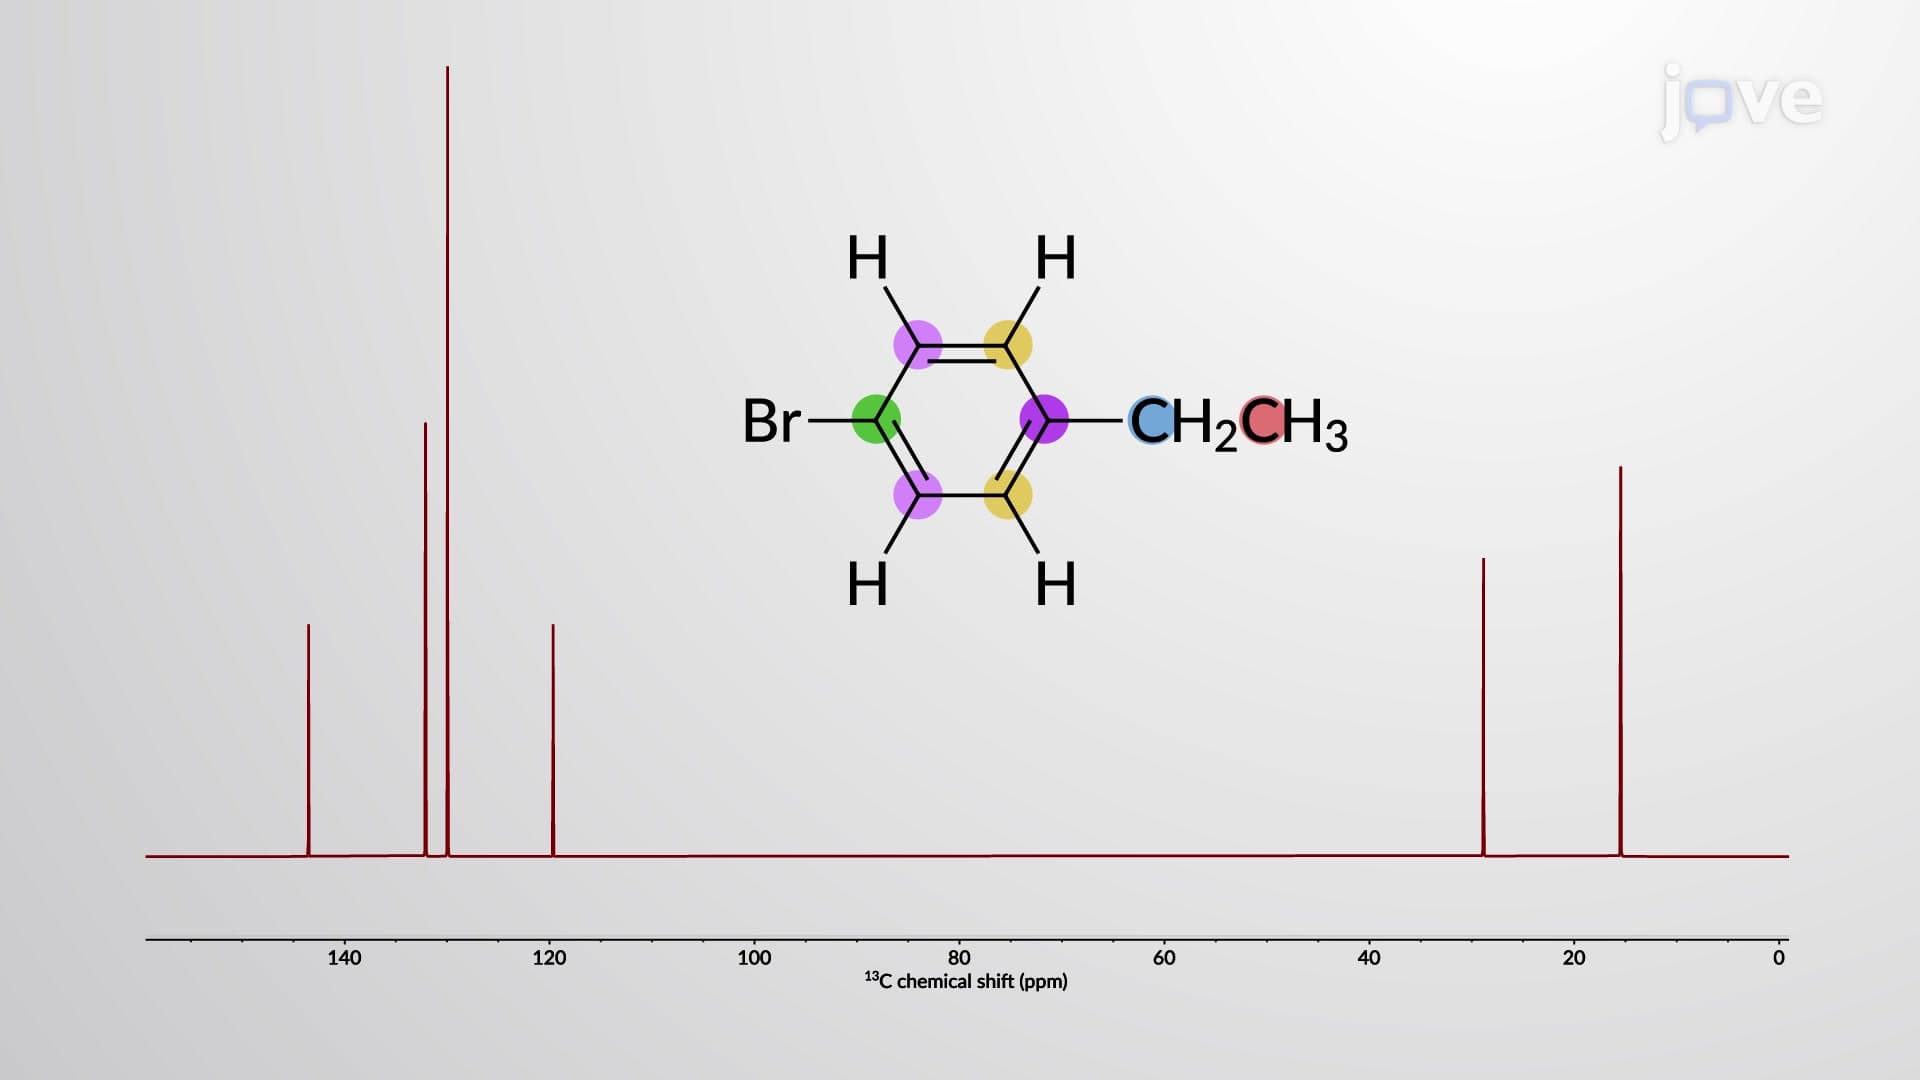
NMR Spectroscopy of Benzene Derivatives
 01:15
Electrophilic Aromatic Substitution: Chlorination and Bromination of Benzene