Related Experiment Video
Updated: Oct 15, 2026
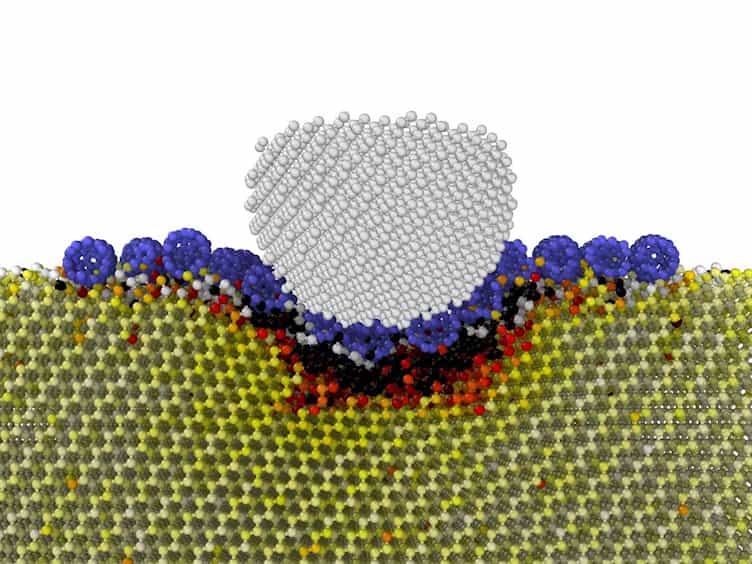
Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Atomic-level engineering of multifunctional BixC16 nanocages for hydrogen storage, thermoelectric energy conversion,
Belqees Hassan1, Manahil E E Mofdal2
1Department of Physics, College of Science, Qassim University, Buraydah, 51452, Saudi Arabia. B.hassan@qu.edu.sa.

Context:
The development of stable carbon-based nanomaterials with tunable electronic, magnetic, thermoelectric, and hydrogen storage properties is important for emerging energy conversion and nanoelectronic technologies. Here, Bi-doped C16 nanocages, BixC16 (x = 1-4), corresponding to Bi concentrations of 5.8, 11.1, 15.7, and 20.0 wt%, were systematically investigated. Negative formation energies, phonon spectra without imaginary modes, and 300 K ab initio molecular dynamics simulations confirm the energetic, dynamical, and thermal stability of the considered nanocages. Increasing Bi incorporation produces progressive structural expansion and bond elongation while maintaining the integrity of the C16 framework. Strong Bi-6p/C-2p hybridization modifies the electronic structure and enhances spin polarization. The magnetic anisotropy energy and Curie temperature increase with Bi concentration, reaching approximately 3.8 meV cell-1 and 920 K, respectively, for BixC16. Bi doping also enhances thermoelectric performance, with a maximum Seebeck coefficient of 340 μV K⁻1 and a peak ZT of 2.78 at 1200 K. Hydrogen storage calculations yield maximum gravimetric and volumetric capacities of 6.7 wt% and 67.4 g H2 L⁻1, respectively. Among the investigated systems, it exhibits the most favorable combination of stability, magnetic properties, thermoelectric performance, and hydrogen storage capability, highlighting its potential for multifunctional energy and nanoelectronic applications.
Methods:
The structural, electronic, magnetic, thermodynamic, hydrogen storage, and thermoelectric properties of Bi-doped C16 nanocage were investigated using spin-polarized density functional theory (DFT) calculations implemented in the CASTEP package [1, 2]. The electron exchange-correlation interactions were described using the generalized gradient approximation (GGA) within the Perdew-Burke-Ernzerhof (PBE) functional. The interaction between valence electrons and ionic cores was treated using ultrasoft pseudopotential, while the electronic wave functions were expanded using a plane-wave basis set with a kinetic energy cutoff of 600 eV, which was confirmed to provide converged total energies and forces [3]. Geometry optimization was performed using the Broyden-Fletcher-Goldfarb-Shanno (BFGS) minimization algorithm until the convergence criteria of 1.0 × 10⁻6 eV atom⁻1 for total energy, 0.01 eV Å⁻1 for maximum force, 0.02 GPa for stress, and 5.0 × 10⁻4 Å for atomic displacement were simultaneously satisfied. The isolated Bi-doped C16 nanocage was modeled within a 20 × 20 × 20 Å3 cubic supercell to eliminate interactions between periodic images. Owing to the molecular nature of the nanocage, Brillouin-zone integration was performed using a Γ-point (1 × 1 × 1 Monkhorst-Pack) sampling grid. Spin polarization was included throughout all calculations because Bi incorporation can induce magnetic moments through orbital hybridization and spin-orbit interactions. Formation energies were calculated to evaluate energetic stability, while optimized bond lengths, cell volume, and mass density were obtained from the fully relaxed structures. Electronic properties were investigated by calculating the band structure, spin-polarized density of states (DOS), and partial density of states (PDOS) along the high-symmetry path of the Brillouin zone. Spin-orbit coupling (SOC) was incorporated in the electronic and magnetic calculations to accurately describe the relativistic effects associated with the heavy Bi atoms. Magnetic properties, including spontaneous magnetization, exchange coupling, magnetic anisotropy energy (MAE), and Curie temperature, were obtained from spin-polarized total energy calculations. The exchange coupling constants were evaluated by comparing different magnetic configurations within the Heisenberg model, whereas the Curie temperatures were estimated using the mean field approximation. Lattice dynamical stability was examined by calculating phonon dispersion curves using density functional perturbation theory (DFPT). The absence of imaginary phonon frequencies was taken as evidence of dynamical stability. Thermodynamic quantities, including the vibrational free energy (Fvib), entropy (S), constant-volume heat capacity (CV), and vibrational enthalpy (Hvib), were evaluated within the quasi-harmonic approximation (QHA) over the temperature range of 0-1200 K. The heat capacity was compared with the classical Dulong-Petit limit to verify the validity of the phonon calculations at elevated temperatures [4]. Thermoelectric transport coefficients were determined by solving the semi-classical Boltzmann transport equation within the constant relaxation time approximation (CRTA) using the BoltzTraP2 code [5, 6]. Temperature-dependent Seebeck coefficient, electrical conductivity, electronic thermal conductivity, and thermoelectric figure of merit (ZT) were calculated between 0 and 1200 K. Hydrogen storage properties were investigated by identifying the energetically favorable H₂ adsorption sites on the Bi-doped nanocage. , denote the total energies of the hydrogen-adsorbed nanocage, the pristine Bi-doped nanocage, and an isolated H2 molecule, respectively. Gravimetric and volumetric hydrogen storage capacities, adsorption isotherms, Van't Hoff desorption behavior, binding enthalpy, and diffusion barriers were subsequently evaluated from the optimized adsorption configurations. Phonon dispersion calculations were performed using the finite displacement method within density functional theory (DFT) to evaluate the dynamical stability of the optimized structures. The phonon frequencies and dispersion relations were obtained throughout the Brillouin zone using an appropriate supercell and k-point sampling, with the absence of imaginary frequencies taken as evidence of dynamical stability. To examine thermal stability under operating conditions, ab initio molecular dynamics (AIMD) simulations were performed in the canonical (NVT) ensemble using the Nosé-Hoover thermostat at 300 K for 10 ps with a time step of 1 fs. The combination of DFT, phonon analysis, Boltzmann transport calculations, hydrogen adsorption modeling, and AIMD simulations provides a comprehensive theoretical framework for evaluating the multifunctional properties of Bi-doped C16 nanocage for applications in hydrogen storage, thermoelectric energy conversion, spintronics, and nanoelectronics.
More Related Videos
 08:40
Synthesis of Metal Nanoparticles Supported on Carbon Nanotube with Doped Co and N Atoms and its Catalytic Applications in Hydrogen Production
Published on: December 6, 2021
 08:07
Ultrahigh Density Array of Vertically Aligned Small-molecular Organic Nanowires on Arbitrary Substrates
Published on: June 18, 2013