Related Experiment Video
Updated: Jun 25, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Quantum Quality with Classical Cost: Ab Initio Nonadiabatic Dynamics Simulations Using the Mapping Approach to
Jonathan R Mannouch1, Aaron Kelly1
1Hamburg Center for Ultrafast Imaging, Universität Hamburg and the Max Planck Institute for the Structure and Dynamics of Matter, Luruper Chaussee 149, 22761 Hamburg, Germany.

Abstract:
Nonadiabatic dynamics methods are an essential tool for investigating photochemical processes. In the context of employing first-principles electronic structure techniques, such simulations can be carried out in a practical manner using semiclassical trajectory-based methods or wave packet approaches. While all approaches applicable to first-principles simulations are necessarily approximate, it is commonly thought that wave packet approaches offer inherent advantages over their semiclassical counterparts in terms of accuracy and that this trait simply comes at a higher computational cost. Here we demonstrate that the mapping approach to surface hopping (MASH), a recently introduced trajectory-based nonadiabatic dynamics method, can be efficiently applied in tandem with ab initio electronic structure. Our results even suggest that MASH may provide more accurate results than on-the-fly wave packet techniques, all at a much lower computational cost.

Related Concept Videos
 02:45
The Quantum-Mechanical Model of an Atom
 03:24
Hybridization of Atomic Orbitals I
 03:35
Hybridization of Atomic Orbitals II
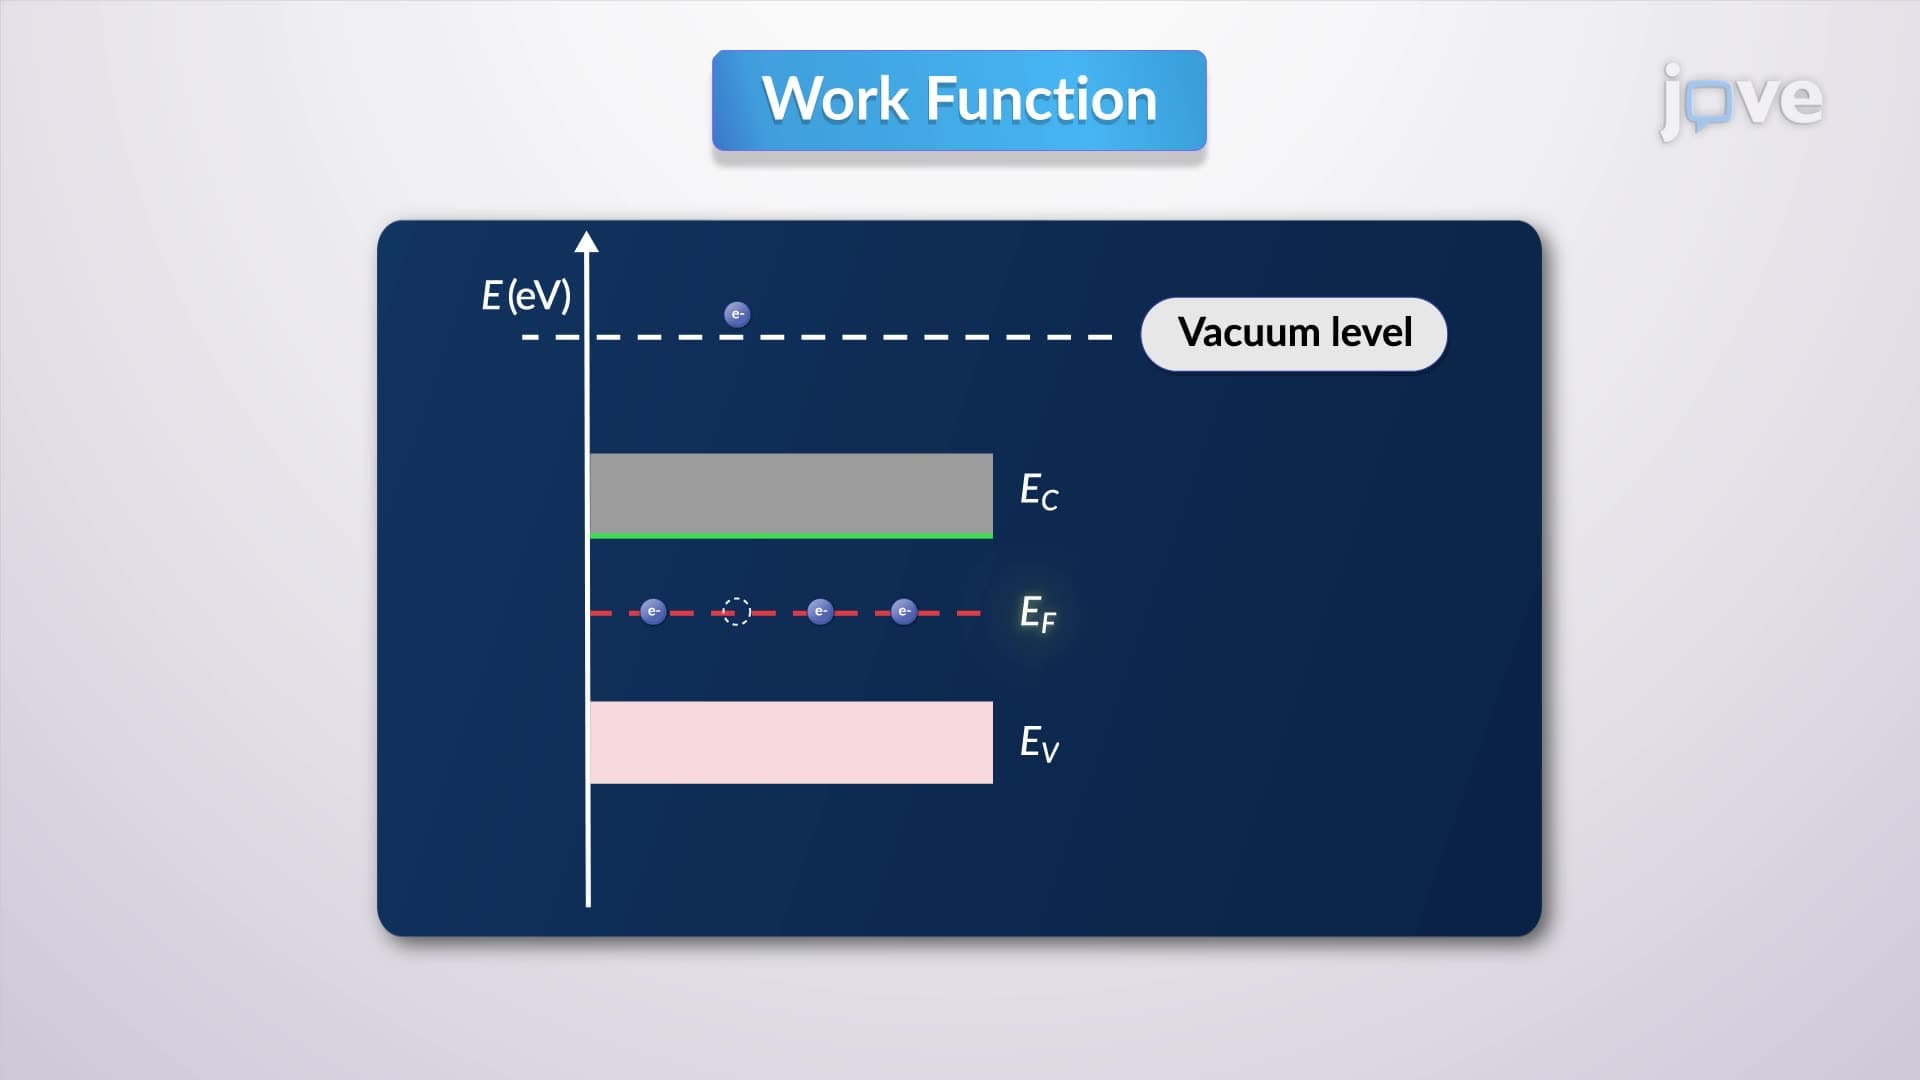 01:12
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
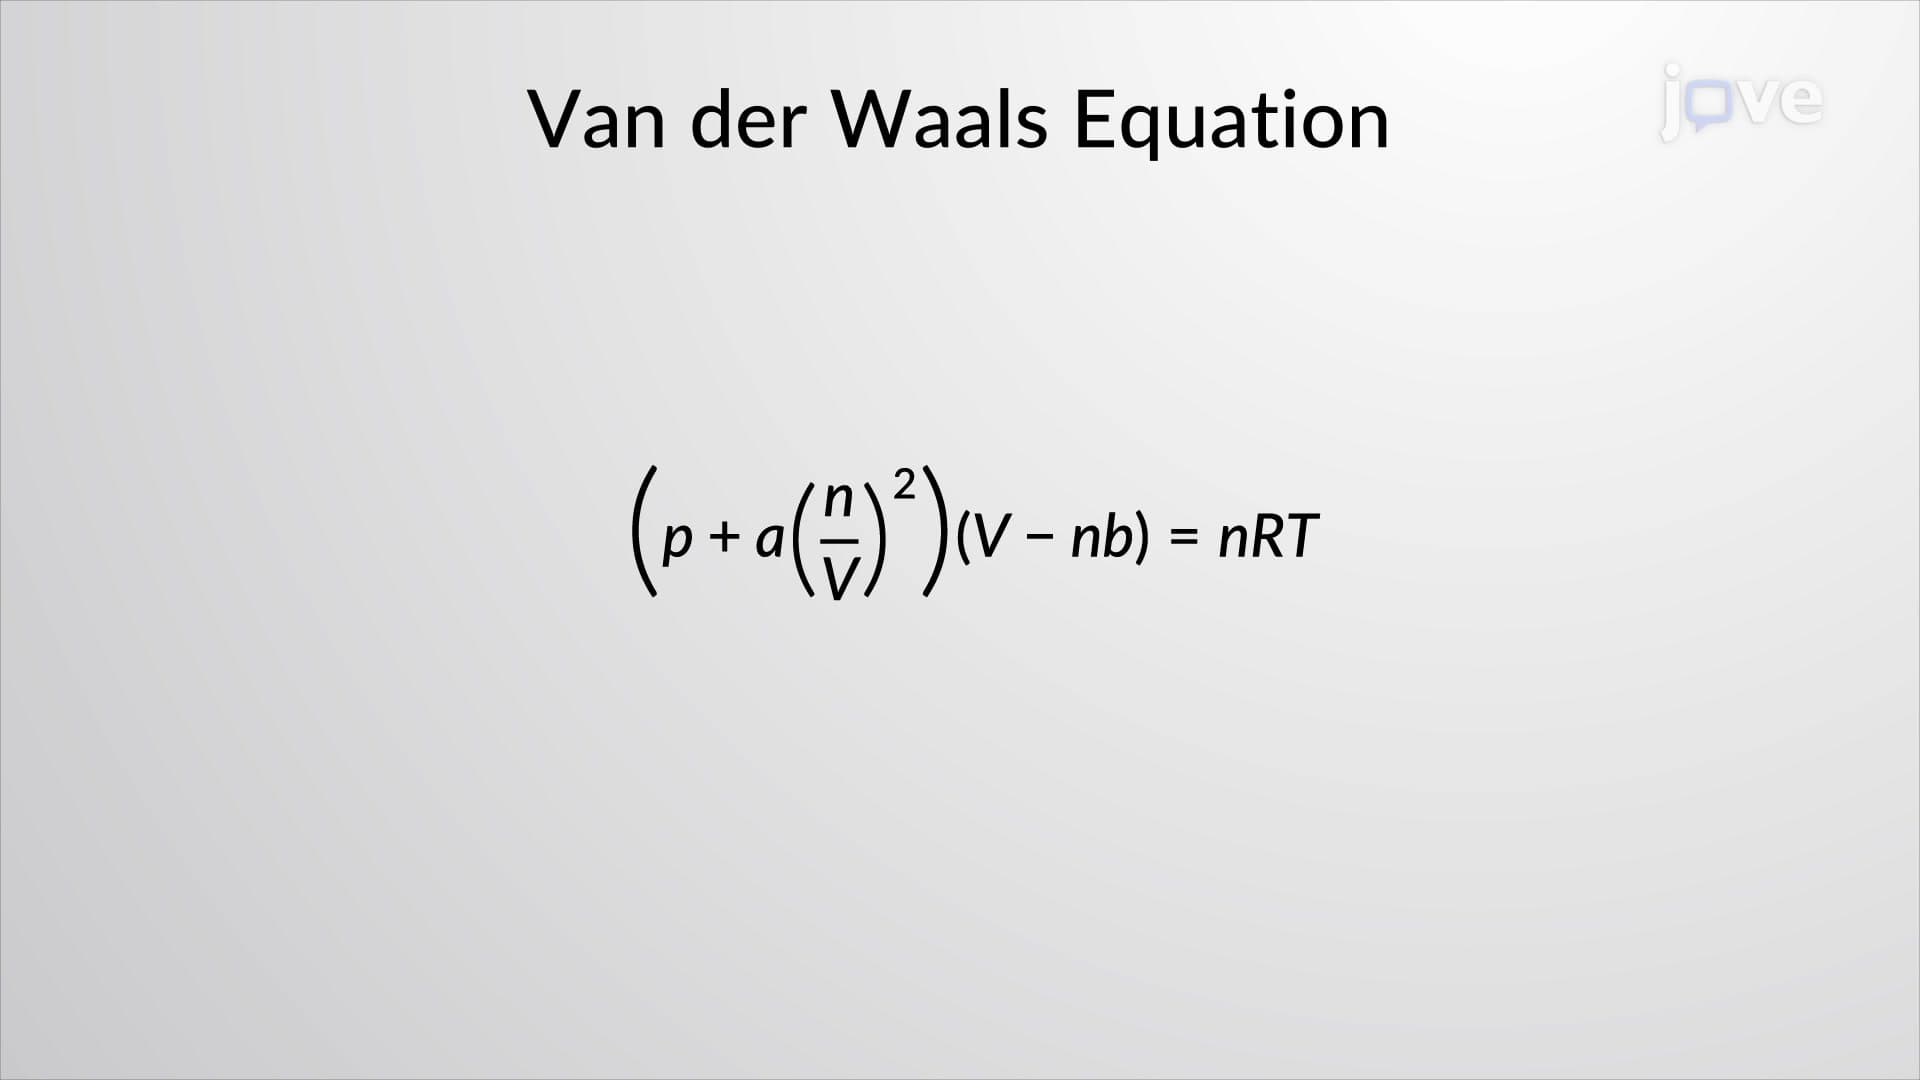 01:10
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:58
Cooperative Allosteric Transitions