Related Experiment Video
Updated: Jun 24, 2025

Synthesis and Performance Characterizations of Transition Metal Single Atom Catalyst for Electrochemical CO2 Reduction
Published on: April 10, 2018
Understanding the coupling of non-metallic heteroatoms to CO2 from a Conceptual DFT perspective
Maxime Ferrer1,2, José Elguero1, Ibon Alkorta3
1Instituto de Química Médica, CSIC, Juan de La Cierva,3, 28006, Madrid, Spain.

Context:
A Conceptual DFT (CDFT) study has been carry out to analyse the coupling reactions of the simplest amine (CH3NH2), alcohol (CH3OH), and thiol (CH3SH) compounds with CO2 to form the corresponding adducts CH3NHCO2H, CH3OCO2H, and CH3SCO2H. The reaction mechanism takes place in a single step comprising two chemical events: nucleophilic attack of the non-metallic heteroatoms to CO2 followed by hydrogen atom transfer (HAT). According to our calculations, the participation of an additional nucleophilic molecule as HAT assistant entails important decreases in activation electronic energies. In such cases, the formation of a six-membered ring in the transition state (TS) reduces the angular stress with respect to the non-assisted paths, characterised by four-membered ring TSs. Through the analysis of the energy and reaction force profiles along the intrinsic reaction coordinate (IRC), the ratio of structural reorganisation and electronic rearrangement for both activation and relaxation energies has been computed. In addition, the analysis of the electronic chemical potential and reaction electronic flux profiles confirms that the highest electronic activity as well as their changes take place in the TS region. Finally, the distortion/interaction model using an energy decomposition scheme based on the electron density along the reaction coordinate has been carried out and the relative energy gradient (REG) method has been applied to identify the most important components associated to the barriers.
Methods:
The theoretical calculation were performed with Gaussian-16 scientific program. The B3LYP-D3(BJ)/aug-cc-pVDZ level was used for optimization of the minima and TSs. IRC calculations has also been carried out connecting the TS with the associated minima. Conceptual-DFT (CDFT) calculations have been carried out with the Eyringpy program and in-house code. The distortion/interaction model along the reaction coordinate have used the decomposition scheme of Mandado et al. and the analysis of the importance of each components have been done with the relative energy gradient (REG) method.
More Related Videos

Related Concept Videos
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...
 02:42
Valence Bond Theory
 01:22
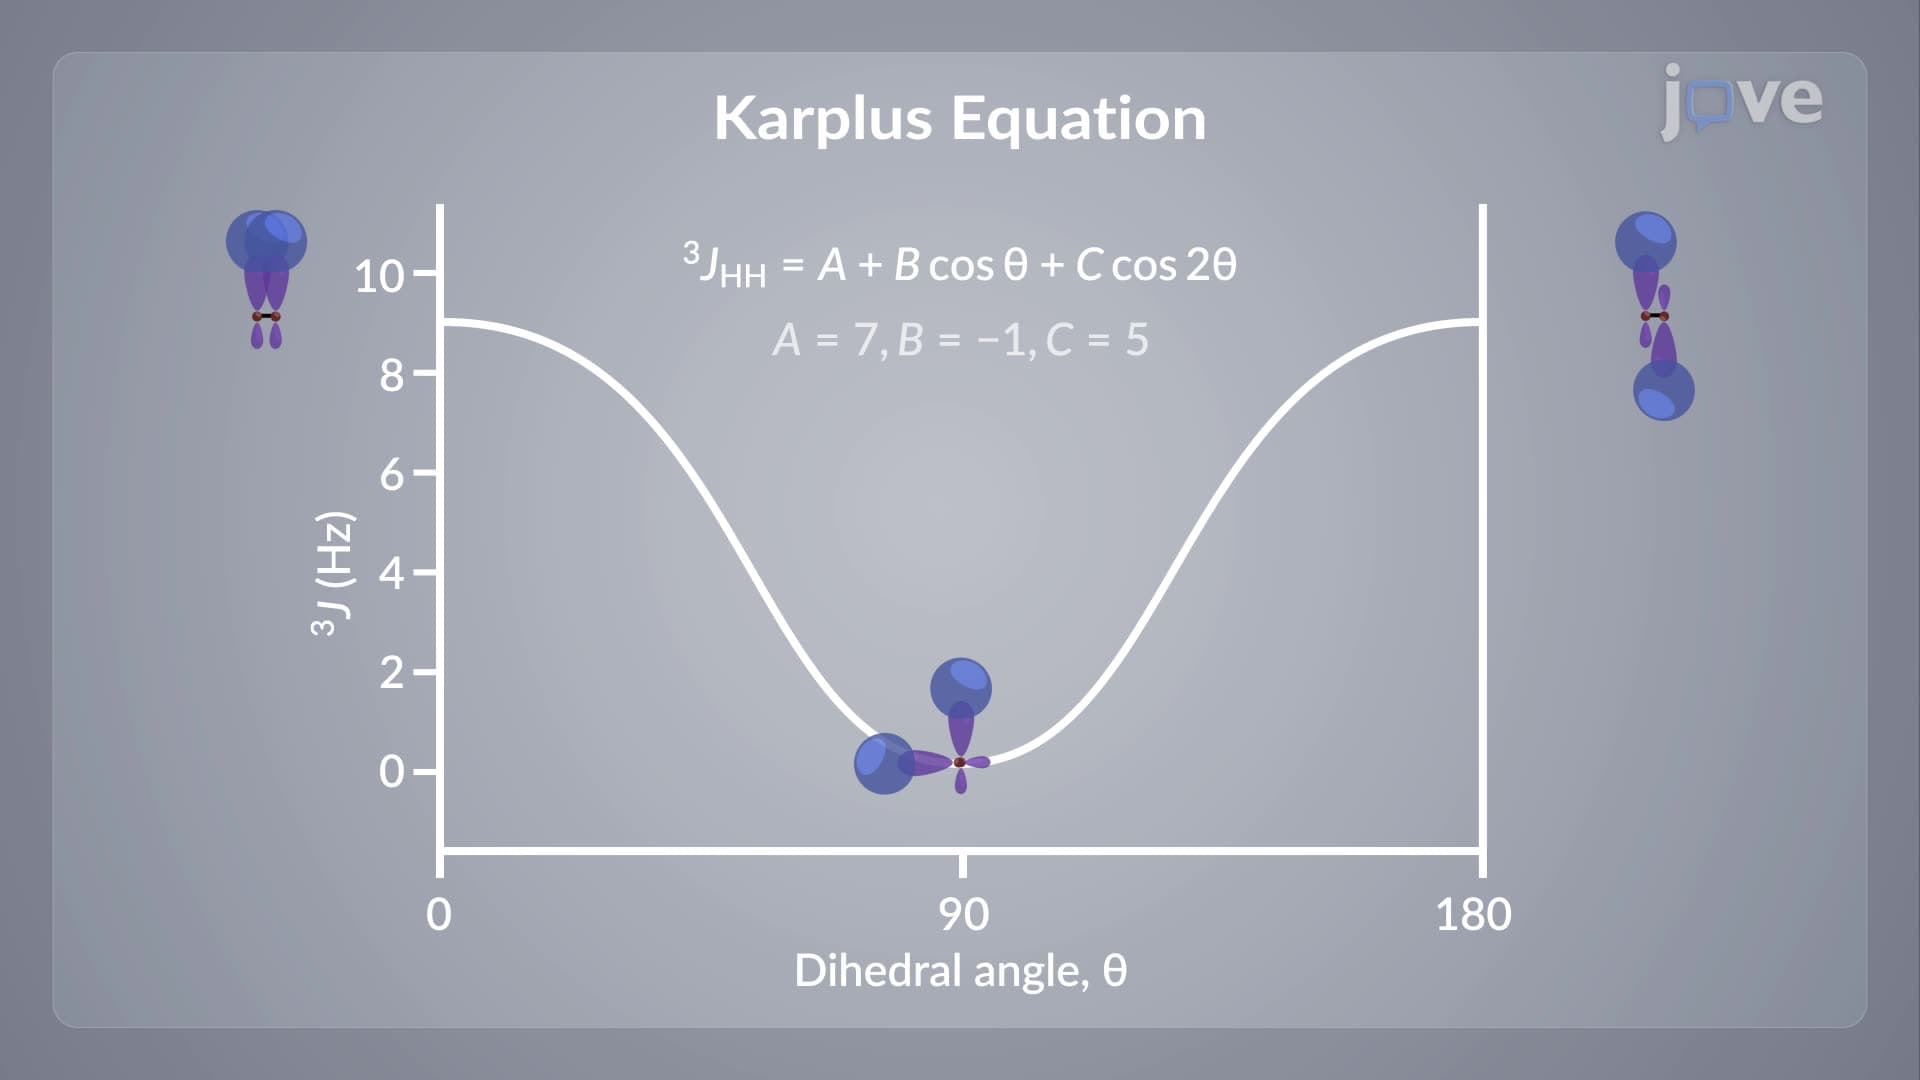
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
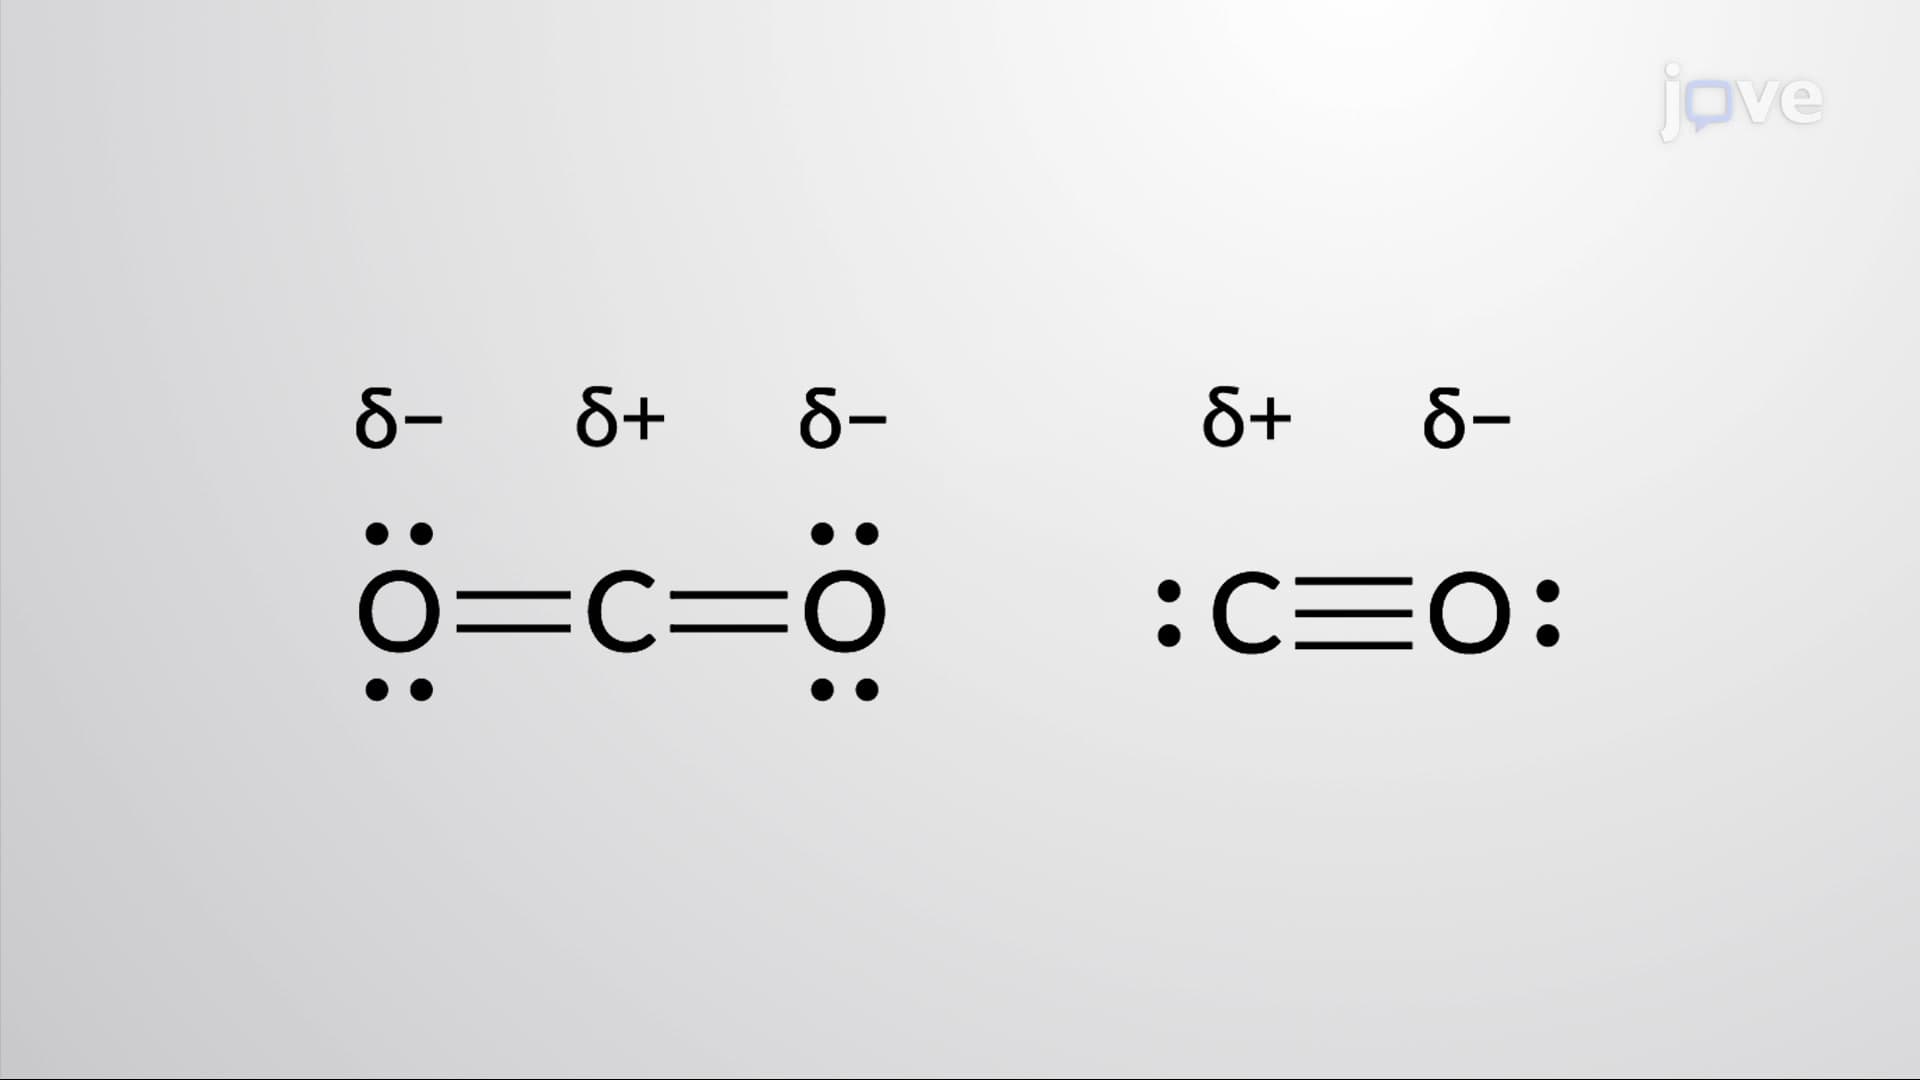 02:24
Polar Covalent Bonds
 02:32
Bonding in Metals