Related Experiment Video
Updated: Jun 24, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Nonadiabatic molecular dynamics with subsystem density functional theory: application to crystalline pentacene
Qingxin Zhang1, Xuecheng Shao2, Wei Li3
1Department of Chemistry, University at Buffalo, The State University of New York, Buffalo, NY 14260, United States of America.

Abstract:
In this work, we report the development and assessment of the nonadiabatic molecular dynamics approach with the electronic structure calculations based on the linearly scaling subsystem density functional method. The approach is implemented in an open-source embedded Quantum Espresso/Libra software specially designed for nonadiabatic dynamics simulations in extended systems. As proof of the applicability of this method to large condensed-matter systems, we examine the dynamics of nonradiative relaxation of excess excitation energy in pentacene crystals with the simulation supercells containing more than 600 atoms. We find that increased structural disorder observed in larger supercell models induces larger nonadiabatic couplings of electronic states and accelerates the relaxation dynamics of excited states. We conduct a comparative analysis of several quantum-classical trajectory surface hopping schemes, including two new methods proposed in this work (revised decoherence-induced surface hopping and instantaneous decoherence at frustrated hops). Most of the tested schemes suggest fast energy relaxation occurring with the timescales in the 0.7-2.0 ps range, but they significantly overestimate the ground state recovery rates. Only the modified simplified decay of mixing approach yields a notably slower relaxation timescales of 8-14 ps, with a significantly inhibited ground state recovery.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
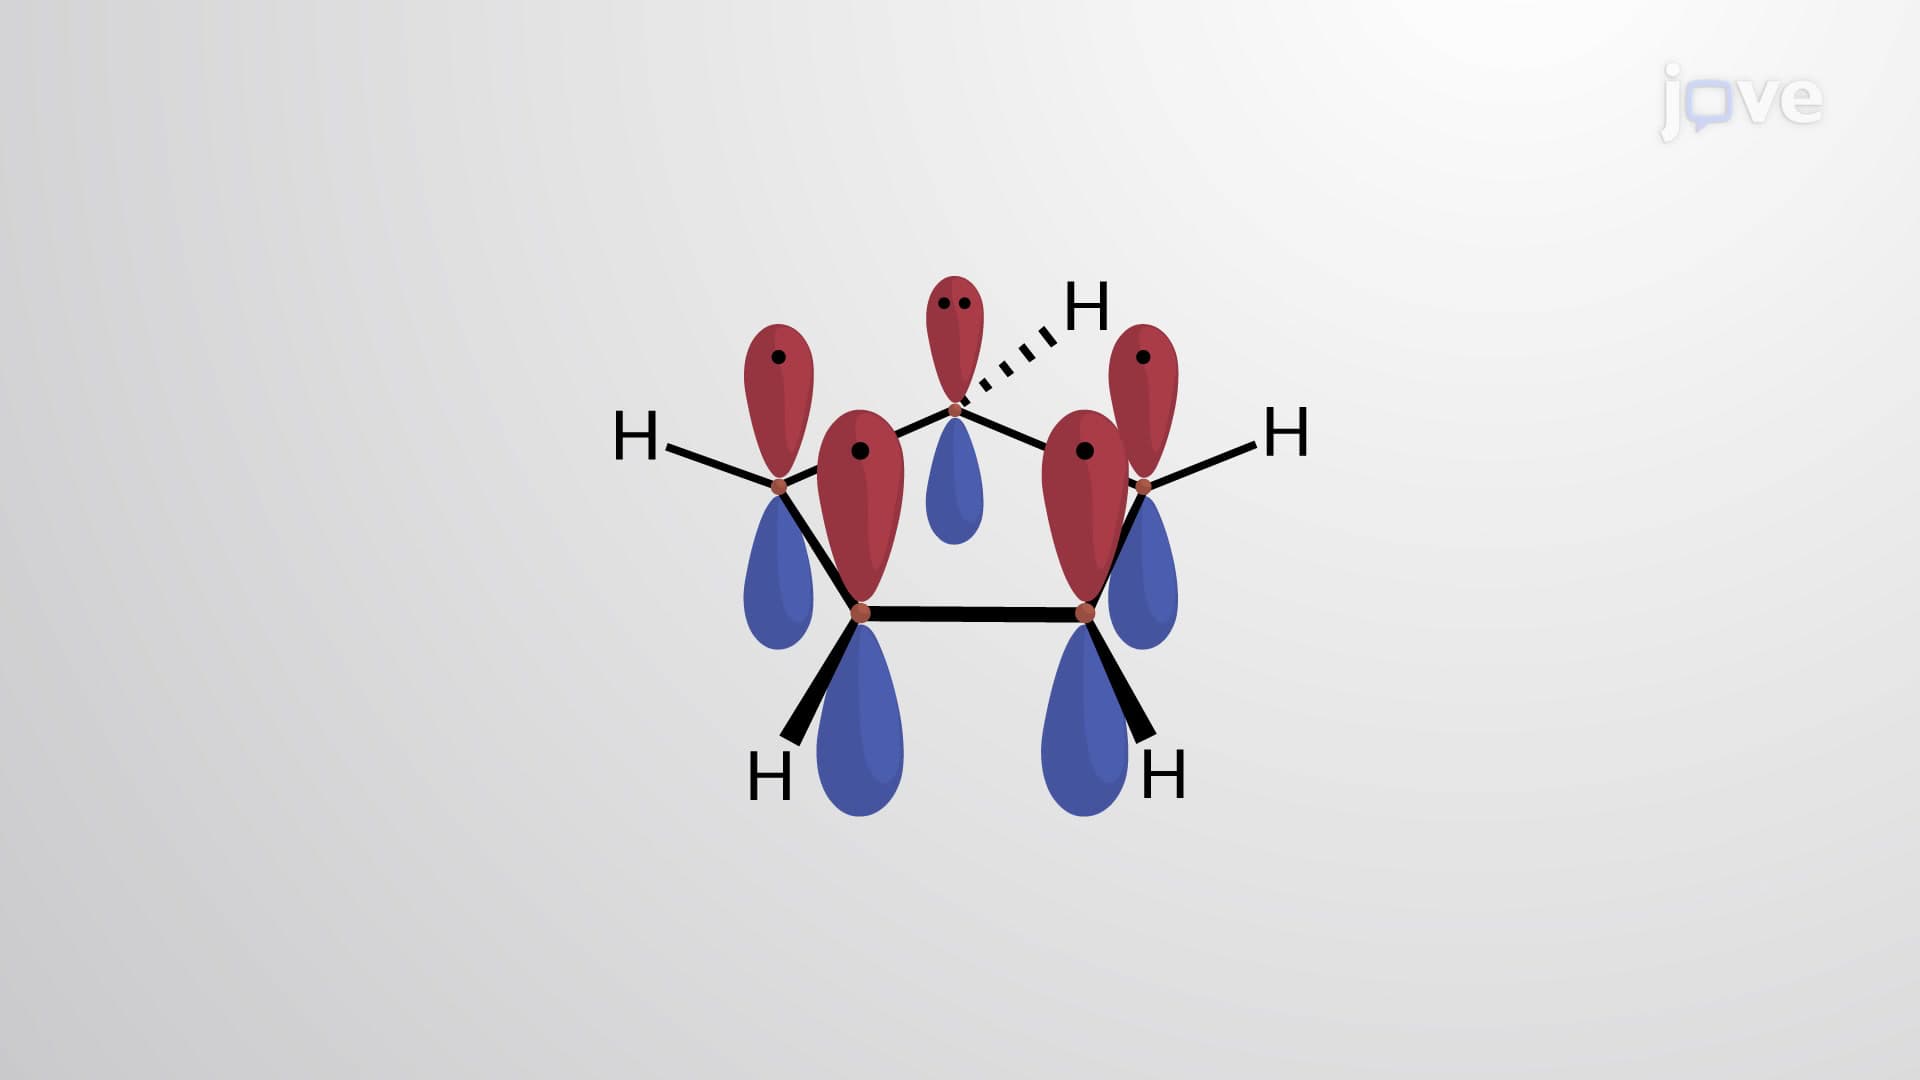
Due to the absence of continuous...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...