Toward Accurate Calculation of Excitation Energies on Quantum Computers with ΔADAPT-VQE: A Case Study of BODIPY Derivatives
Related Experiment Video
Updated: Jun 22, 2025

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Anton Nykänen1, Leander Thiessen1, Elsi-Mari Borrelli1
1Algorithmiq Ltd., Kanavakatu 3C, FI-00160 Helsinki, Finland.

Abstract:
Quantum chemistry simulations offer a cost-effective way to computationally design BODIPY photosensitizers. However, accurate predictions of excitation energies pose a challenge for time-dependent density functional theory and equation-of-motion coupled-cluster singles and doubles methods. By contrast, reliable predictions can be achieved by multireference quantum chemistry methods; unfortunately, their computational cost increases exponentially with the number of electrons. Alternatively, quantum computing holds potential for an exact simulation of the photophysical properties in a computationally more efficient way. Herein, we introduce the state-specific ΔUCCSD-VQE (unitary coupled-cluster singles and doubles-variational quantum eigensolver) and ΔADAPT-VQE methods in which the electronically excited state is calculated via a non-Aufbau configuration. We show for six BODIPY derivatives that the proposed methods predict accurate excitation energies that are in good agreement with those from experiments. Due to its performance and simplicity, we believe that ΔADAPT will become a useful approach for the simulation of BODIPY photosensitizers on near-term quantum devices.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
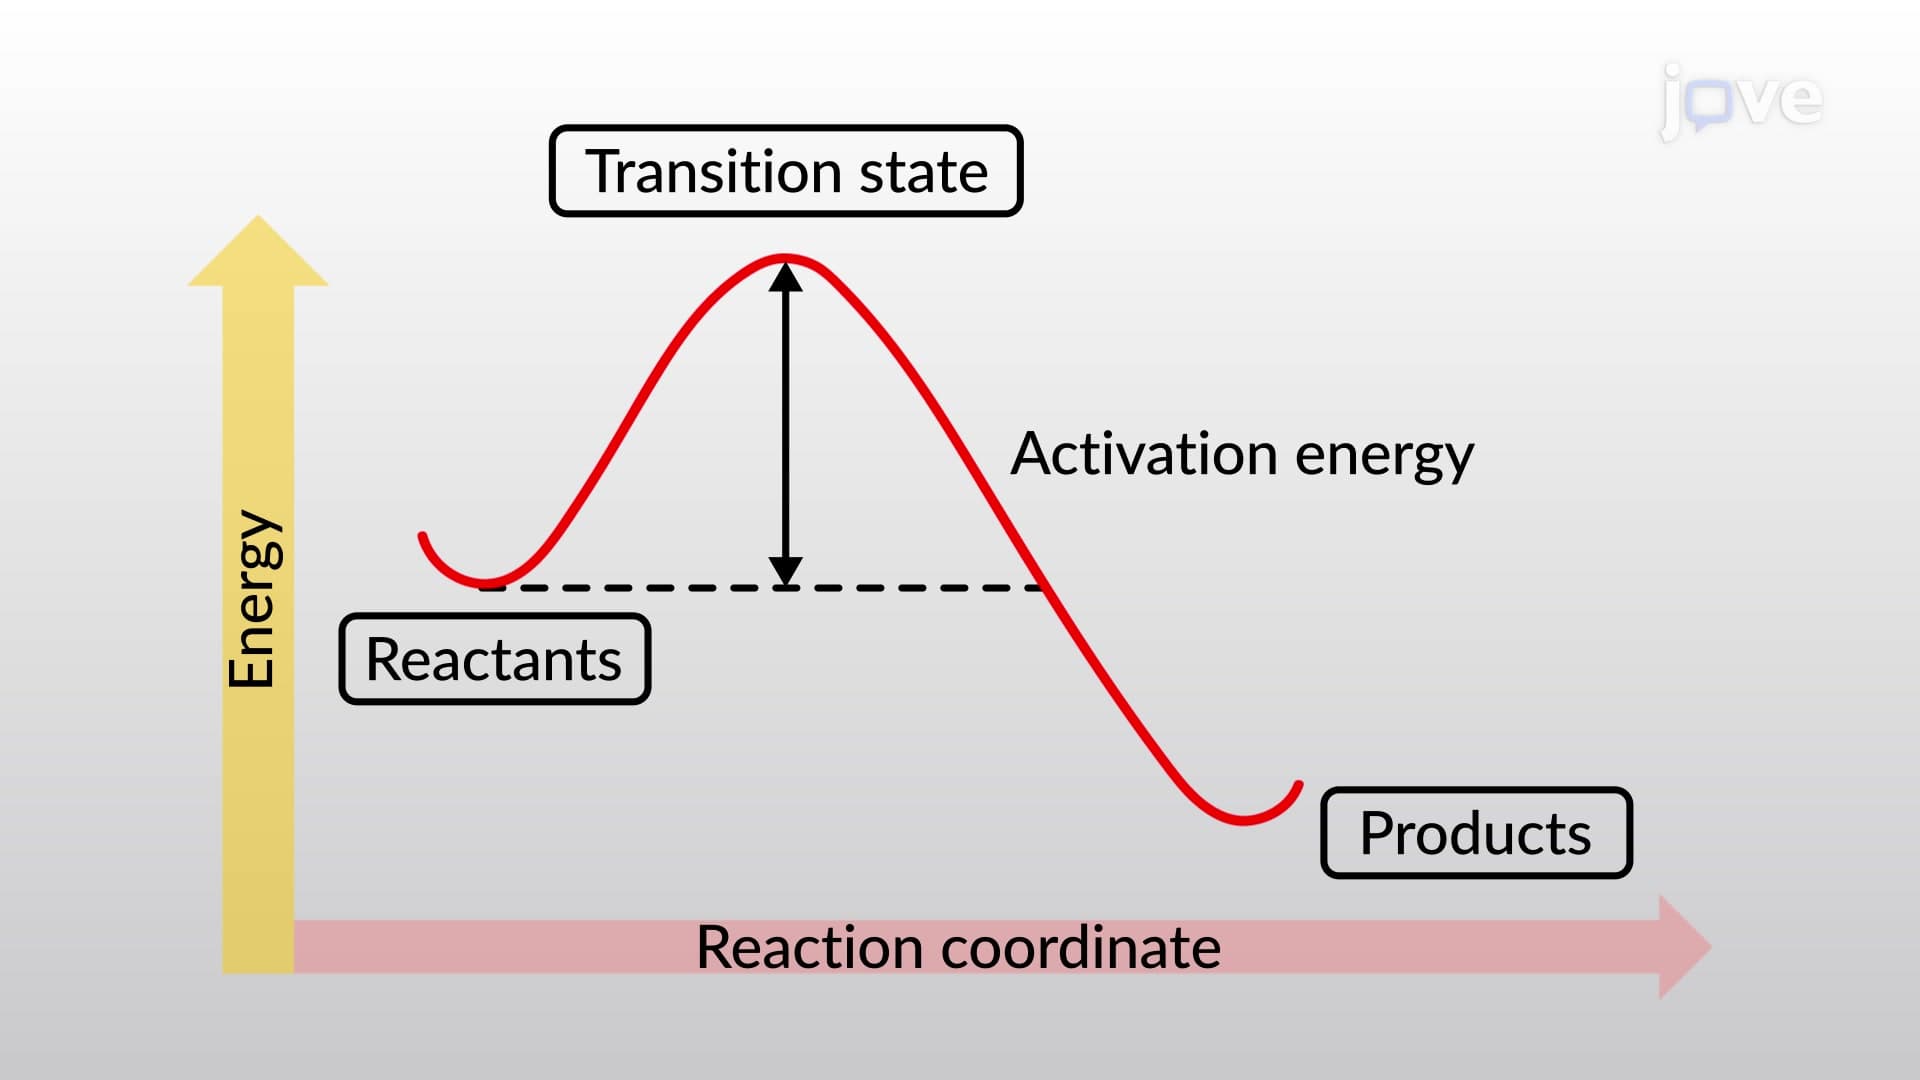 02:13
Bond Dissociation Energy and Activation Energy
 01:27
π Electron Effects on Chemical Shift: Overview
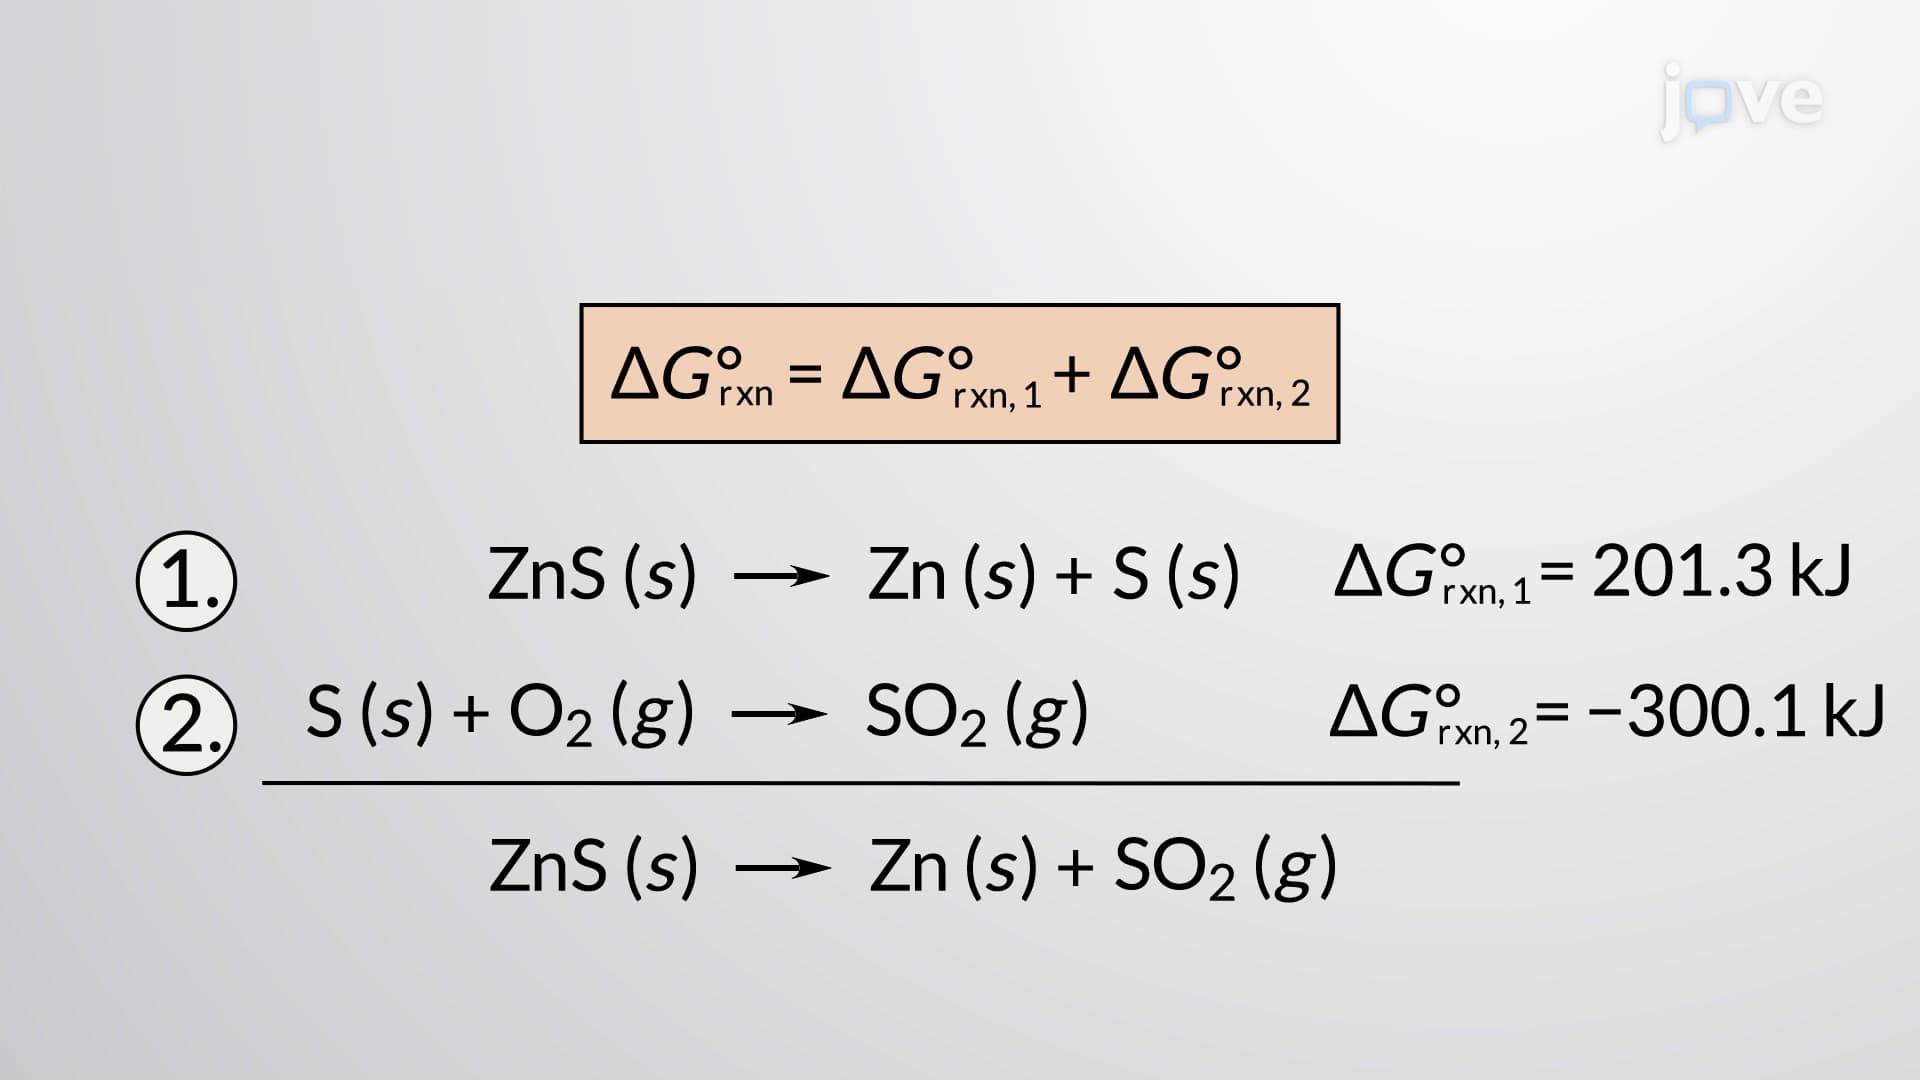 02:49
Calculating Standard Free Energy Changes
 02:45
The Quantum-Mechanical Model of an Atom
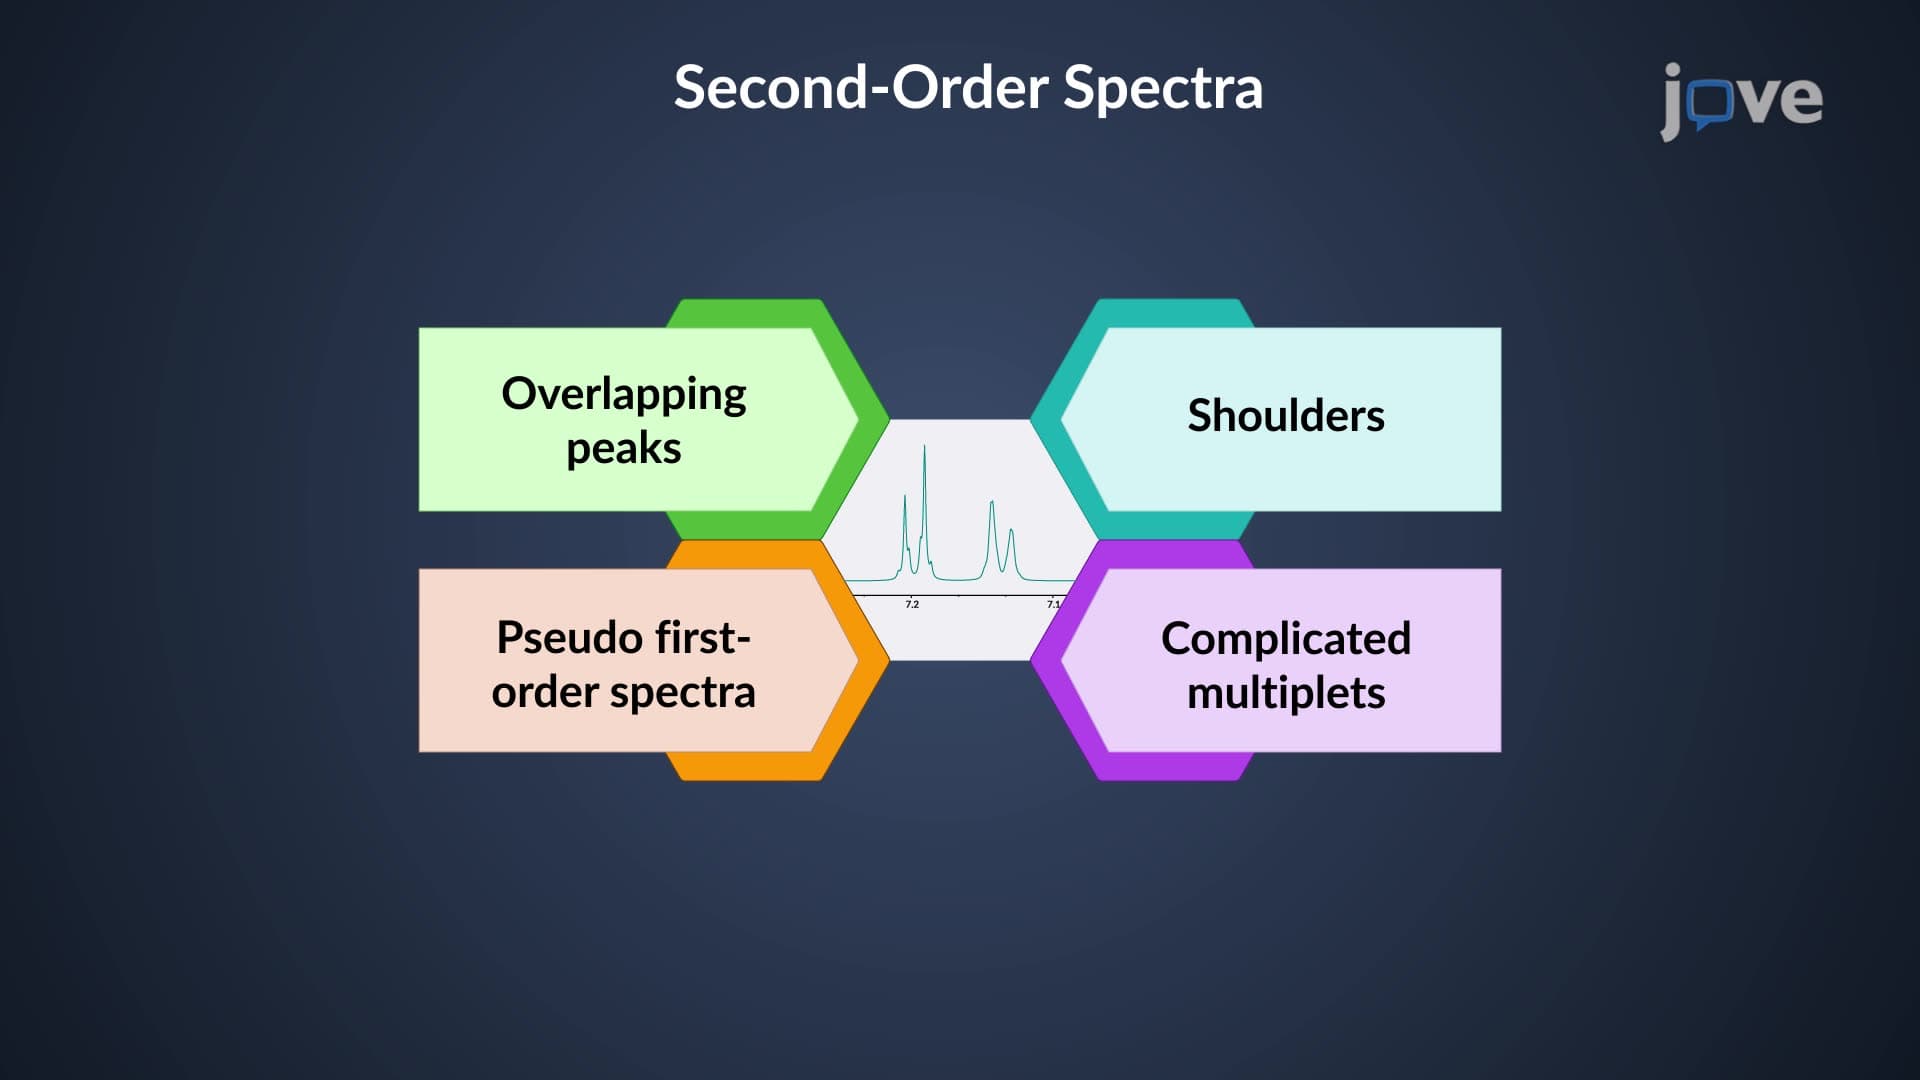 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds