Related Experiment Video
Updated: Jun 15, 2025

A Hyperandrogenic Mouse Model to Study Polycystic Ovary Syndrome
Published on: October 2, 2018
46,XX males with congenital adrenal hyperplasia: a clinical and biochemical description
Bas P H Adriaansen1,2, Agustini Utari3,4, Dineke Westra5
1Department of Pediatrics, Division of Pediatric Endocrinology, Amalia Children's Hospital, Radboud University Medical Center, Nijmegen, Netherlands.

Introduction:
Congenital adrenal hyperplasia (CAH) due to 21-hydroxylase deficiency (21OHD) or 11-hydroxylase deficiency (11OHD) is characterized by underproduction of cortisol and overproduction of adrenal androgens. These androgens lead to a variable degree of virilization of the female external genitalia in 46,XX individuals. Especially in developing countries, diagnosis is often delayed and 46,XX patients might be assigned as males. This study aims to describe the clinical and biochemical characteristics of a unique cohort of untreated male-reared 46,XX classic CAH patients from Indonesia and discusses treatment challenges.
Methods:
Nine untreated classic CAH patients with 46,XX genotype and 21OHD (n=6) or 11OHD (n=3), aged 3-46 years old, were included. Biometrical parameters, clinical characteristics, and biochemical measurements including glucocorticoids, renin, androgens, and the pituitary-gonadal axis were evaluated.
Results:
All patients had low early morning serum cortisol concentrations (median 89 nmol/L) without significant increase after ACTH stimulation. Three patients with salt wasting 21OHD reported one or more periods with seizures and/or vomiting in their past until the age of 6, but not thereafter. The remaining patients reported no severe illness or hospitalization episodes, despite their decreased capacity to produce cortisol. In the 21OHD patients, plasma renin levels were elevated compared to the reference range, and in 11OHD patients renin levels were in the low-normal range. All adult patients had serum testosterone concentrations within the normal male reference range. In 21OHD patients, serum 11-oxygenated androgens comprised 41-60% of the total serum androgen concentrations. Glucocorticoid treatment was offered to all patients, but they refused after counseling as this would reduce their endogenous androgen production and they did not report complaints of their low cortisol levels.
Discussion:
We describe a unique cohort of untreated classic 46,XX male CAH patients without overt clinical signs of cortisol deficiency despite their cortisol underproduction and incapacity to increase cortisol levels after ACTH stimulation. The described adolescent and adult patients produce androgen levels within or above the normal male reference range. Glucocorticoid treatment will lower these adrenal androgen concentrations. Therefore, in 46,XX CAH patients reared as males an individual treatment approach with careful counseling and clear instructions is needed.
Related Concept Videos
 01:27
Adrenal Gland Disorders
Adrenal insufficiency, characterized by insufficient cortisol and aldosterone production, leads to conditions like Addison's disease. This disorder, affecting the adrenal cortex, exhibits symptoms such as skin bronzing, dehydration, low blood pressure, fatigue, and weight loss. Congenital adrenal hyperplasia, a genetic ailment causing...
 01:43
Sex-linked Disorders
 01:31
Hormones of the Adrenal Glands
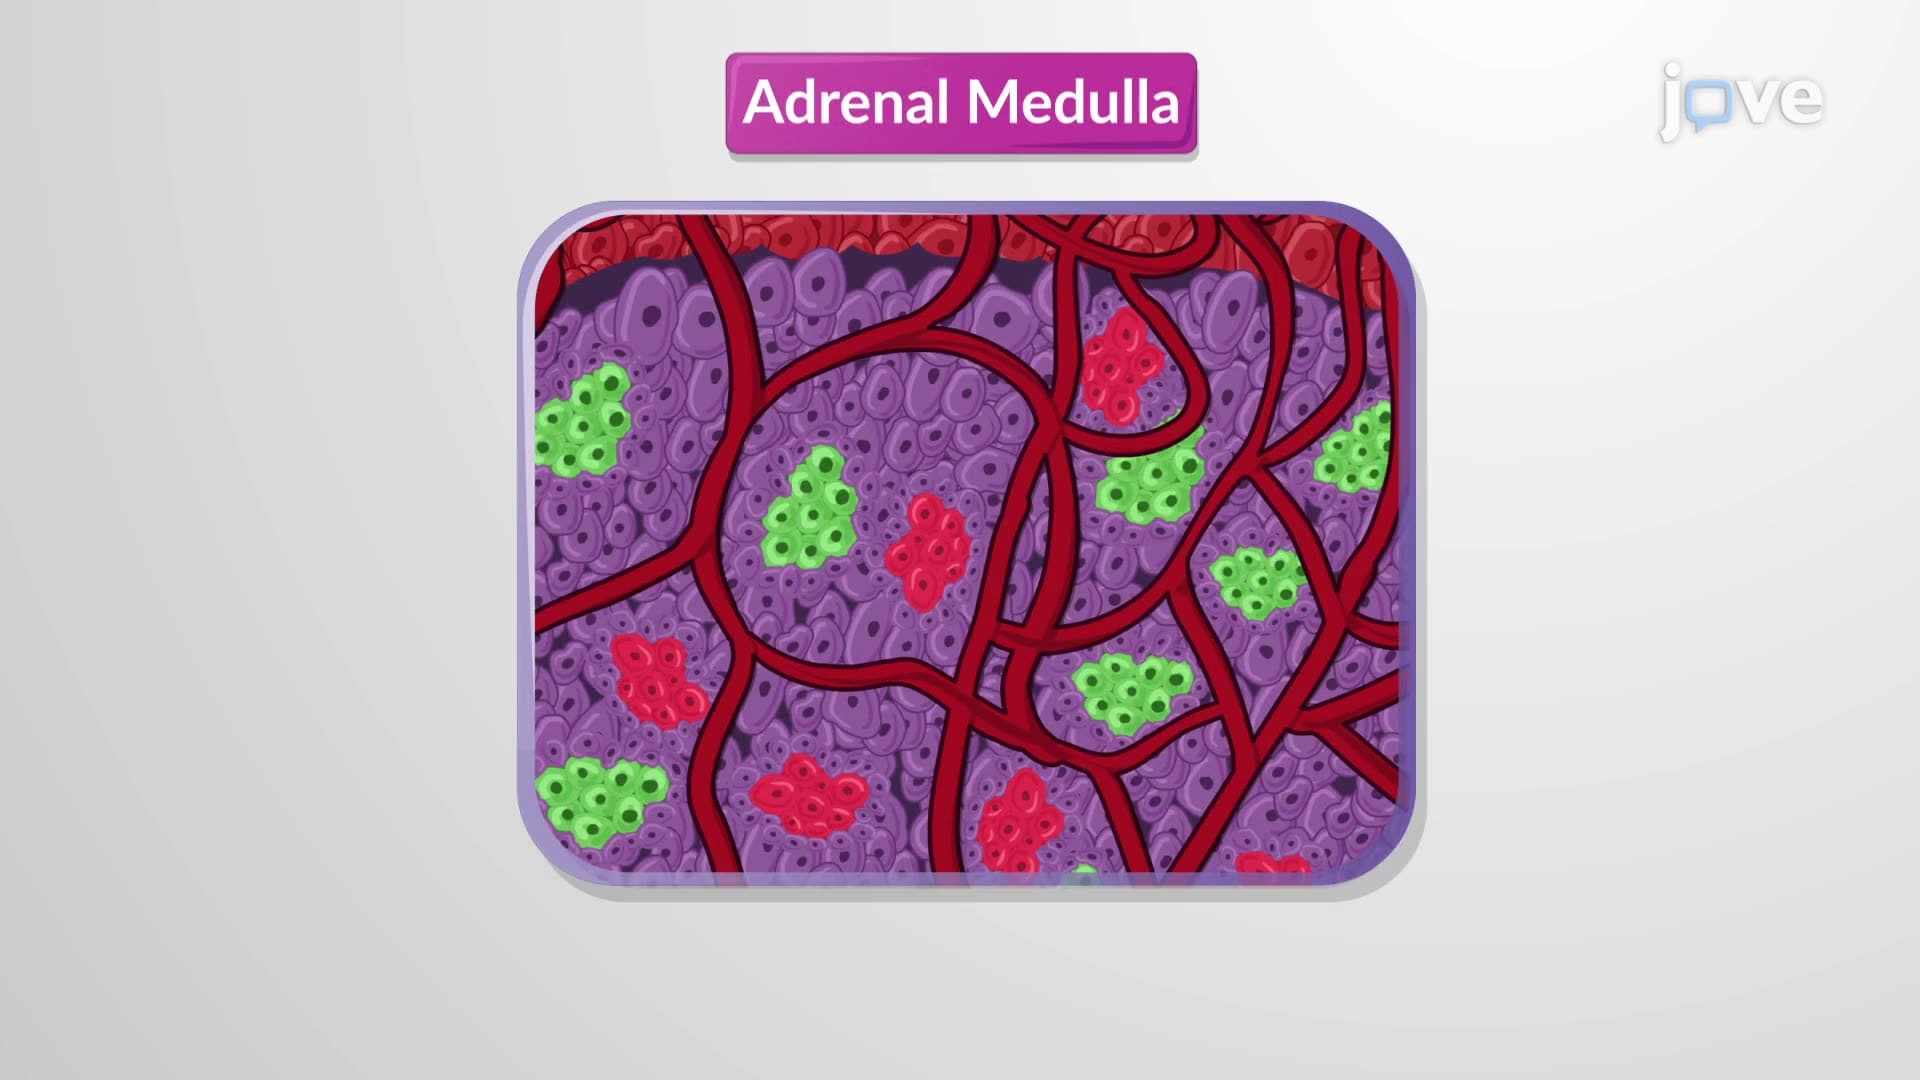
The adrenal cortex, a powerhouse of hormone synthesis, generates over two dozen corticosteroid hormones. The zona glomerulosa produces mineralocorticoids, exemplified by aldosterone, influencing the electrolyte composition of body fluids. The synthesis of glucocorticoids such as cortisol and...
 01:21
Smooth Endoplasmic Reticulum
The ER provides optimal conditions for synthesizing steroid hormones and lipids, such as phospholipids and triglycerides. Traditionally, lipid metabolism was considered to be a smooth ER function. However, there is no direct evidence to prove that rough ER is completely excluded from lipid...
 01:35
Pedigree Analysis
 02:19
The Y Chromosome Determines Maleness
Evolution
Around 300 million years ago, the two sex chromosomes diverged from two identical autosomal chromosomes. Over time, the Y chromosome has lost most of its genes, shrinking in size....