Related Experiment Video
Updated: Jun 11, 2025


Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Janus Monolayer of 1T-TaSSe: A Computational Study
1Department of Solid State Physics, Faculty of Physics and Applied Informatics, University of Lodz, Ulica Pomorska 149/153, 90-236 Lodz, Poland.

Abstract:
Materials exhibiting charge density waves are attracting increasing attention owing to their complex physics and potential for applications. In this paper, we present a computational, first principles-based study of the Janus monolayer of 1T-TaSSe transition metal dichalcogenide. We extensively compare the results with those obtained for parent compounds, TaS2 and TaSe2 monolayers, with confirmed presence of 13×13 charge density waves. The structural and electronic properties of the normal (undistorted) phase and distorted phase with 13×13 periodic lattice distortion are discussed. In particular, for a normal phase, the emergence of dipolar moment due to symmetry breaking is demonstrated, and its sensitivity to an external electric field perpendicular to the monolayer is investigated. Moreover, the appearance of imaginary energy phonon modes suggesting structural instability is shown. For the distorted phase, we predict the presence of a flat, weakly dispersive band related to the appearance of charge density waves, similar to the one observed in parent compounds. The results suggest a novel platform for studying charge density waves in two-dimensional transition metal dichalcogenides.
More Related Videos
 09:55
Preparation of Janus Particles and Alternating Current Electrokinetic Measurements with a Rapidly Fabricated Indium Tin Oxide Electrode Array
Published on: June 23, 2017
 08:18
Microscopic Visualization of Porous Nanographenes Synthesized through a Combination of Solution and On-Surface Chemistry
Published on: March 4, 2021
Related Concept Videos
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:10
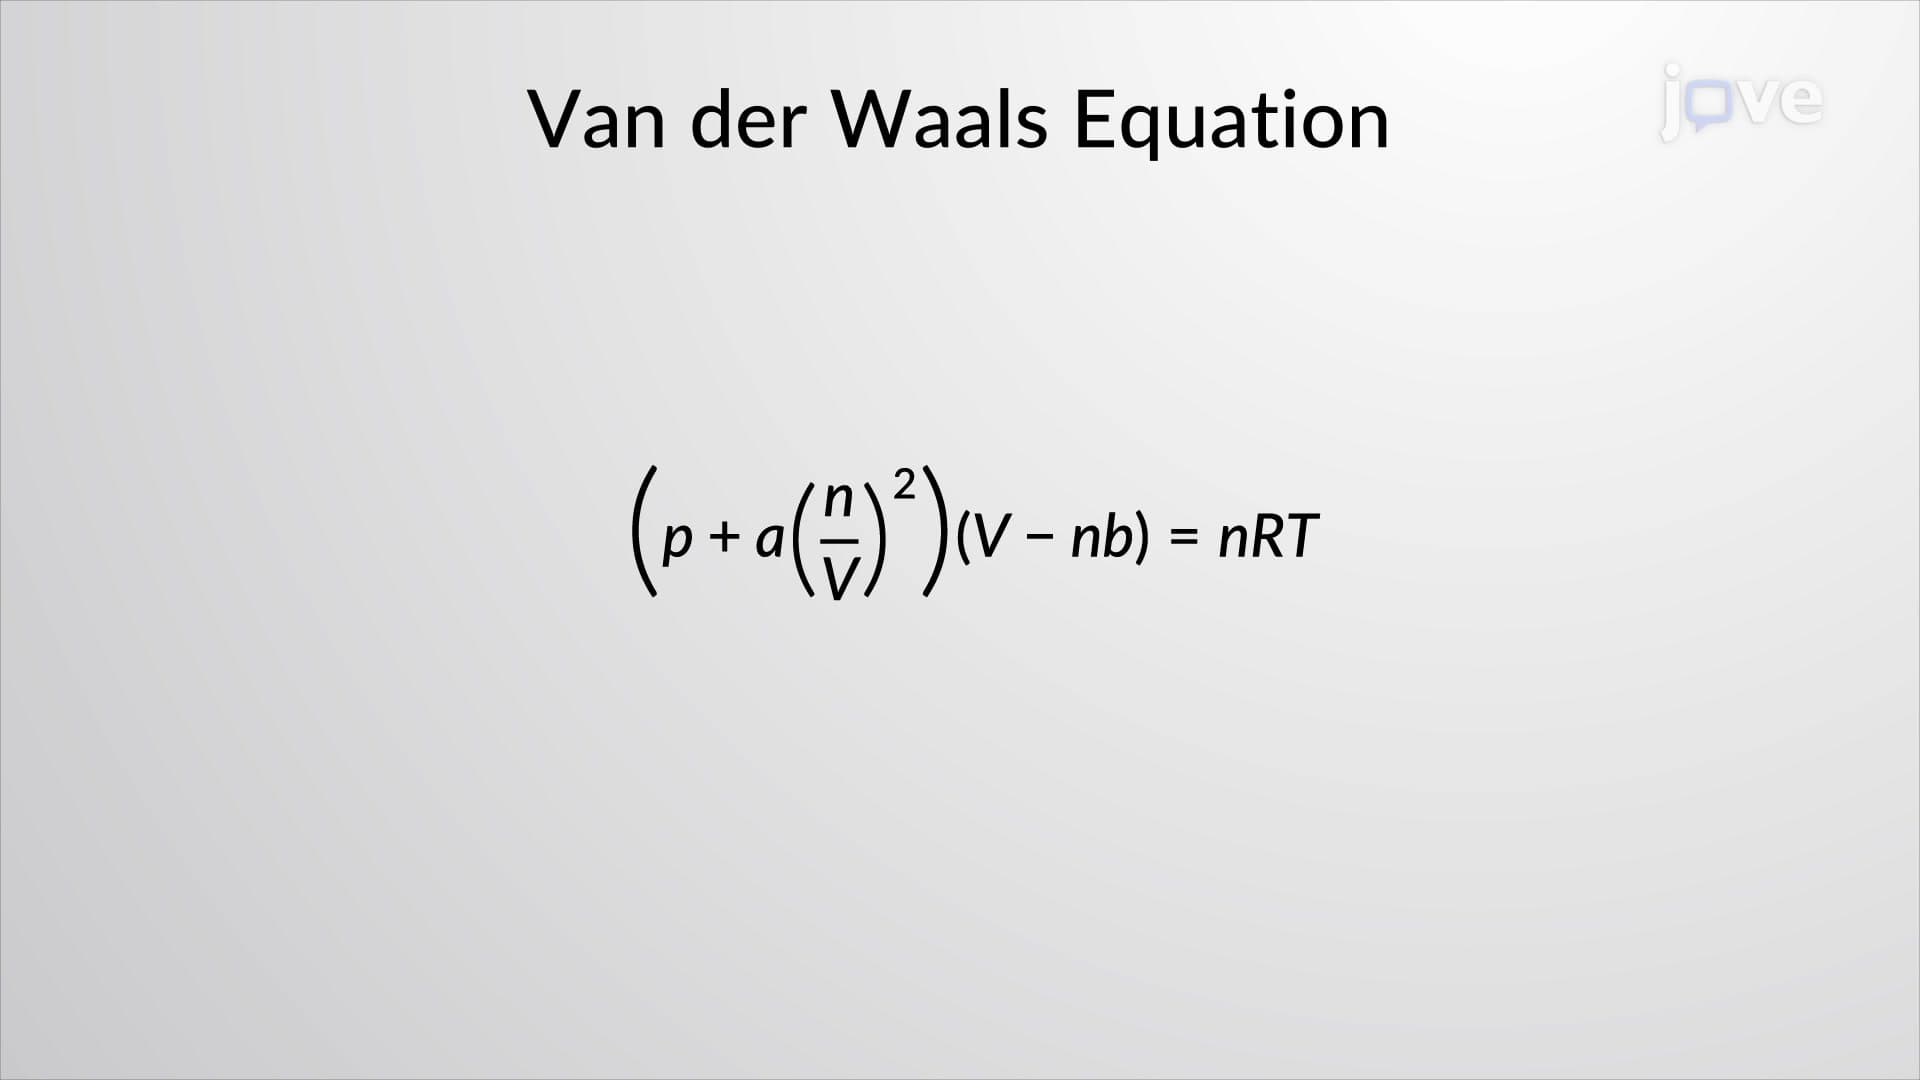
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:54
Trends in Lattice Energy: Ion Size and Charge