Related Experiment Video
Updated: Jun 9, 2025

Kinase Inhibitor Screening In Self-assembled Human Protein Microarrays
Published on: October 23, 2019
Ibrutinib Promotes Atrial Fibrillation by Disrupting A-Kinase Anchoring Protein 1-Mediated Mitochondrial Quality
Yukun Li1,2, Xinmeng Liu1,2, Rong Lin1,2
1Department of Cardiology, Beijing Anzhen Hospital, Capital Medical University, Beijing 100012, China.

Background: Ibrutinib, a potent Bruton's tyrosine kinase inhibitor with marked efficacy against hematological malignancies, is associated with the heightened risk of atrial fibrillation (AF). Although ibrutinib-induced AF is linked to enhanced oxidative stress, the underlying mechanisms remain unclear. Objective: This research aimed to explore the molecular mechanism and regulatory target in ibrutinib-induced AF. Methods: We performed in vivo electrophysiology studies using ibrutinib-treated mice, and then employed proteomic and single-cell transcriptomic analyses to identify the underlying targets and mechanisms. The effects of A-kinase anchoring protein 1 (AKAP1) depletion on mitochondrial quality surveillance (MQS) were evaluated using both in vivo and ex vivo AKAP1 overexpression models. Results: Atrial AKAP1 expression was significantly reduced in ibrutinib-treated mice, leading to inducible AF, atrial fibrosis, and mitochondrial fragmentation. These pathological changes were effectively mitigated in an overexpression model of ibrutinib-treated mice injected with an adeno-associated virus carrying Akap1. In ibrutinib-treated atrial myocytes, AKAP1 down-regulation promoted dynamin-related protein 1 (DRP1) translocation into mitochondria by facilitating DRP1 dephosphorylation at Ser637, thereby mediating excessive mitochondrial fission. Impaired MQS was also suggested by defective mitochondrial respiration, mitochondrial metabolic reprogramming, and suppressed mitochondrial biogenesis, accompanied by excessive oxidative stress and inflammatory activation. The ibrutinib-mediated MQS disturbance can be markedly improved with the inducible expression of the AKAP1 lentiviral system. Conclusions: Our findings emphasize the key role of AKAP1-mediated MQS disruption in ibrutinib-induced AF, which explains the previously observed reactive oxygen species overproduction. Hence, AKAP1 activation can be employed to prevent and treat ibrutinib-induced AF.
Background: Ibrutinib, a potent Bruton's tyrosine kinase inhibitor with marked efficacy against hematological malignancies, is associated with the heightened risk of atrial fibrillation (AF). Although ibrutinib-induced AF is linked to enhanced oxidative stress, the underlying mechanisms remain unclear. Objective: This research aimed to explore the molecular mechanism and regulatory target in ibrutinib-induced AF. Methods: We performed in vivo electrophysiology studies using ibrutinib-treated mice, and then employed proteomic and single-cell transcriptomic analyses to identify the underlying targets and mechanisms. The effects of A-kinase anchoring protein 1 (AKAP1) depletion on mitochondrial quality surveillance (MQS) were evaluated using both in vivo and ex vivo AKAP1 overexpression models. Results: Atrial AKAP1 expression was significantly reduced in ibrutinib-treated mice, leading to inducible AF, atrial fibrosis, and mitochondrial fragmentation. These pathological changes were effectively mitigated in an overexpression model of ibrutinib-treated mice injected with an adeno-associated virus carrying Akap1. In ibrutinib-treated atrial myocytes, AKAP1 down-regulation promoted dynamin-related protein 1 (DRP1) translocation into mitochondria by facilitating DRP1 dephosphorylation at Ser637, thereby mediating excessive mitochondrial fission. Impaired MQS was also suggested by defective mitochondrial respiration, mitochondrial metabolic reprogramming, and suppressed mitochondrial biogenesis, accompanied by excessive oxidative stress and inflammatory activation. The ibrutinib-mediated MQS disturbance can be markedly improved with the inducible expression of the AKAP1 lentiviral system. Conclusions: Our findings emphasize the key role of AKAP1-mediated MQS disruption in ibrutinib-induced AF, which explains the previously observed reactive oxygen species overproduction. Hence, AKAP1 activation can be employed to prevent and treat ibrutinib-induced AF.
More Related Videos
 08:03
Hybrid Cell Analysis System to Assess Structural and Contractile Changes of Human iPSC-Derived Cardiomyocytes for Preclinical Cardiac Risk Evaluation
Published on: October 20, 2022
 08:05
Optimization of Transesophageal Atrial Pacing to Assess Atrial Fibrillation Susceptibility in Mice
Published on: June 29, 2022
Related Concept Videos
 01:12
Antiarrhythmic Drugs: Class III Agents as Potassium Channel Blockers
 01:17
Intracellular Signaling Affects Focal Adhesions
Some...
 01:24
Antiarrhythmic Drugs: Class II Agents as β-Adrenergic Blockers
 01:20
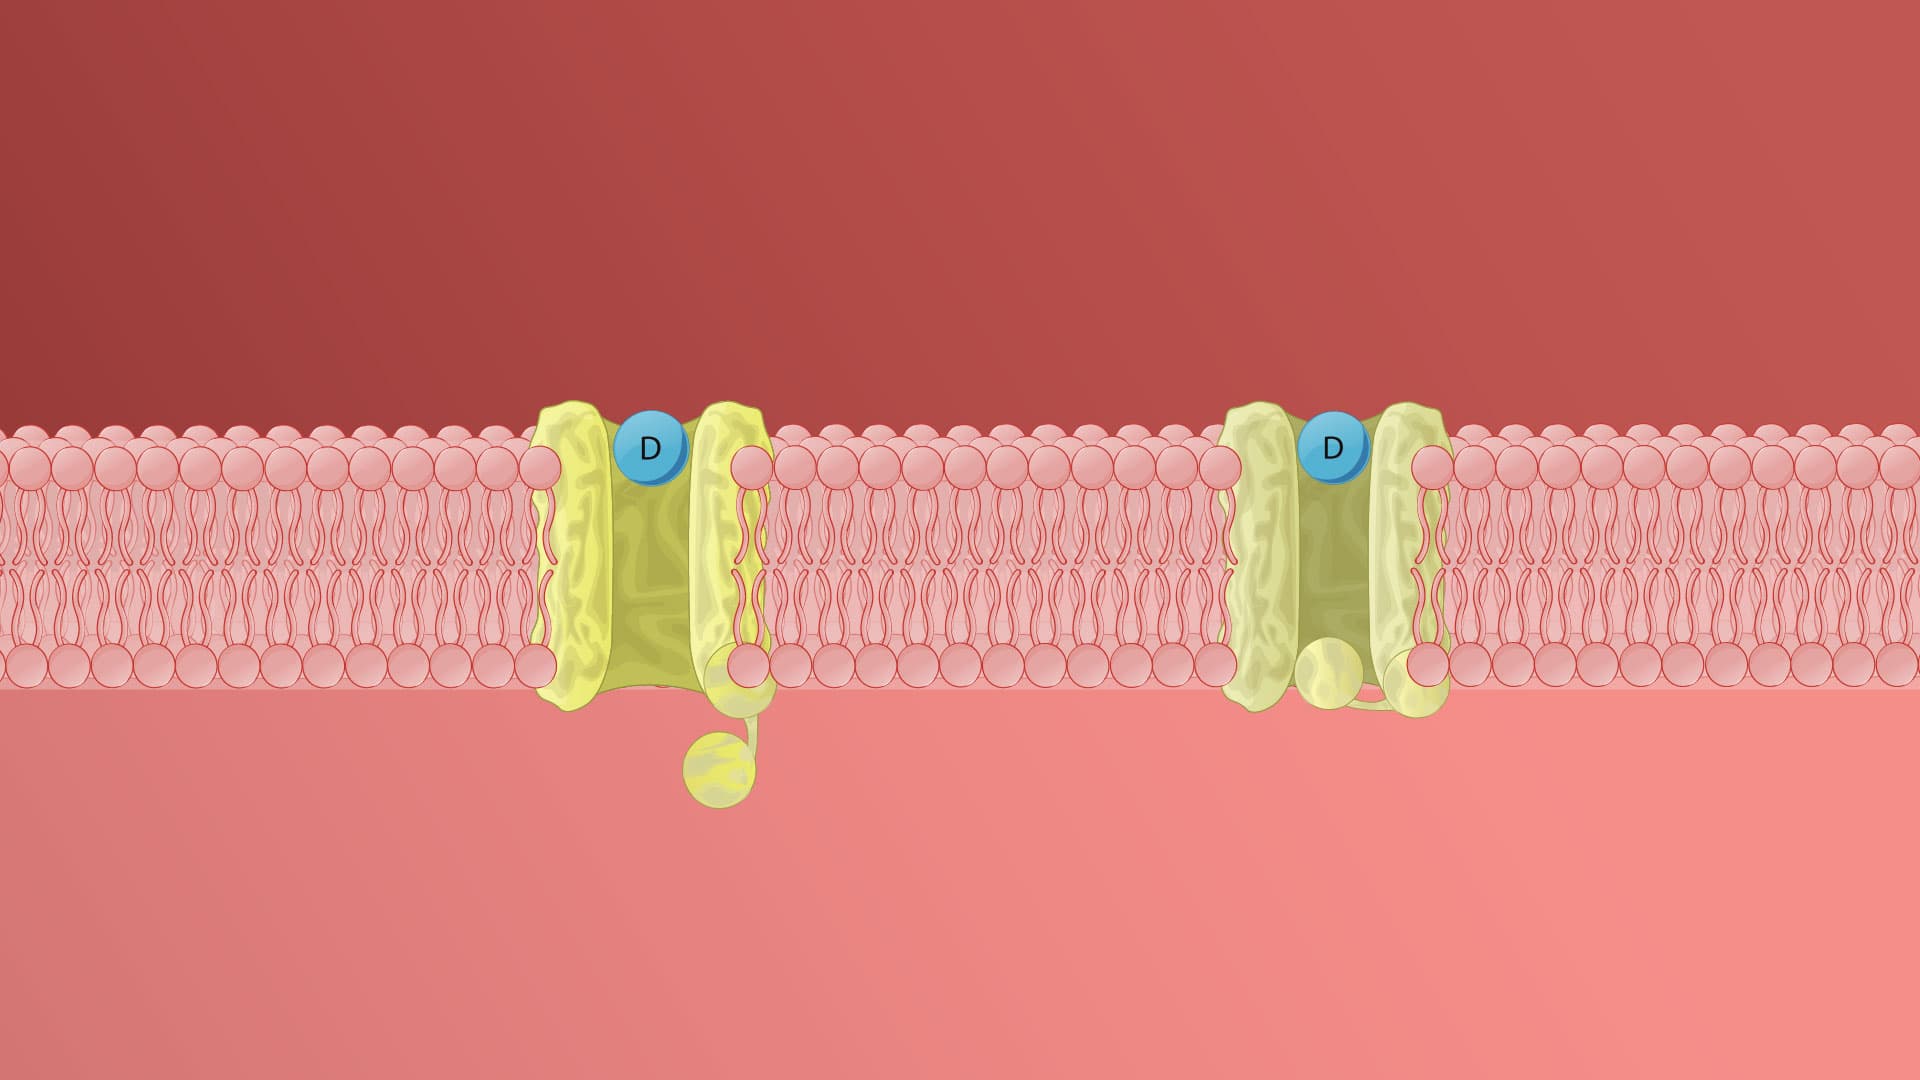
Antiarrhythmic Drugs: Class IV Agents as Calcium Channel Blockers
Verapamil, a calcium channel blocker, inhibits calcium movement across myocardial cell membranes and vascular smooth muscle. This results in the dilation of coronary and...
 01:26
Heart Failure Drugs: Inhibitors of Renin-Angiotensin System
 02:57
Targeted Cancer Therapies
There are several types of targeted therapies against...