Related Experiment Video
Updated: Jun 9, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
"Best" Iterative Coupled-Cluster Triples Model? More Evidence for 3CC
Nakul K Teke1, Ajay Melekamburath1, Bimal Gaudel1
1Department of Chemistry, Virginia Tech, Blacksburg, Virginia 24061, United States.

Abstract:
To follow up on the unexpectedly good performance of several coupled-cluster models with approximate inclusion of 3-body clusters [Rishi, V.; Valeev, E. F. J. Chem. Phys. 2019, 151, 064102.] we performed a more complete assessment of the 3CC method [Feller, D. . J. Chem. Phys. 2008, 129, 204105.] for accurate computational thermochemistry in the standard HEAT framework. New spin-integrated implementation of the 3CC method applicable to closed- and open-shell systems utilizes a new automated toolchain for derivation, optimization, and evaluation of operator algebra in many-body electronic structure. We found that with a double-ζ basis set the 3CC correlation energies and their atomization energy contributions are almost always more accurate (with respect to the CCSDTQ reference) than the CCSDT model as well as the standard CCSD(T) model. The mean absolute errors in cc-pVDZ {3CC, CCSDT, and CCSD(T)} electronic (per valence electron) and atomization energies relative to the CCSDTQ reference for the HEAT data set [Tajti, A. . J. Chem. Phys. 2004, 121, 11599-11613.], were {24, 70, 122} μEh/e and {0.46, 2.00, 2.58} kJ/mol, respectively. The mean absolute errors in the complete-basis-set limit {3CC, CCSDT, and CCSD(T)} atomization energies relative to the HEAT model reference, were {0.52, 2.00, and 1.07} kJ/mol, The significant and systematic reduction of the error by the 3CC method and its lower cost than CCSDT suggests it as a viable candidate for post-CCSD(T) thermochemistry applications, as well as the preferred alternative to CCSDT in general.

Related Concept Videos
 01:22
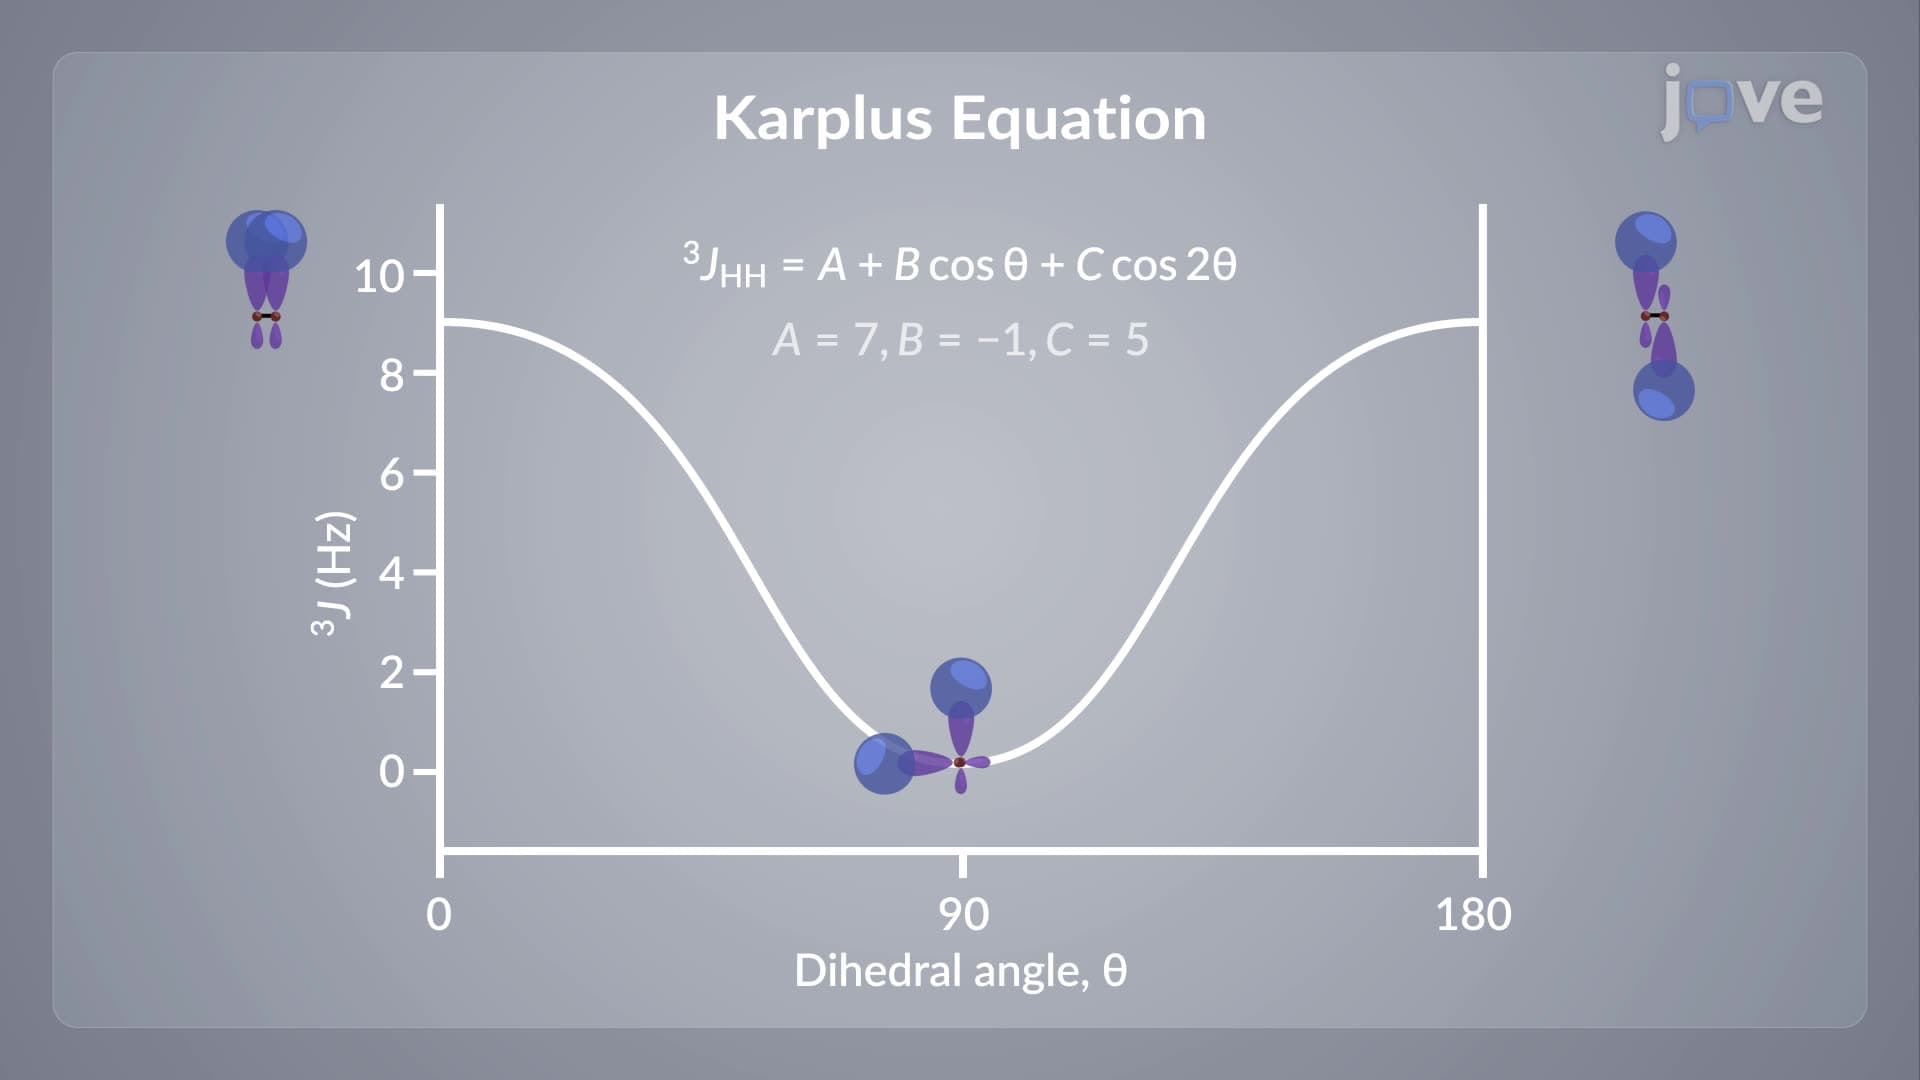
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...