Related Experiment Video
Updated: Jun 6, 2025

Simultaneous Affinity Enrichment of Two Post-Translational Modifications for Quantification and Site Localization
Published on: February 27, 2020
Assessment of Data-Independent Acquisition Mass Spectrometry (DIA-MS) for the Identification of Single Amino Acid
Ivo Fierro-Monti1,2, Klemens Fröhlich2, Christian Schori2
1European Molecular Biology Laboratory's European Bioinformatics Institute (EMBL-EBI), Hinxton CB10 1SD, Cambridgeshire, UK.

Abstract:
Proteogenomics integrates genomic and proteomic data to elucidate cellular processes by identifying variant peptides, including single amino acid variants (SAAVs). In this study, we assessed the capability of data-independent acquisition mass spectrometry (DIA-MS) to identify SAAV peptides in HeLa cells using various search engine pipelines. We developed a customised sequence database (DB) incorporating SAAV sequences from the HeLa genome and conducted searches using DIA-NN, Spectronaut, and Fragpipe-MSFragger. Our evaluation focused on identifying true positive SAAV peptides and false positives through entrapment DBs. This study revealed that DIA-MS provides reproducible and comprehensive coverage of the proteome, identifying a substantial proportion of SAAV peptides. Notably, the DIA-MS searches maintained consistent identification of SAAV peptides despite varying sizes of the entrapment DB. A comparative analysis showed that Fragpipe-MSFragger (FP-DIA) demonstrated the most conservative and effective performance, exhibiting the lowest false discovery match ratio (FDMR). Additionally, integrating DIA and data-dependent acquisition (DDA) MS data search outputs enhanced SAAV peptide identification, with a lower false discovery rate (FDR) observed in DDA searches. The validation using stable isotope dilution and parallel reaction monitoring (SID-PRM) confirmed the SAAV peptides identified by DIA-MS and DDA-MS searches, highlighting the reliability of our approach. Our findings underscore the effectiveness of DIA-MS in proteogenomic workflows for identifying SAAV peptides, offering insights into optimising search engine pipelines and DB construction for accurate proteomics analysis. These methodologies advance the understanding of proteome variability, contributing to cancer research and the identification of novel proteoform therapeutic targets.
More Related Videos
 12:49
Quantification of Site-specific Protein Lysine Acetylation and Succinylation Stoichiometry Using Data-independent Acquisition Mass Spectrometry
Published on: April 4, 2018
 17:12
Profiling of Methyltransferases and Other S-adenosyl-L-homocysteine-binding Proteins by Capture Compound Mass Spectrometry CCMS
Published on: December 20, 2010
Related Concept Videos
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:19
MALDI-TOF Mass Spectrometry
Matrix-assisted laser desorption ionization (MALDI) is a commonly...
 01:19
Mass Spectrometry of Amines
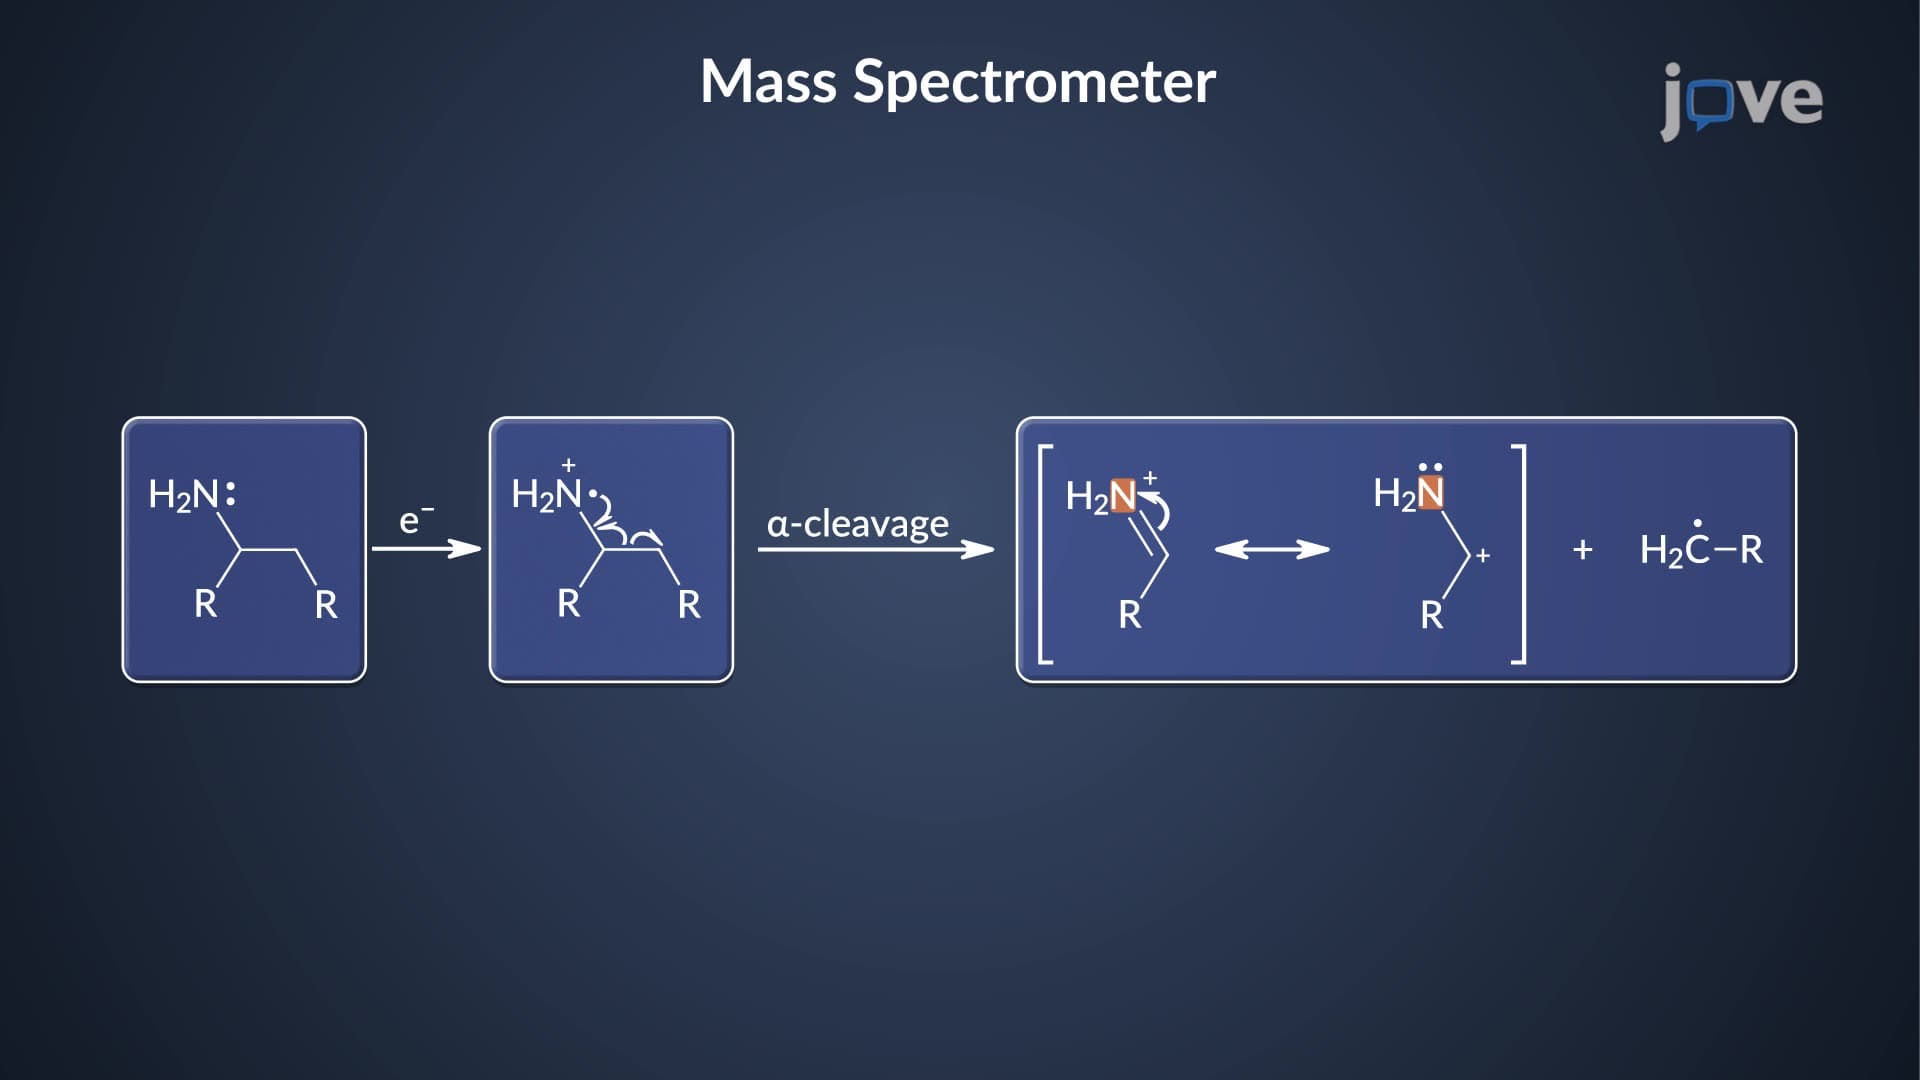
Amines undergo fragmentation through α cleavage, producing nitrogen-containing cations—iminium ions—and alkyl radicals. Mass spectra of aromatic and cyclic aliphatic amines exhibit...
 00:55
Mass Spectrometry: Amine Fragmentation
In amines, the number of nitrogen atoms affects the mass of the molecular ion, which is described by the nitrogen rule of mass spectrometry. This rule states that a compound containing...
 01:21
Mass Spectrometry: Complex Analysis
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
 01:19
Mass Spectrometry: Overview