DrugAssist: a large language model for molecule optimization
Related Experiment Video
Updated: May 7, 2025
Curation of Computational Chemical Libraries Demonstrated with Alpha-Amino Acids
Published on: April 13, 2022
Geyan Ye1, Xibao Cai2, Houtim Lai1
1Tencent AI Lab, Tencent, Shenzhen 518057, China.

Abstract:
Recently, the impressive performance of large language models (LLMs) on a wide range of tasks has attracted an increasing number of attempts to apply LLMs in drug discovery. However, molecule optimization, a critical task in the drug discovery pipeline, is currently an area that has seen little involvement from LLMs. Most of existing approaches focus solely on capturing the underlying patterns in chemical structures provided by the data, without taking advantage of expert feedback. These non-interactive approaches overlook the fact that the drug discovery process is actually one that requires the integration of expert experience and iterative refinement. To address this gap, we propose DrugAssist, an interactive molecule optimization model which performs optimization through human-machine dialogue by leveraging LLM's strong interactivity and generalizability. DrugAssist has achieved leading results in both single and multiple property optimization, simultaneously showcasing immense potential in transferability and iterative optimization. In addition, we publicly release a large instruction-based dataset called 'MolOpt-Instructions' for fine-tuning language models on molecule optimization tasks. We have made our code and data publicly available at https://github.com/blazerye/DrugAssist, which we hope to pave the way for future research in LLMs' application for drug discovery.
Related Concept Videos
 02:00
Molecular Models
 02:40
Ligand Binding Sites
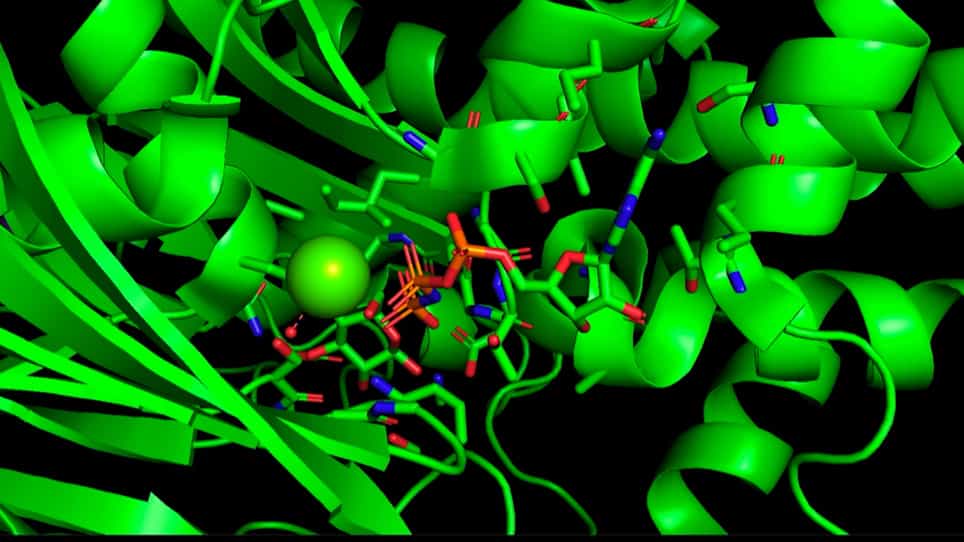
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:27
Predicting Molecular Geometry
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:26
Drug Discovery: Overview