Related Experiment Video
Updated: Jun 1, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accelerating Quantum Anharmonic Vibrational Calculations by Atom-Specific Hybrid Basis Set-Based Potential Energy
Mokshi Sharma1, Dhiksha Sharma1, Tapta Kanchan Roy1
1Department of Chemistry and Chemical Sciences, Central University Jammu, Samba 181143, Jammu and Kashmir, India.

Abstract:
The development of accurate yet fast quantum mechanical methods to calculate the anharmonic vibrational spectra of large molecules is one of the major goals of ongoing developments in this field. This study extensively explores and validates a hybrid electronic basis set approach for anharmonic vibrational calculations, where the molecule is segregated into different computational layers, and such layers are then treated with different levels of electronic basis sets. Following the system-bath model, the atoms corresponding to the active sites are treated in more accurate but computationally slower, large basis set and the rest of the atoms in less accurate but computationally faster, small basis set to construct the anharmonic hybrid potential energy surface (PES). Such a hybrid protocol for constructing an anharmonic PES is named as the atom-specific hybrid basis set (ASHBS) approach. The accuracy of the ASHBS approach is tested and established by evaluating the harmonic and anharmonic frequencies of a set of four prototype molecules. Following the ASHBS approach, for a chosen active site, the transitions of the corresponding modes are found to be closer to that of a high basis set, with a majority of target modes displaying a mean absolute error of around 3.3 cm-1, while achieving the computational acceleration of 2-3 times. This study also provides insights into determining the optimal layer size to balance computational efficiency and accuracy, offering a suitable alternative, particularly for large molecules.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 03:35
Hybridization of Atomic Orbitals II
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:10
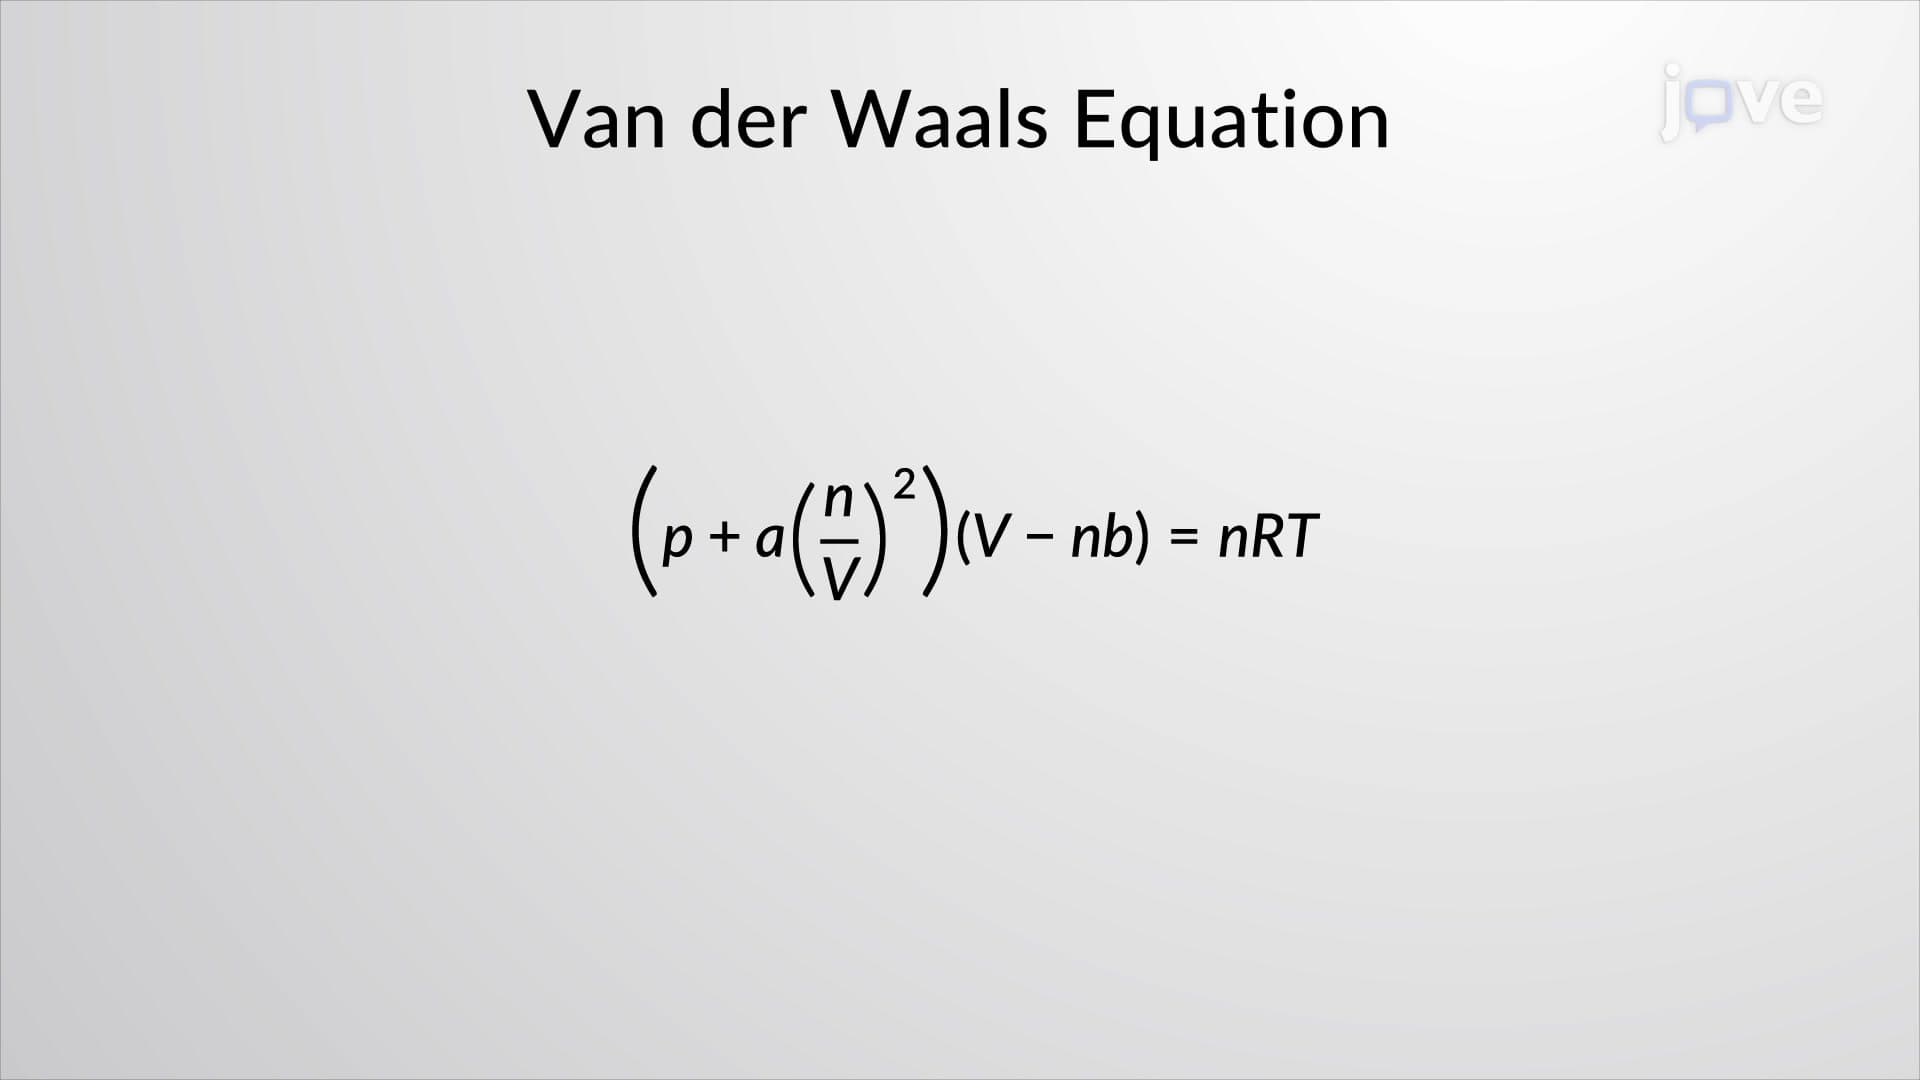
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:45
The Quantum-Mechanical Model of an Atom
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...