Related Experiment Video
Updated: May 20, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Constrained dipole moment density functional theory for the calculation of the charge-transfer energy in non-covalent
Eduardo Zúñiga-Rivera1, Javier Carmona-Espíndola2, José L Gázquez1
1Departamento de Química, Universidad Autónoma Metropolitana-Iztapalapa, Av. San Rafael Atlixco 186, Ciudad de México 09340, México.

Abstract:
The original constrained dipole moment density functional theory allows one to control the magnitude of the molecular dipole moment in a variational and non-empirical way. In this work, we extend this methodology to control the three Cartesian components of the molecular dipole moment. The new theoretical development is suitable for the calculation of the charge-transfer energy contributions to the total interaction energies in non-covalent complexes. To test the reliability of the theoretical development, we form three sets of non-covalent complexes from the literature with a total of fifty-one systems. The former set of complexes includes many different types of non-covalent interactions, the second set consists of prototypical non-covalent complexes and three biologically relevant interactions between DNA base pairs, and the third set comprises halogen bonding complexes. We determined the charge-transfer energy contributions and the total interaction energies of all these complexes. The calculated charge-transfer energies are in very good agreement with the ones calculated using the fragment-based Hirshfeld methodology, which has been proven to be reliable. Nevertheless, the new procedure relies on the molecular dipole moment, which is observable, while the fragment-based Hirshfeld methodology relies on a definition of a population analysis.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2ây2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2ây2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:36
Molecular Geometry and Dipole Moments
 02:48
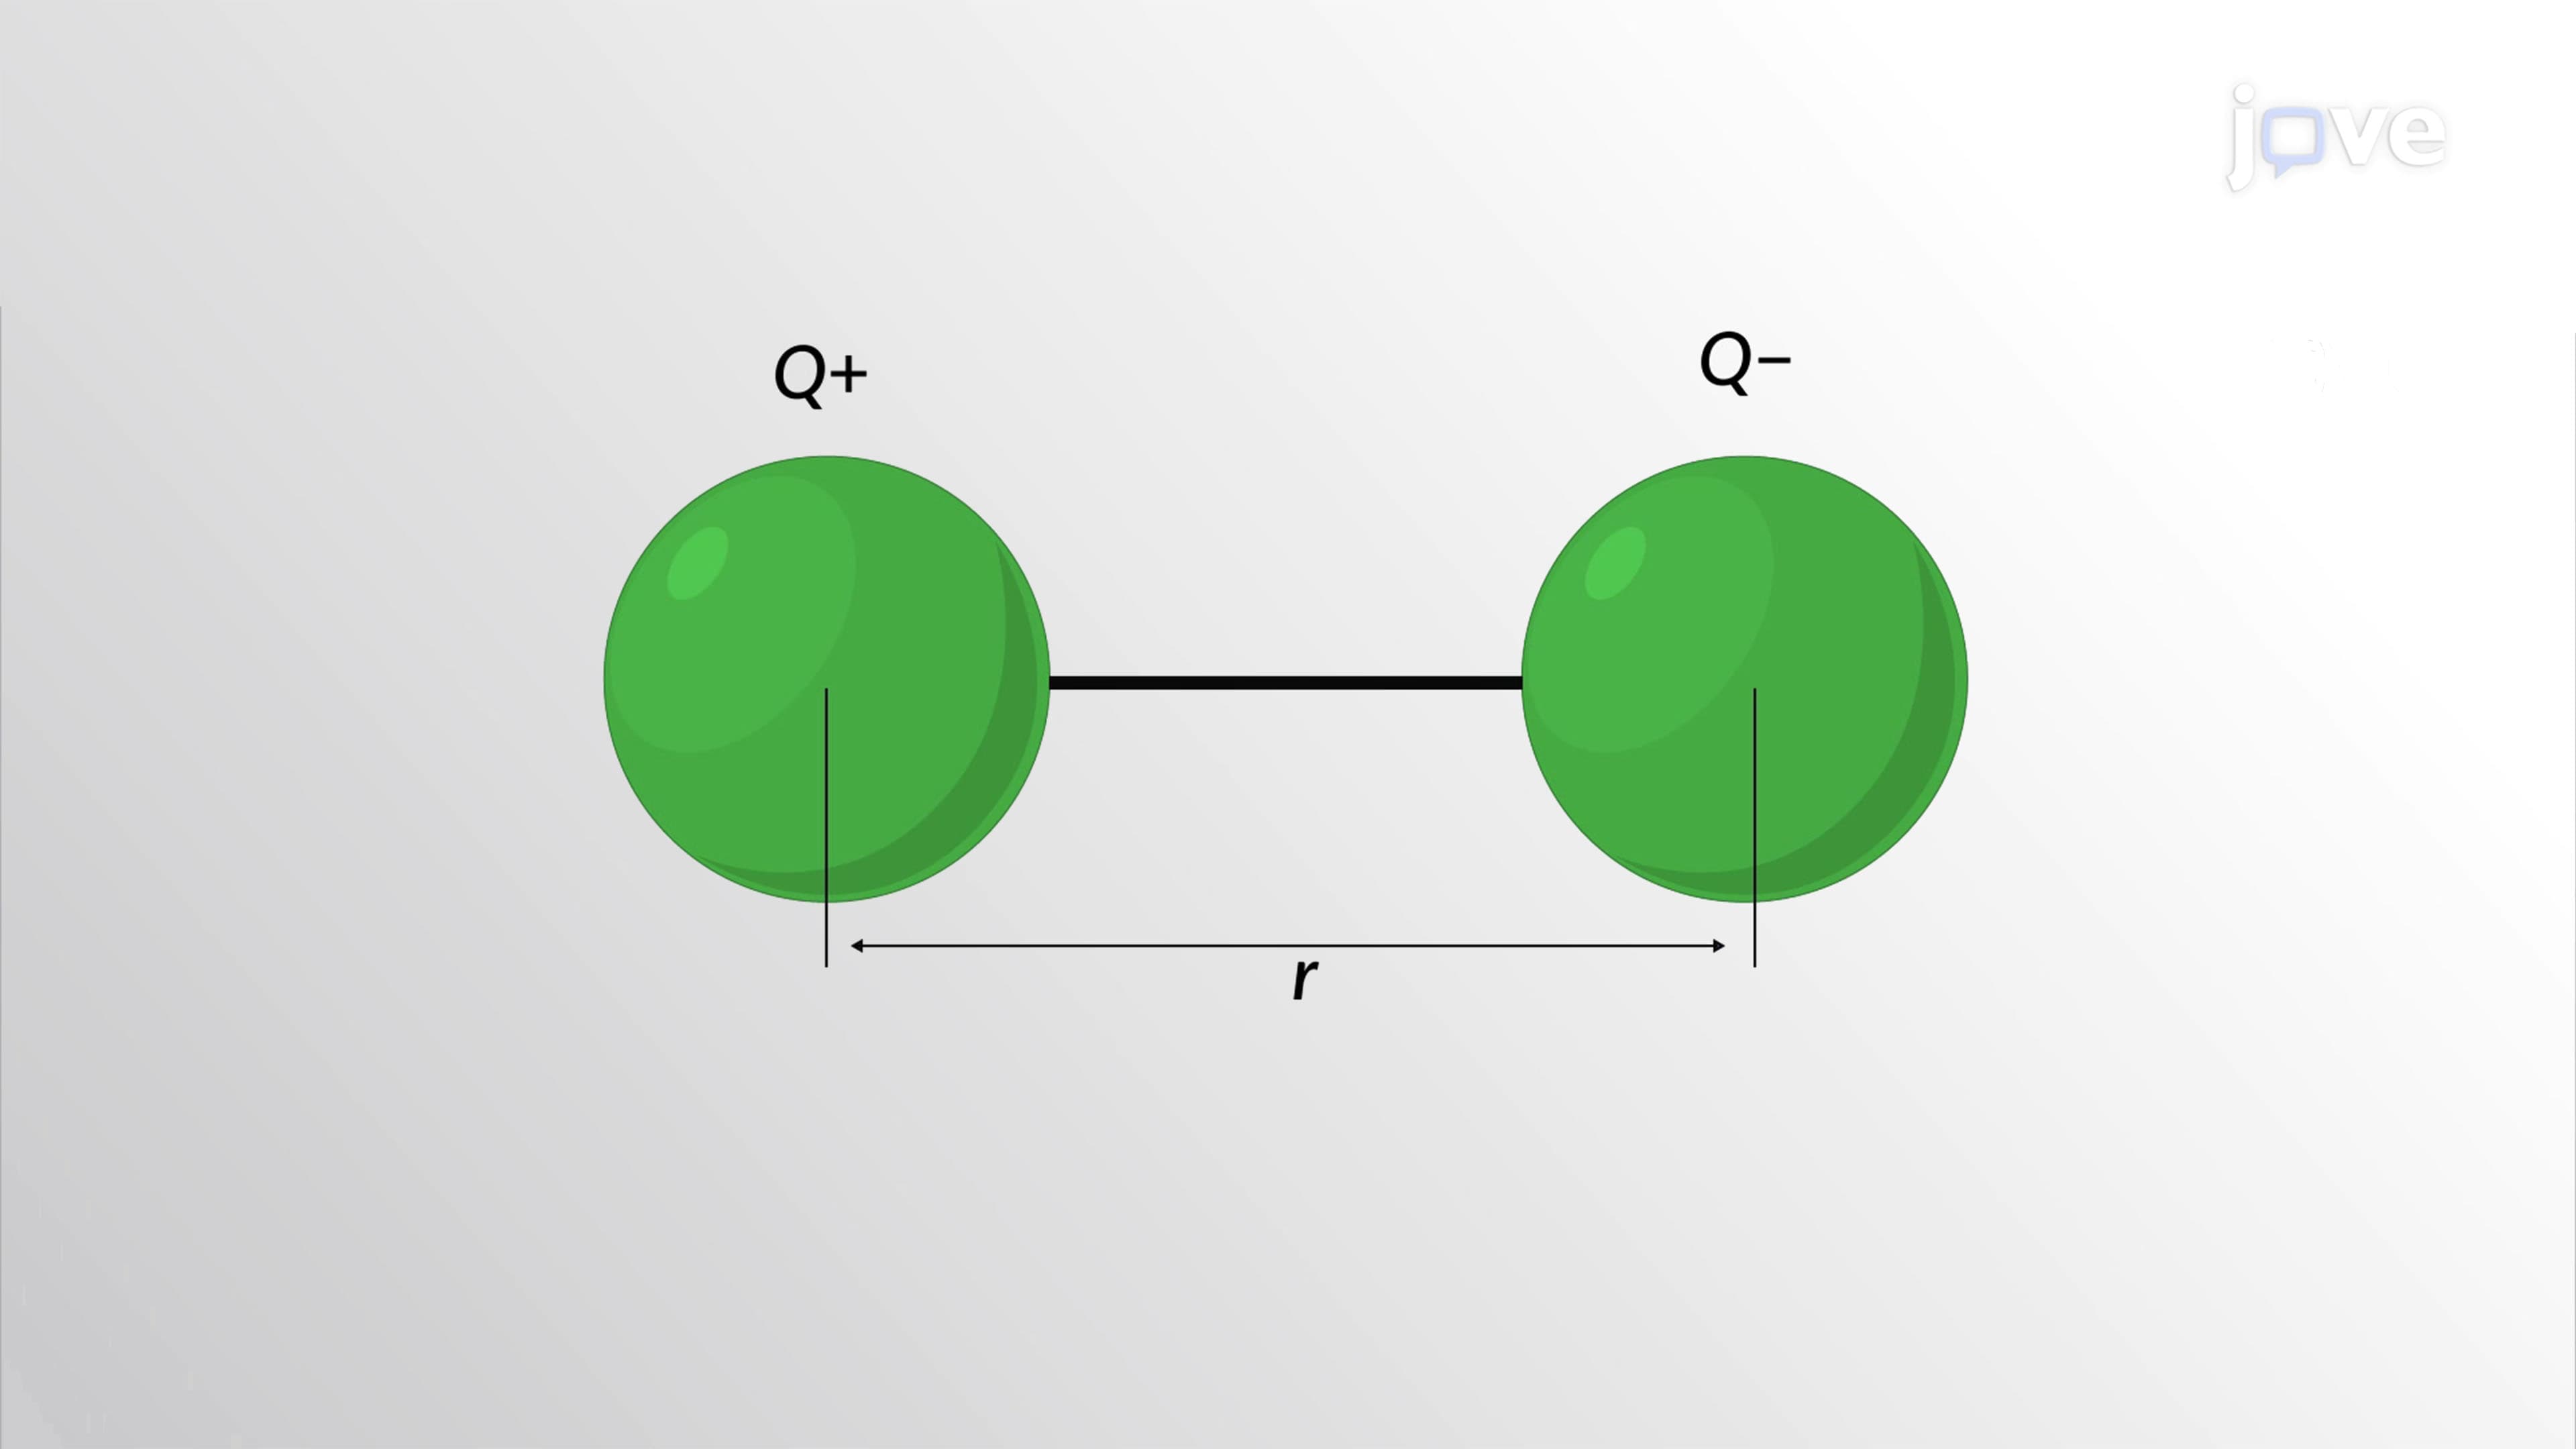
Bond Polarity, Dipole Moment, and Percent Ionic Character
 01:27
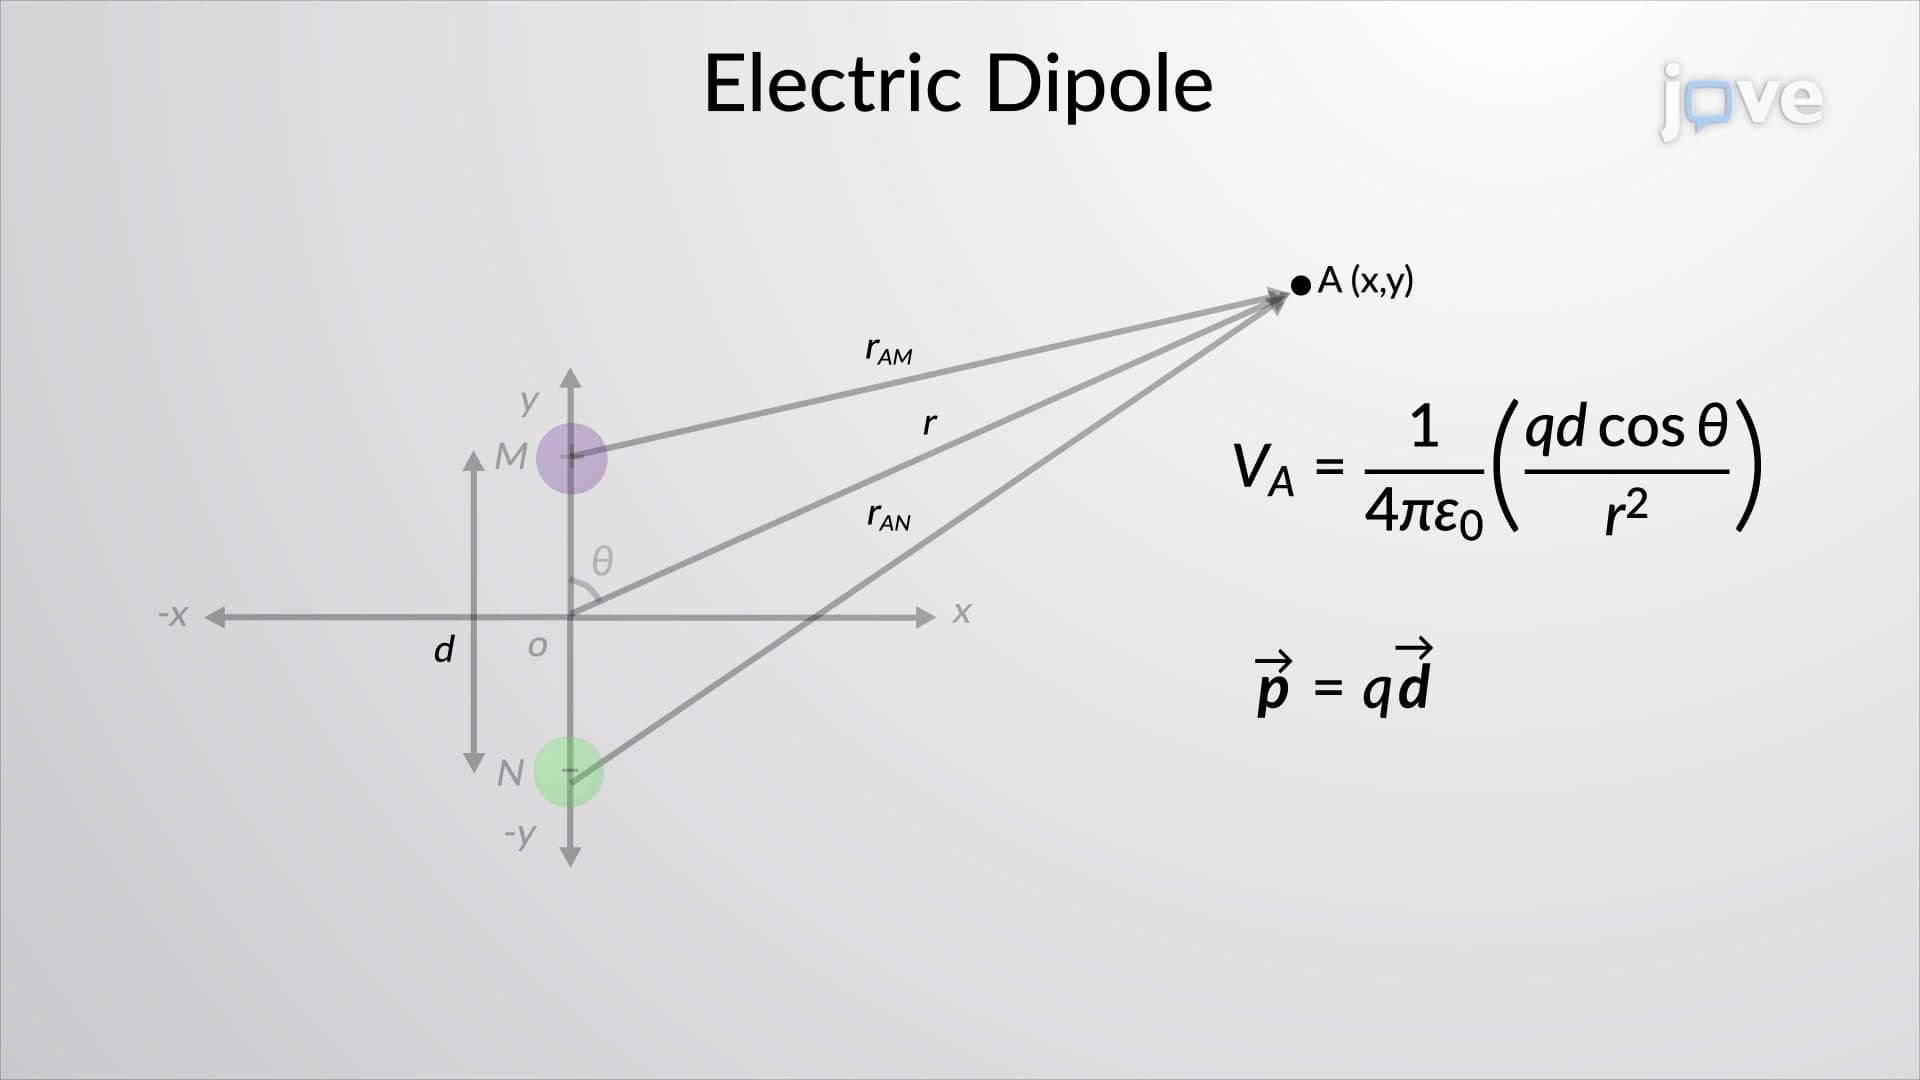
Calculations of Electric Potential II
Consider a...
 01:30
Electric Dipoles and Dipole Moment
Theoretically, studying electric dipoles leads to understanding why the resultant electric forces around us are weak. Since electric forces are strong, remnant net charges are rare. Hence, the interaction between dipoles helps us understand electrical interactions in...