Related Experiment Video
Updated: May 15, 2025

Fabrication of Three-Dimensional Graphene-Based Polyhedrons via Origami-Like Self-Folding
Published on: September 23, 2018
Gold Clusters on Graphene/Graphite-Structure and Energy Landscape
Manoj Settem1, Melisa M Gianetti2, Roberto Guerra3
1Dipartimento di Ingegneria Meccanica e Aerospaziale Sapienza Università di Roma via Eudossiana 18 00184 Roma Italy.

Abstract:
Adopting an advanced microscopic model of the Au-graphite interaction, a systematic study of Au nanoclusters (up to sizes of 11 238 atoms) on graphene and on graphite is carried out to explore their structure and energy landscape. Using parallel tempering molecular dynamics, structural distribution as a function of temperature is calculated in the entire temperature range. Low-energy structures are identified through a combination of structural optimization and Wulff-Kaischew construction which are then used to explore the energy landscape. The potential energy surface (PES), which is energy as a function of translation and rotation, is calculated for a few Au nanoclusters along specific directions on carbon lattice. Minimum-energy pathways are identified on the PES indicating a reduced barrier for pathways involving simultaneous rotation and translation. Diffusion simulations of Au233 on graphite show that diffusion mechanism is directly related to the PES, and the information of the cluster pinning events is already present in the PES. Finally, a comparison of various interaction models highlights the importance of reasonably correct Au-C interactions which is crucial for studying the energy landscape and cluster sliding.
Related Concept Videos
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...
 02:54
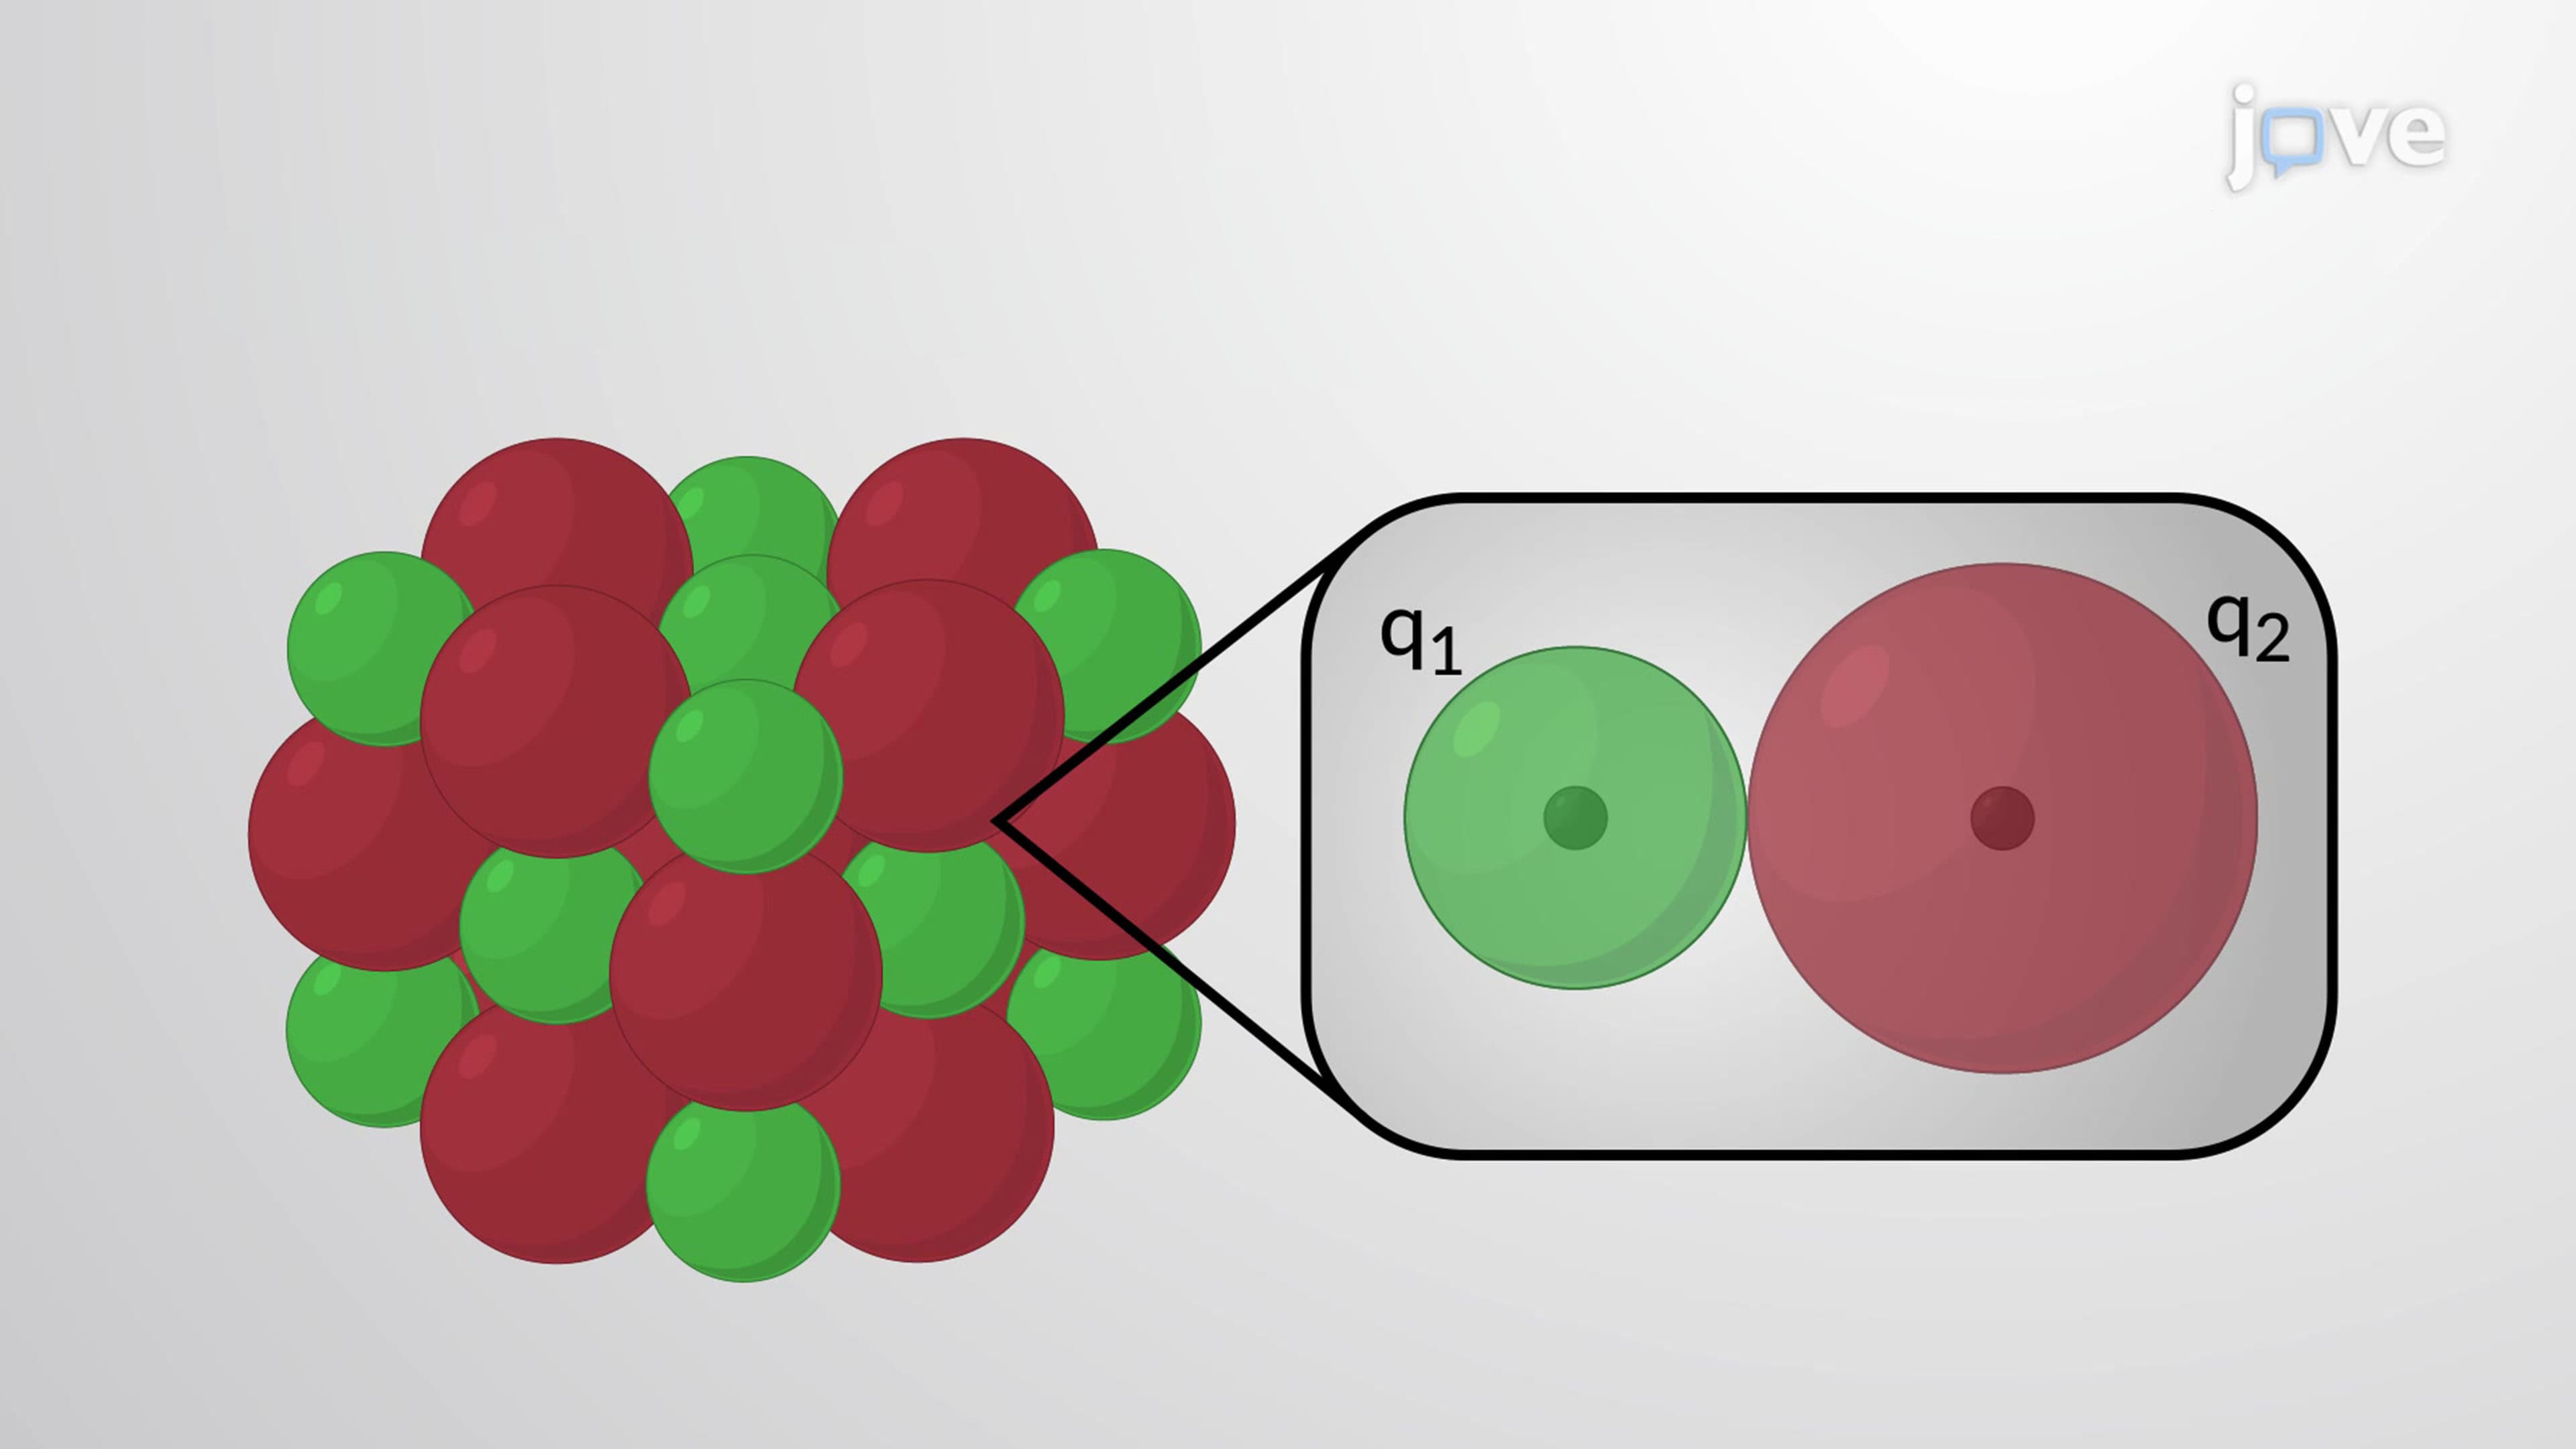
Trends in Lattice Energy: Ion Size and Charge