Related Experiment Video
Updated: Jul 13, 2026
Monitoring Protein Adsorption with Solid-state Nanopores
Published on: December 2, 2011
From slit pores to 3D frameworks: Advances in molecular modeling of adsorption in nanoporous carbons
Nicholas J Corrente1, Alexander V Neimark1
1Department of Chemical and Biochemical Engineering, Rutgers, The State University of New Jersey, Piscataway, NJ 08854, United States.

Abstract:
Recent advances in computational capabilities have revolutionized the modeling of nanoporous carbons, enabling a transition from idealized pore descriptions to versatile three-dimensional molecular models. This review traces the evolution from traditional continuous potential methods and simple pore models to modern simulation techniques that generate realistic carbon structures incorporating surface heterogeneity, pore connectivity, and framework flexibility. We examine various approaches including Hybrid Reverse Monte Carlo, Quench Molecular Dynamics, and Annealed Molecular Dynamics methods, discussing their respective strengths and limitations. Particular attention is given to the choice of interatomic potentials and their impact on structural predictions. The development of million-atom models captures long-range ordering effects previously inaccessible to simulation. Applications of the 3D models demonstrate their ability to quantitatively predict adsorption behavior and provide the improved characterization of practical carbons using novel methods such as 3D-VIS and APDM. Recent hybrid MD/MC approaches, which incorporate the effects of structure flexibility, offer new insights into adsorbate-induced structural changes. This review highlights how advancing computational methods are bridging the gap between molecular-level understanding and practical applications in the carbon materials design and modeling of adsorption processes.
More Related Videos
Related Concept Videos
 01:28
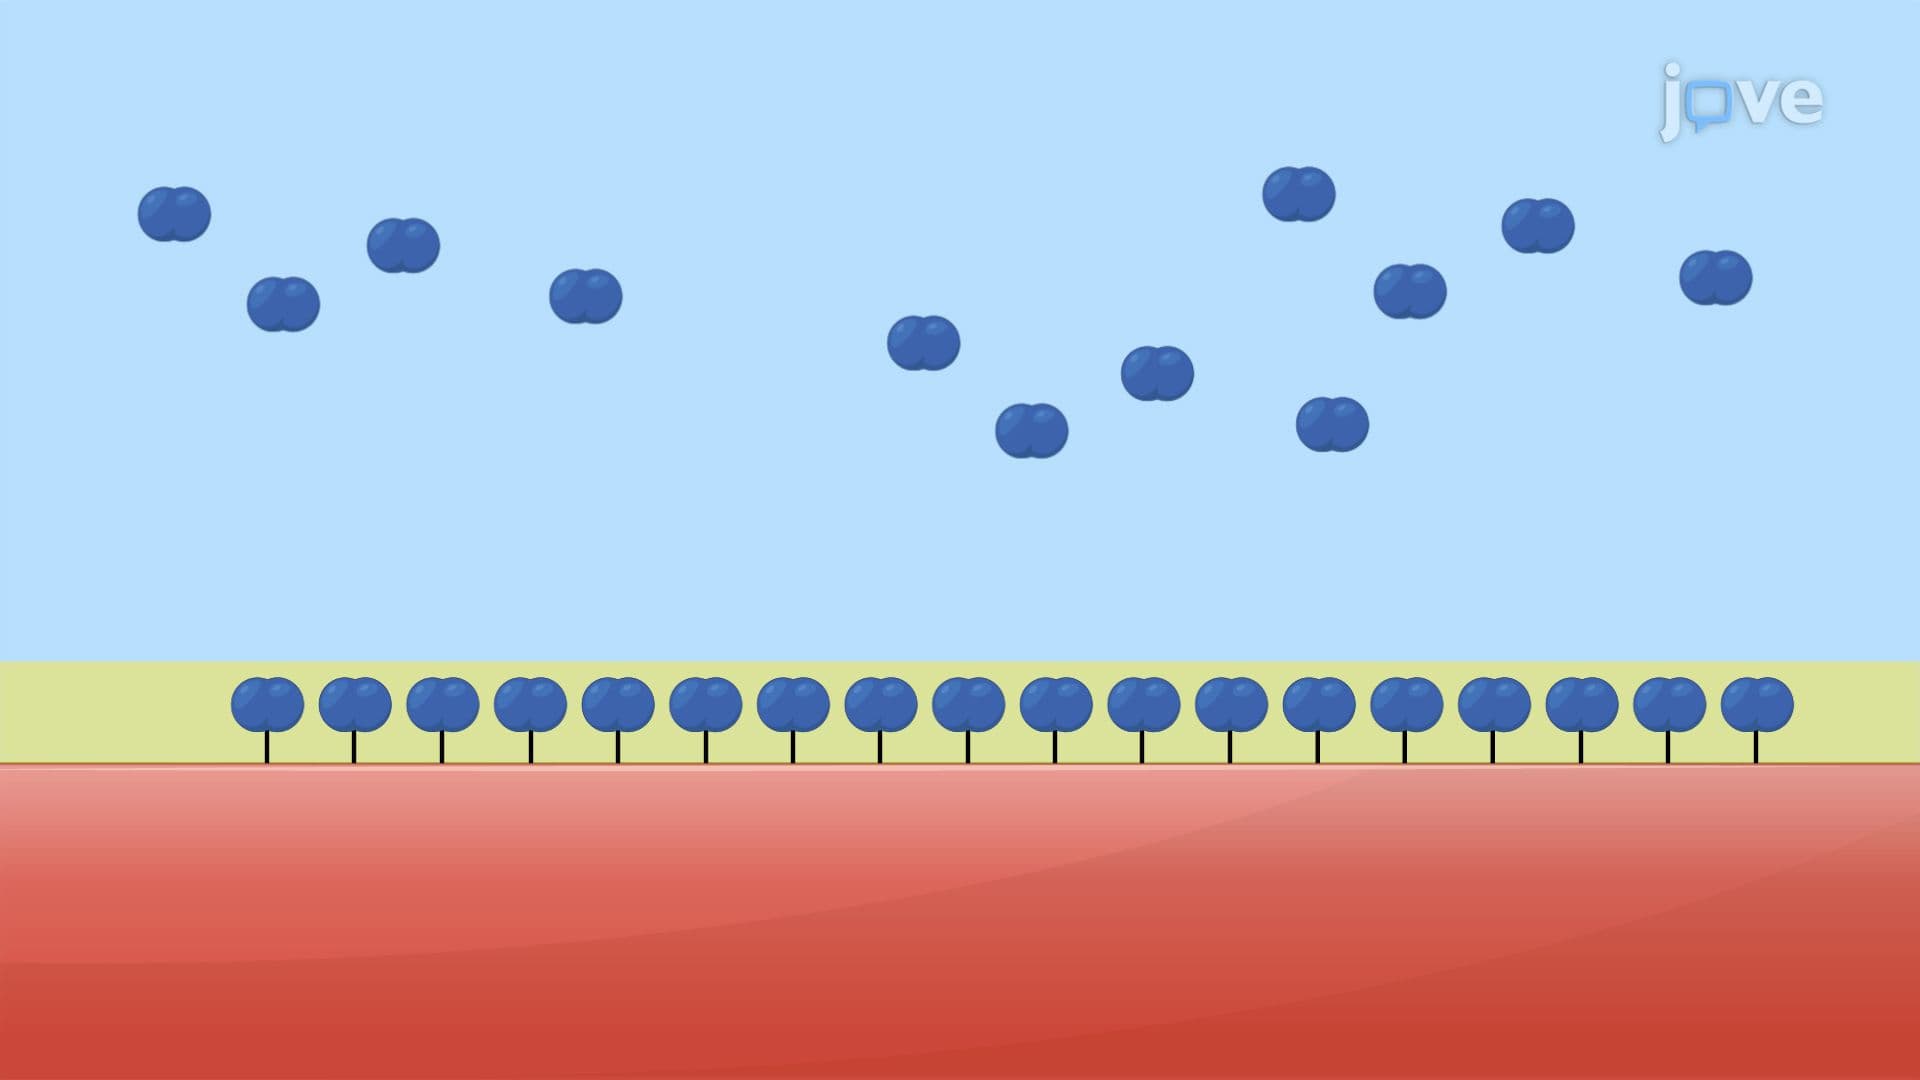
Adsorption of Gases on Solids
 01:29
Adsorption Isotherms I
 01:25
Adsorption Isotherms II