Related Experiment Video
Updated: May 14, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Diagonalization without Diagonalization: A Direct Optimization Approach for Solid-State Density Functional Theory
Tianbo Li1, Min Lin1, Stephen G Dale2
1SEA AI Lab, Singapore 138522, Singapore.

Abstract:
We present a novel approach to address the challenges of variable occupation numbers in direct optimization of density functional theory (DFT). By parametrizing both the eigenfunctions and the occupation matrix, our method minimizes the free energy with respect to these parameters. As the stationary conditions require the occupation matrix and the Kohn-Sham Hamiltonian to be simultaneously diagonalizable, this leads to the concept of "self-diagonalization," where, by assuming a diagonal occupation matrix without loss of generality, the Hamiltonian matrix naturally becomes diagonal at stationary points. Our method incorporates physical constraints on both the eigenfunctions and the occupations into the parametrization, transforming the constrained optimization into an fully differentiable unconstrained problem, which is solvable via gradient descent. Implemented in JAX, our method was tested on aluminum and silicon, confirming that it achieves efficient self-diagonalization, produces the correct Fermi-Dirac distribution of the occupation numbers and yields band structures consistent with those obtained with SCF eigensolver methods in Quantum Espresso.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:10
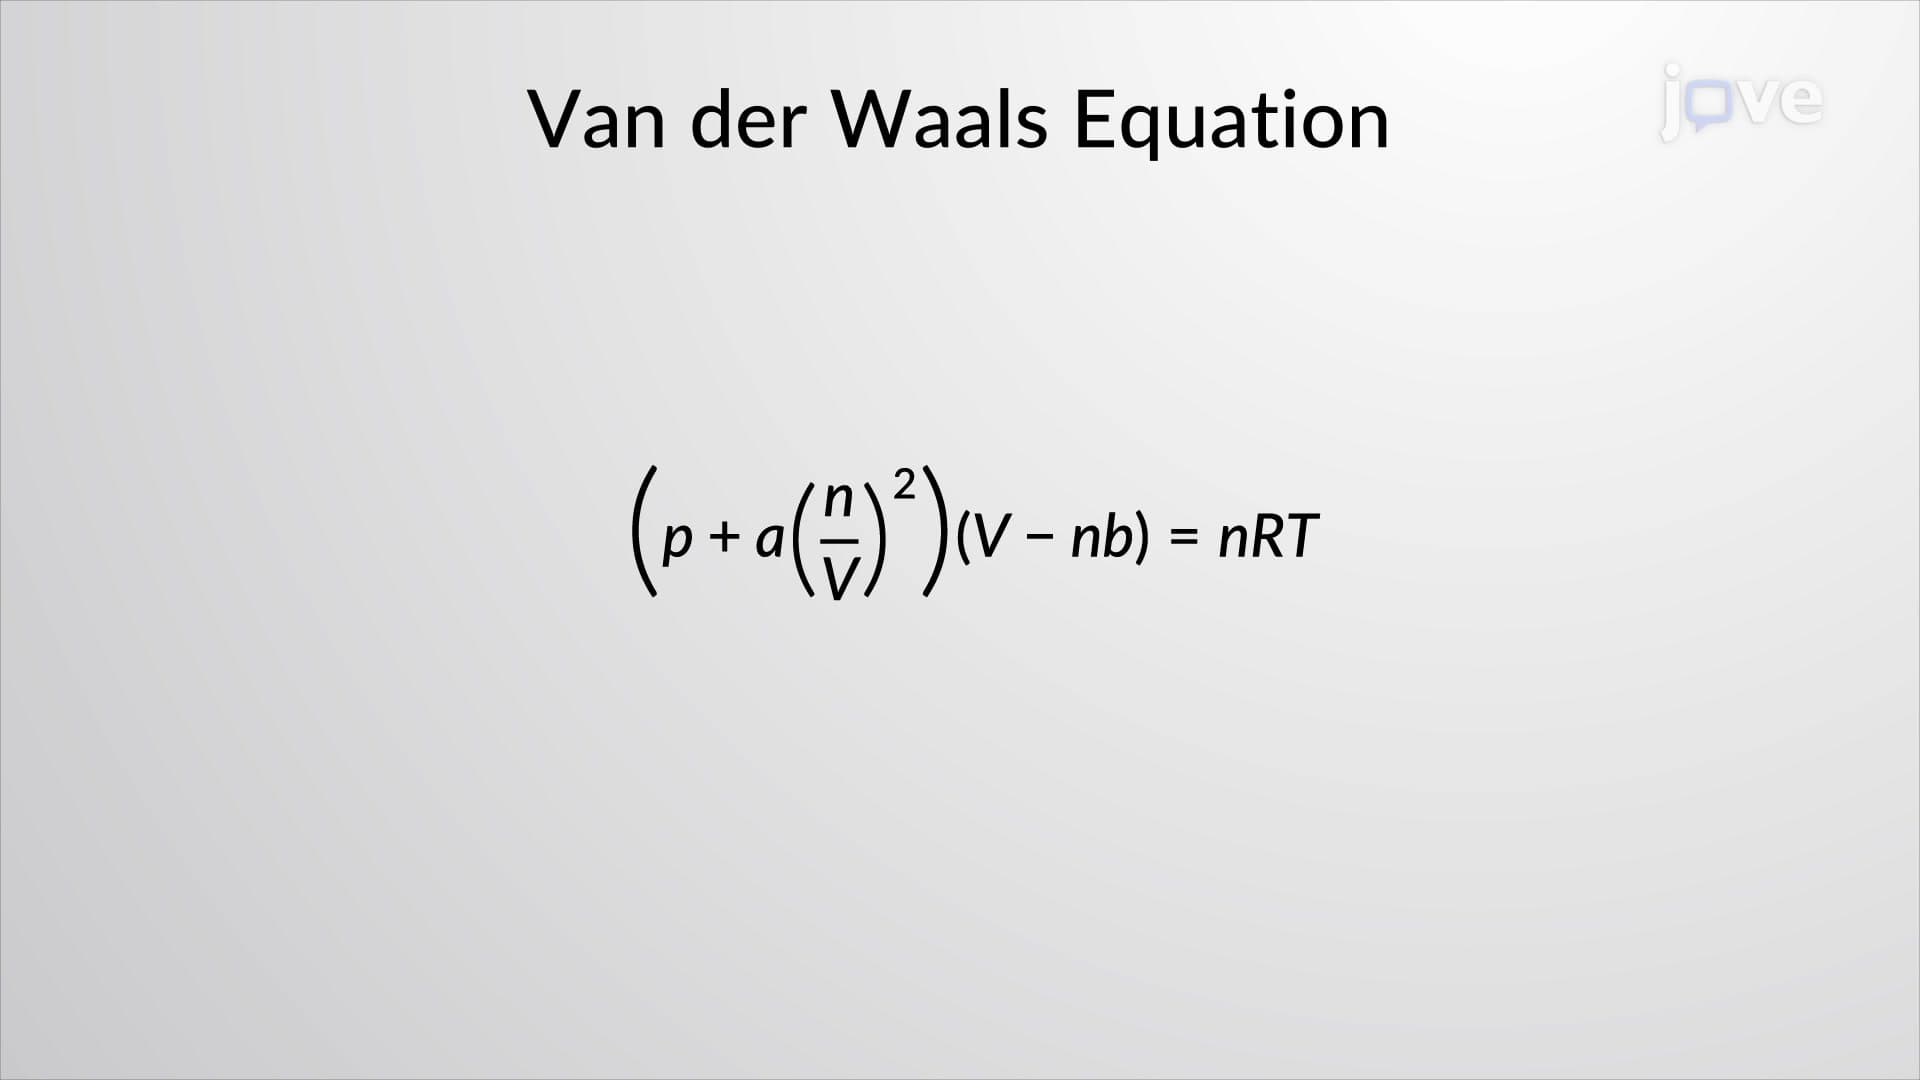
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...
 01:12
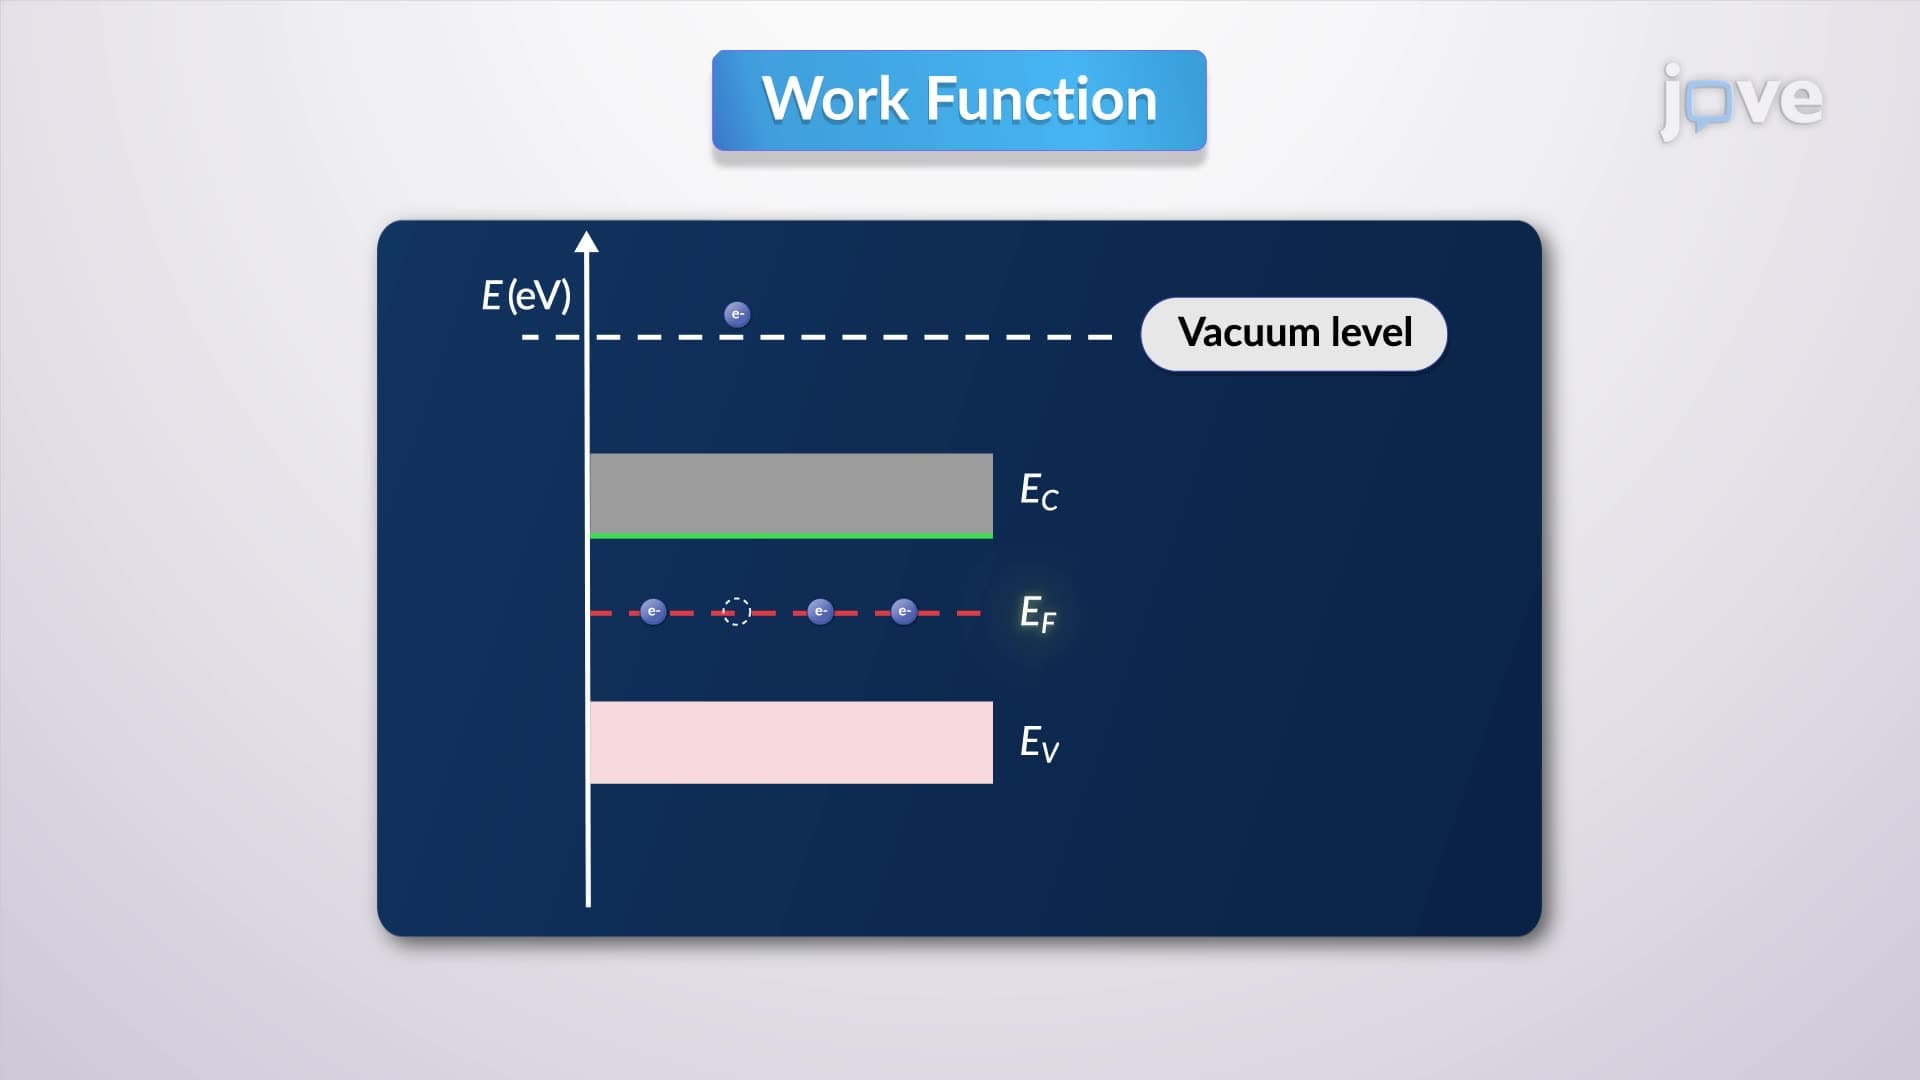
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 01:26
Thermodynamic Potentials
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...