Related Experiment Video
Updated: May 12, 2025

Sequence-specific Labeling of Nucleic Acids and Proteins with Methyltransferases and Cofactor Analogues
Published on: November 22, 2014
METTL3-dependent m6A modification of SNAP29 induces "autophagy-mitochondrial crisis" in the ischemic microenvironment
Ningning Yang1,2,3, Yingying Lai1,2,4, Gaoxiang Yu1,2,4
1Department of Orthopaedics, The Second Affiliated Hospital and Yuying Children's Hospital of Wenzhou Medical University, Wenzhou, China.

Abstract:
Necrosis at the ischemic distal end of flap transplants increases patients' pain and economic burden. Reactive oxygen species (ROS) and mitochondrial damage are crucial in regulating parthanatos, but the mechanisms linking disrupted macroautophagic/autophagic flux to parthanatos in ischemic flaps remain unclear. The results of western blotting, immunofluorescence staining, and a proteomic analysis revealed that the autophagic protein SNAP29 was deficient in ischemic flaps, resulting in disrupted autophagic flux, increased ROS-induced parthanatos, and aggravated ischemic flap necrosis. The use of AAV vector to restore SNAP29 in vivo mitigated the disruption of autophagic flux and parthanatos. Additionally, quantification of the total m6A level and RIP-qPCR, MeRIP-qPCR, and RNA stability assessments were performed to determine differential Snap29 mRNA m6A methylation levels and mRNA stability in ischemic flaps. Various in vitro and in vivo tests were conducted to verify the ability of METTL3-mediated m6A methylation to promote SNAP29 depletion and disrupt autophagic flux. Finally, we concluded that restoring SNAP29 by inhibiting METTL3 and YTHDF2 reversed the "autophagy-mitochondrial crisis", defined for the first time as disrupted autophagic flux, mitochondrial damage, mitochondrial protein leakage, and the occurrence of parthanatos. The reversal of this crisis ultimately promoted the survival of ischemic flaps.Abbreviations: AAV = adeno-associated virus; ACTA2/α-SMA = actin alpha 2, smooth muscle, aorta; AIFM/AIF = apoptosis-inducing factor, mitochondrion-associated; ALKBH5 = alkB homolog, RNA demythelase; Baf A1 = bafilomycin A1; CQ = chloroquine; DHE = dihydroethidium; ECs = endothelial cells; F-CHP = 5-FAM-conjugated collagen-hybridizing peptide; GO = gene ontology; HUVECs = human umbilical vein endothelial cells; KEGG = Kyoto Encyclopedia of Genes and Genomes; LC-MS/MS = liquid chromatography-tandem mass spectrometry; LDBF = laser doppler blood flow; m6A = N6-methyladenosine; MAP1LC3/LC3 = microtubule-associated protein 1 light chain 3; MeRIP = methylated RNA immunoprecipitation; METTL3 = methyltransferase 3, N6-adenosine-methyltransferase complex catalytic subunit; NAC = N-acetylcysteine; OGD = oxygen glucose deprivation; PAR = poly (ADP-ribose); PARP1 = poly (ADP-ribose) polymerase family, member 1; PECAM1/CD31 = platelet/endothelial cell adhesion molecule 1; ROS = reactive oxygen species; RT-qPCR = reverse transcription quantitative polymerase chain reaction; RIP = RNA immunoprecipitation; SNAP29 = synaptosomal-associated protein 29; SNARE = soluble N-ethylmaleimide-sensitive factor attachment protein receptor; SQSTM1 = sequestosome 1; SRAMP = sequence-based RNA adenosine methylation site predicting; STX17 = syntaxin 17; TMT = tandem mass tag; TUNEL = terminal deoxynucleotidyl transferase dUTP nick end labeling; VAMP8 = vesicle-associated membrane protein 8; WTAP = WT1 associating protein; YTHDF2 = YTH N6-methyladenosine RNA binding protein 2; 3' UTR = 3'-untranslated region.
Insights
Restoring synaptosomal-associated protein 29 (SNAP29) in ischemic flaps prevents necrosis by reversing the autophagy-mitochondrial crisis. This involves inhibiting METTL3-mediated m6A methylation to restore autophagic flux and reduce parthanatos.
Area of Science:
- Molecular Biology
- Cellular Biology
- Regenerative Medicine
Background:
- Ischemic flap necrosis increases patient pain and healthcare costs.
- Mechanisms linking disrupted autophagy to parthanatos in ischemic flaps are unclear.
- Reactive oxygen species (ROS) and mitochondrial damage contribute to parthanatos.
Purpose of the Study:
- To investigate the role of SNAP29 in ischemic flap necrosis.
- To elucidate the link between disrupted autophagic flux, m6A methylation, and parthanatos.
- To identify therapeutic targets for promoting ischemic flap survival.
Main Methods:
- Western blotting, immunofluorescence staining, and proteomic analysis identified SNAP29 deficiency.
- Adeno-associated virus (AAV) vector was used to restore SNAP29 in vivo.
- RNA immunoprecipitation (RIP-qPCR), methylated RNA immunoprecipitation (MeRIP-qPCR), and RNA stability assays assessed m6A methylation of Snap29 mRNA.
- In vitro and in vivo experiments verified METTL3's role in SNAP29 depletion and autophagic flux disruption.
Main Results:
- SNAP29 deficiency in ischemic flaps disrupted autophagic flux, increased ROS-induced parthanatos, and aggravated necrosis.
- In vivo SNAP29 restoration via AAV mitigated autophagic flux disruption and parthanatos.
- METTL3-mediated m6A methylation of Snap29 mRNA promoted its depletion and disrupted autophagic flux.
- Inhibiting METTL3 and YTHDF2 restored SNAP29, reversed the 'autophagy-mitochondrial crisis', and promoted flap survival.
Conclusions:
- SNAP29 deficiency is a key driver of ischemic flap necrosis via the autophagy-mitochondrial crisis.
- METTL3-mediated m6A methylation of Snap29 mRNA is a critical regulatory mechanism.
- Targeting METTL3 and YTHDF2 to restore SNAP29 presents a promising therapeutic strategy for ischemic flap survival.
Related Concept Videos
 01:22
PI3K/mTOR/AKT Signaling Pathway
 02:23
Master Transcription Regulators
 01:19
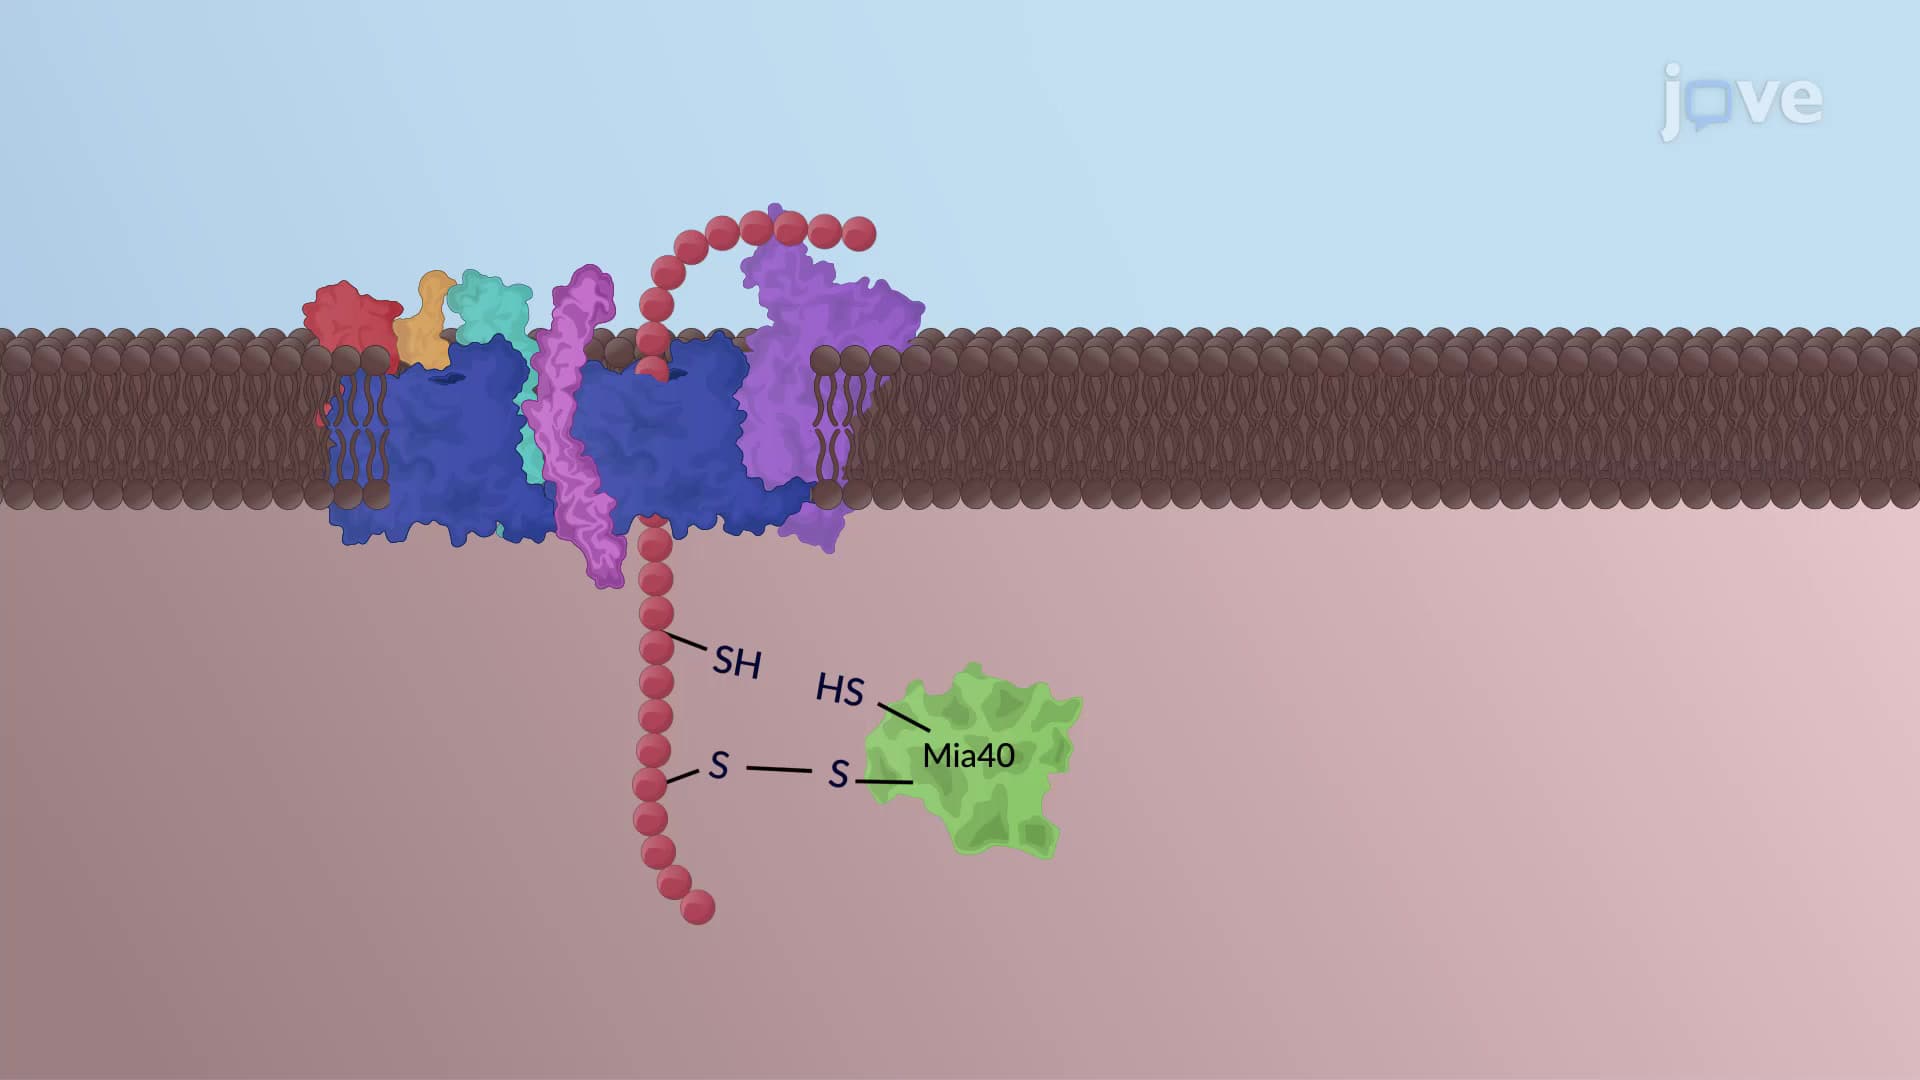
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 03:03
mTOR Signaling and Cancer Progression
The mTOR pathway or the...
 01:07
MAPK Signaling Cascades
 01:16
Calmodulin-dependent Signaling
The Ca2+-CaM complex does not have enzymatic activity by itself. Instead, the complex binds downstream target proteins, including membrane proteins or enzymes,...