Related Experiment Video
Updated: May 12, 2025


Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
Case Report: Unusual Neurological Features of Leigh Syndrome due to m.8993T>G Pathogenic Variant in the MT-ATP6 Gene
Ramya Treitel1, Julie McLaughlin2, Marta Frigeni2,3
1Department of Pediatrics, Division of Pediatric Neurology, Zucker School of Medicine at Hofstra/Northwell, New York, New York, USA.

Abstract:
The MT-ATP6 gene m.8993T>G pathogenic variant has been associated with Leigh syndrome, especially in patients exhibiting a high degree of heteroplasmy. Although patients may present with a wide phenotypic spectrum, characteristic findings include bilateral, symmetric hyperintensities in the basal ganglia and brainstem on brain MRI, particularly on T2-weighted and fluid-attenuated inversion recovery sequences. Additionally, the biochemical phenotype associated with this pathogenic variant often mimics that of multiple carboxylase deficiency and proximal urea cycle disorders. This report describes a male infant with an atypical neurological presentation of Leigh syndrome. At 2 months of age, he presented with status epilepticus of left temporal origin that was refractory to treatment. Initial brain MRI revealed a large region of non-enhancing signal abnormality in the left temporal lobe, raising concern for an infectious etiology. However, biochemical testing revealed hypocitrullinemia, elevated 3-hydroxyisovalerylcarnitine, elevated propionylcarnitine, and urinary excretion of lactate and pyruvate, prompting further investigation for MT-ATP6 mitochondrial disease. Mitochondrial DNA analysis confirmed the presence of a homoplasmic m.8993T>G pathogenic variant in the MT-ATP6 gene. Despite treatment with citrulline and high-dose biotin, the patient died 5 weeks later due to cardiorespiratory failure following a severe respiratory infection. Retrospective review of his newborn screening revealed two screens positive for low citrulline that were ultimately cleared on a third screen, delaying the diagnosis. This case underscores the importance of considering MT-ATP6 mitochondrial disease in the differential diagnosis of patients presenting with atypical neurological symptoms and biochemical abnormalities. It also highlights the value of newborn screening in identifying potential mitochondrial disorders, where early diagnosis and timely intervention may improve outcomes, even in severe cases.


Related Concept Videos
 01:18
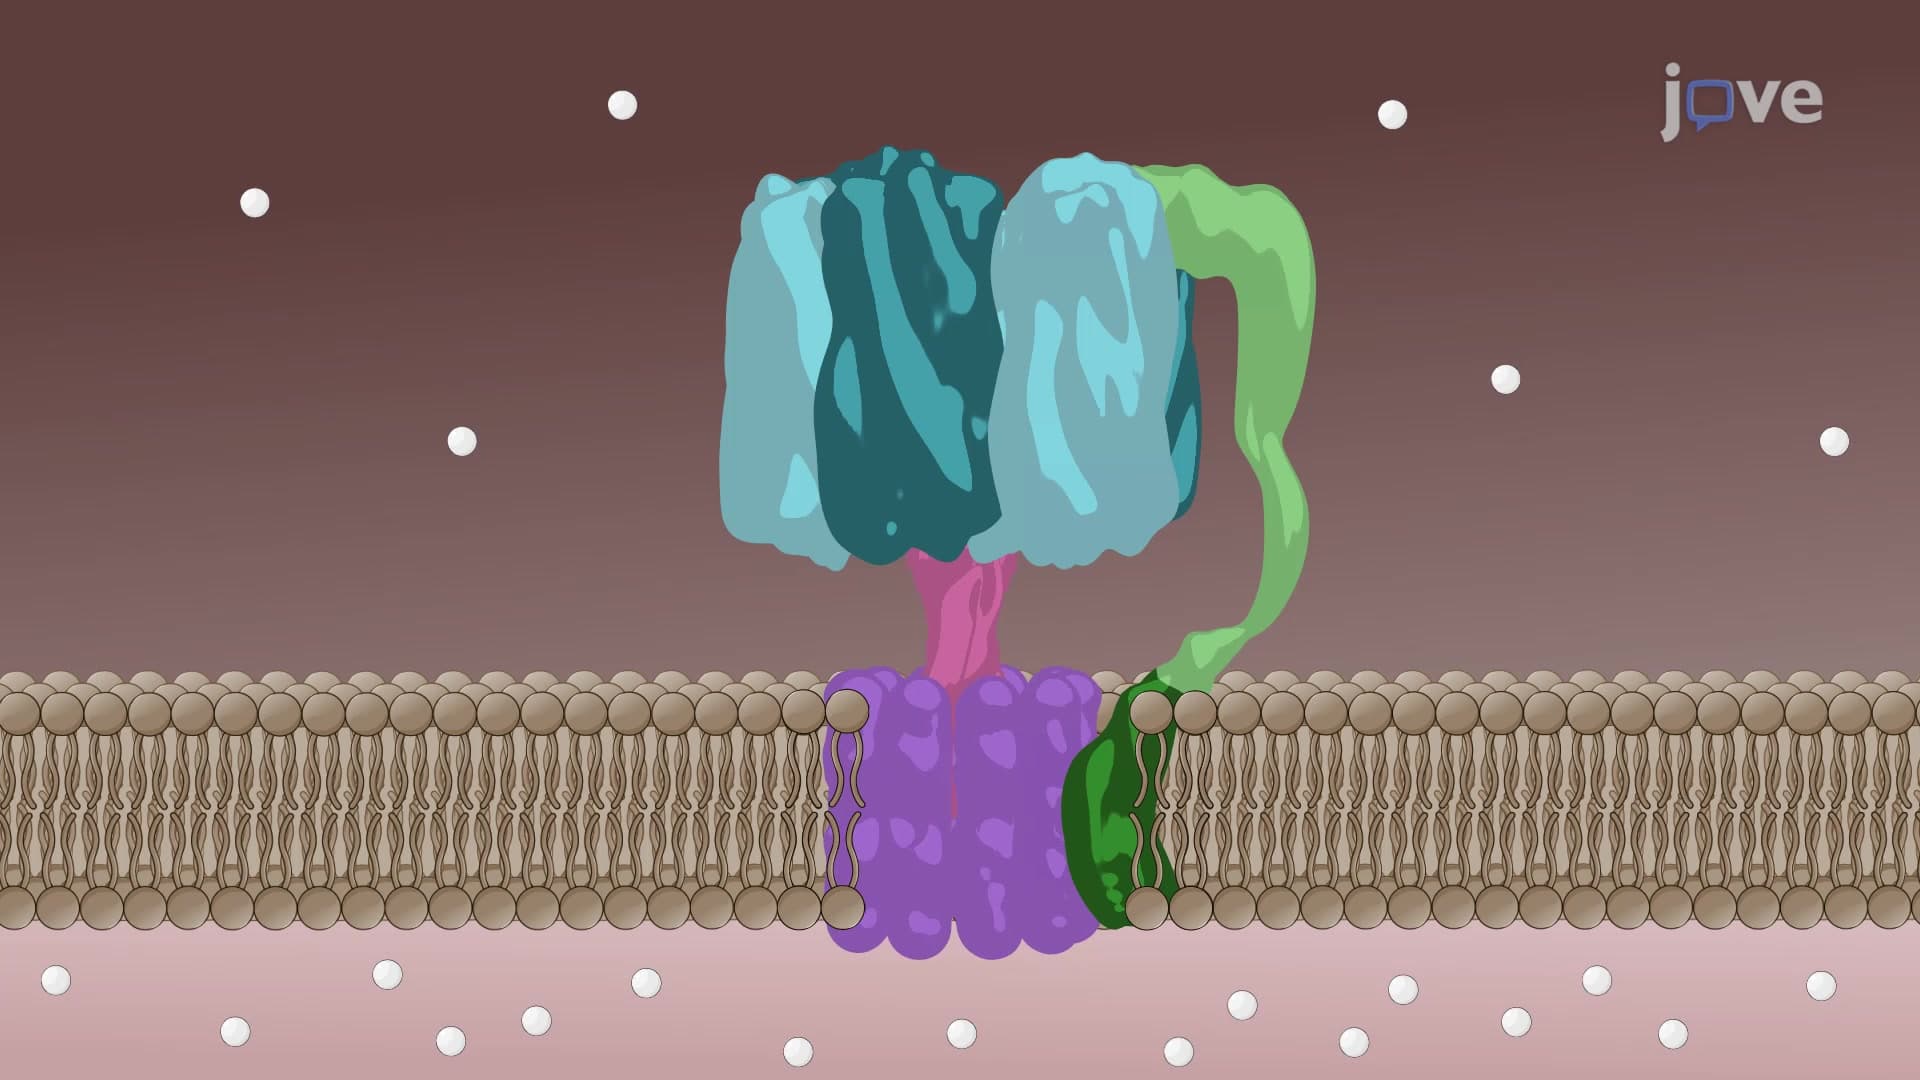
ATP Synthase: Structure
 01:48
ATP Synthase: Mechanism
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
 02:59
Animal Mitochondrial Genetics
 01:33
Pleiotropy