Related Experiment Video
Updated: May 17, 2025

Evaluation of Vascular Control Mechanisms Utilizing Video Microscopy of Isolated Resistance Arteries of Rats
Published on: December 5, 2017
Deficiency of NPR-C triggers high salt-induced thoracic aortic dissection by impairing mitochondrial homeostasis
Jin-Qiu Wei1,2, Yi Yang3, Wen-Hui Zhai1
1Department of Cardiovascular Medicine, Department of Hypertension, Ruijin Hospital and State Key Laboratory of Medical Genomics, Shanghai Key Laboratory of Hypertension, Shanghai Institute of Hypertension, Shanghai Jiao Tong University School of Medicine, Ruijin 2nd Road 197, Shanghai 200025, China.

Aims:
Thoracic aortic dissection (TAD) is a highly fatal disease lacking effective pharmacologic interventions in clinical practice. Emerging evidence indicates that the natriuretic peptide receptor C (NPR-C) plays a crucial role in the regulation of cardiovascular diseases. However, the precise involvement of NPR-C in TAD remains elusive. In this study, the role and molecular mechanisms of NPR-C in the pathogenesis of TAD were investigated.
Methods And Results:
Through integrated analyses of human TAD transcriptome and single-cell sequencing data sets, we identified that NPR-C was downregulated in the aortas of acute TAD patients and in beta-aminopropionitrile (BAPN)-treated mice. Intriguingly, vascular smooth muscle cell (VSMC)-specific NPR-C knockout (NPR-CSMKO) mice, rather than endothelial cell-specific NPR-C knockout mice, developed TAD after treated with angiotensin II (Ang II) plus high salt diet (HSD), but not Ang II alone. Loss of NPR-C function promoted extracellular matrix degeneration, VSMCs apoptosis, and inflammation. RNA-sequencing analysis revealed that mitochondrial fatty acid oxidation (FAO) genes were significantly downregulated in the thoracic aortas of NPR-CSMKO mice treated with Ang II plus HSD. Notably, the expression of HADHB, a subunit of mitochondrial trifunctional protein (MTP) responsible for FAO, was obviously decreased in NPR-CSMKO mice treated with Ang II plus HSD. Mechanistically, knockdown of NPR-C activated ERK1/2 pathway, which decreased the expression and activity of peroxisome proliferator-activated receptor γ (PPARγ) and inhibited HADHB expression. Furthermore, NPR-C agonist, C-ANP4-23, mitigated the progression of TAD in BAPN-treated mice. Activation of MTP by spermidine (SPD) effectively prevented TAD formation in NPR-CSMKO mice treated with Ang II plus HSD.
Conclusion:
Our data highlight a critical role of HSD in triggering TAD and a previously unrecognized role of NPR-C that protects against TAD through regulating mitochondrial homeostasis. Therefore, NPR-C activation and SPD supplementation could be new prevention and treatment strategies for TAD.
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:08
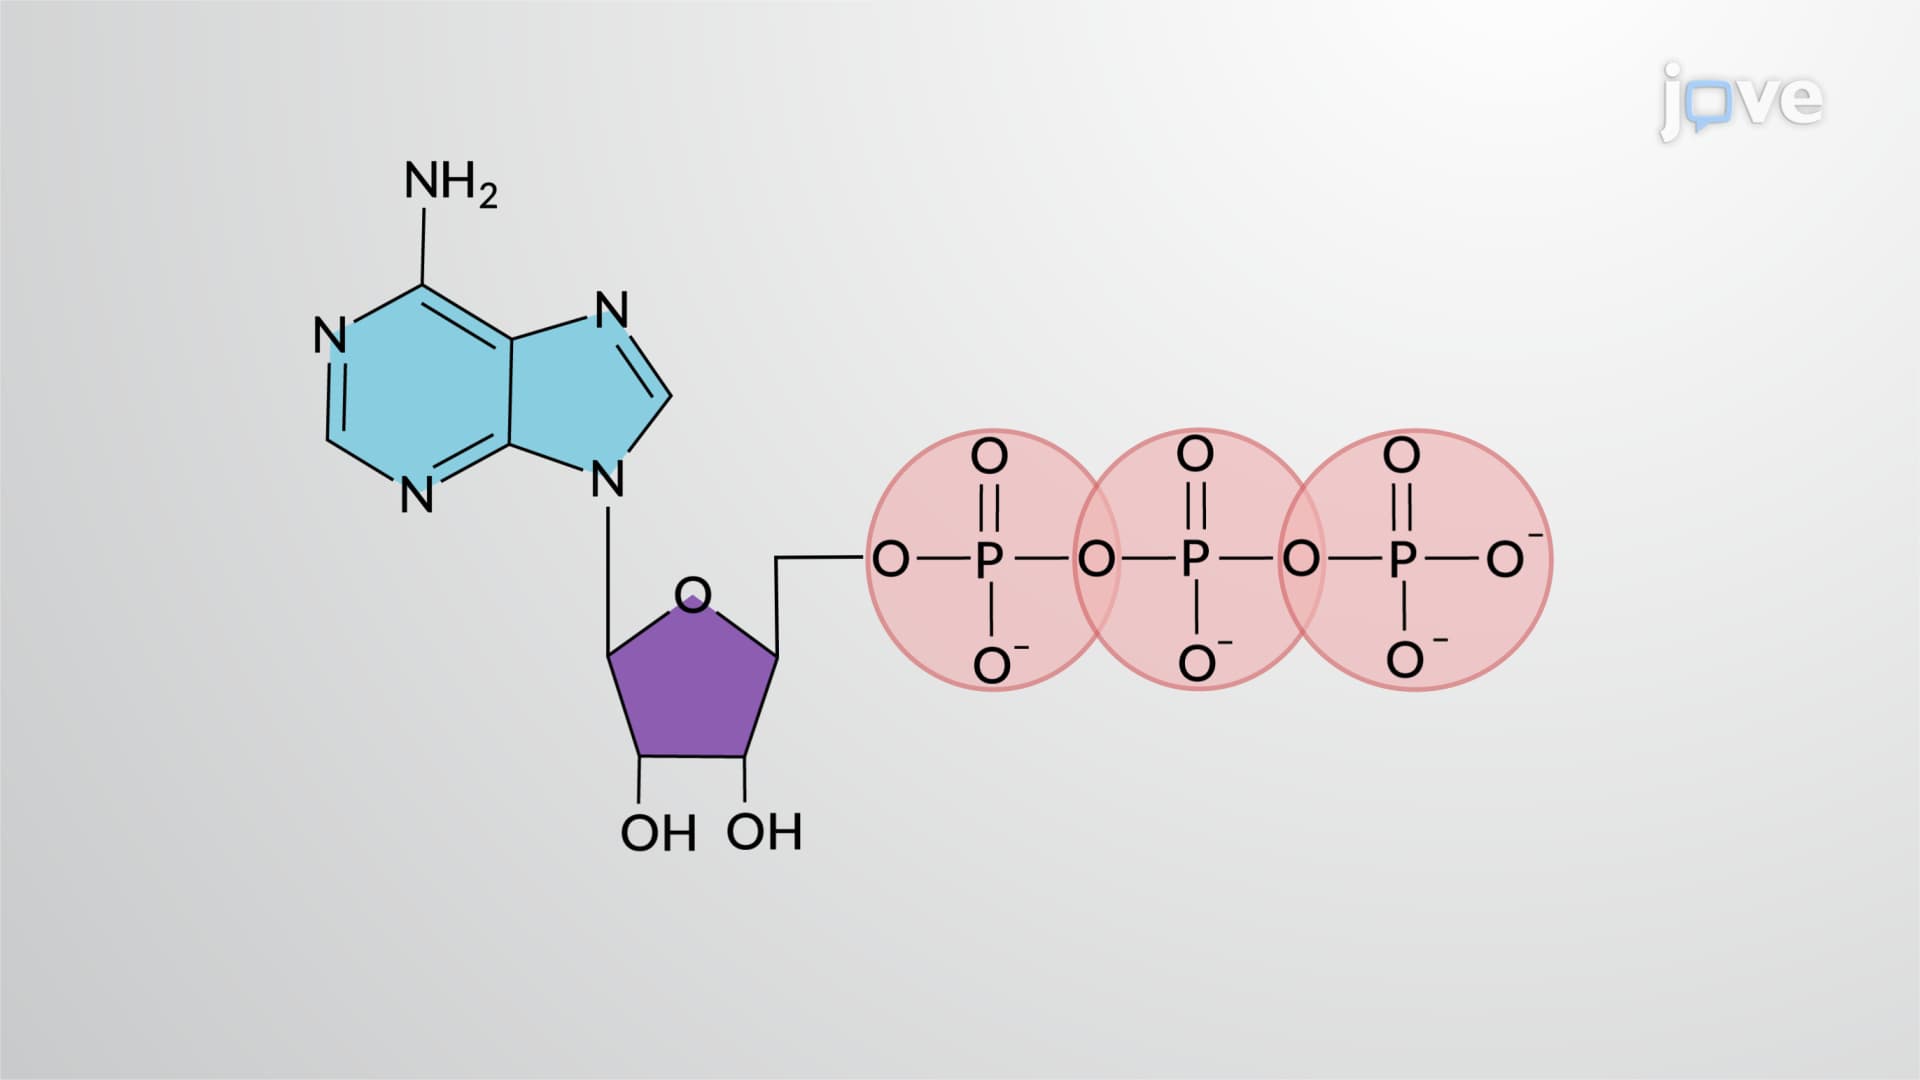
Hydrolysis of ATP
If one phosphate group is removed, a molecule of ADP—adenosine diphosphate—remains, along with inorganic phosphate. ADP can be further hydrolyzed to AMP—adenosine...
 01:18
Disorders of the Autonomic Nervous System
Raynaud's disease, also known as Raynaud's...
 01:26
Heart Failure Drugs: Inhibitors of Renin-Angiotensin System