Related Experiment Video
Updated: May 7, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Investigating Ionic Diffusivity in Amorphous LiPON using Machine-Learned Interatomic Potentials
Aqshat Seth1, Rutvij Pankaj Kulkarni1, Gopalakrishnan Sai Gautam1
1Department of Materials Engineering, Indian Institute of Science, Bengaluru 560012, India.

Abstract:
Due to its immense importance as an amorphous solid electrolyte in thin-film devices, lithium phosphorus oxynitride (LiPON) has garnered significant scientific attention. However, investigating Li+ transport within the LiPON framework, especially across a Li||LiPON interface, has proven challenging due to its amorphous nature and varying stoichiometry, necessitating large supercells and long time scales for computational models. Notably, machine-learned interatomic potentials (MLIPs) can combine the computational speed of classical force fields with the accuracy of density functional theory (DFT), making them the ideal tool for modeling such amorphous materials. Thus, in this work, we train and validate the neural equivariant interatomic potential (NequIP) framework on a comprehensive DFT-based data set consisting of 13,454 chemically relevant structures to describe LiPON. With optimized training (validation) energy and force mean absolute errors of 5.5 (6.1) meV/atom and 13.6 (13.2) meV/Å, respectively, we employ the trained potential to model Li transport in both bulk LiPON and across Li||LiPON interfaces. Amorphous LiPON structures generated by the optimized potential resemble those generated by ab initio molecular dynamics, with N being incorporated on nonbridging apical and bridging sites. Subsequent analysis of Li+ diffusivity in the bulk LiPON structures indicates broad agreement with prior computational and experimental literature. Further, we investigate the anisotropy in Li+ transport across the Li(110)||LiPON and Li(111)||LiPON interface, where we observe Li transport across the interface to be one order of magnitude slower than Li motion within the bulk Li and LiPON phases. Nevertheless, we note that this anisotropy of Li transport across the interface is minor, and we do not expect it to cause any significant impedance buildup. Finally, our work highlights the efficiency of MLIPs in enabling high-fidelity modeling of complex noncrystalline systems over large length and time scales.
More Related Videos
 11:27
A Package of Established Analytical Tools to Investigate the Solid-State Alteration of Lipid-Based Excipients
Published on: August 9, 2022
 07:31
Author Spotlight: Advancing Cell Membrane Biophysics - Exploring Interactions and Challenges Through Experimental and Computational Approaches
Published on: September 1, 2023
Related Concept Videos
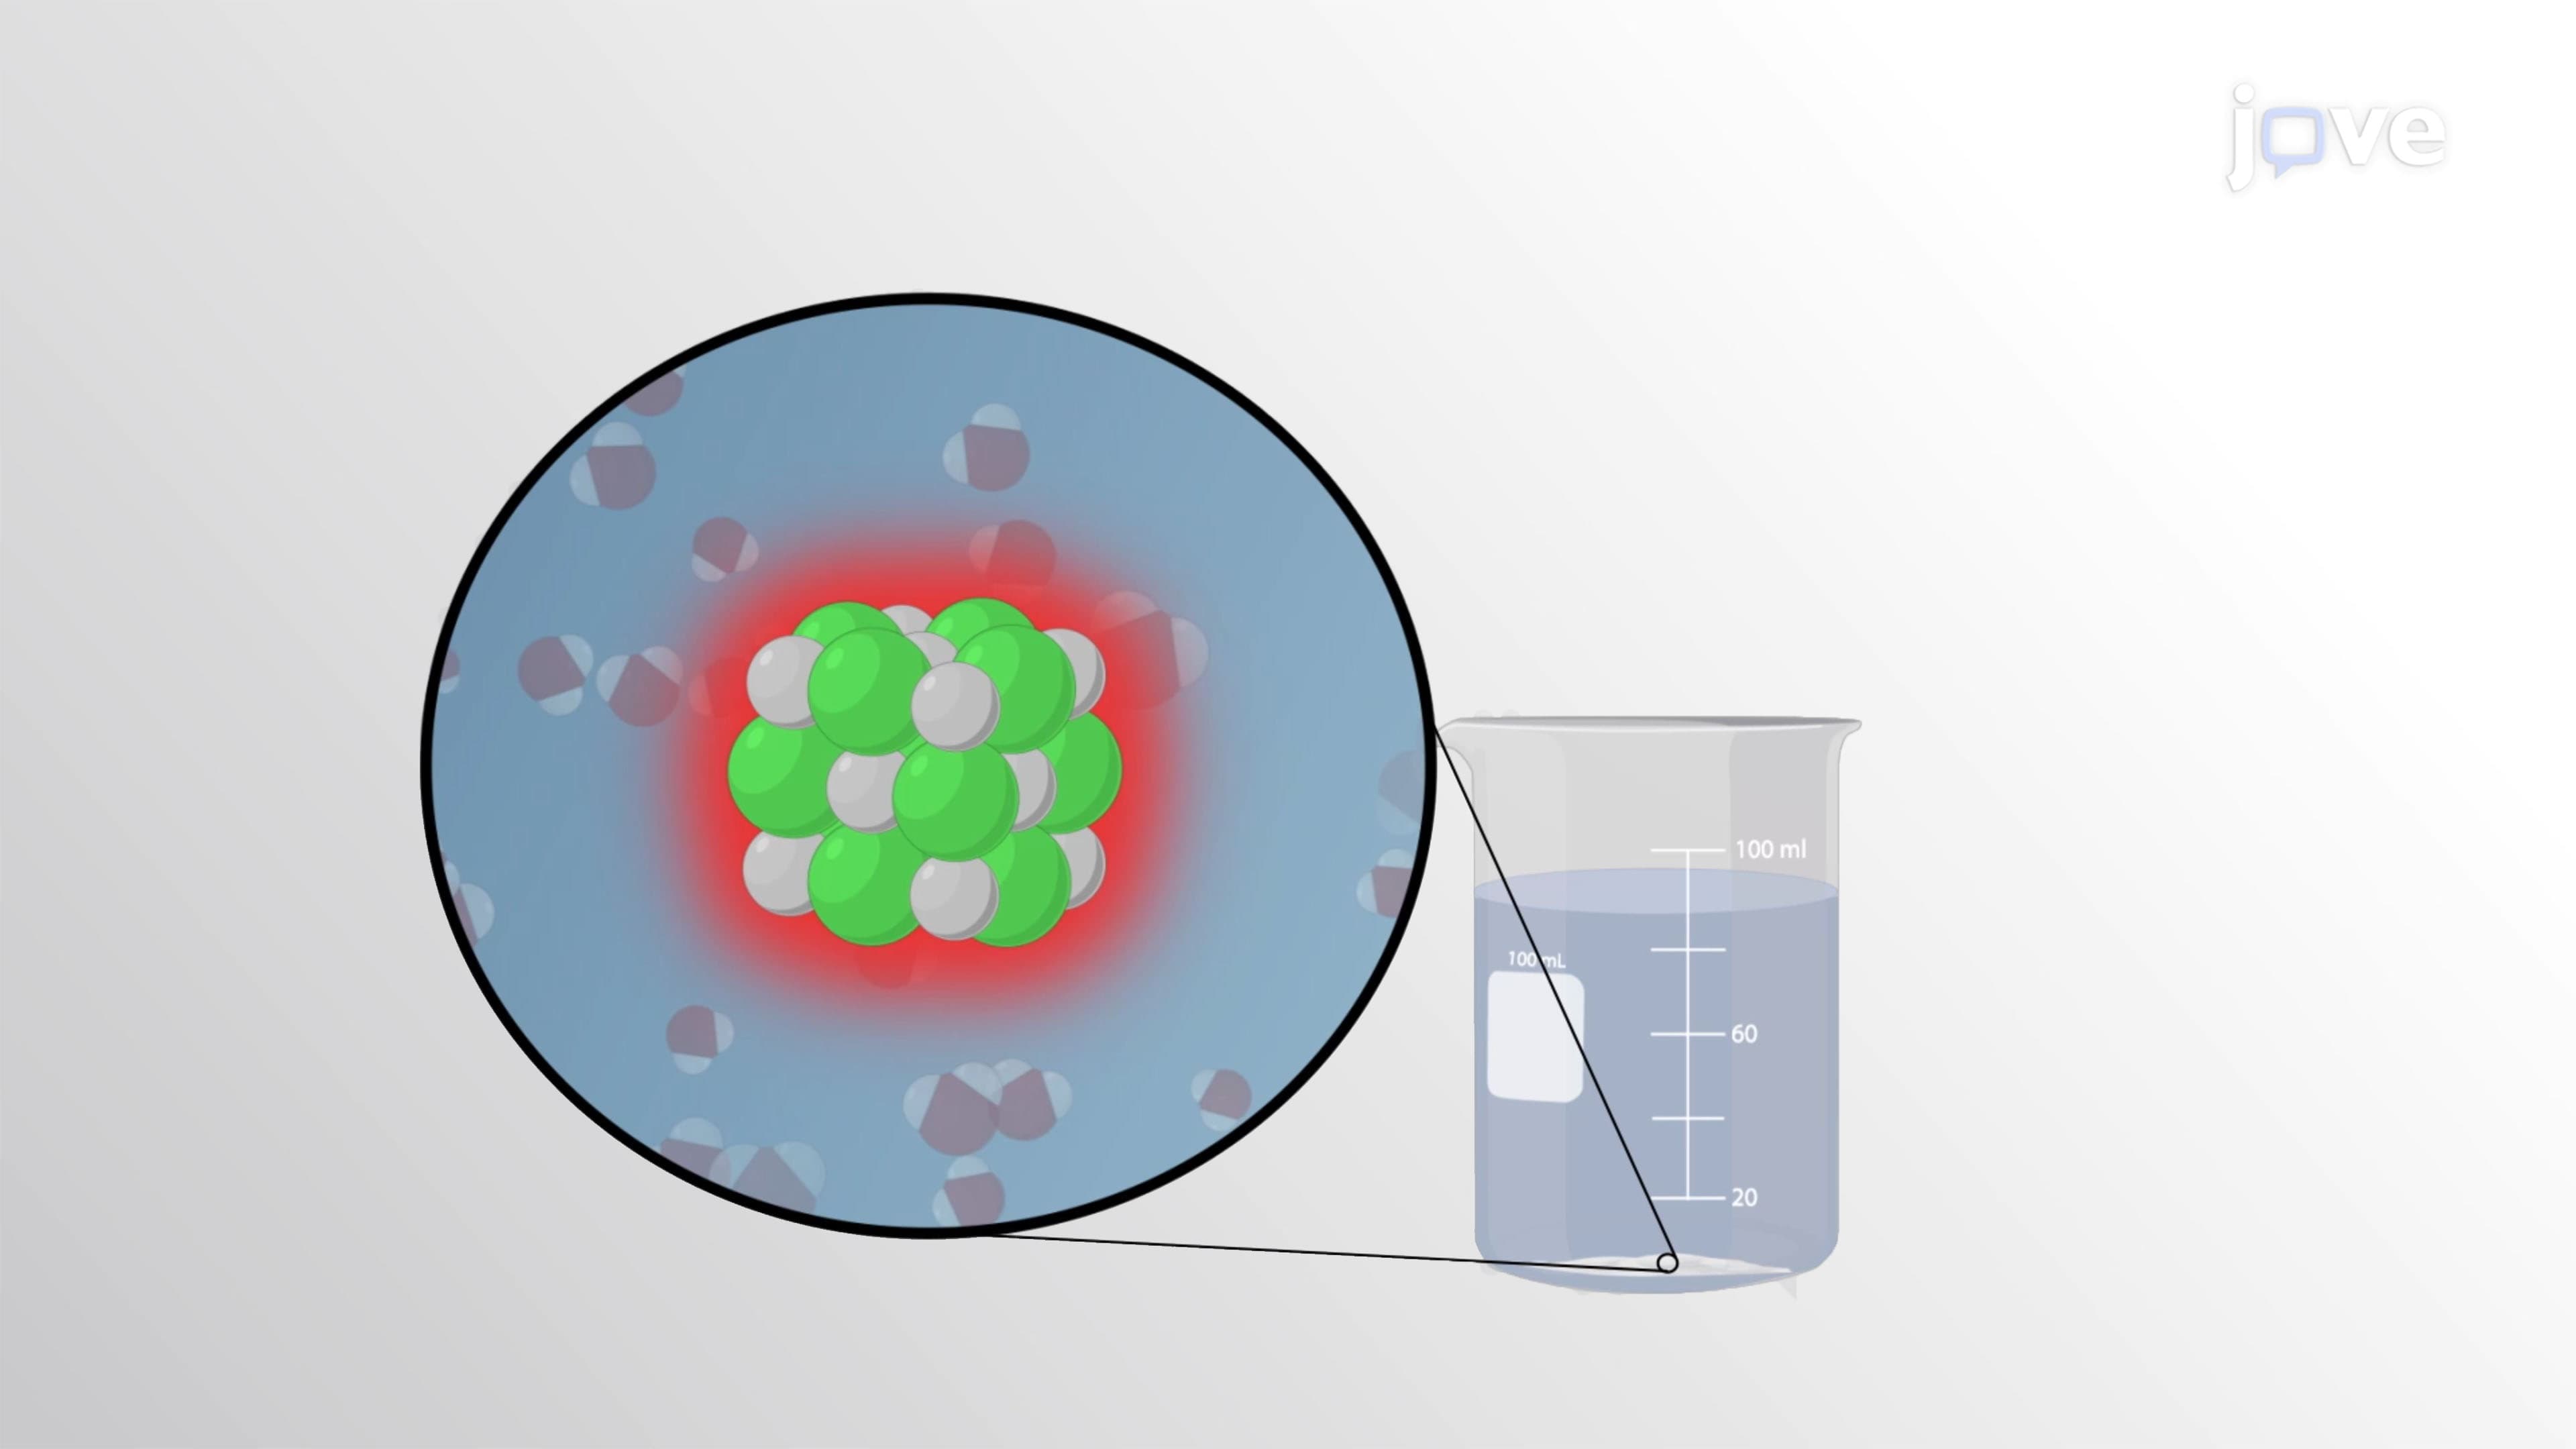 02:55
Solubility of Ionic Compounds
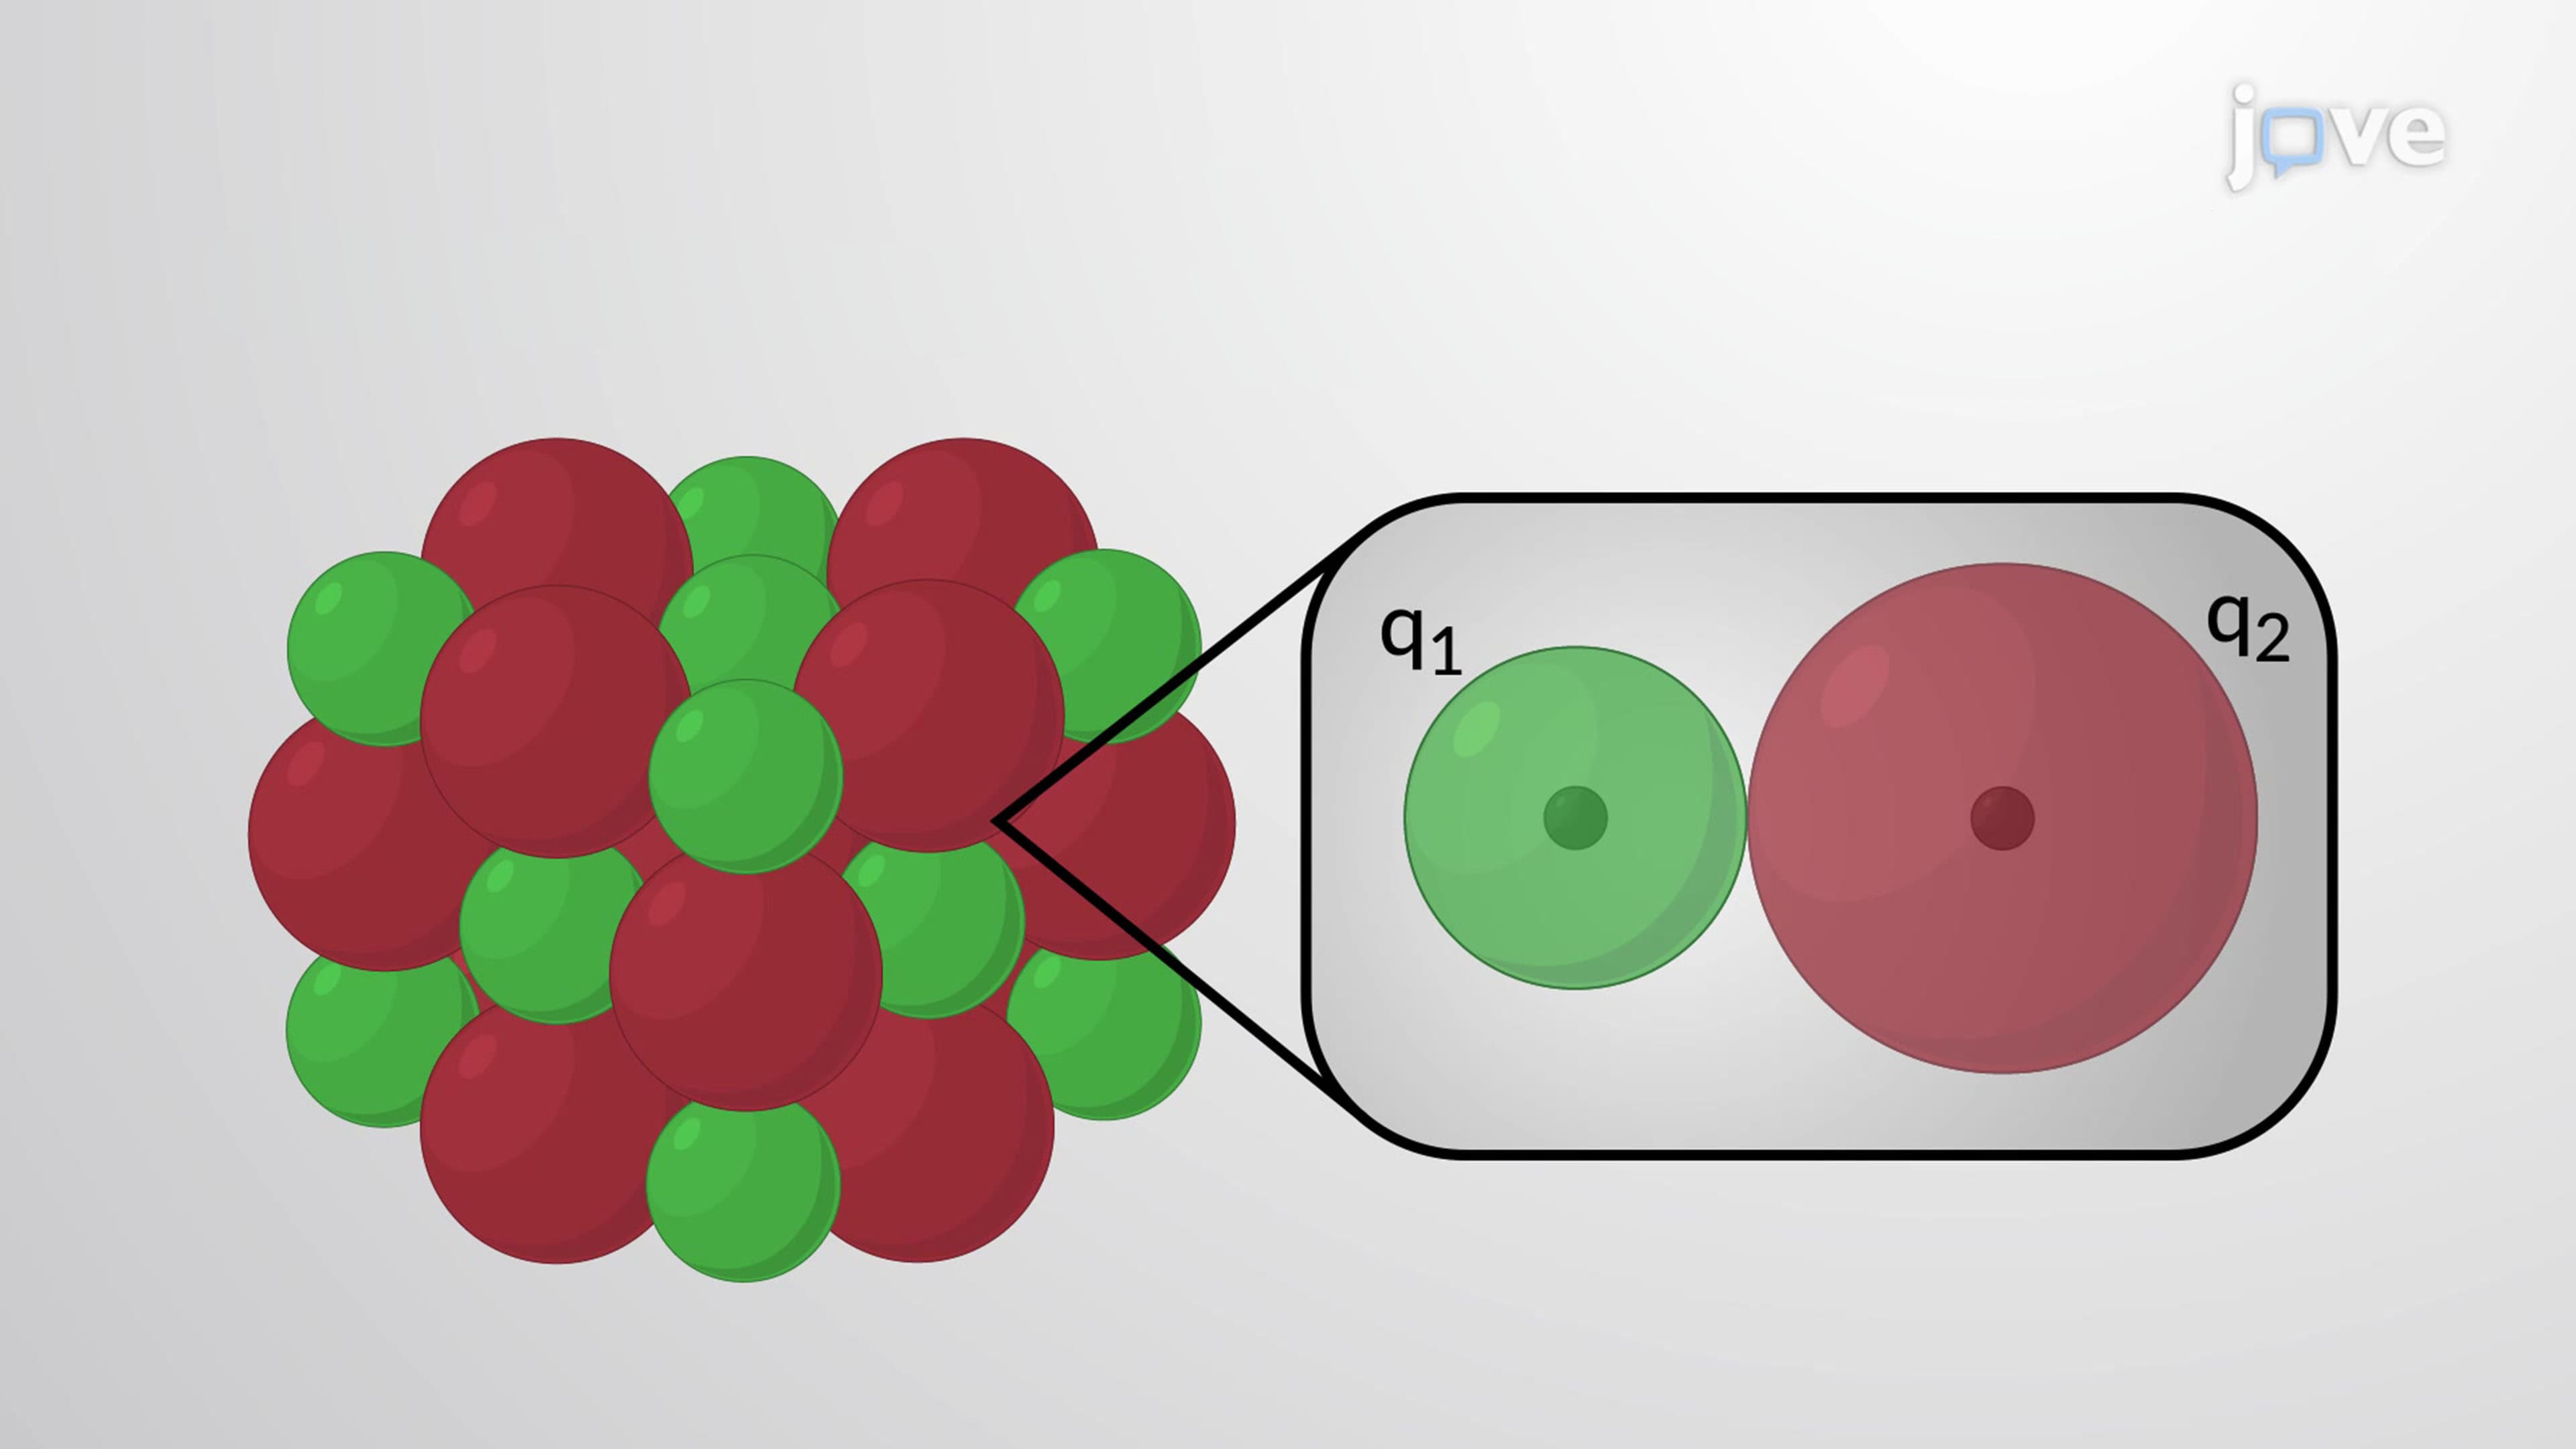 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 02:56
Intermolecular Forces and Physical Properties
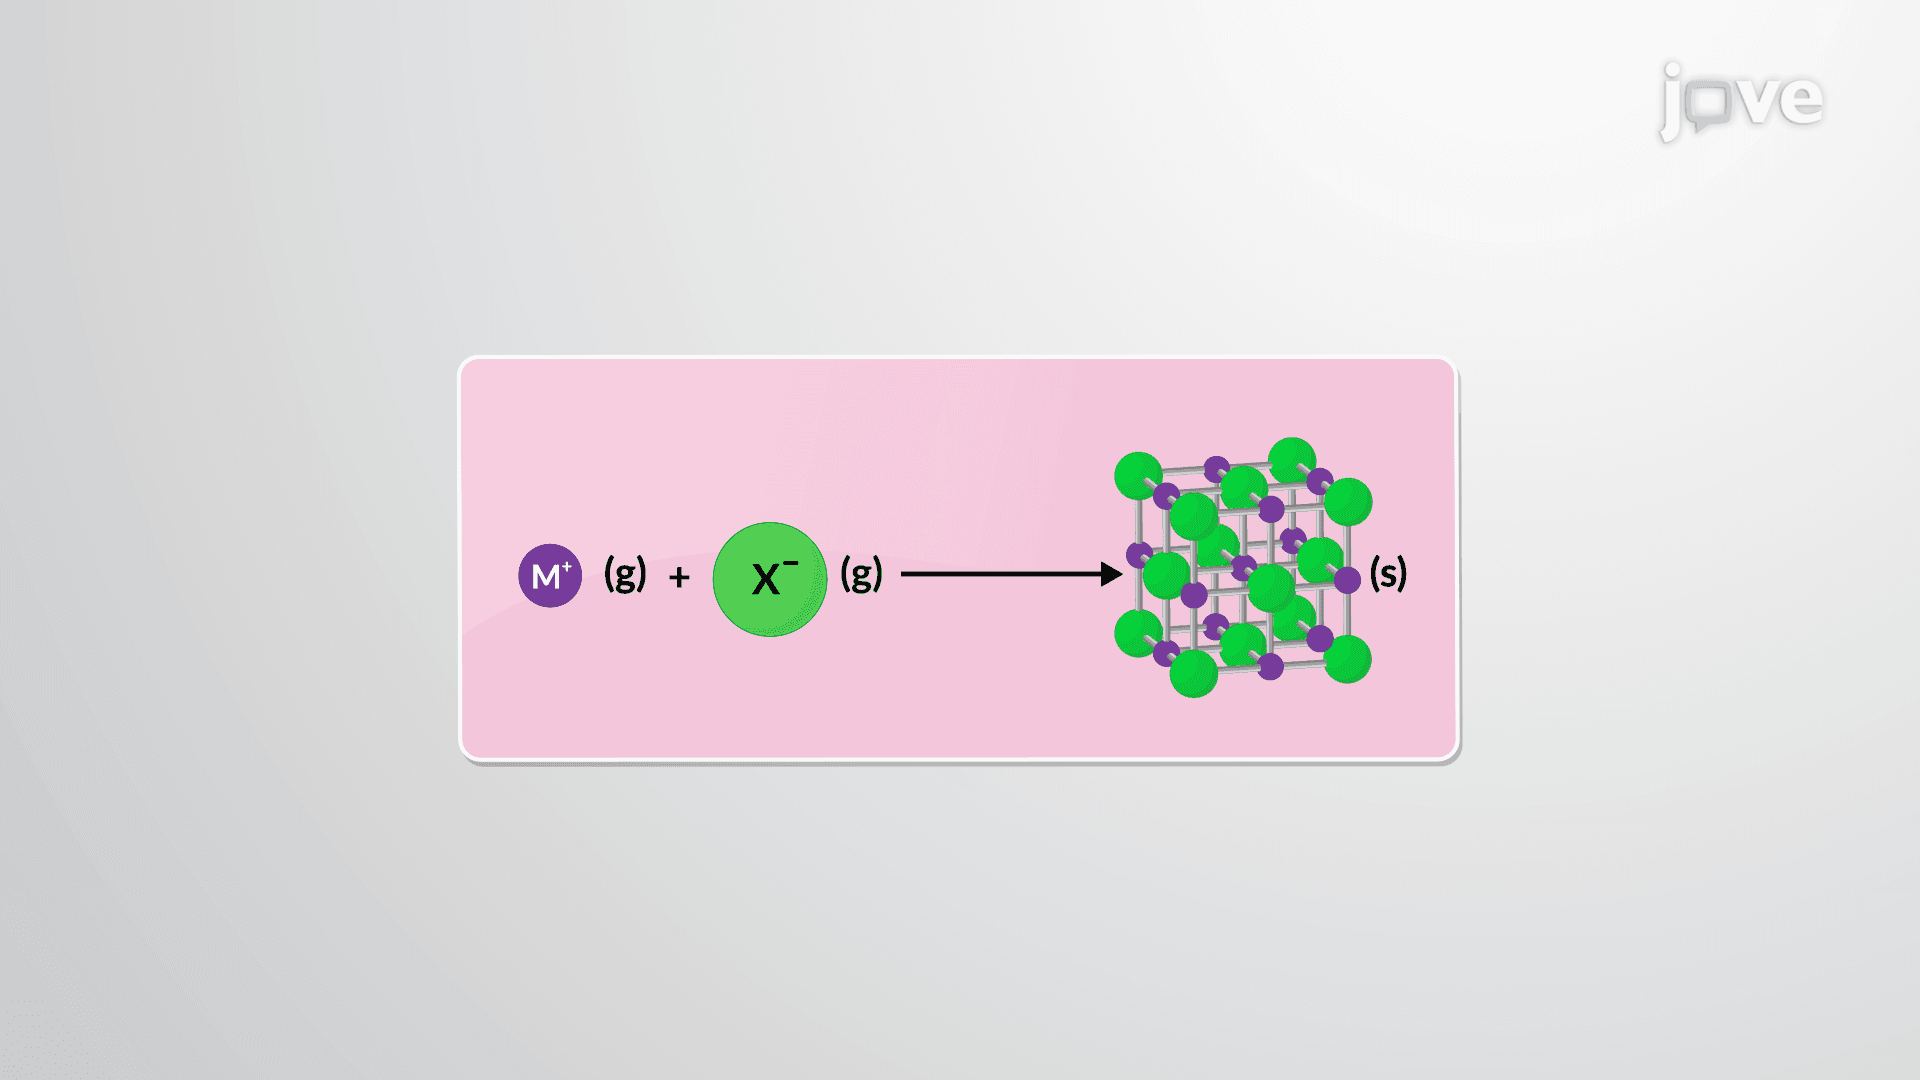 01:27
Lattice Energies of Ionic Crystals