Related Experiment Video
Updated: Jun 13, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Random Phase Approximation Correlation Energy Using Real-Space Density Functional Perturbation Theory
Boqin Zhang1, Shikhar Shah2, John E Pask3
1College of Engineering, Georgia Institute of Technology, Atlanta, Georgia 30332, United States.

Abstract:
We present a real-space method for computing the random phase approximation (RPA) correlation energy within Kohn-Sham density functional theory, leveraging the low-rank nature of the frequency-dependent density response operator. In particular, we employ a cubic-scaling formalism based on density functional perturbation theory that circumvents the calculation of the response function matrix, instead relying on the ability to compute its product with a vector through the solution of the associated Sternheimer linear systems. We develop a large-scale parallel implementation of this formalism using the subspace iteration method in conjunction with the spectral quadrature method while employing the Kronecker product-based method for the application of the Coulomb operator and the conjugate orthogonal conjugate gradient method for the solution of the linear systems. We demonstrate convergence with respect to key parameters and verify the method's accuracy by comparing with plane-wave results. We show that the framework achieves good strong scaling to many thousands of processors, reducing the time to solution for a lithium hydride system with 128 electrons to around 150 s on 4608 processors.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:45
The Quantum-Mechanical Model of an Atom
 01:21
Energy Associated With a Charge Distribution
 01:10
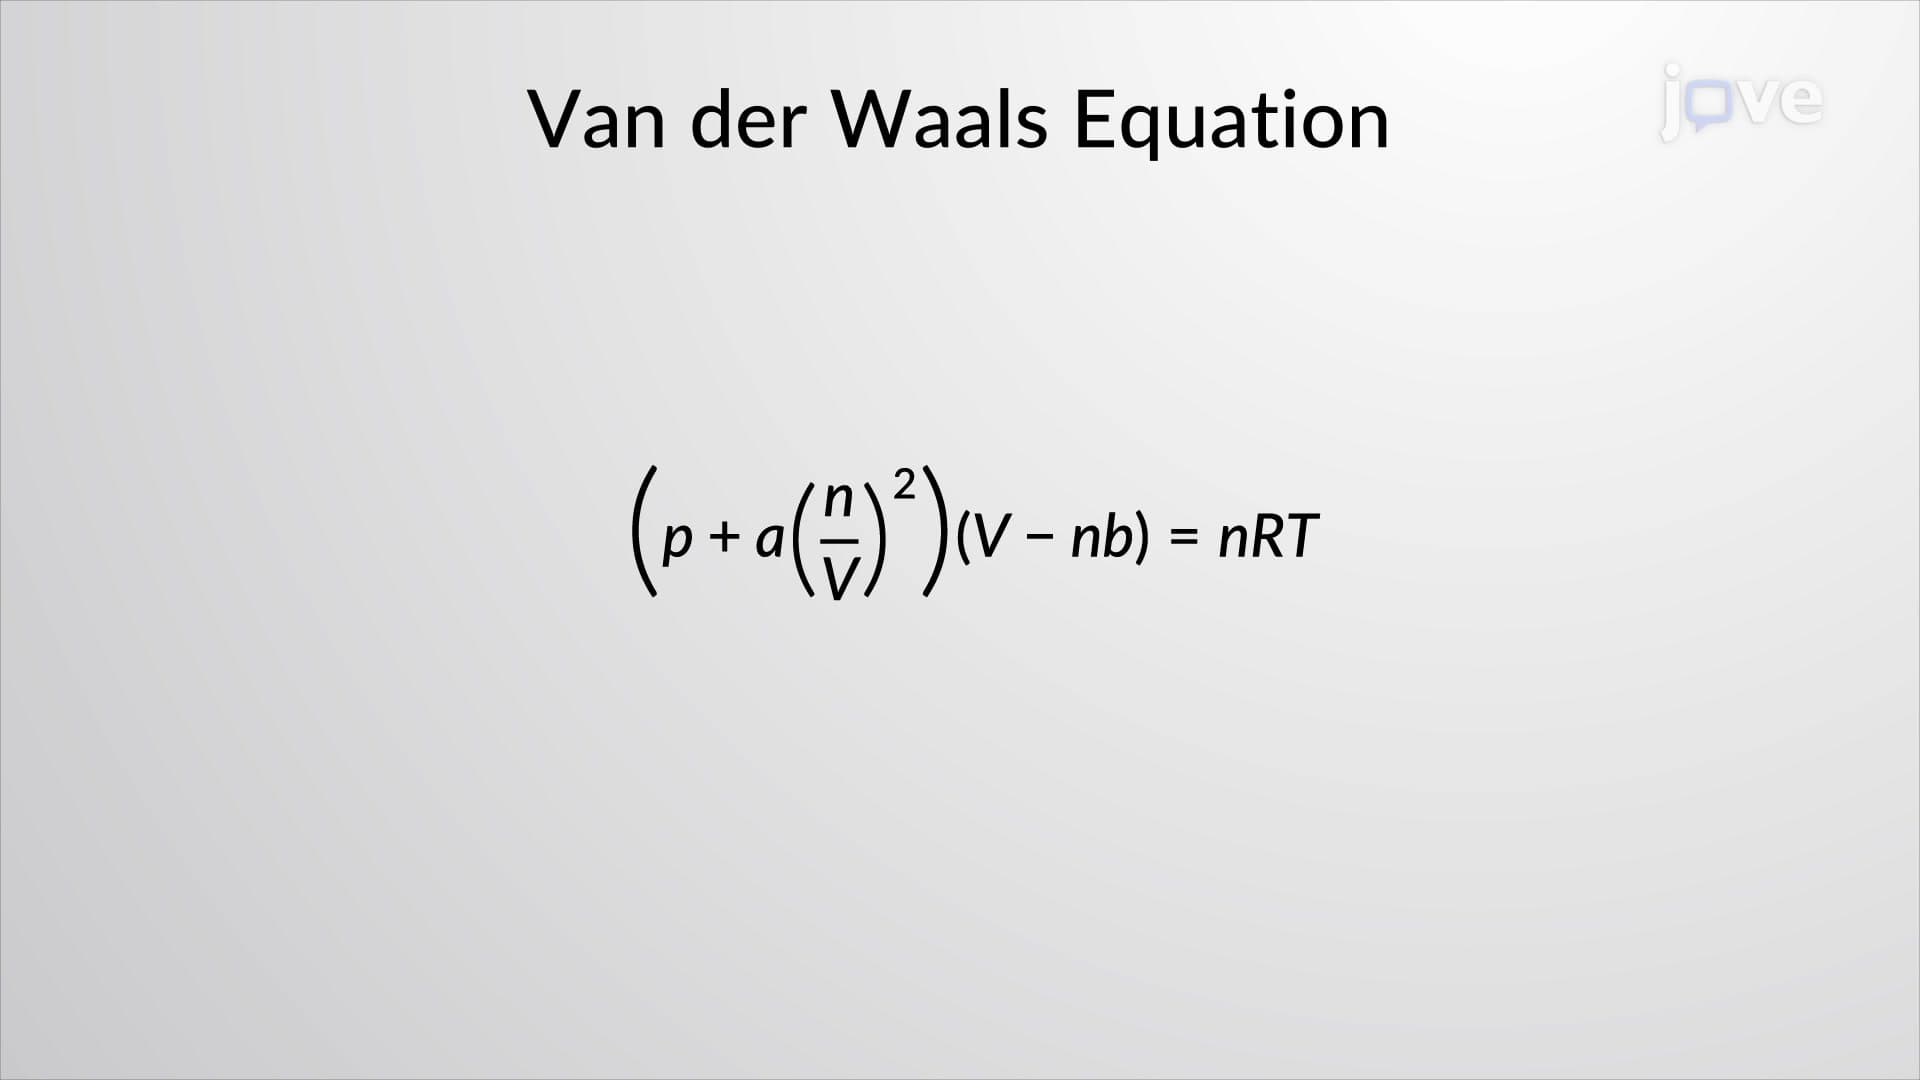
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 03:21
The Energies of Atomic Orbitals