Related Experiment Video
Updated: Sep 19, 2025
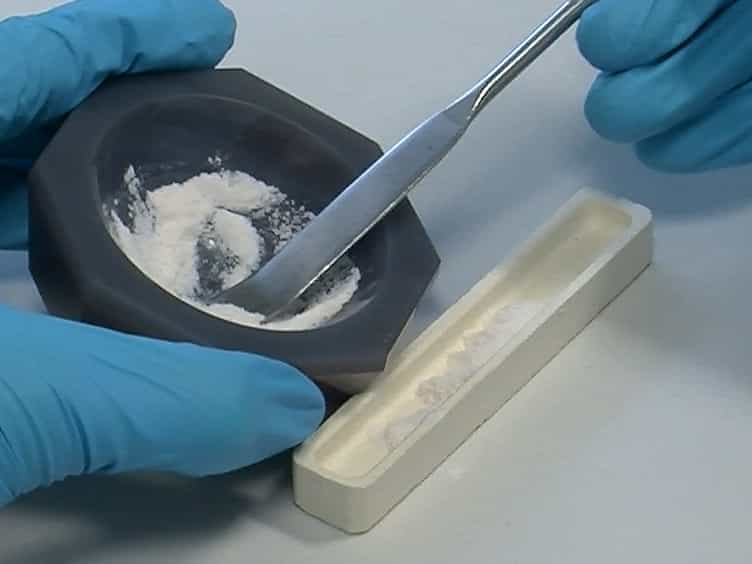

Rapid Mix Preparation of Bioinspired Nanoscale Hydroxyapatite for Biomedical Applications
Published on: February 23, 2017
Influence of Ion Substitution on the Properties of Apatite-Based Materials: Computational Predictions Using Density
Henrique S Marques1, Albert F B Bittencourt2,3, Juarez L F Da Silva3
1Institute of Mathematics and Computer Sciences, University of São Paulo, 13566-590 São Carlos, SP, Brazil.

Abstract:
Apatite-based materials have attracted recognition as promising candidates for catalytic applications because of their tunable properties that can be achieved through ionic substitutions and their compatibility with sustainability goals for environmentally friendly catalysts. However, a thorough understanding of their physicochemical properties at the atomic level remains insufficient. In this study, calculations based on density functional theory combined with Spearman's correlation are used to investigate the effects of cationic and anionic substitutions on the structural, energetic, and electronic properties of materials similar to apatite with Ca/P ratios ranging from 0.50 to 2.00. Our results reveal that substitutions with d-block elements, such as Zn and Cd, reduce the energy gap at the Γ-point and decrease the ionic character of the materials, leading to reduced stability. Additionally, d-p orbital hybridization within the PO4 3-, AsO4 3-, and VO4 3- groups significantly influences structural stability. Using Spearman's correlation analysis, we identified significant trends, specifically indicating a strong correlation between net atomic charges, energy gaps, and cohesive energy. These results offer critical insights into how ionic substitutions influence the tunable characteristics of materials resembling apatite.
More Related Videos
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:54
Trends in Lattice Energy: Ion Size and Charge
 01:19
Ionic Strength: Effects on Chemical Equilibria
In this solution, the primary...
 01:31
Essential Minerals for Bone Health
Calcium and Phosphorus
Calcium is a critical component of bones, especially in the form of calcium phosphate and calcium carbonate. Since the body cannot make calcium, it must be obtained from the diet. However, calcium cannot be absorbed from the small intestine without...