Related Experiment Video
Updated: Sep 14, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Deciphering the Role of Hydrogen Bonding and Water Dynamics in Hole Transfer on Anatase TiO2(101): An ab Initio
Xiaodan Yan1, Xiao Han1, Jinlu He1
1Inner Mongolia Key Laboratory of Rare Earth Catalysis, College of Chemistry and Chemical Engineering, Inner Mongolia University, Hohhot 010021, People's Republic of China.

Abstract:
Using time-dependent density functional theory (TD-DFT) and nonadiabatic molecular dynamics (NAMD) simulations, we elucidate how hydrogen bonding and water dynamics regulate hole transfer at the anatase TiO2(101)/water interface. Compared to low-density water (LW), moderate-density water (MW) enhances hydrogen bonding between surface- and nonsurface-adsorbed water, restricting interfacial water mobility. This suppresses nonadiabatic coupling and slows the hole transfer. Conversely, higher-density water (HW) near the vacuum destabilizes hydrogen bonding networks, freeing interfacial water and amplifying thermal motion. Enhanced disorder strengthens nonadiabatic coupling, accelerating the hole transfer. Temperature-dependent simulations show that thermal energy overcomes hydrogen bonding constraints: elevated temperatures intensify water dynamics and nonadiabatic coupling, accelerating hole transfer. These results establish hydrogen bonding and thermal fluctuations as key regulators of charge dynamics at the semiconductor/liquid interfaces.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:28
Probing the Structure and Dynamics of Interfacial Water with Scanning Tunneling Microscopy and Spectroscopy
Published on: May 27, 2018
Related Concept Videos
 01:04
Hydrogen Bonds
 01:01
Introduction to Chemical Bonds
The electrons of the outermost energy level determine the energetic stability of the atom and its tendency to form chemical bonds with other atoms. The innermost electron shell has a maximum capacity of two electrons, but the next two electron shells can each have a maximum of eight electrons. This is known as the octet rule, which states that, with the exception of the innermost shell, atoms are most stable energetically when they have eight electrons in their valence shell, the...
 03:51
Molecular Orbital Theory II
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:20
Aldehydes and Ketones with Water: Hydrate Formation
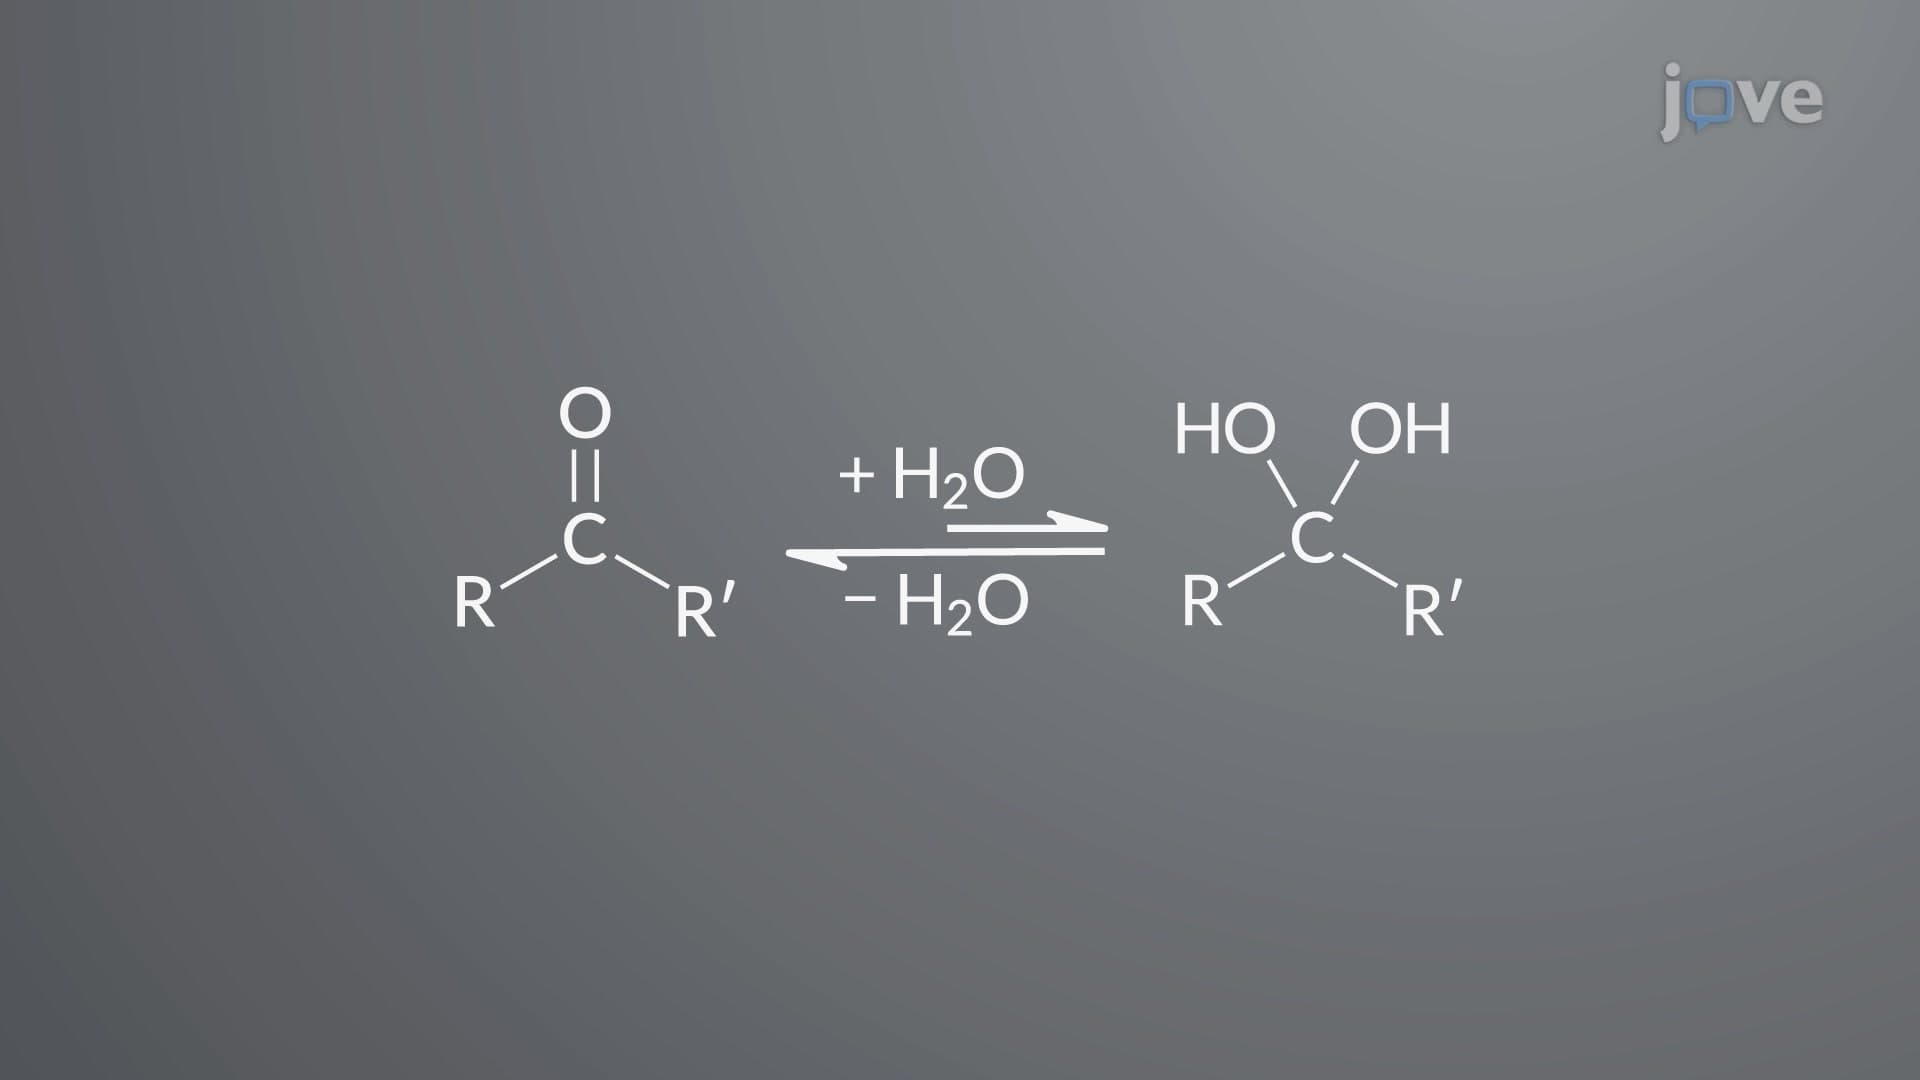
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...