MEGDTA: multi-modal drug-target affinity prediction based on protein three-dimensional structure and ensemble graph neural network
Related Experiment Video
Updated: Sep 11, 2025

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Zhanwei Hou1, Yijun Li1, Haixia Zhai1
1School of Software, Henan Polytechnic University, Jiaozuo, 454000, China.

Background:
Drug development is a time-consuming and costly endeavor, and utilizing computer-aided methods to predict drug-target affinity (DTA) can significantly accelerate this process. The key to accurate DTA prediction lies in selecting appropriate computational models to effectively extract features from drug molecular structures and target protein structures. Existing methods usually ignore the features of the protein three-dimensional structure.
Results:
This paper proposes a multi-modal drug-target affinity prediction model based on protein three-dimensional structure and ensemble graph neural networks (MEGDTA). This model aims to capture diverse features from drug and target structure using neural network architectures, especially for protein three-dimensional structure. First, one drug is represented into two forms by a molecular graph and a Morgan Fingerprint, and their features are extracted by constructing a graph feature space and a fully connected network, respectively. Second, for a protein, a residue graph is constructed based on its three-dimensional structure. And, the protein sequence and residue graph are processed using a long short-term memory (LSTM) network and multiple parallel graph neural networks (GNNs) with variant modules to learn the latent features of proteins. Third, a cross-attention mechanism fuses the extracted features of the drug and protein, followed by fully connected layers to finalize the prediction. The source code of MEGDTA is available from https://github.com/liyijuncode/MEGDTA .
Conclusions:
MEGDTA is validated on three publicly available benchmark datasets, Davis, KIBA and Metz. A comparative study is conducted with other existing models. The results show that MEGDTA performs strongly in terms of mean squared error (MSE) and concordance index (CI), and r2m, which demonstrate the effectiveness of MEGDTA.

Related Concept Videos
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:15
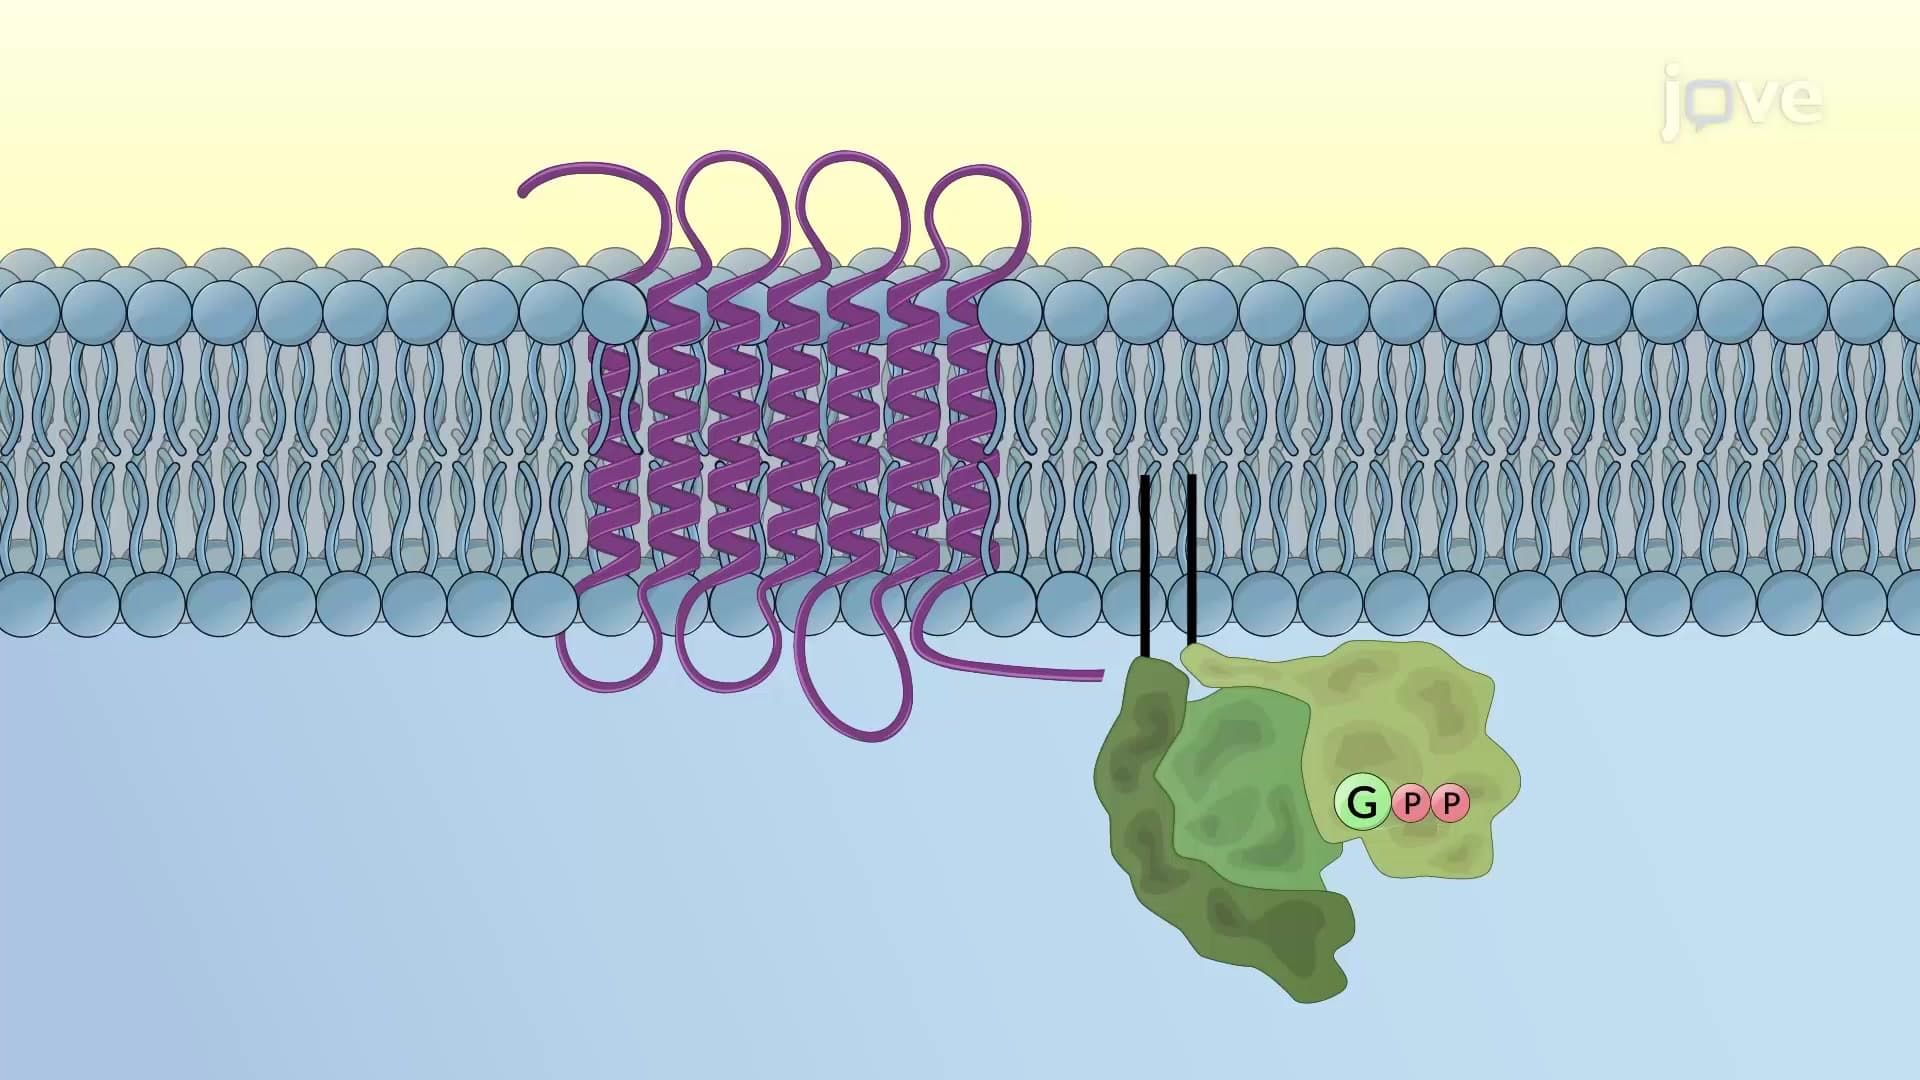
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:22
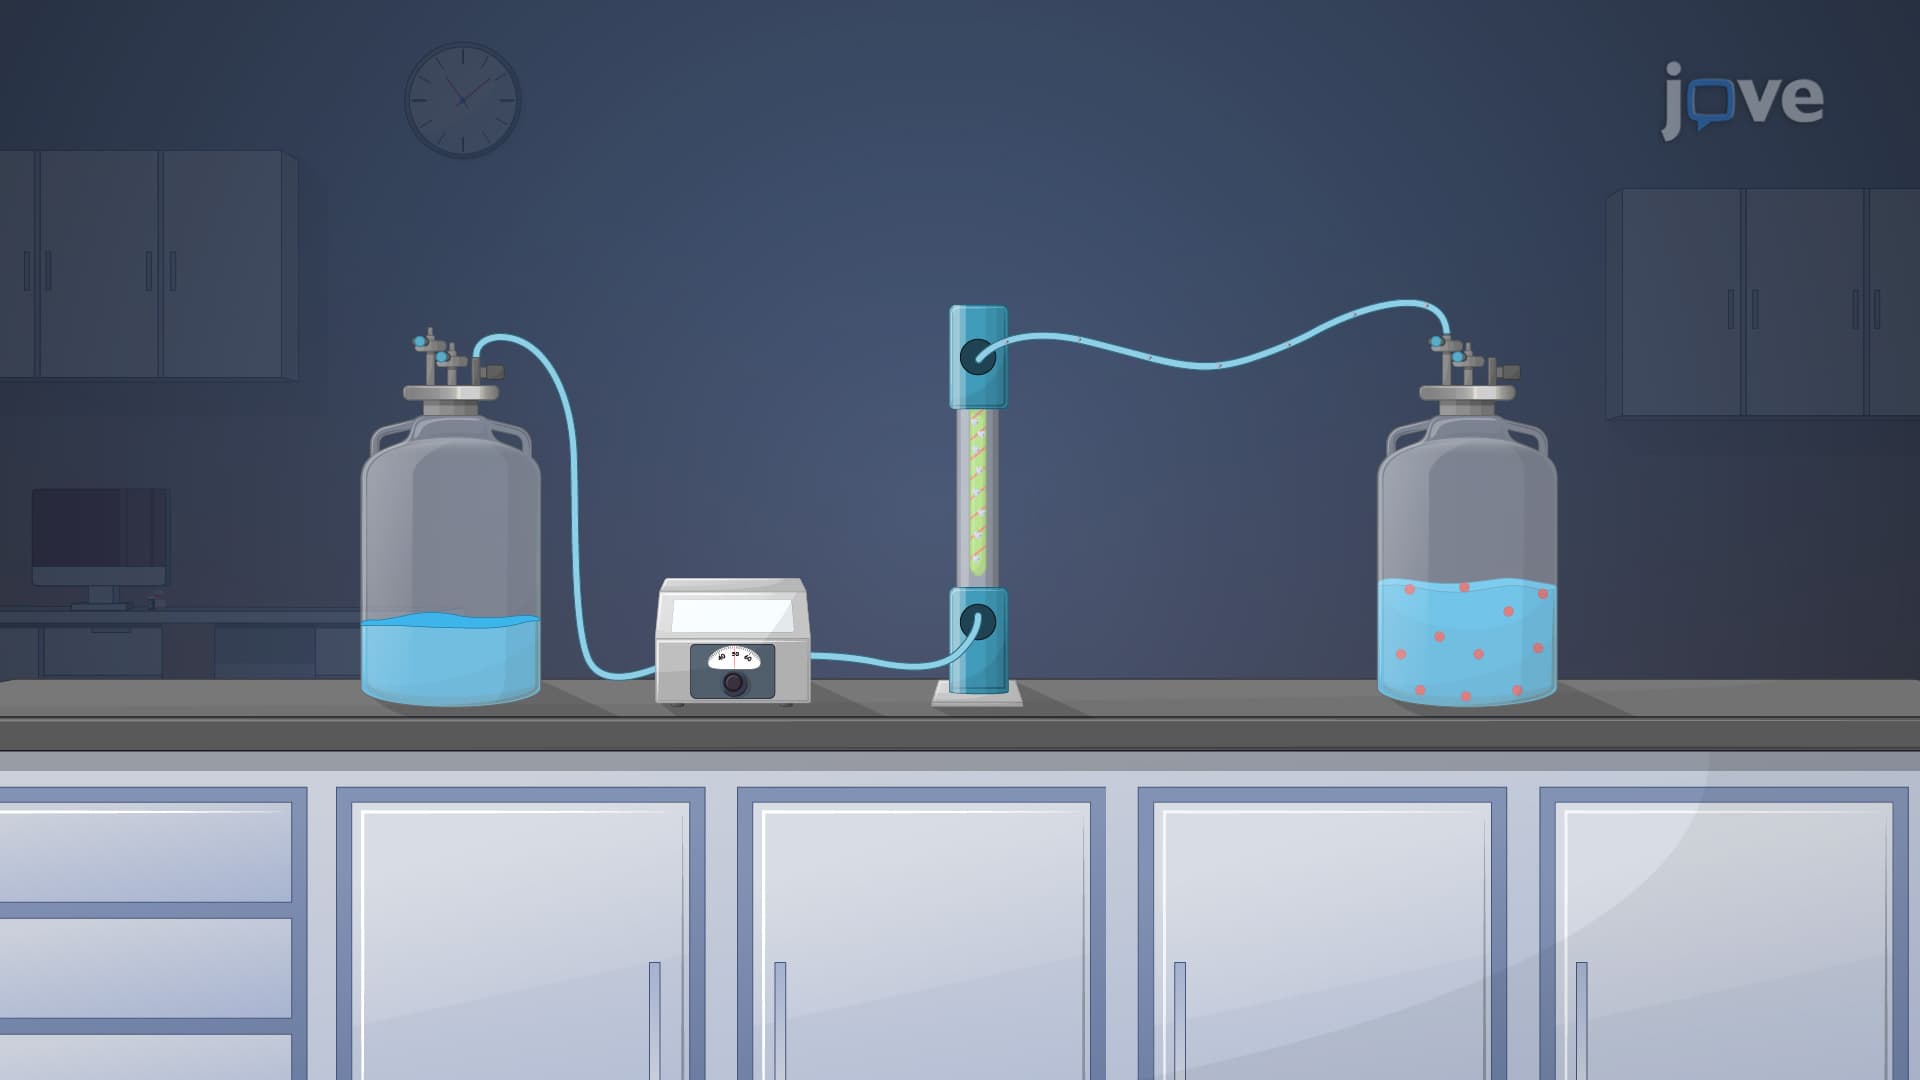
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...