Related Experiment Video
Updated: Sep 11, 2025


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Multiscale Motif-Aware Relation Graph Structure for Drug-Target Binding Affinity Prediction

Abstract:
Exploring drug-target binding affinity (DTA) is essential for drug discovery. Numerous works rely on the one-dimensional SMILES representation of drugs for predicting drug-target affinity, but ignore the crucial structural information of drug molecules. Considering structural information is an important factor in determining the affinity properties of drugs, we propose using the multiscale motif-aware relation graph (MMRG) rather than the SMILES representation to build drug descriptors. MMRG explicitly provides crucial motif-level structural and topological information of drugs, thereby ameliorating the predictive power of models. With this idea, we propose a novel MMRG construction approach for drugs, including a multiscale motif-aware learning for extracting motif-level structural information from motifs with different sizes and a relation graph construction approach for extracting topological information from chemical bonds. We implement a graph convolutional network to learn from MMRGs, and the learned latent features are used to predict drug-target affinity. The experiment results on the Davis and KIBA dataset report that our model can significantly outperform existing methods in drug-target affinity prediction, with an average improvement of 15.61% and 8.50% respectively. We further explain the principle of MMRG to improve the drug-target affinity prediction accuracy by comparing its generative function with the state-of-the-art method.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 02:04
Protein-protein Interfaces
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
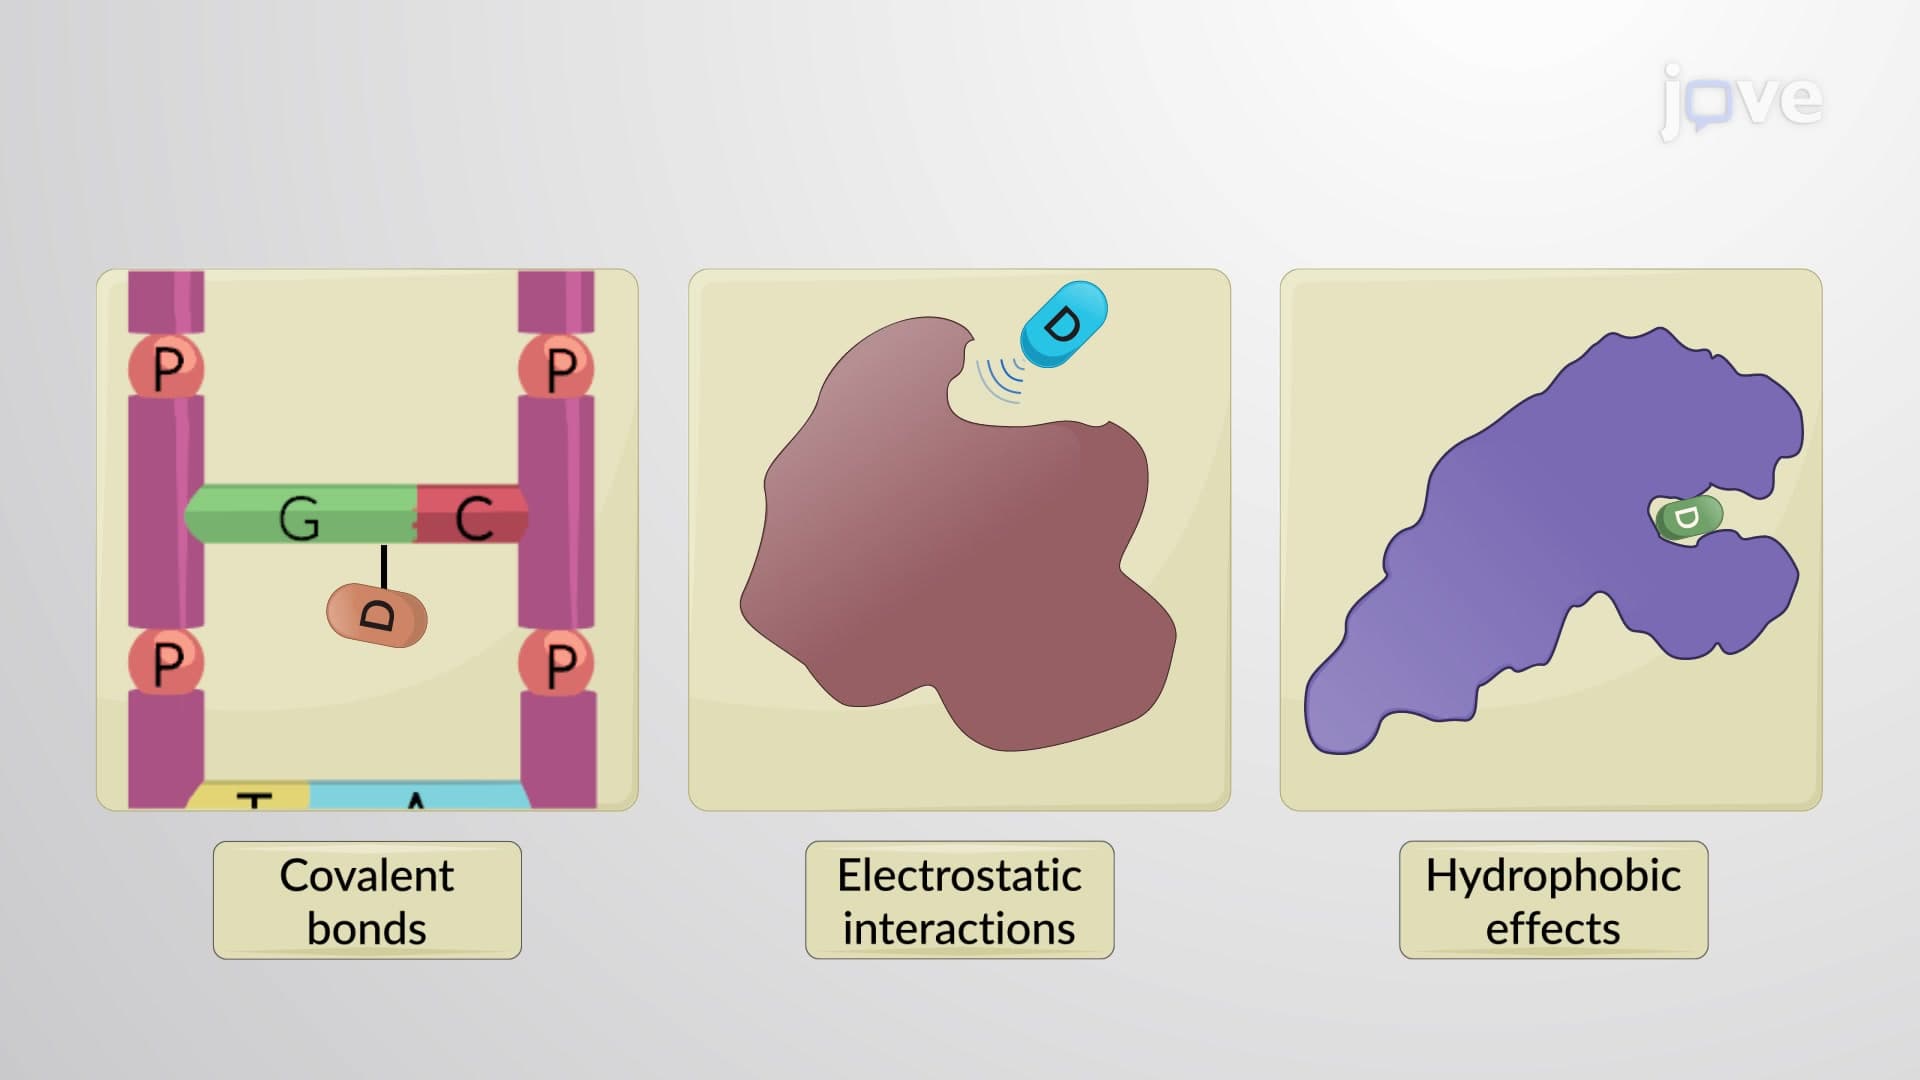 01:25
Drug-Receptor Bonds
In...
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...