

Systematic Embedding of All-Atom Reactive Molecular Dynamics into a Coarse-Grained Environment
Kuntal Ghosh1, Da Teng2, Gregory A Voth1
1Department of Chemistry, Chicago Center for Theoretical Chemistry, James Franck Institute, and Institute for Biophysical Dynamics, The University of Chicago, Chicago, Illinois 60637, United States.

Abstract:
Quantum mechanics/molecular mechanics (QM/MM) simulations are widely used for modeling chemical reactivity in complex environments. In the QM/MM method, the reactive center under study is typically treated using explicit quantum chemical calculations, while the nonreactive environment is treated using classical molecular mechanics. However, for large systems, even with the use of classical force fields for the MM part, QM/MM simulations are very computationally costly, particularly for systems with complex MM dynamics. In this work, we propose a much faster alternative method for studying reactivity denoted by multiscale reactive molecular dynamics/coarse-grained molecular mechanics (MS-RMD/CG-MM). MS-RMD has been shown to be a powerful method for modeling reactions involving proton transport and in principle other reactions. This all-atom reactive MD model, systematically parametrized from constrained DFT calculations, is embedded in a CG environment in this work, where the CG force fields are derived using the multiscale coarse-graining (MS-CG) method. We apply this scheme to organic SN2 reactions in a CG polar solvent (acetone) as examples.

Related Concept Videos
 02:00
Molecular Models
 01:17
Coupled Reactions
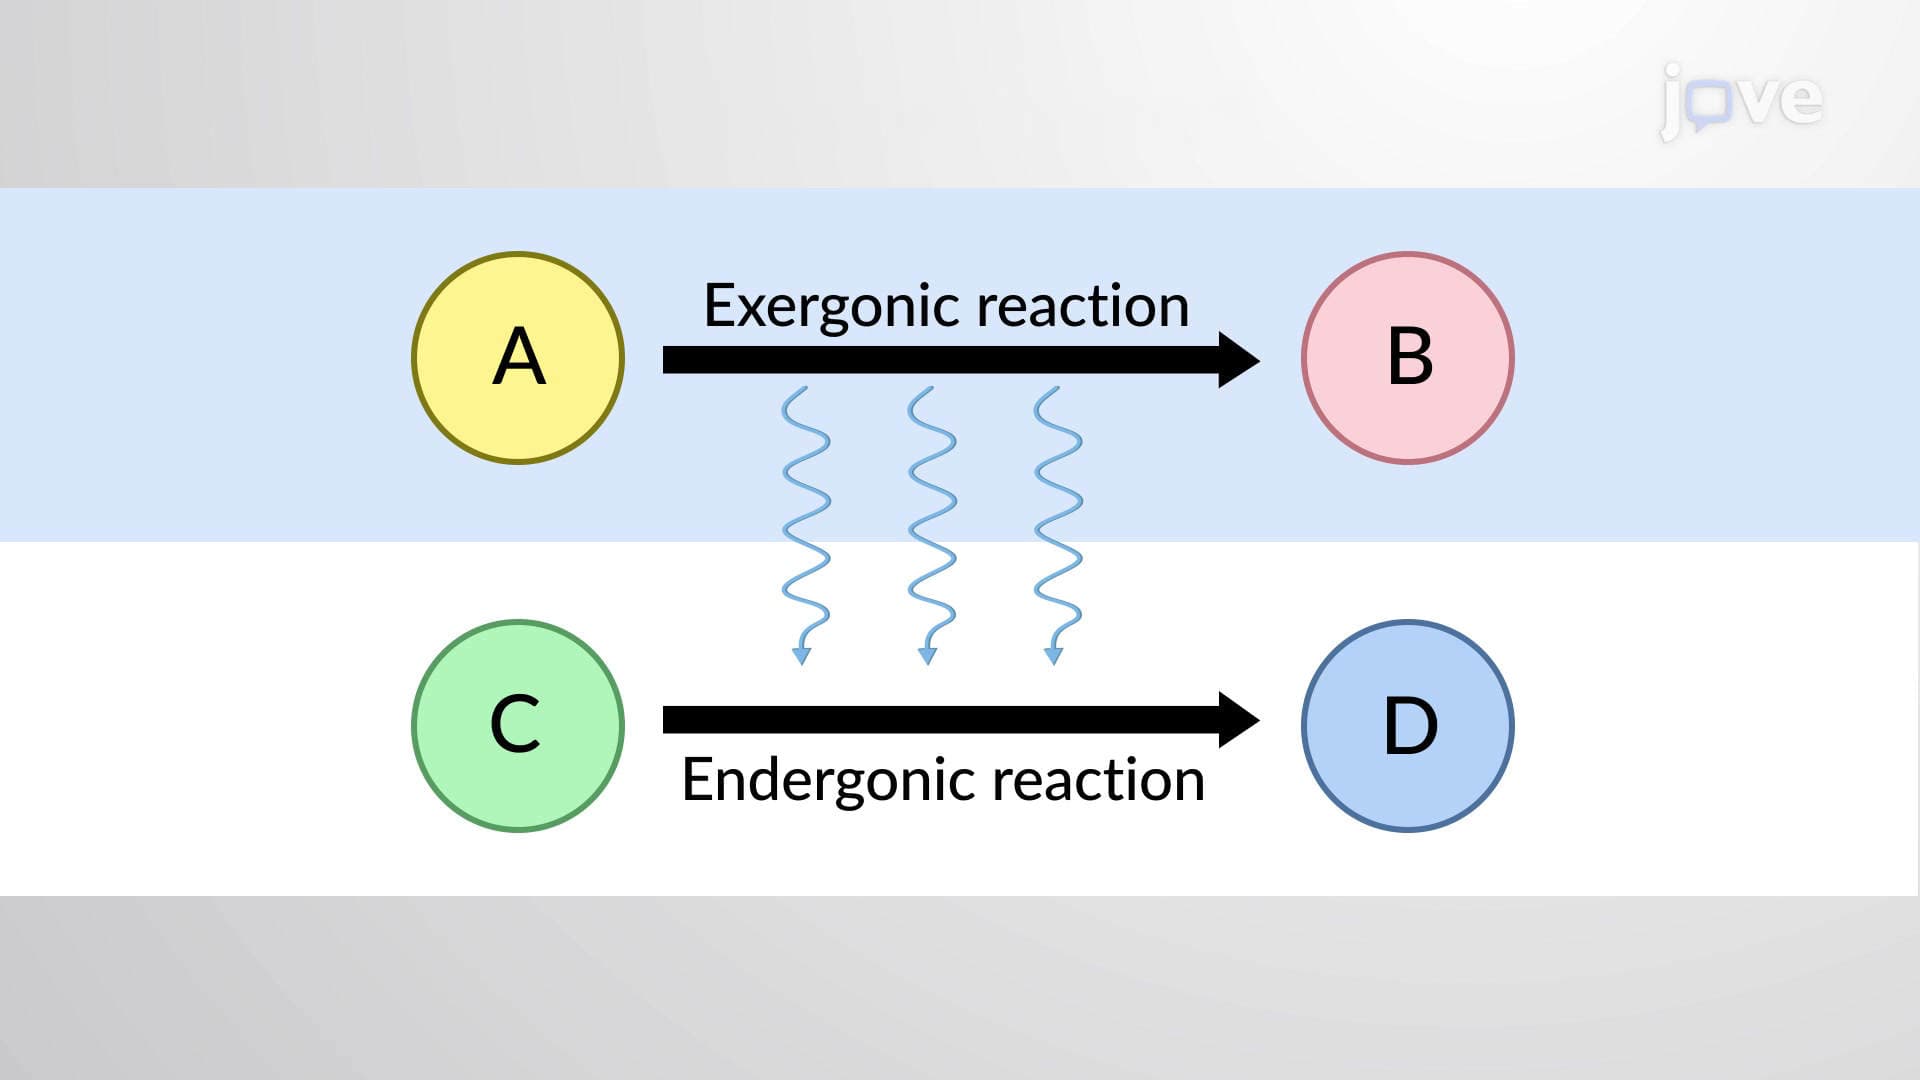
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions....
 01:23
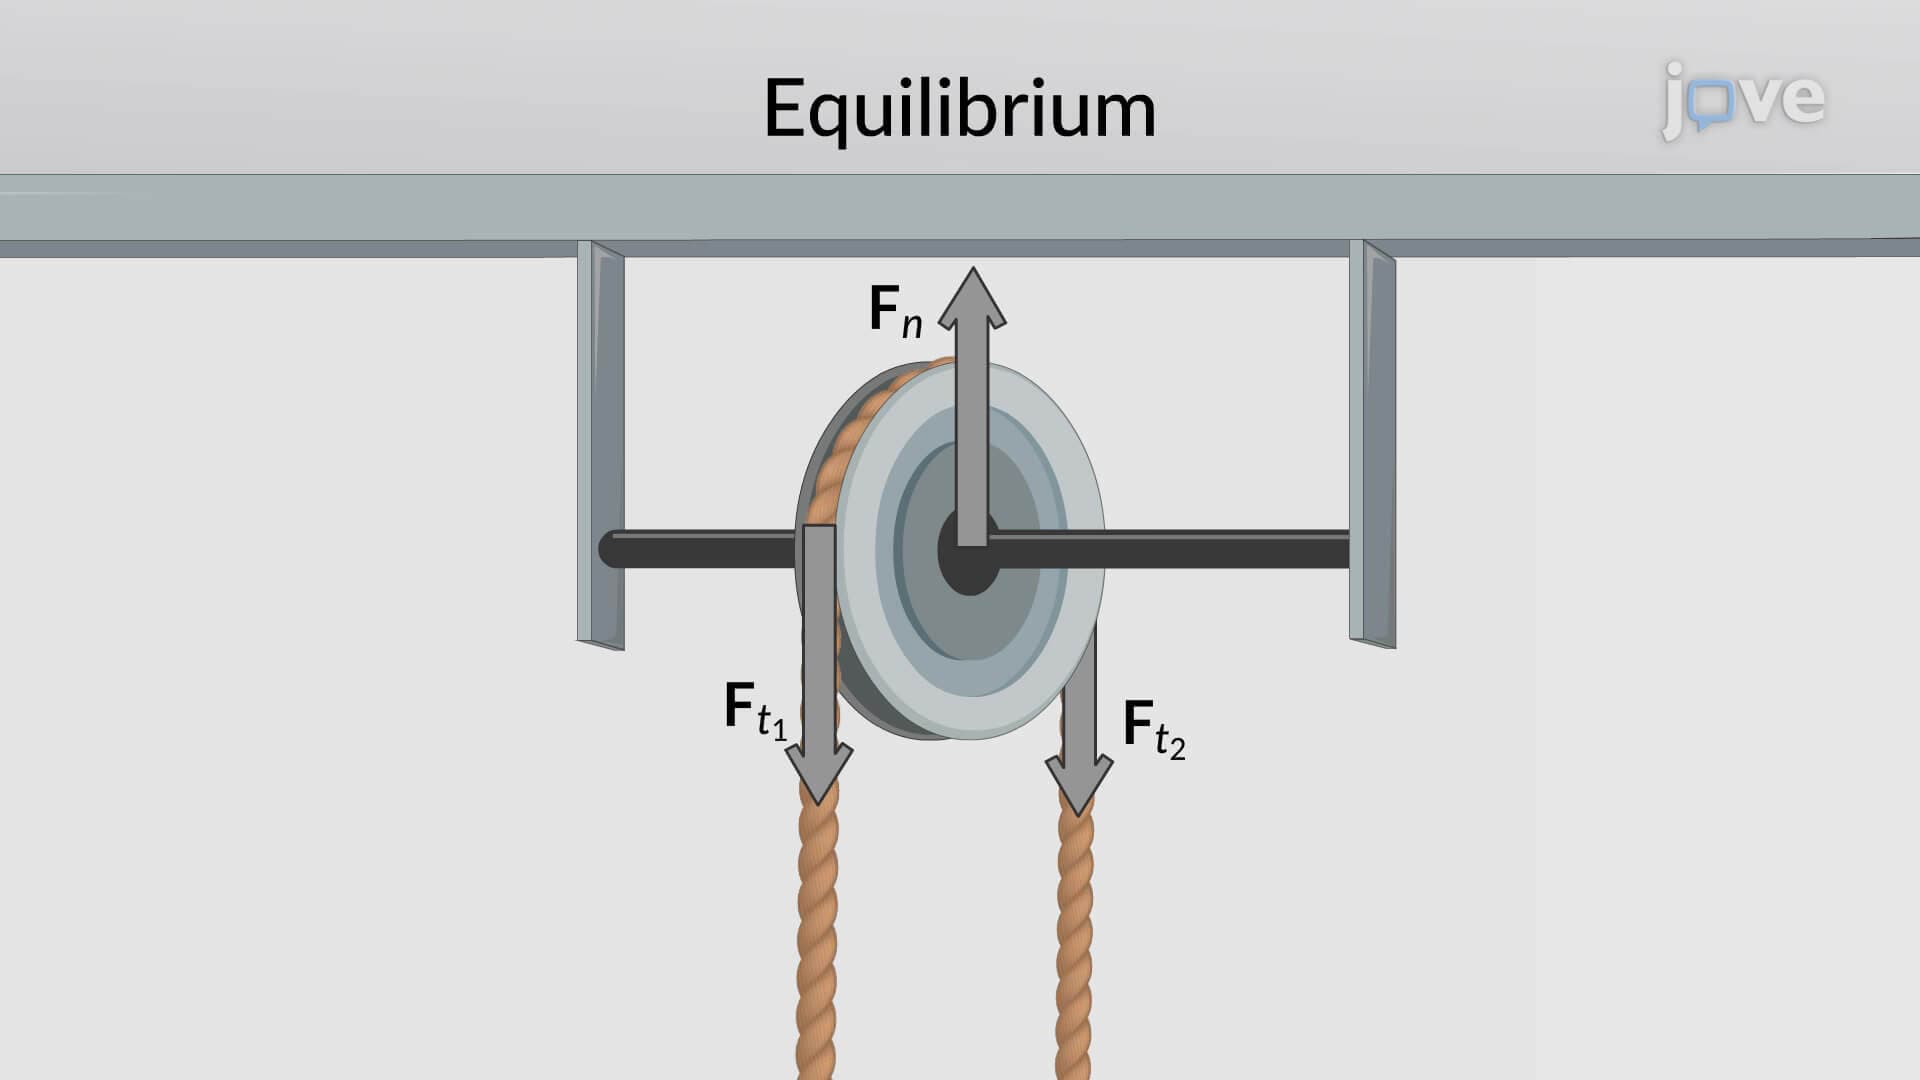
Equilibrium Conditions for a Particle
To understand the concept of equilibrium, let us first consider the forces acting on an object. When different forces act on an object, they can...