Related Experiment Video
Updated: Jan 15, 2026
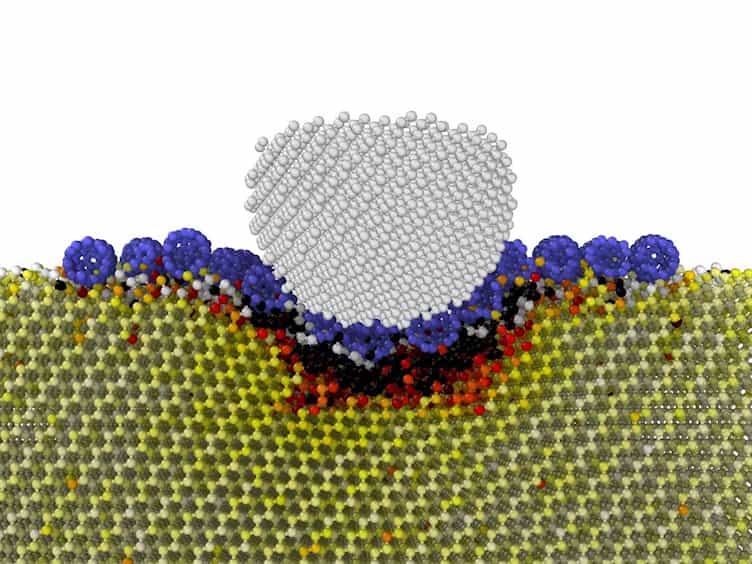

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
First-principles investigation of 6H-SiC dominated by strong covalent bonding: electronic structure, mechanical
Jing-Yi Xia1, Zheng-Tang Liu2, Qi-Jun Liu3
1Bond and Band Engineering Group, School of Physical Science and Technology, Southwest Jiaotong University, Chengdu, 610031, People's Republic of China. xia.jing.yi.xnjd@my.swjtu.edu.cn.

Context:
Silicon carbide (SiC), a third-generation semiconductor, is renowned for its wide bandgap, exceptional thermal conductivity and high breakdown field. The unique ABCACB stacking atomic arrangement of hexagonal SiC (6H-SiC) induces direction-dependent electronic, optical, and mechanical responses, which are crucial for emerging applications. Using first-principles calculations, we comprehensively characterize these properties of 6H-SiC. Our findings reveal dual bandgaps (2.82 eV indirect and 4.16 eV direct), with dispersive band edges that are conducive to carrier transport. Further calculations indicate that the carrier effective mass along the (001) direction is smaller than (100) direction, and one key factor causing this anisotropy is the directional changes in sp3 hybridized orbitals due to the unique atomic stacking. DOS, Mulliken population and charge density studies collectively reveal the covalent-dominated bonding nature, which underpins its dispersive band edges, hard texture and brittleness. The wide direct bandgap and unique electronic structure contribute to its broad spectral transparency and low optical loss. Moreover, A strong directional dependence is observed in both the optical and mechanical properties of 6H-SiC, where the (001) direction demonstrates higher compressive stiffness and lower optical absorption and loss.
Methods:
All calculations were conducted using density functional theory (DFT) as implemented in the CASTEP code, with norm-conserving pseudopotentials employed. For geometry optimization, we utilized the generalized gradient approximation with the Perdew-Burke-Ernzerhof (GGA-PBE) functional, whereas the electronic structure and optical characteristics were determined using the Heyd-Scuseria-Ernzerhof (HSE06) hybrid functional.
More Related Videos

Related Concept Videos
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 02:40
MO Theory and Covalent Bonding
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Electron Configurations
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...
 03:51
Molecular Orbital Theory II