Related Experiment Video
Updated: Jan 12, 2026

A Rapid and Quantitative Fluorimetric Method for Protein-Targeting Small Molecule Drug Screening
Published on: October 16, 2015
Drug-biomolecular interaction: Spectroscopic and computational insights into esomeprazole binding with human serum
Anna Tanuja Safala Bodapati1, Ragaiahgari Srinivas Reddy2, Kandikonda Lavanya3
1Chemistry Division, BS&H Department, BVRIT Hyderabad, College of Engineering for Women, Hyderabad 500090, India.

Abstract:
Cellular process relies on specific binding of drug molecules with desired biological receptors. Biomolecular receptors and drugs exhibit their biological functions upon their interactions. Understanding the binding parameters and energetics of the binding provides a plethora of information that may be helpful in drug design and discovery. Further, drug interaction with carrier proteins affects the pharmacokinetics and dynamics of other exogenous and endogenous drugs. In this work, we report the binding behavior of esomeprazole with human serum albumin using several biophysical techniques. Qualitative and quantitative aspects of the binding along with the binding energetics has been focused in extracting the thermodynamics parameters and key interaction force. The binding constants (Kb)obtained from Scatchard analysis are-1.17 × 10⁵, 7.49 × 10⁴, and 3.23 × 10⁴ M-1 at 298, 303, 308 K respectively. Esomeprazole binds preferentially at site 1 in subdomain IIA of human serum albumin and was confirmed, supported by site-specific marker displacement studies and docking simulations. Negative value of change in free energy (∆G0) -32 kJ/mol at 298 K showed thermodynamically favourable interaction and outweigh of entropic factor (T∆S0 = 230.76 ± 3 kJ for T = 298 K) over the enthalpic contribution (∆H0 = -261.99 kJ/mol) revealed an entropy-driven process. Binding of esomeprazole affected the helical structure of human serum albumin. Molecular docking studies (theoretical) shows the binding pocket of ESM at site1(IIA), which is in accordance with the experimental result. Further, the interface residues involved in the binding were analysed from the 2D diagram and ligplot of the docked complex.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
Related Concept Videos
 01:29
Drug Distribution: Plasma Protein Binding
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:30
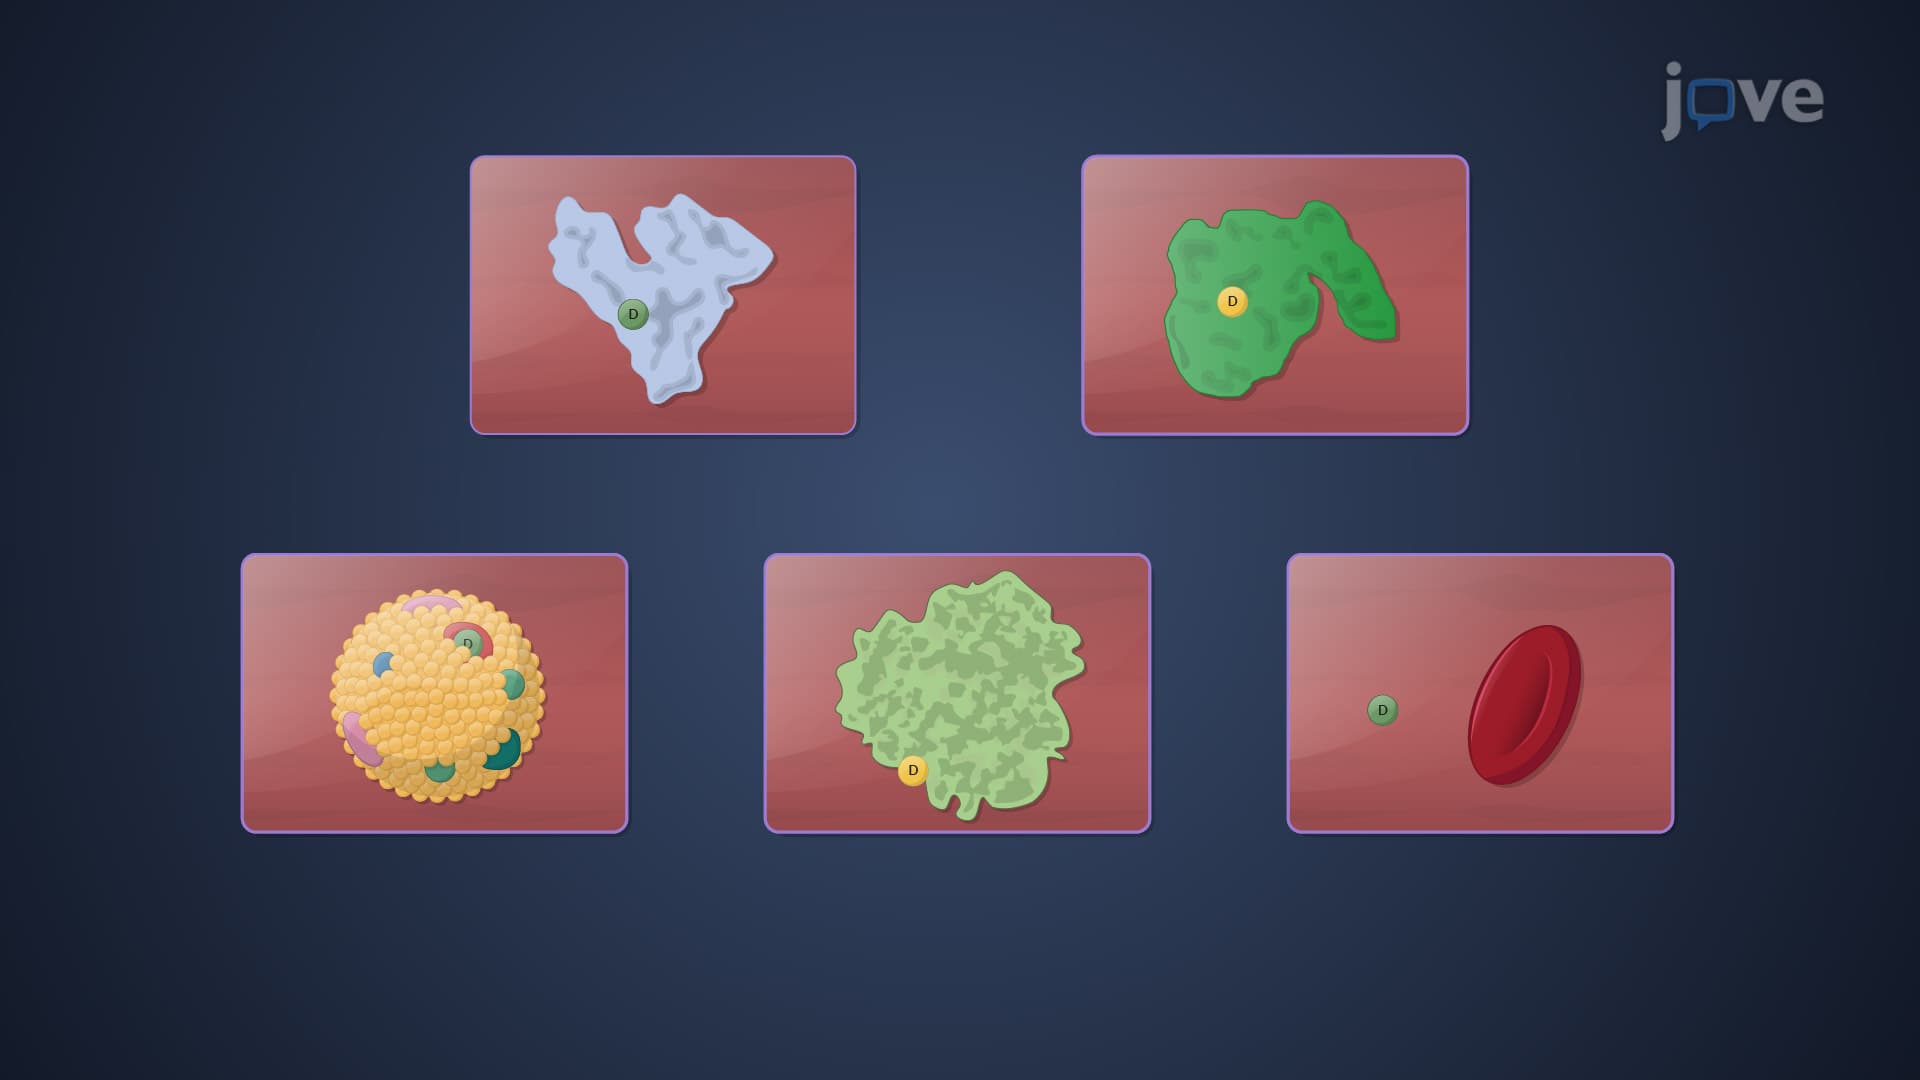
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are...
 01:20
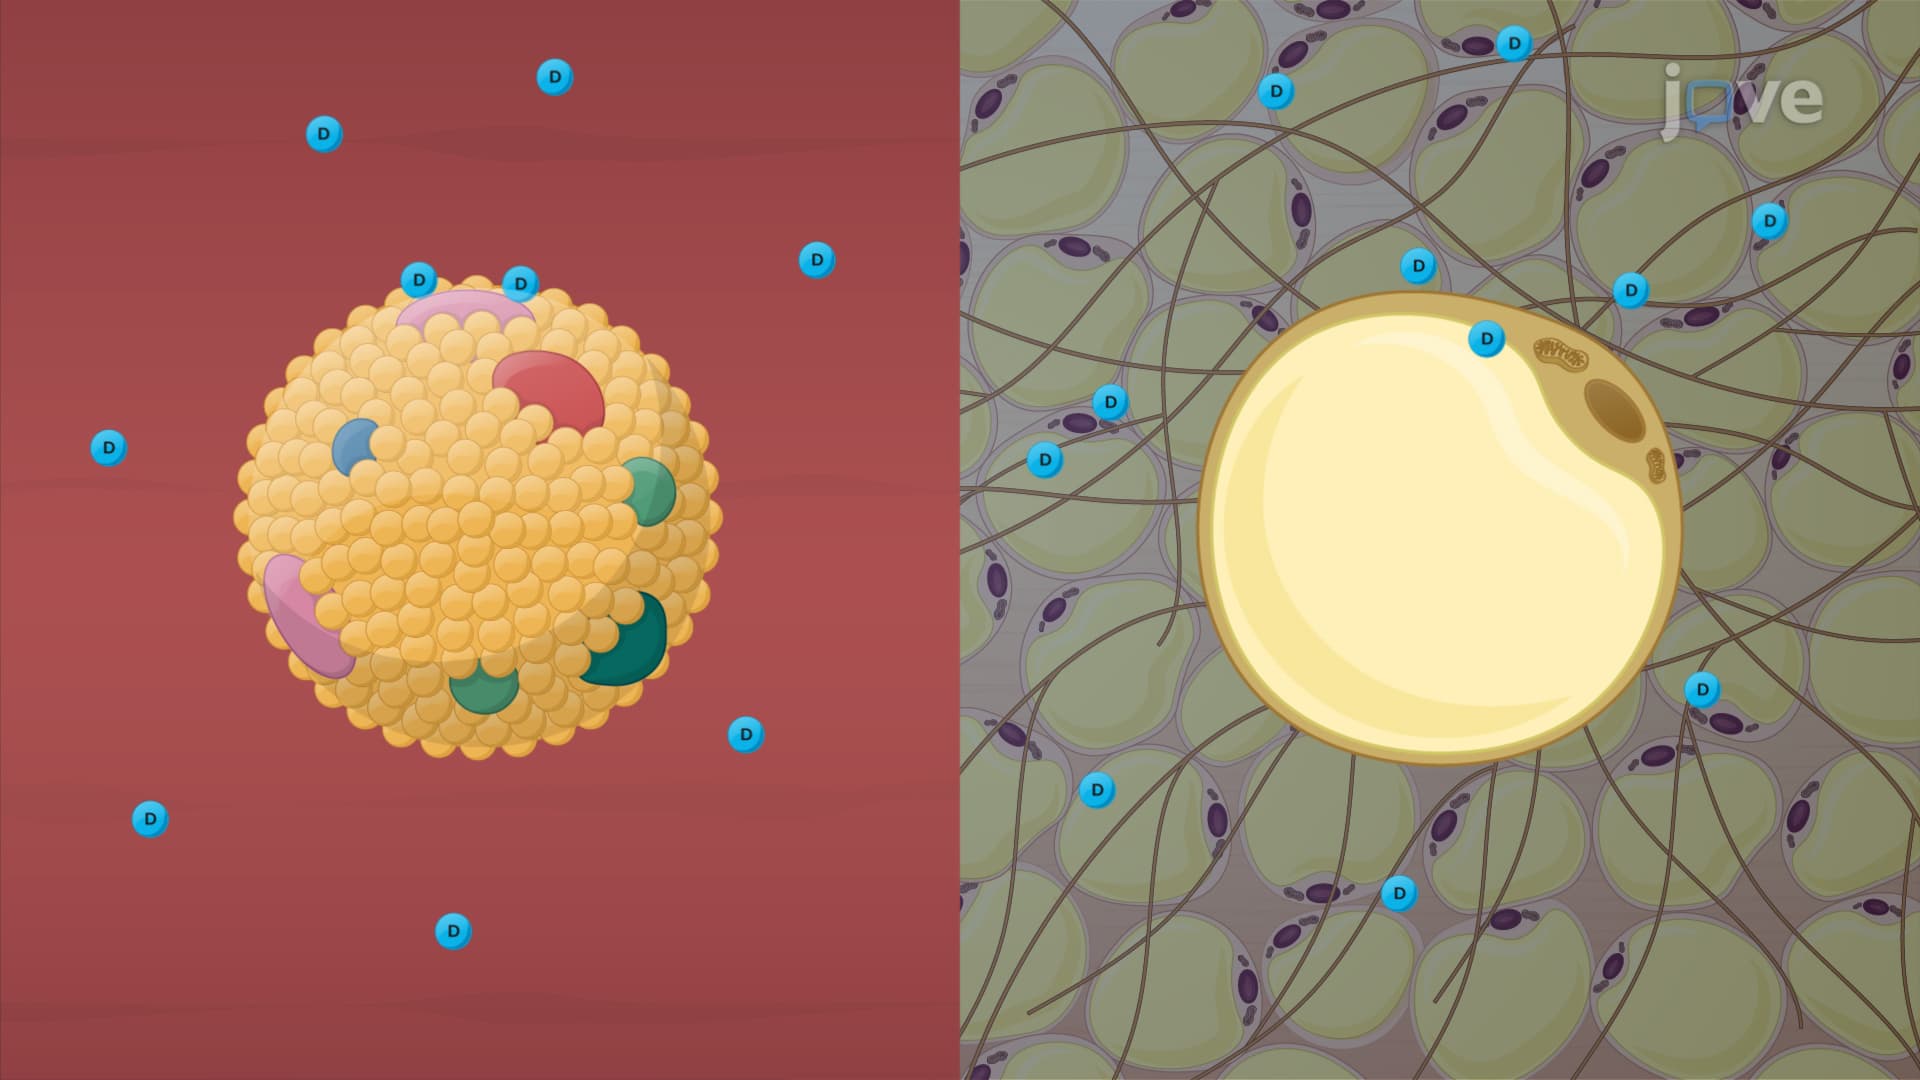
Factors Affecting Protein-Drug Binding: Protein-Related Factors
The physicochemical properties of a drug play a significant role in its ability to bind to proteins. Lipophilic drugs, which dissolve in fats, oils, and lipids, can be...
 01:18
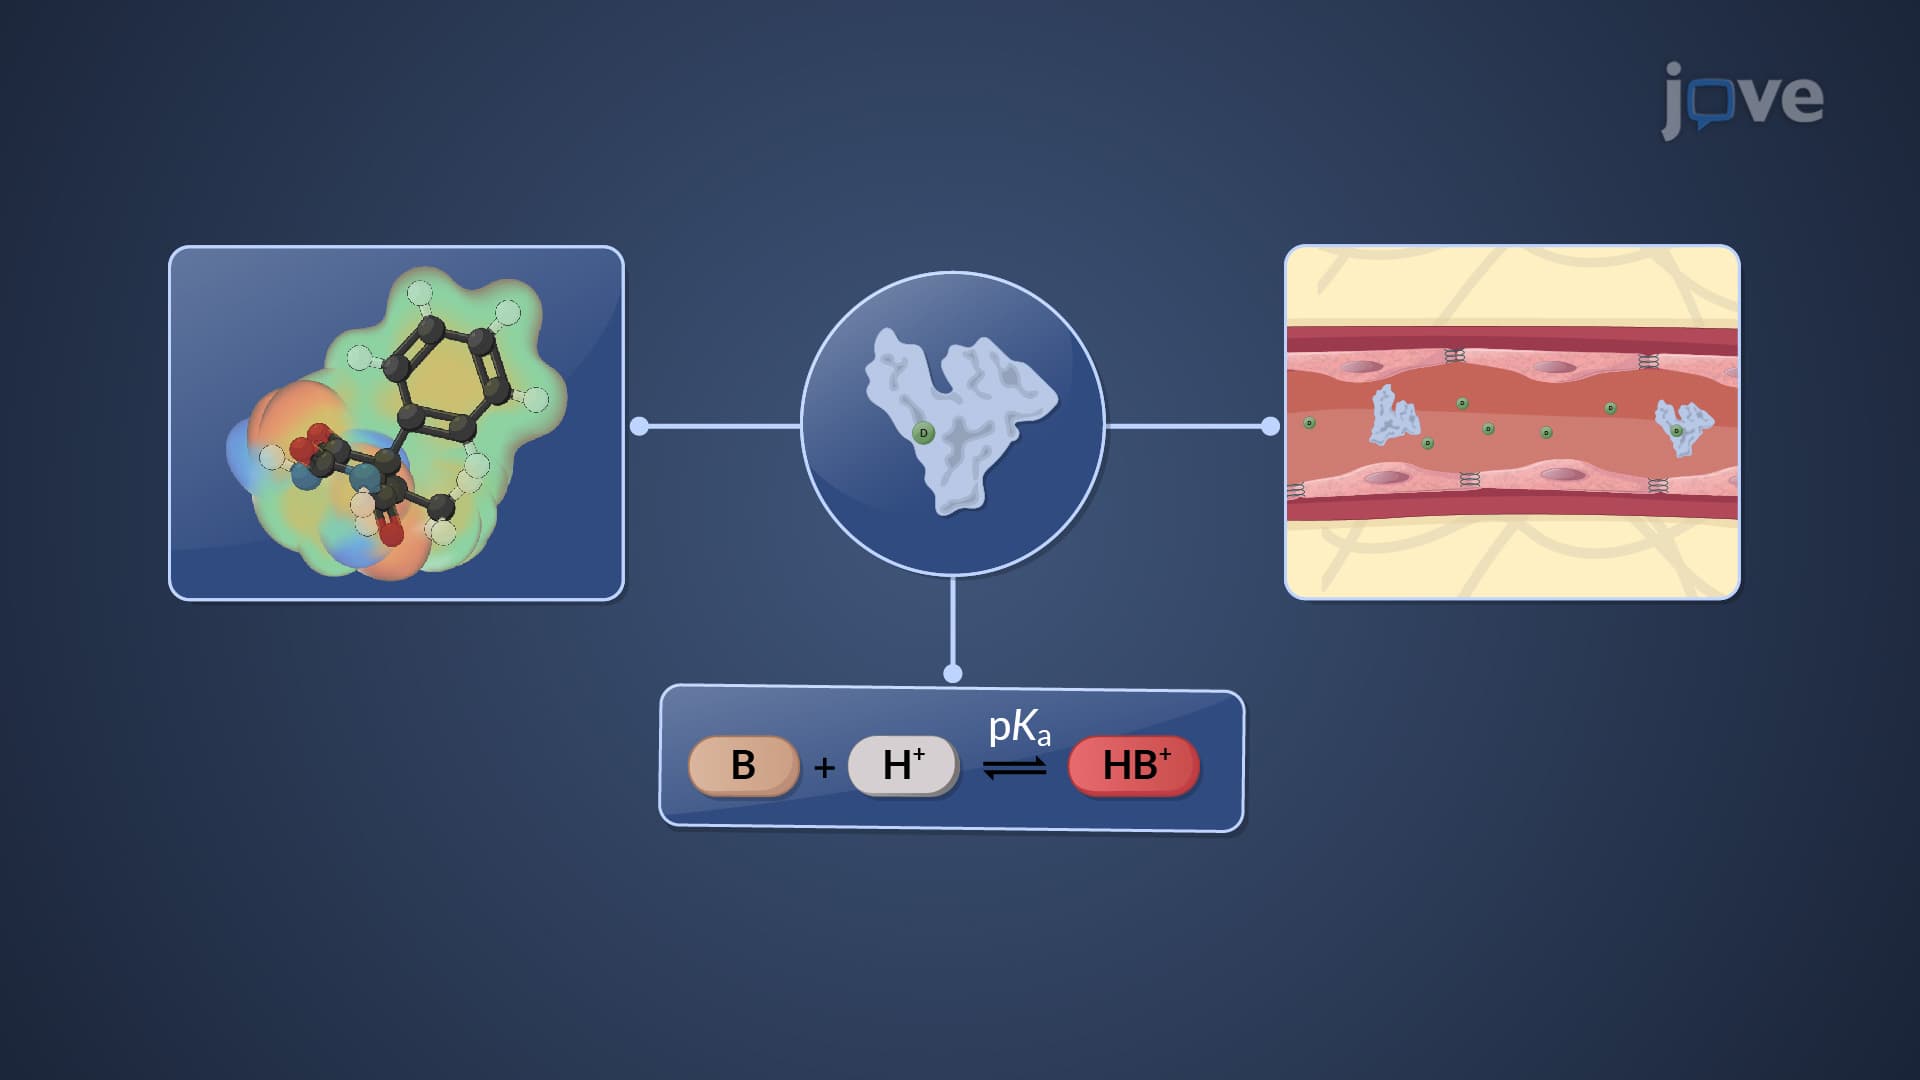
Factors Affecting Protein-Drug Binding: Drug-Related Factors
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...
 01:23
Factors Affecting Protein-Drug Binding: Drug Interactions
Displacement interactions can have varying outcomes, ranging from toxicity to virtually...