Palladium-catalyzed asymmetric propargylic sulfonylation of propargyl esters with sulfonyl hydrazides
Zhi-Xiong Ou1, Bu-Hong Zhang1, Jia-Xin Song1
1Key Laboratory of Organic Synthesis of Jiangsu Province, College of Chemistry, Chemical Engineering and Materials Science, Soochow University, Suzhou 215123, P. R. China. wangxw@suda.edu.cn.

Abstract:
Palladium-catalyzed enantioselective propargylic sulfonylation between racemic propargyl esters and readily available sulfonyl hydrazides with an N-benzylamino-bridged bidentate phosphoramidite (N-Bn-Me-BIPAM) ligand was reported. The corresponding reactions provided a series of enantiomerically rich propargylic sulfones in good yields with high enantioselectivity under mild reaction conditions.
More Related Videos
 19:58
Palladium N-Heterocyclic Carbene Complexes: Synthesis from Benzimidazolium Salts and Catalytic Activity in Carbon-carbon Bond-forming Reactions
Published on: July 30, 2017
 11:44
Mizoroki-Heck Cross-coupling Reactions Catalyzed by Dichloro{bis[1,1',1''-phosphinetriyltripiperidine]}palladium Under Mild Reaction Conditions
Published on: March 20, 2014
Related Concept Videos
 02:26
Preparation and Reactions of Sulfides
 01:12
Acid Halides to Esters: Alcoholysis
 01:08
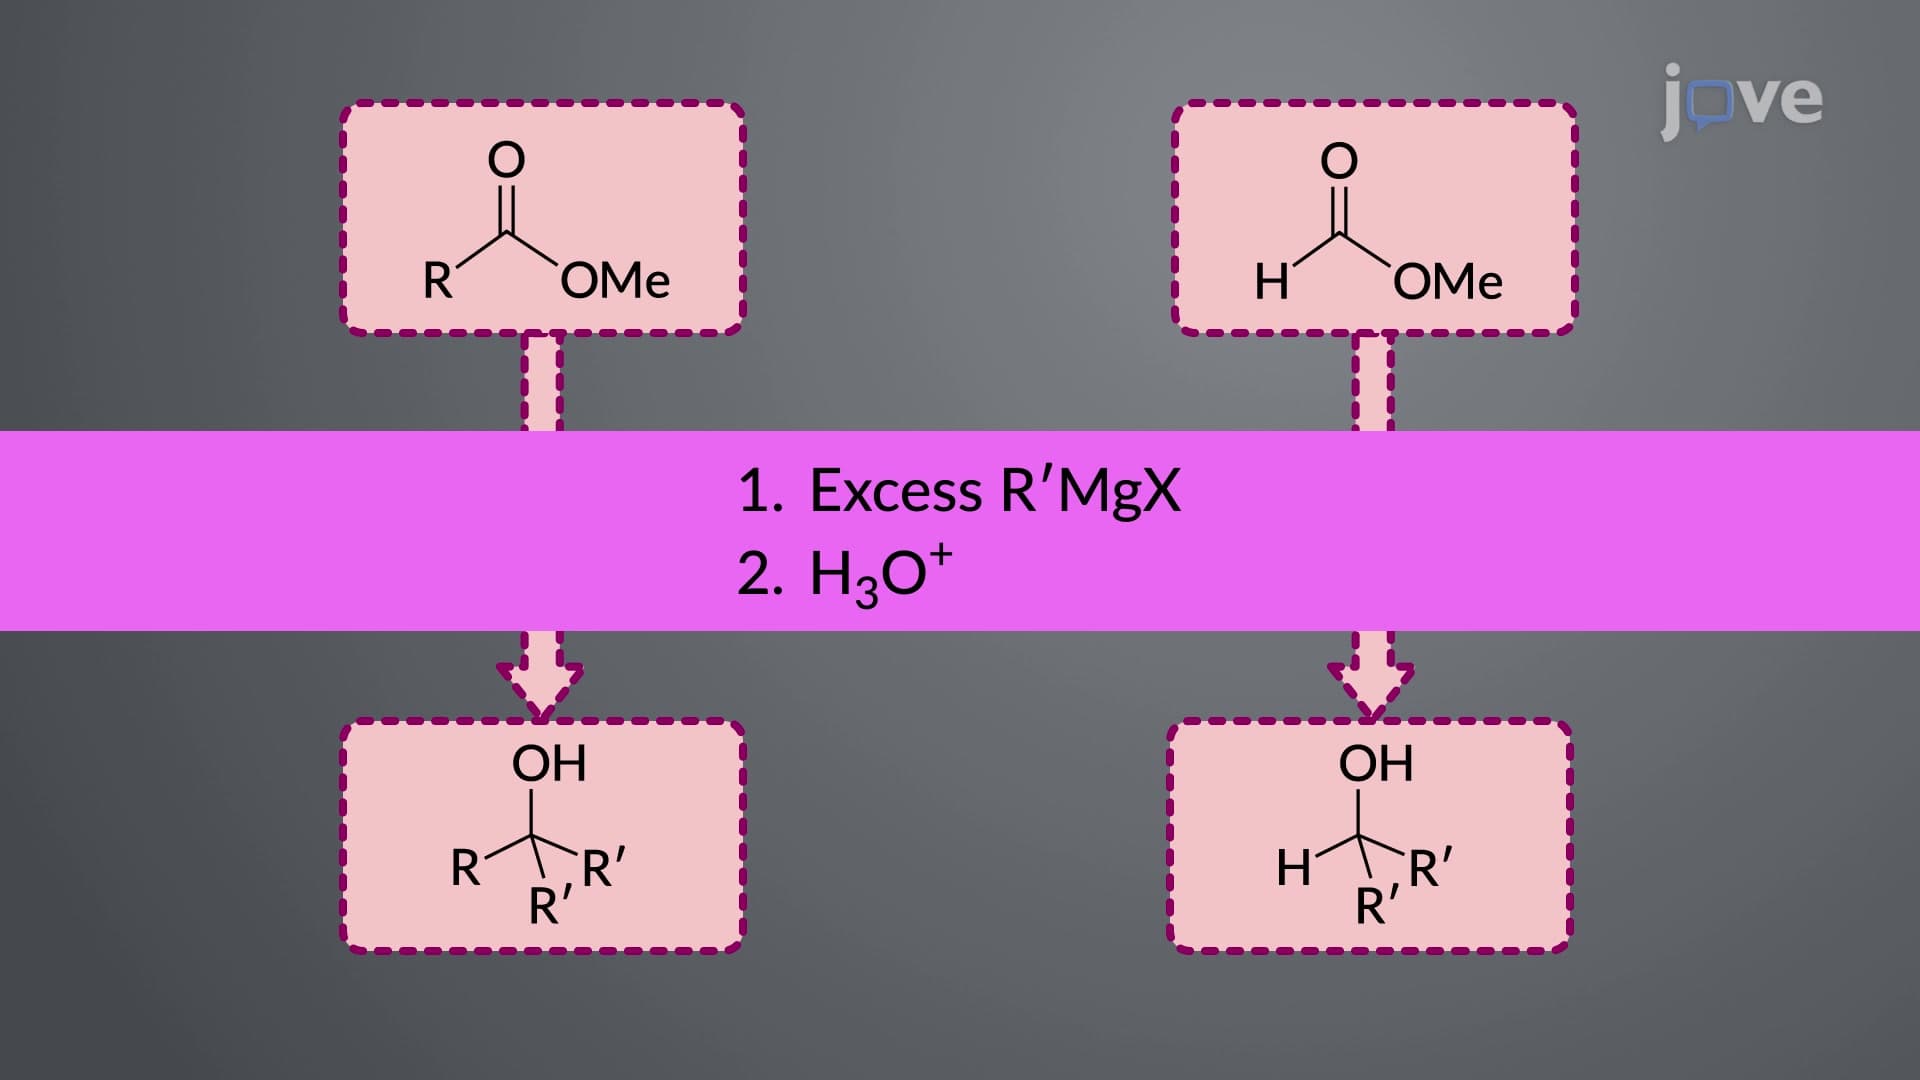
Esters to Alcohols: Grignard Reaction
The reaction requires two equivalents of the Grignard reagent and introduces two identical alkyl groups, derived from the Grignard reagent, bonded to the hydroxyl-bearing carbon of the alcohol.
The reaction follows the typical nucleophilic acyl substitution mechanism. The Grignard...
 01:27
Radical Substitution: Allylic Bromination
 01:15
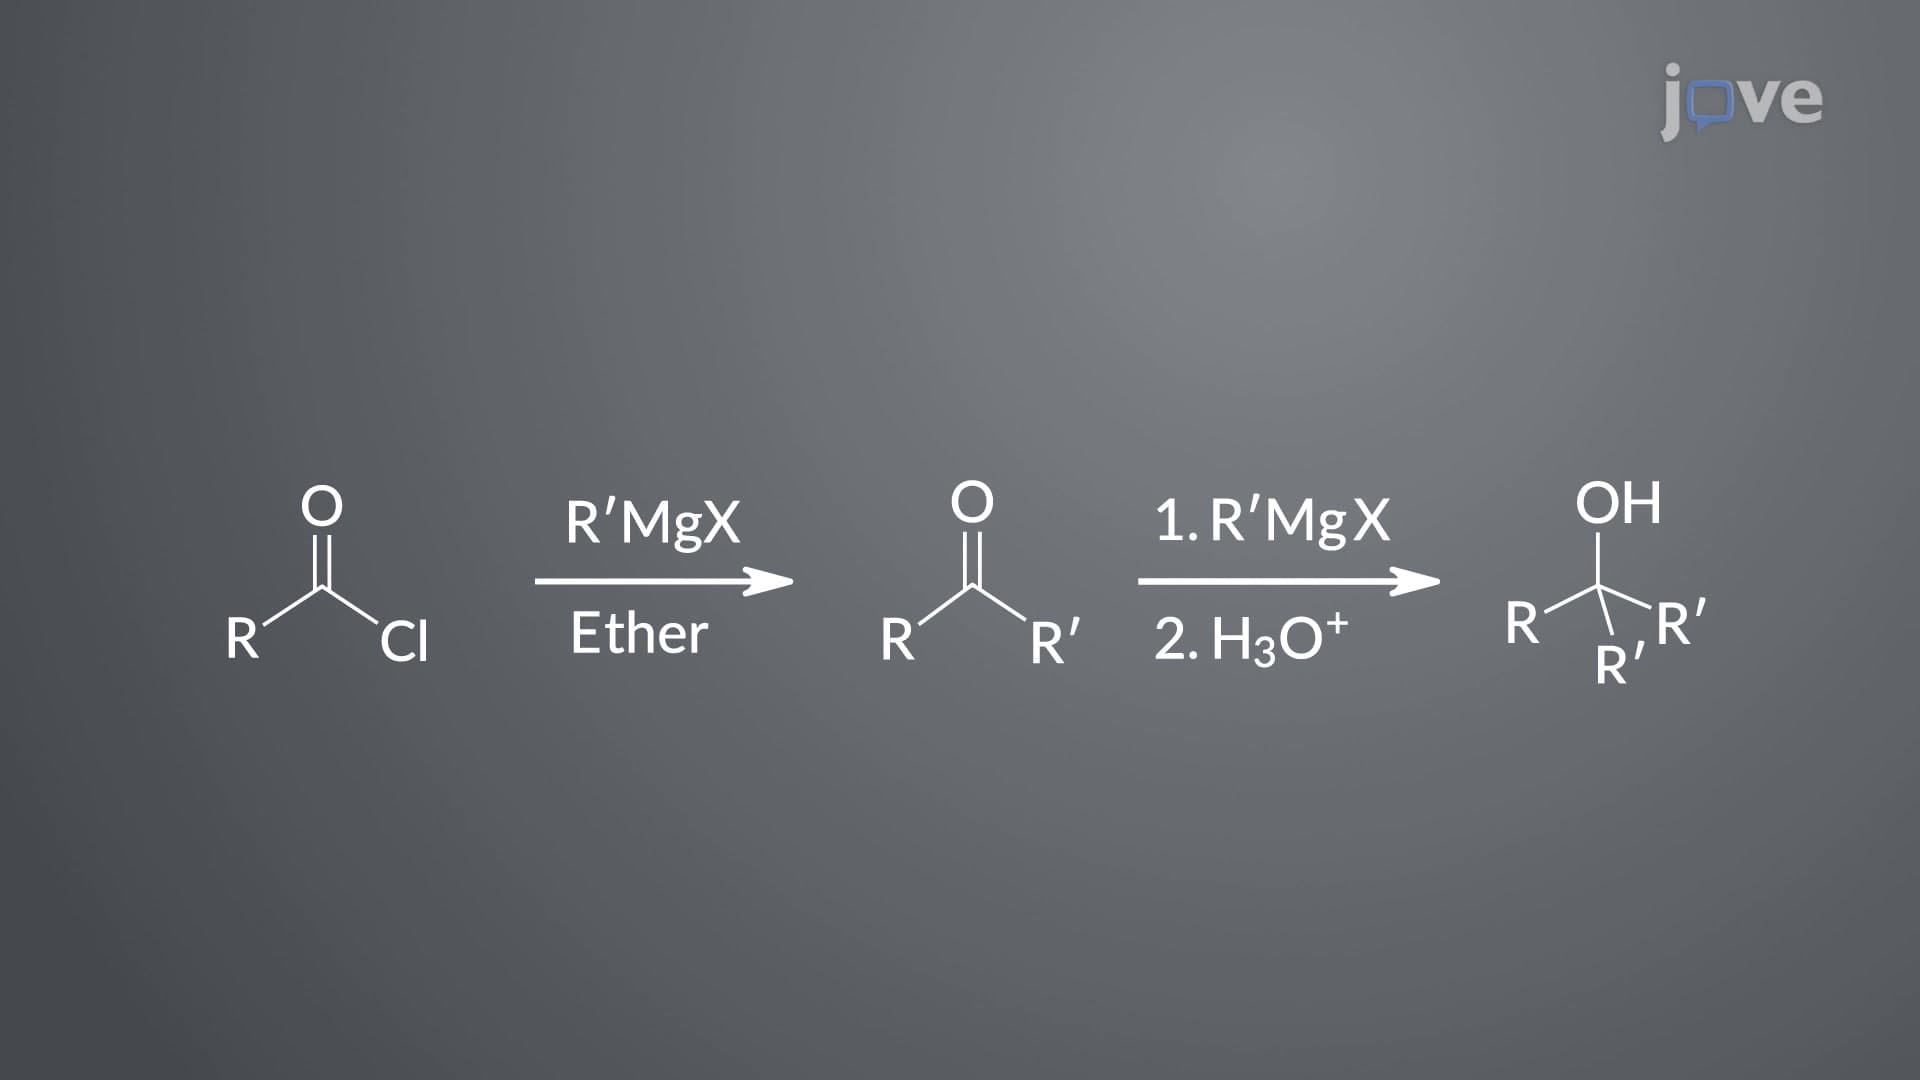
Acid Halides to Alcohols: Grignard Reaction
Grignard reagents are a source of carbanions and function as nucleophiles. The mechanism begins with the nucleophilic attack by the carbanion at the carbonyl carbon of the acid halide to form a tetrahedral intermediate. Next, the carbonyl group is re-formed, and the halide ion departs,...
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.