Related Experiment Video
Updated: Jul 5, 2026

Use of Ultra-high Field MRI in Small Rodent Models of Polycystic Kidney Disease for In Vivo Phenotyping and Drug Monitoring
Published on: June 23, 2015
Inherited Type-1 renal tubular acidosis with short stature: a rare case report from Nepal
Dipesh Kumar Singh1, Avinas Rai1, Robin Maskey1
1Department of Internal Medicine, B.P. Koirala Institute of Health Sciences, Dharan, Nepal.

Introduction:
Distal renal tubular acidosis (dRTA), or Type-1 RTA, is a rare disorder characterized by the failure of alpha-intercalated cells in the distal nephron to excrete hydrogen ions. This leads to a normal anion gap metabolic acidosis, hypokalemia, hypercalciuria, and persistently alkaline urine. Complications include nephrocalcinosis, growth retardation, and skeletal abnormalities. Hereditary forms, often involving mutations in genes like SLC4A4, typically present in early life and may include systemic features such as sensorineural hearing loss. The present case indeed reports and summarizes both clinical symptoms and diagnosis, long-term outcomes, genetic inheritance, epidemiology, and current treatment options, with the aim of shedding more light onto this rare disorder, with a specific focus on the diagnostic challenges posed by a delayed presentation and the importance of genetic evaluation.
Case Presentation:
We report a 27-year-old female with progressive lower limb weakness and chronic muscle cramps. She had a longstanding history of renal calculi since adolescence and marked growth delay. Laboratory tests revealed normal anion gap metabolic acidosis, urinary pH >7.0, hypokalemia, hypophosphatemia, and vitamin D deficiency. Imaging showed bilateral nephrocalcinosis. Genetic testing identified a pathogenic SLC4A4 mutation, confirming inherited dRTA. Treatment with potassium citrate, sodium bicarbonate, vitamin D, calcium, and iron supplementation led to significant clinical improvement.
Discussion:
This case highlights the diagnostic challenges of hereditary dRTA and its systemic burden. Nephrolithiasis, short stature, and persistent metabolic acidosis should prompt consideration of RTA. Although SLC4A4 is commonly linked with proximal defects, its mutation can present with distal tubular dysfunction, emphasizing the role of genetic analysis.
Conclusion:
Hereditary dRTA is a rare but treatable condition. Early diagnosis and lifelong alkali therapy can significantly reduce complications, including chronic kidney disease and skeletal deformities. This case reinforces the need for heightened awareness among clinicians to consider dRTA in patients presenting with chronic fatigue, muscle weakness, short stature, and recurrent nephrolithiasis. Furthermore, it underscores the value of genetic evaluation in uncovering the precise etiology, allowing for tailored management and genetic counseling.
Related Concept Videos
 01:10
Nature and Nurture
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:20
Inborn Errors of Metabolism
 01:24
Renal Tubule and Collecting Duct
Proximal Convoluted Tubule (PCT):
The PCT is the initial segment of the renal tubule, extending from the Bowman's capsule that encloses the glomerulus. Its convoluted structure and microvilli-lined cells increase the surface area for reabsorption. The PCT reabsorbs glucose, amino acids, sodium, and water from the filtrate, ensuring essential...
 01:29
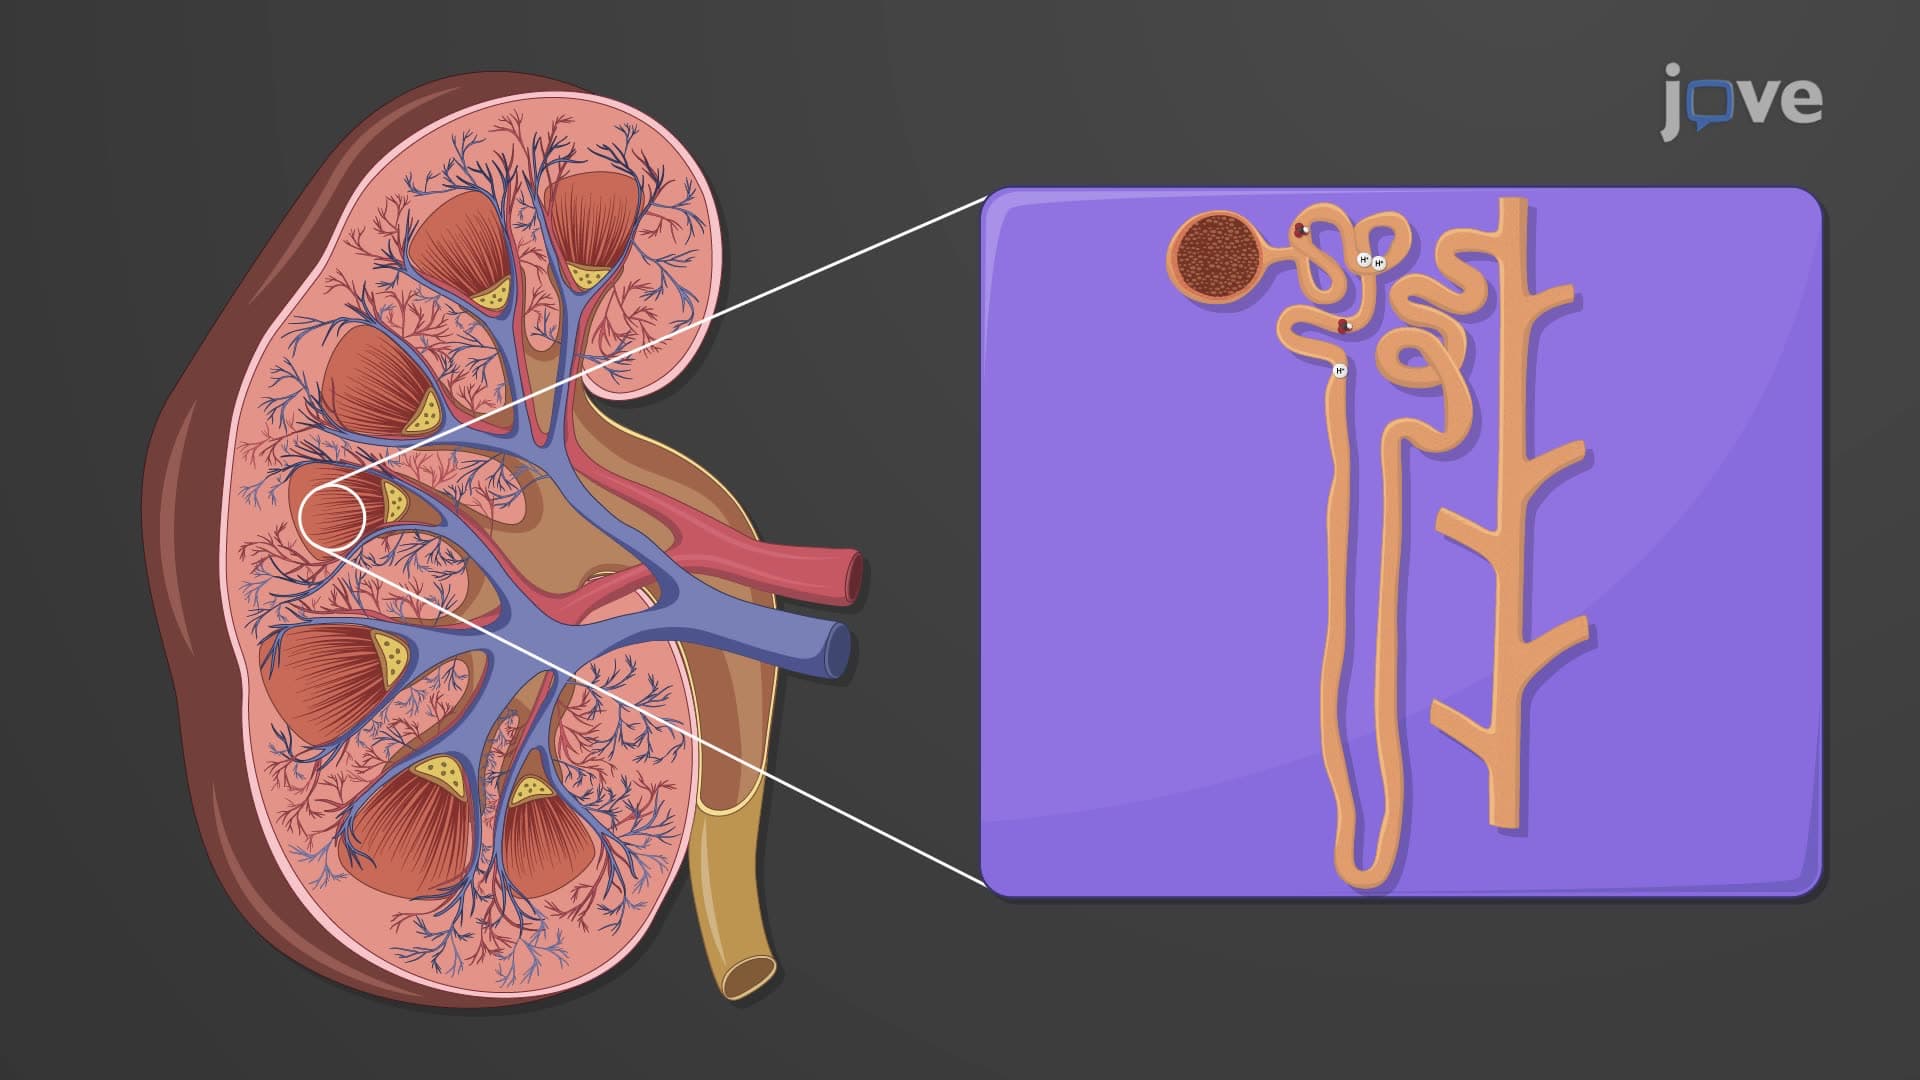
Renal Regulation of Acid-Base Balance
In the kidneys, cells within the proximal convoluted tubules (PCT) and the collecting ducts secrete hydrogen ions (H+) into the tubular fluid. Specifically, in the PCT, Na+/H+ antiporters secrete H+ while reabsorbing Na+.
However, the intercalated cells in...
 01:25
Chronic Kidney Disease I: Introduction