Related Experiment Video
Updated: Feb 5, 2026
Rapid Deletion Production in Fungi via Agrobacterium Mediated Transformation of OSCAR Deletion Constructs
Published on: June 12, 2017
Many faces of non‑deletional α‑thalassaemia variants in neonate and early childhood
Patcharee Komvilaisak1, Napat Laoaroon2, Pongsatorn Paopongsawan2
1Department of Pediatrics, Faculty of Medicine, Khon Kaen University, Khon Kaen, Thailand. patkomwi@gmail.com.

Non-deletional α-thalassemia variants, particularly hemoglobin Constant Spring (Hb CS) and hemoglobin Pakse, are prevalent in northeastern Thailand and neighboring regions. Homozygous Hb CS and compound heterozygous Hb CS/Hb Pakse can present with a wide clinical spectrum. In this study, 49 patients with these α-globin variants-with or without co-inheritance of beta-thalassemia-were evaluated. Genotypes included homozygous Hb CS (n = 27), homozygous Hb CS with Hb E (n = 8), compound heterozygous Hb CS/Hb Pakse (n = 10), compound heterozygous Hb CS/Hb Pakse with Hb E (n = 3), and homozygous Hb Pakse (n = 1). Fetal anemia with hydrops fetalis occurred in 25 patients; 21 underwent intrauterine transfusions (IUTs), resulting in favorable postnatal outcomes in most cases. Two patients with homozygous Hb CS died from complications of severe anemia and prematurity. Early neonatal jaundice occurred in 27 patients, most requiring phototherapy. Direct hyperbilirubinemia was observed in 10 patients; most recovered with supportive care and transfusion, though one died from progressive cholestasis. Twenty four patients required 1-2 postnatal transfusions, with only two requiring later transfusion. Four patients with childhood anemia were initially misdiagnosed with iron deficiency anemia before confirmation of α-thalassemia variants. After the first few months of life, patients typically exhibit mild hypochromic microcytic anemia, elevated red cell distribution width, and reticulocytosis, with basophilic stippling more prominent in Hb CS. During follow-up (3-160 months, median 66), no patients developed hepatosplenomegaly, iron overload, or gallstones. These α-thalassemia variants may cause severe fetal anemia requiring IUT but generally evolve into transfusion-independent mild anemia. Early detection and targeted intervention can improve outcomes in high-prevalence regions.
Non-deletional α-thalassemia variants, particularly hemoglobin Constant Spring (Hb CS) and hemoglobin Pakse, are prevalent in northeastern Thailand and neighboring regions. Homozygous Hb CS and compound heterozygous Hb CS/Hb Pakse can present with a wide clinical spectrum. In this study, 49 patients with these α-globin variants-with or without co-inheritance of beta-thalassemia-were evaluated. Genotypes included homozygous Hb CS (n = 27), homozygous Hb CS with Hb E (n = 8), compound heterozygous Hb CS/Hb Pakse (n = 10), compound heterozygous Hb CS/Hb Pakse with Hb E (n = 3), and homozygous Hb Pakse (n = 1). Fetal anemia with hydrops fetalis occurred in 25 patients; 21 underwent intrauterine transfusions (IUTs), resulting in favorable postnatal outcomes in most cases. Two patients with homozygous Hb CS died from complications of severe anemia and prematurity. Early neonatal jaundice occurred in 27 patients, most requiring phototherapy. Direct hyperbilirubinemia was observed in 10 patients; most recovered with supportive care and transfusion, though one died from progressive cholestasis. Twenty four patients required 1-2 postnatal transfusions, with only two requiring later transfusion. Four patients with childhood anemia were initially misdiagnosed with iron deficiency anemia before confirmation of α-thalassemia variants. After the first few months of life, patients typically exhibit mild hypochromic microcytic anemia, elevated red cell distribution width, and reticulocytosis, with basophilic stippling more prominent in Hb CS. During follow-up (3-160 months, median 66), no patients developed hepatosplenomegaly, iron overload, or gallstones. These α-thalassemia variants may cause severe fetal anemia requiring IUT but generally evolve into transfusion-independent mild anemia. Early detection and targeted intervention can improve outcomes in high-prevalence regions.
Related Concept Videos
 02:30
Histone Variants at the Centromere
 01:28
Erikson's Theory on Socioemotional Development during Childhood
The first four of Erikson's eight...
 01:25
Piaget's Theory of Cognitive Development from Childhood into Adulthood
Schemata: Building Blocks of Knowledge
 01:10
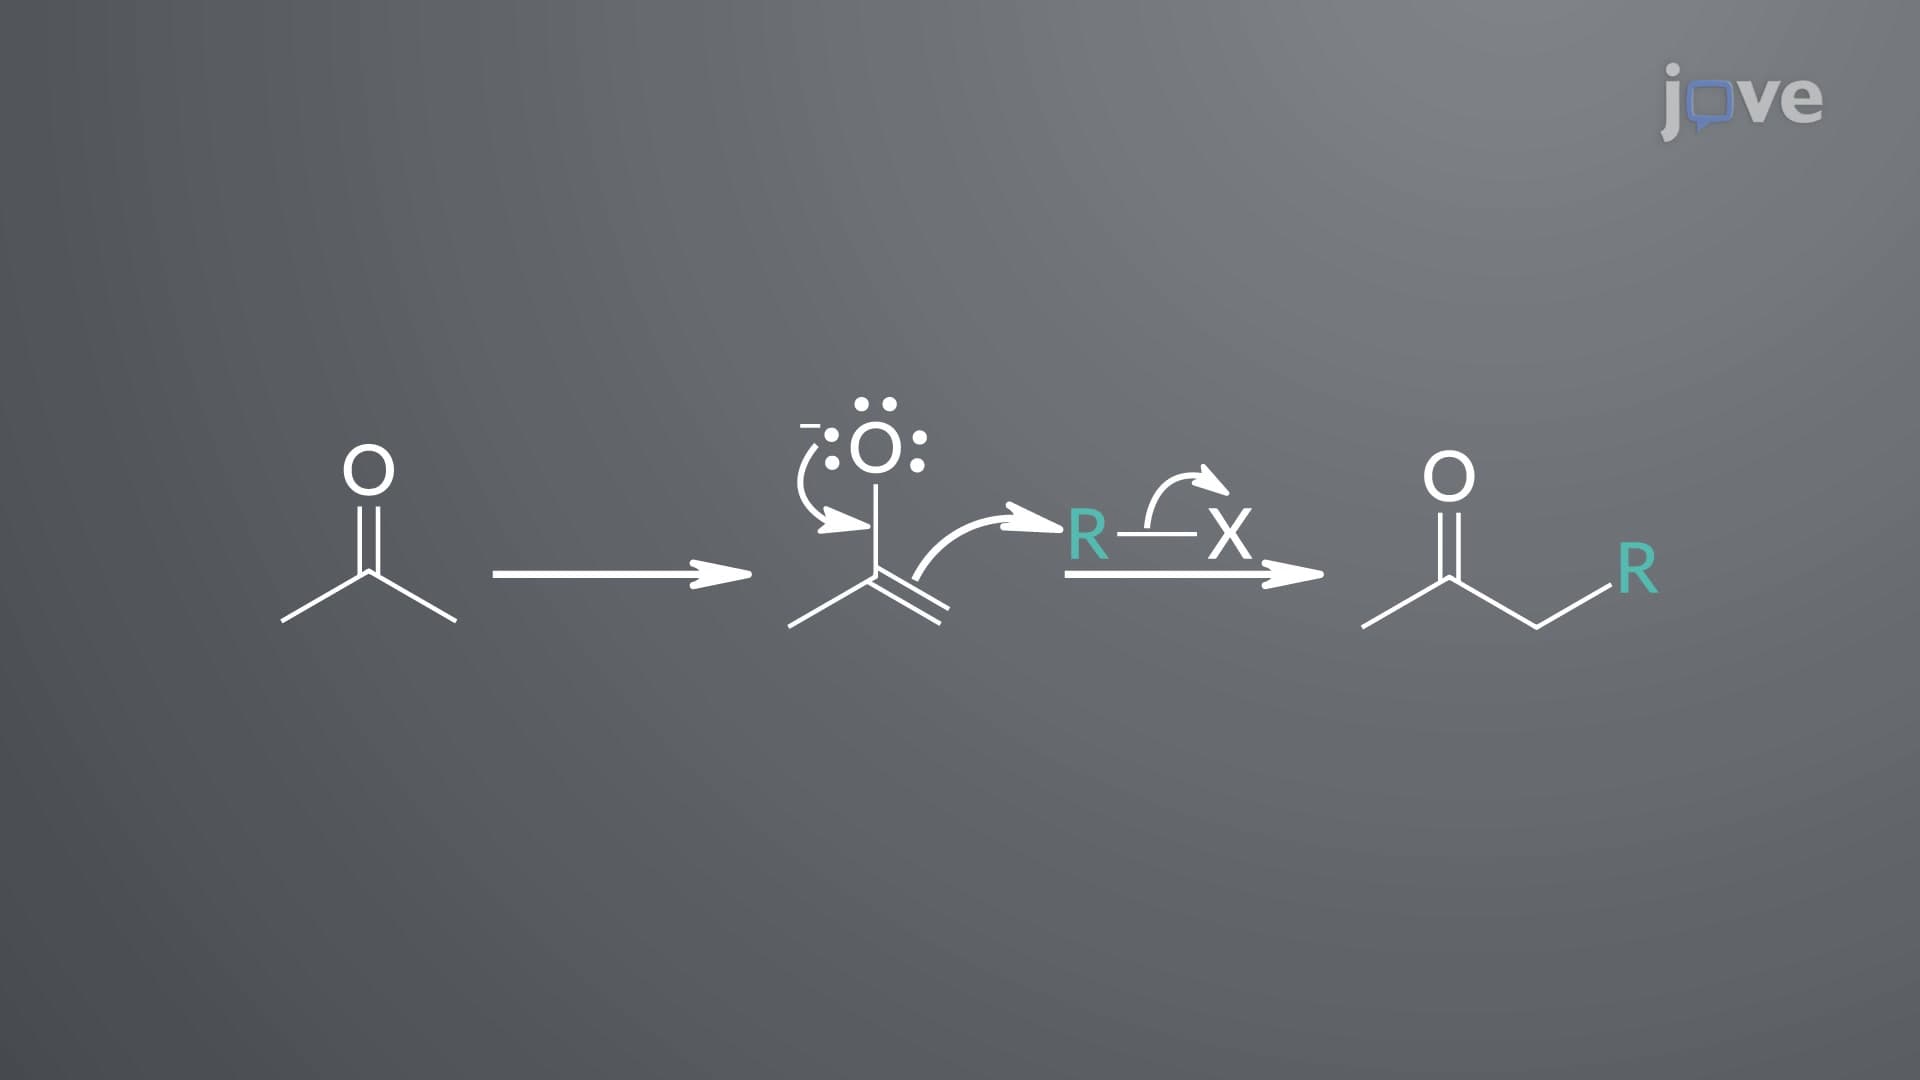
α-Alkylation of Ketones via Enolate Ions
 01:10
Factors Affecting α-Alkylation of Ketones: Choice of Base
The reaction involving bases like EtO− whose conjugate acid EtOH (pKa = 15.9) is stronger than the ketone (pKa = 19.2) results in an equilibrium mixture with higher ketone concentration. As a consequence,...
 01:17
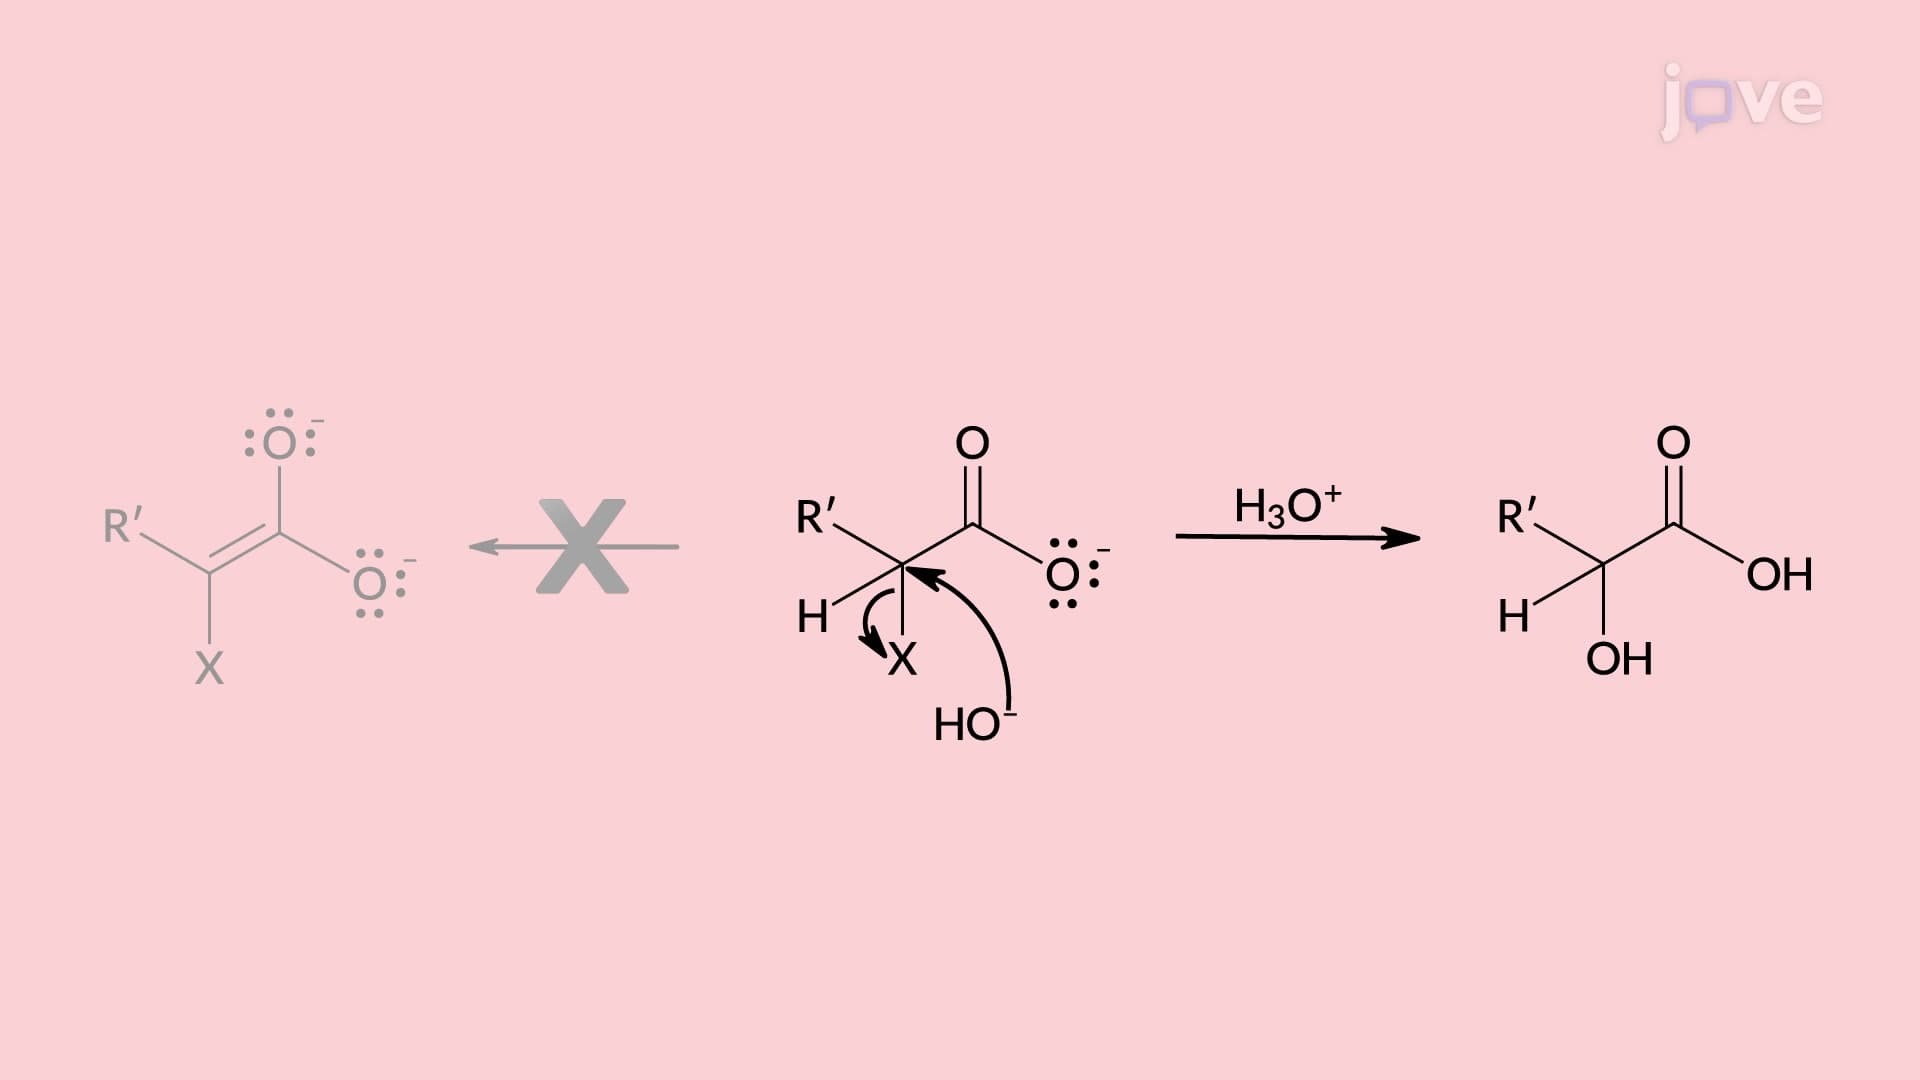
Reactions of α-Halocarbonyl Compounds: Nucleophilic Substitution