A novel tRNASer(AGY) 12244G > a variant impairs mitochondrial function and presents with classical MELAS phenotype
Related Experiment Video
Updated: Feb 17, 2026
Modeling Mitochondrial Disease Using Brain Organoids: A Focus on Mitochondrial Encephalomyopathy, Lactic Acidosis, and Stroke-like Episodes
Published on: October 10, 2025
Xingyu Zhuang1, Jiayin Wang1, Jianing Wang1
1Department of Neurology, Shandong Key Laboratory of Mitochondrial Medicine and Rare Diseases, Research Institute of Neuromuscular and Neurodegenerative Diseases, Qilu Hospital of Shandong University, Jinan, Shandong, China.

Background:
Mitochondrial disorders are a group of heterogeneous diseases marked by deficiencies in oxidative phosphorylation (OXPHOS). A common subtype, MELAS (mitochondrial encephalopathy, lactic acidosis, and stroke-like episodes [SLEs]), is primarily linked to variants in mitochondrial transfer RNA (mt-tRNA) genes, yet the molecular mechanisms underlying many of these variants remain poorly understood.
Methods:
We performed a comprehensive assessment of a 14-year-old male patient, including clinical evaluation, genetic testing, histopathology, and functional biochemical analyses of muscle tissue. A systematic literature review was conducted to compare previously reported MT-TS2 variants and their associated phenotypes.
Results:
We identified a rare m.12244G > A variant in the tRNASer(AGY) gene associated with classical MELAS phenotype. Functional analysis demonstrated impaired mitochondrial translation and OXPHOS dysfunction. Histological findings revealed COX-negative and ragged red fibers, while western blotting indicated downregulation of key mitochondrial proteins. Literature review showed that MT-TS2 variants are associated with variable phenotypes including encephalopathy, myopathy, deafness, diabetes, and retinopathy.
Conclusion:
Our study provides the first experimental validation of the pathogenicity of the m.12244G > A variant, confirming its deleterious impact on mitochondrial function. This finding expands the genotype spectrum of MELAS and highlights the importance of functional validation for rare mtDNA variants.
Insights
A rare mitochondrial DNA variant, m.12244G>A in the MT-TS2 gene, causes MELAS (mitochondrial encephalopathy, lactic acidosis, and stroke-like episodes) by impairing mitochondrial function and translation.
Area of Science:
- Genetics
- Molecular Biology
- Biochemistry
Background:
- Mitochondrial disorders are complex diseases affecting oxidative phosphorylation (OXPHOS).
- MELAS (mitochondrial encephalopathy, lactic acidosis, and stroke-like episodes) is a common subtype often caused by mitochondrial tRNA gene variants.
- The precise mechanisms of many MELAS-associated variants are not fully understood.
Purpose of the Study:
- To investigate the pathogenicity of a rare m.12244G>A variant in the MT-TS2 gene.
- To elucidate the molecular mechanisms underlying the variant's effect on mitochondrial function.
- To expand the understanding of genotype-phenotype correlations in MELAS.
Main Methods:
- Comprehensive clinical evaluation, genetic testing, and muscle tissue analysis (histopathology, biochemical assays) of a patient.
- Functional assessments of mitochondrial translation and OXPHOS.
- Systematic literature review of MT-TS2 variants and associated phenotypes.
Main Results:
- Identified the m.12244G>A variant in the tRNASer(AGY) gene, linked to a classical MELAS phenotype.
- Demonstrated impaired mitochondrial translation and OXPHOS dysfunction due to the variant.
- Observed COX-negative fibers, ragged red fibers, and reduced mitochondrial protein levels; literature review revealed diverse phenotypes for MT-TS2 variants.
Conclusions:
- Provided experimental validation for the pathogenicity of the m.12244G>A variant.
- Confirmed the variant's detrimental impact on mitochondrial function.
- Expanded the known genetic causes of MELAS and emphasized the need for functional studies of mtDNA variants.
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:48
ATP Synthase: Mechanism
 01:19
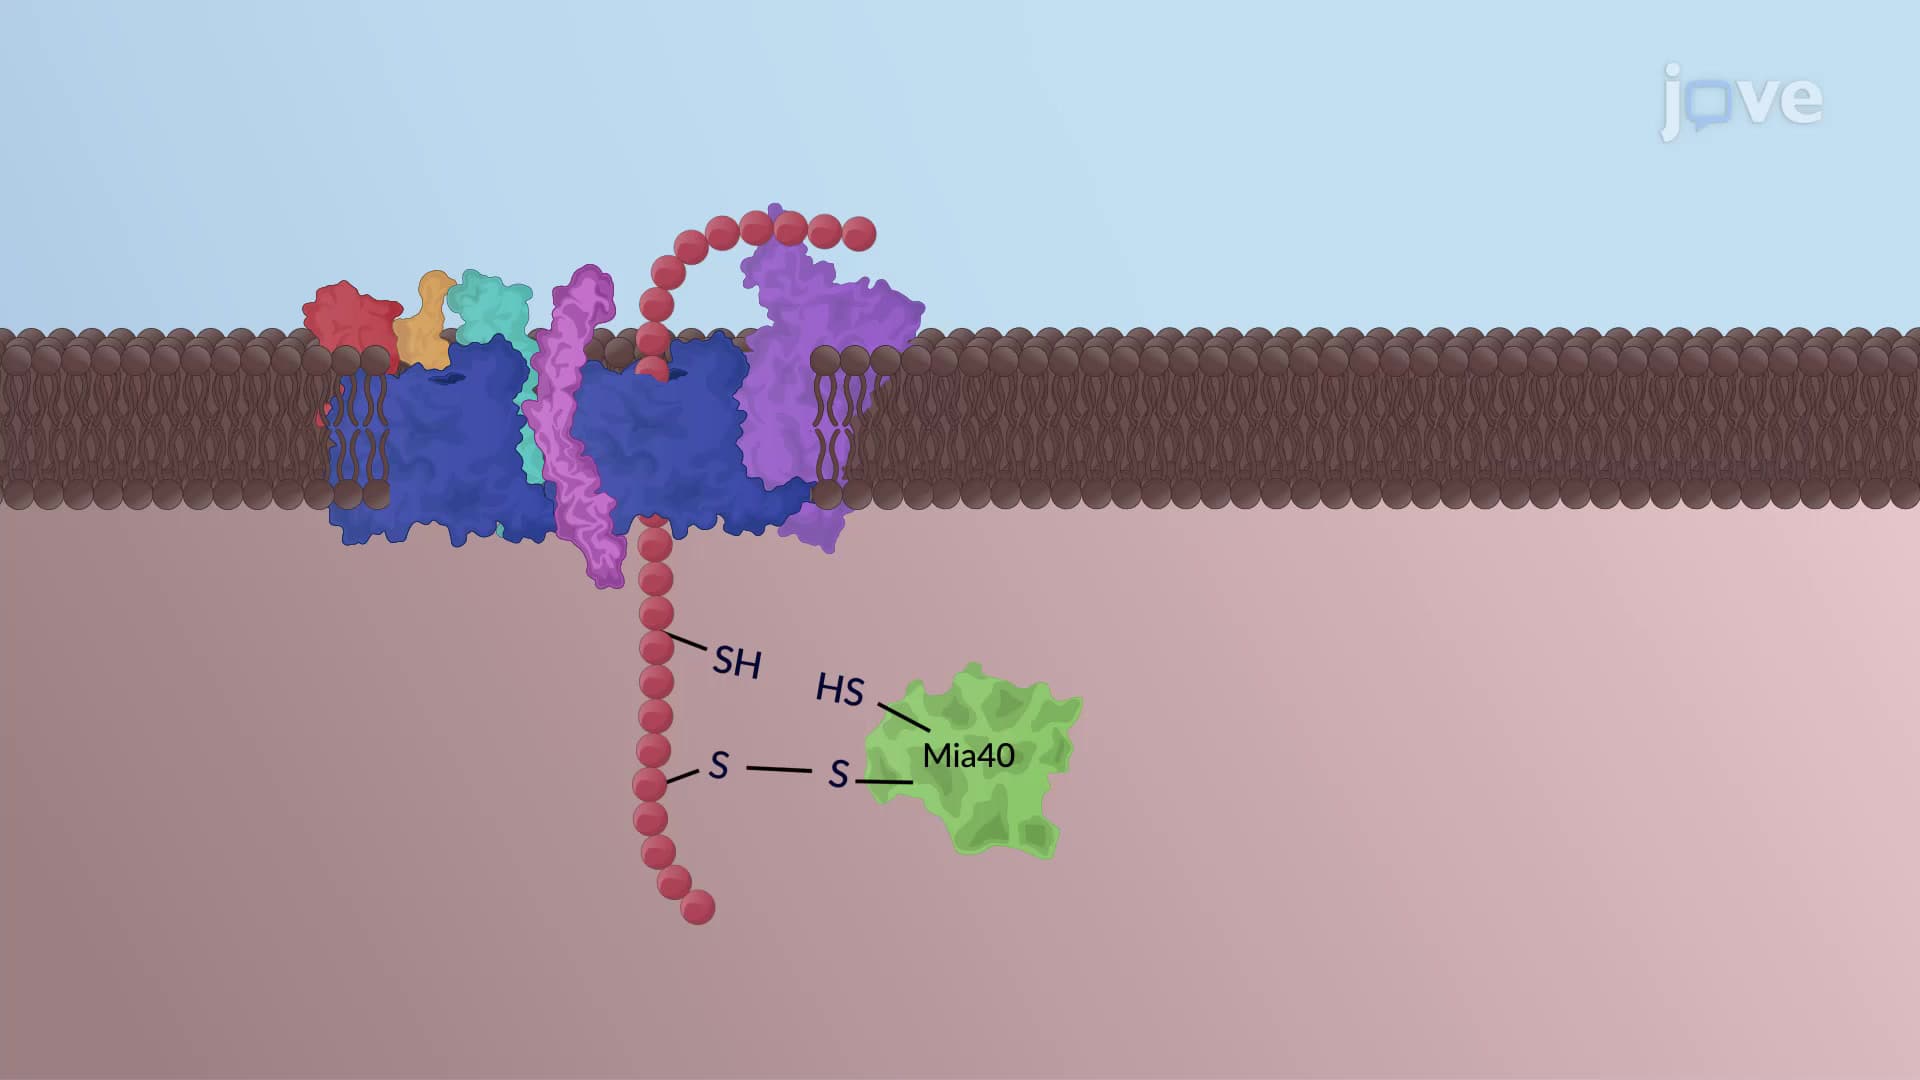
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...