Related Experiment Video
Updated: Mar 29, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Free energy calculations in molecular modeling: from classical methods to machine learning
1, Mumbai, India. apoorva.pur@gmail.com.

Context:
Free energy calculations have evolved from specialized theoretical tools to essential components of modern molecular design, particularly in pharmaceutical research where alchemical binding free energy methods now guide lead optimization decisions. Despite significant methodological advances over the past two decades, including statistically optimal estimators, enhanced sampling techniques, multi-dimensional nonequilibrium protocols, and improved force fields, current methods achieve reliable accuracy only for favorable systems involving rigid proteins and uncharged ligands. Challenges include inadequate sampling of protein flexibility, systematic biases from classical force fields for charged species, and convergence failures for systems with slow conformational transitions, limiting broader application to challenging but industrially relevant targets.
Methods:
This review provides a comprehensive analysis of free energy calculation methodologies, from foundational approaches (thermodynamic integration, Bennett acceptance ratio) to advanced enhanced sampling strategies (replica exchange, metadynamics) and cutting-edge developments including machine learning integration. Current accuracy benchmarks are assessed through systematic analysis of community challenges and industrial validation studies, identifying specific failure modes and their underlying physical origins. This review emphasizes practical method selection criteria and realistic accuracy expectations while also examining how machine learning approaches-including neural network potentials, automated collective variable discovery, and active learning protocols-address limitations in sampling efficiency and force field accuracy.
More Related Videos
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
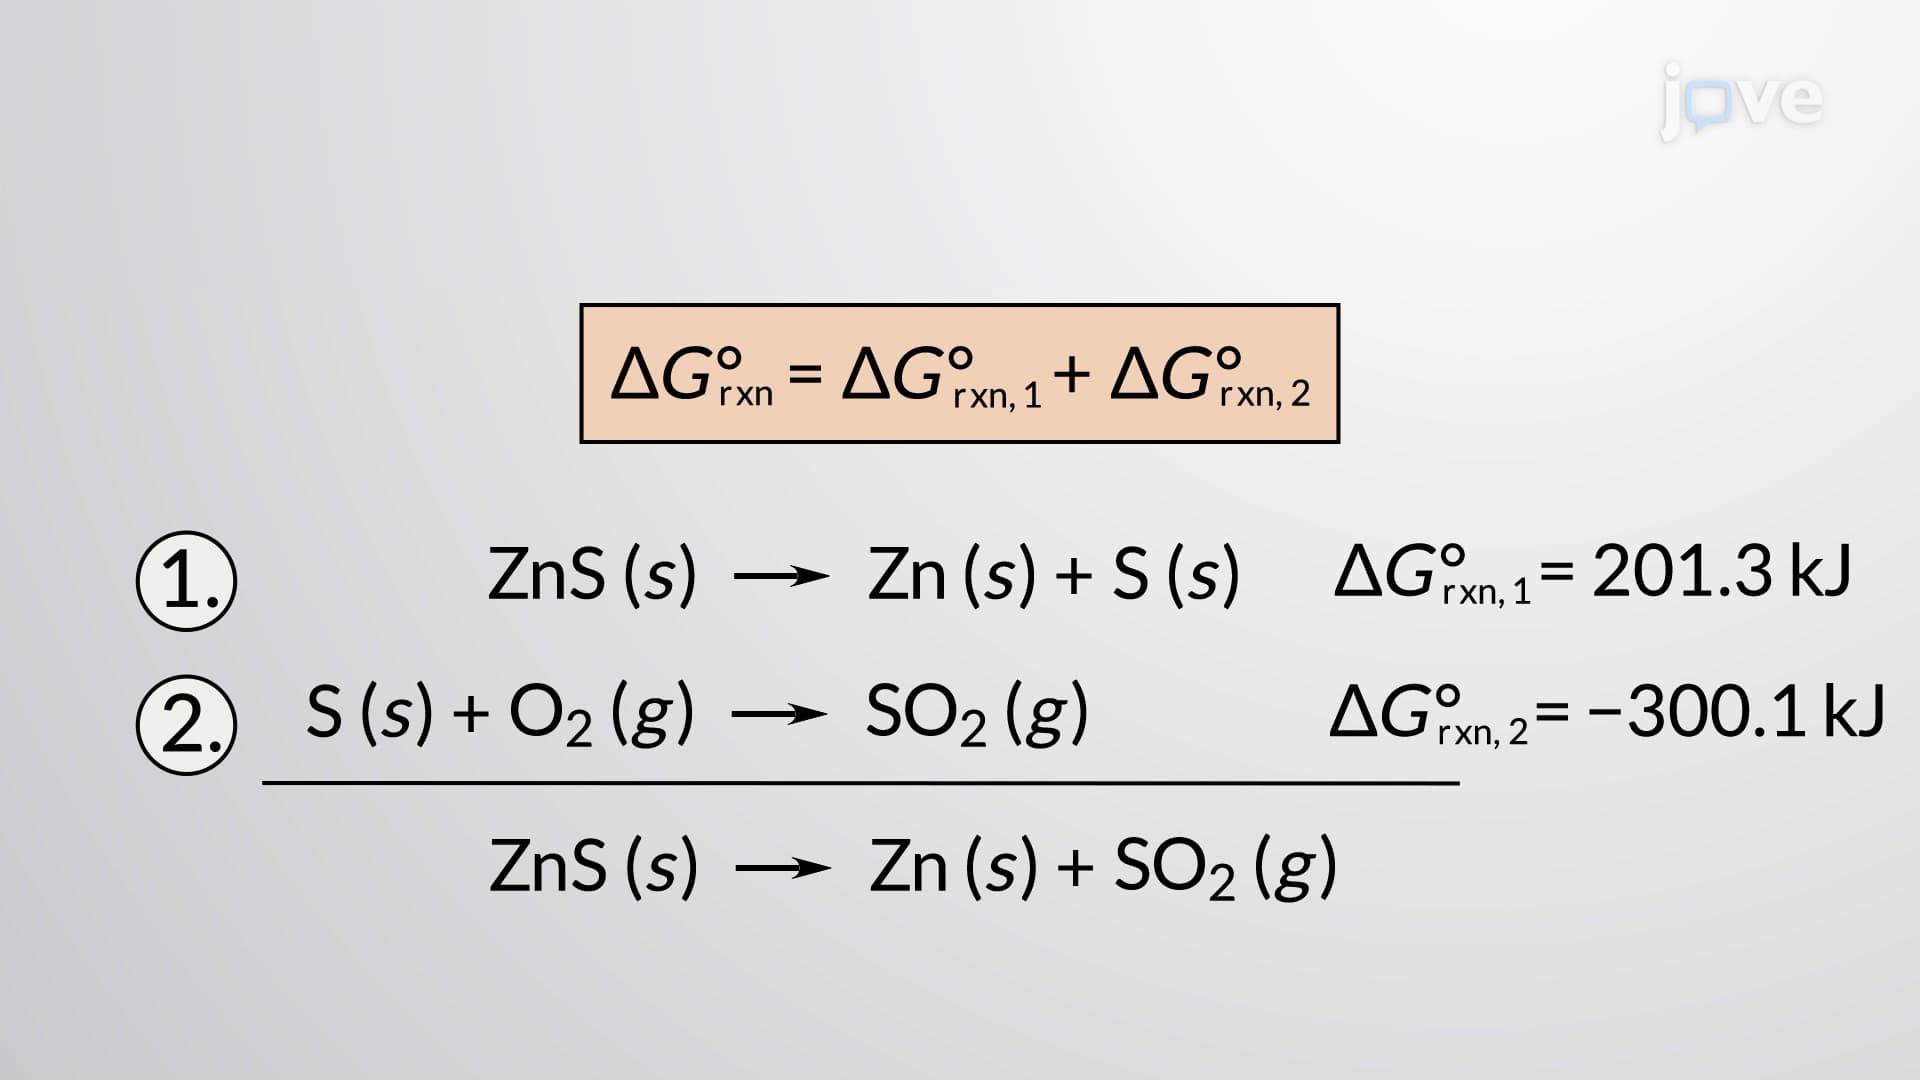 02:49
Calculating Standard Free Energy Changes
 02:27
Predicting Molecular Geometry
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
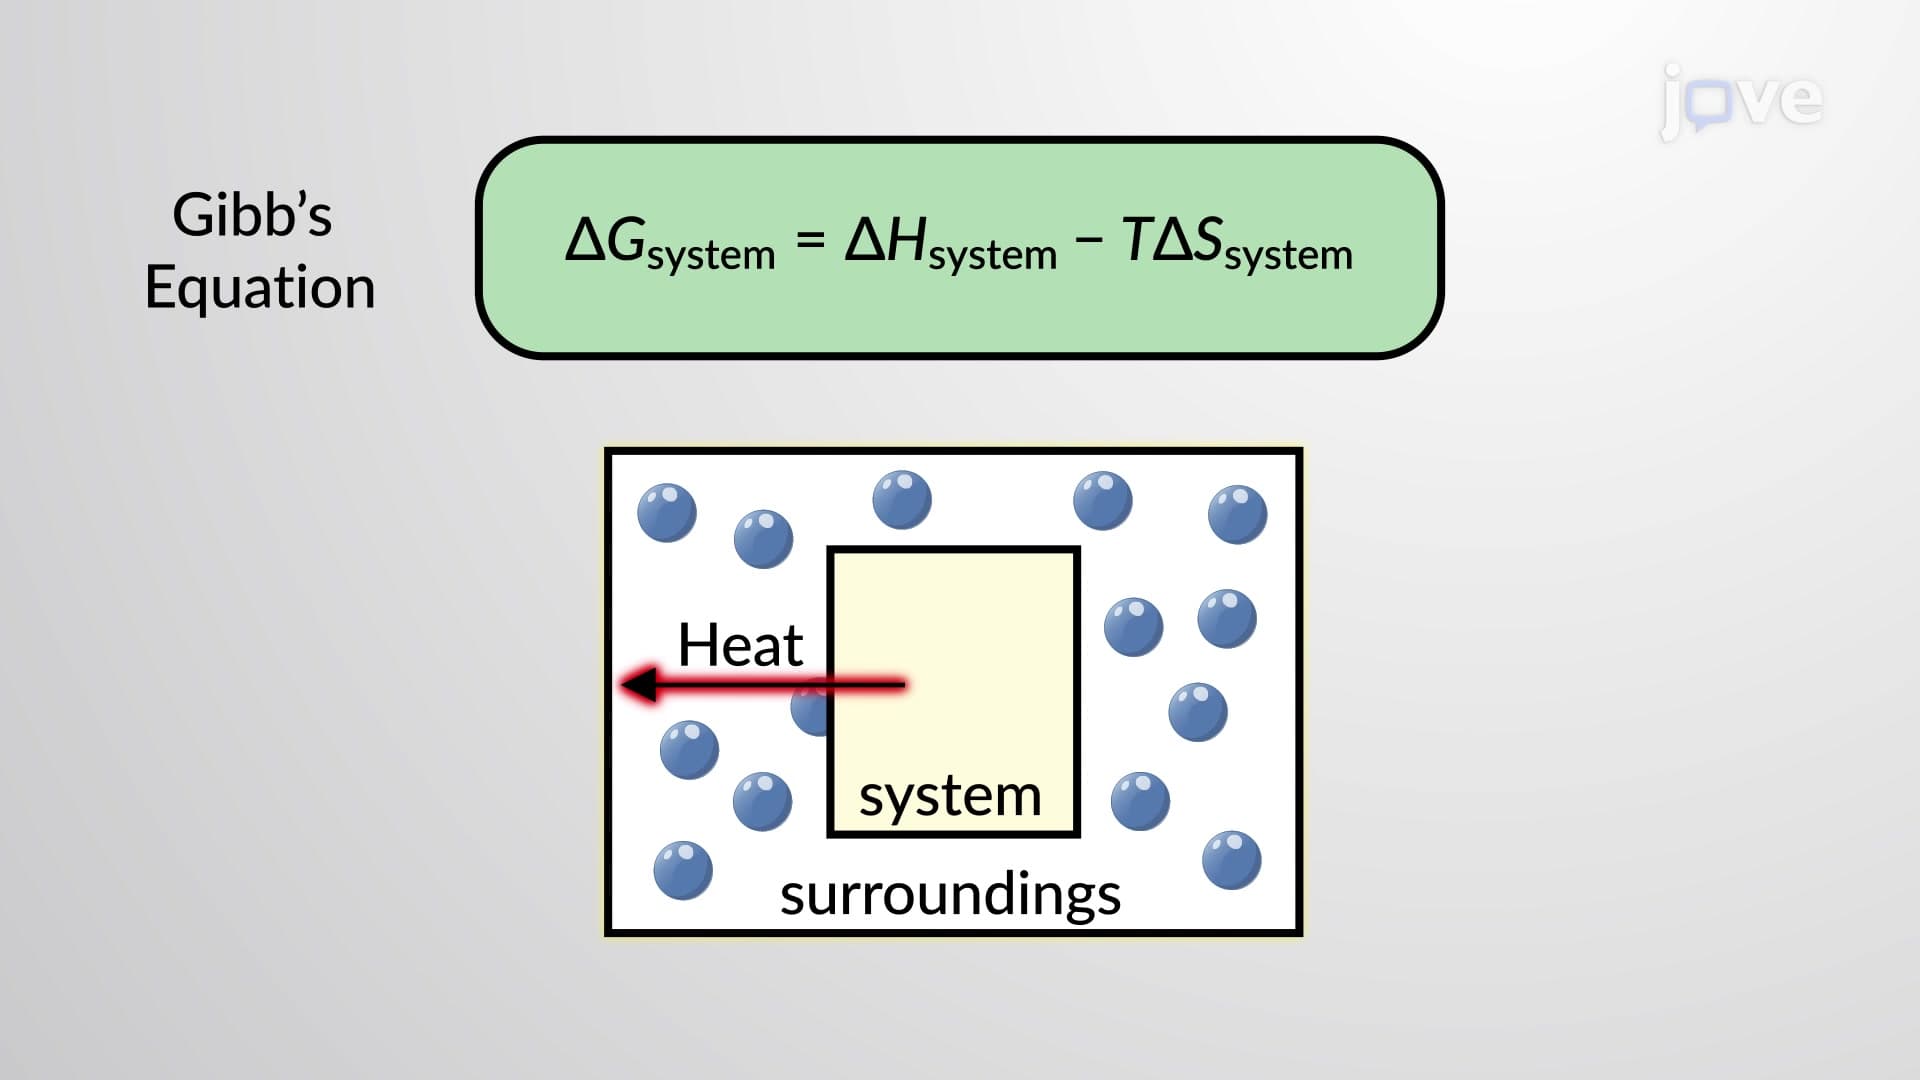 01:05
An Introduction to Free Energy
 02:00
Molecular Models
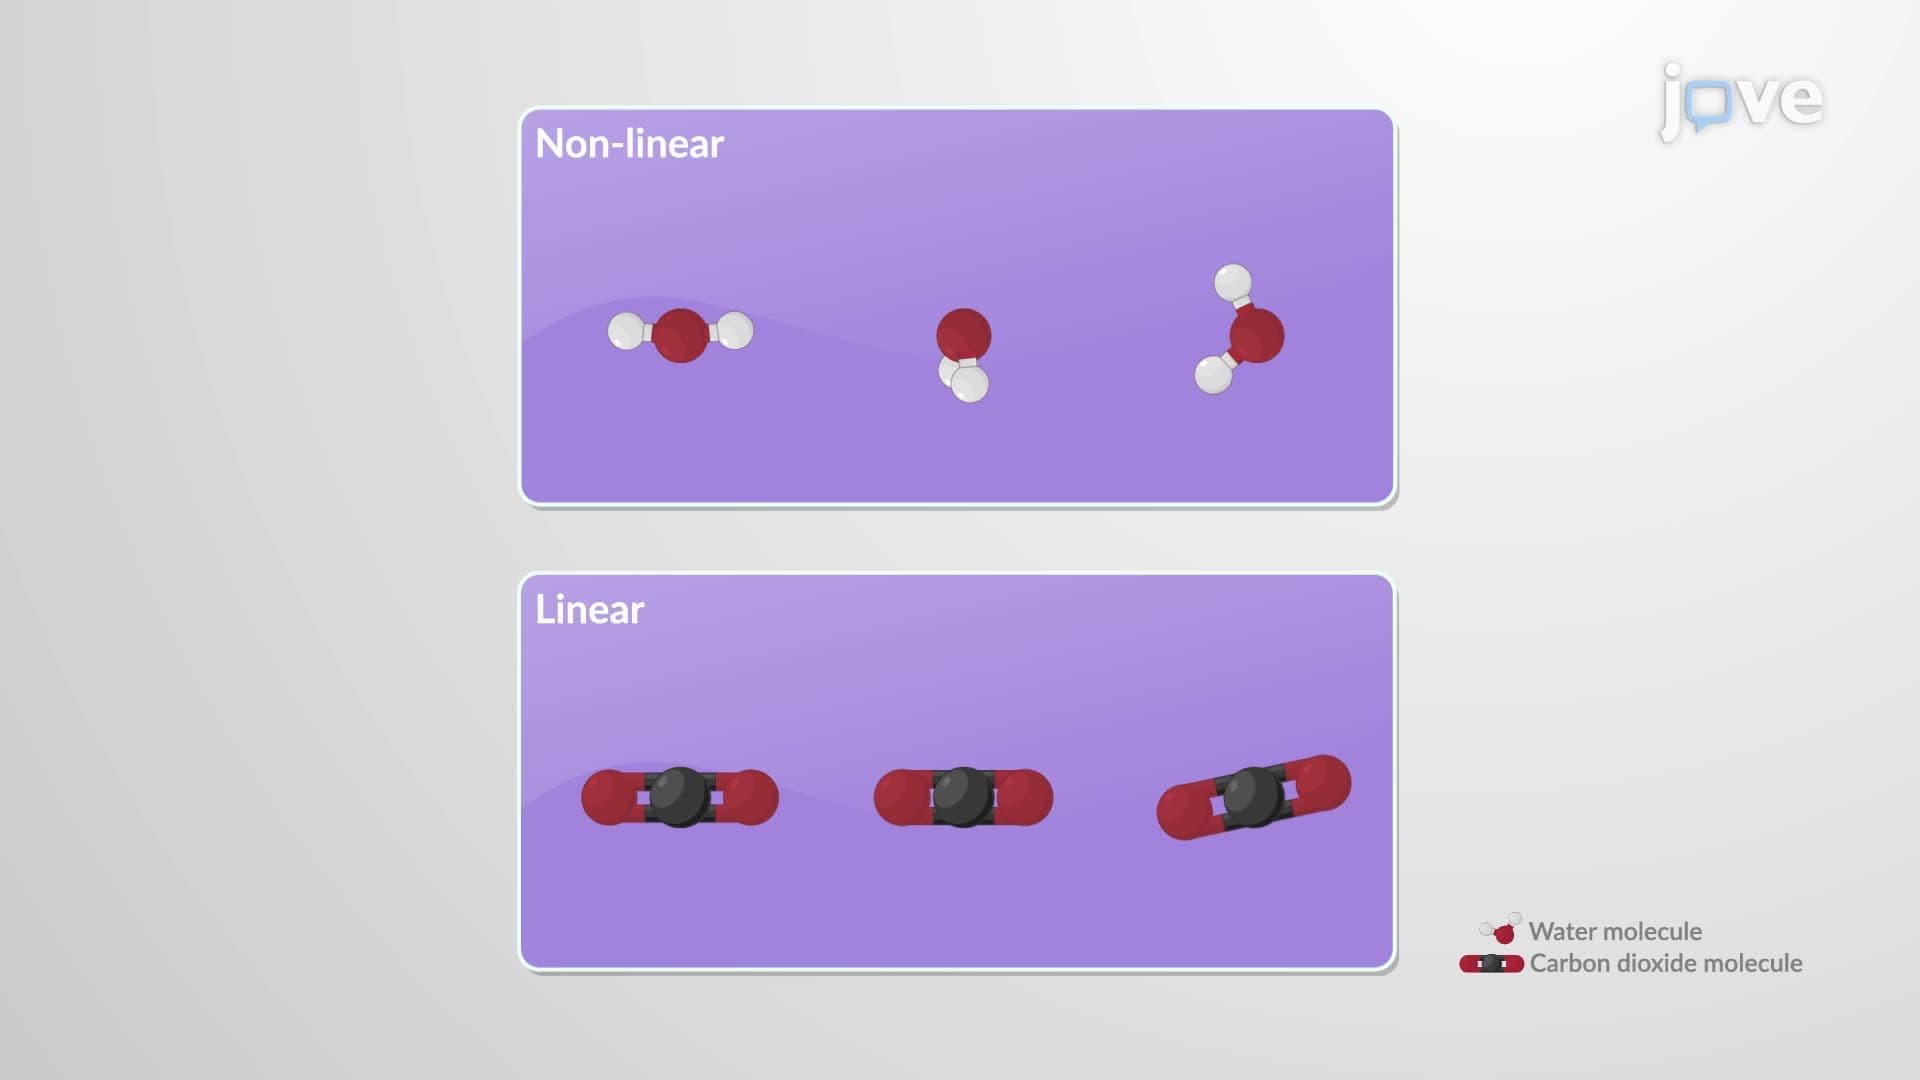 01:27
The Molecular Nature of Internal Energy