Related Experiment Video
Updated: Apr 22, 2026

Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Machine-Learned Electrostatic Potentials for Accurate Hydration Free Energy Calculations
Mathias Hilfiker1,2, Leonardo Medrano Sandonas3, Alexandre Tkatchenko1
1Department of Physics and Materials Science, University of Luxembourg, Luxembourg CityL-1511, Luxembourg.

Abstract:
Free energy calculations are widely used tools in computational chemistry, but their dependence on the assignment of partial charges during force field parametrization reduces their accuracy and reproducibility. In this work, we highlight the direct connection between the low accuracy of AM1-BCC charges on polar species and the poor accuracy of corresponding hydration free energy calculations. We then propose an XGBoost regressor trained on atomic descriptors to rapidly predict charges obtained with high-fidelity density functional theory calculations at PBE0-D3(BJ)/def2-TZVP level. The more accurate electrostatic description results in more reliable free energy calculations than those obtained with semiempirical AM1-BCC charges. Finally, we leverage this predictive model in combination with a 1 ns gas-phase molecular dynamics simulation to propose the Boltzmann Percentile method for assigning charges representative of the conformational ensemble of a molecule. Charges obtained with this method are robust to different input conformations, and the resulting free energies, calculated on a subset of the FreeSolv data set, show a root mean squared error of 1.69 kcal/mol against the 3.05 kcal/mol obtained with semiempirical charges as well as a significantly better ranking. We provide a trained model, easily integrable into traditional workflows, enabling free energy calculations at the same computational cost as AM1-BCC charges. These two aspects make it a realistic tool for enhancing already expensive free energy calculations, and more in general, molecular dynamics simulations in condensed phase.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:42
Aqueous Solutions and Heats of Hydration
When ionic compounds dissolve in water, the ions in the solid separate and disperse uniformly throughout the solution because water molecules surround and solvate the ions, reducing the strong electrostatic forces between them. This process...
 01:27
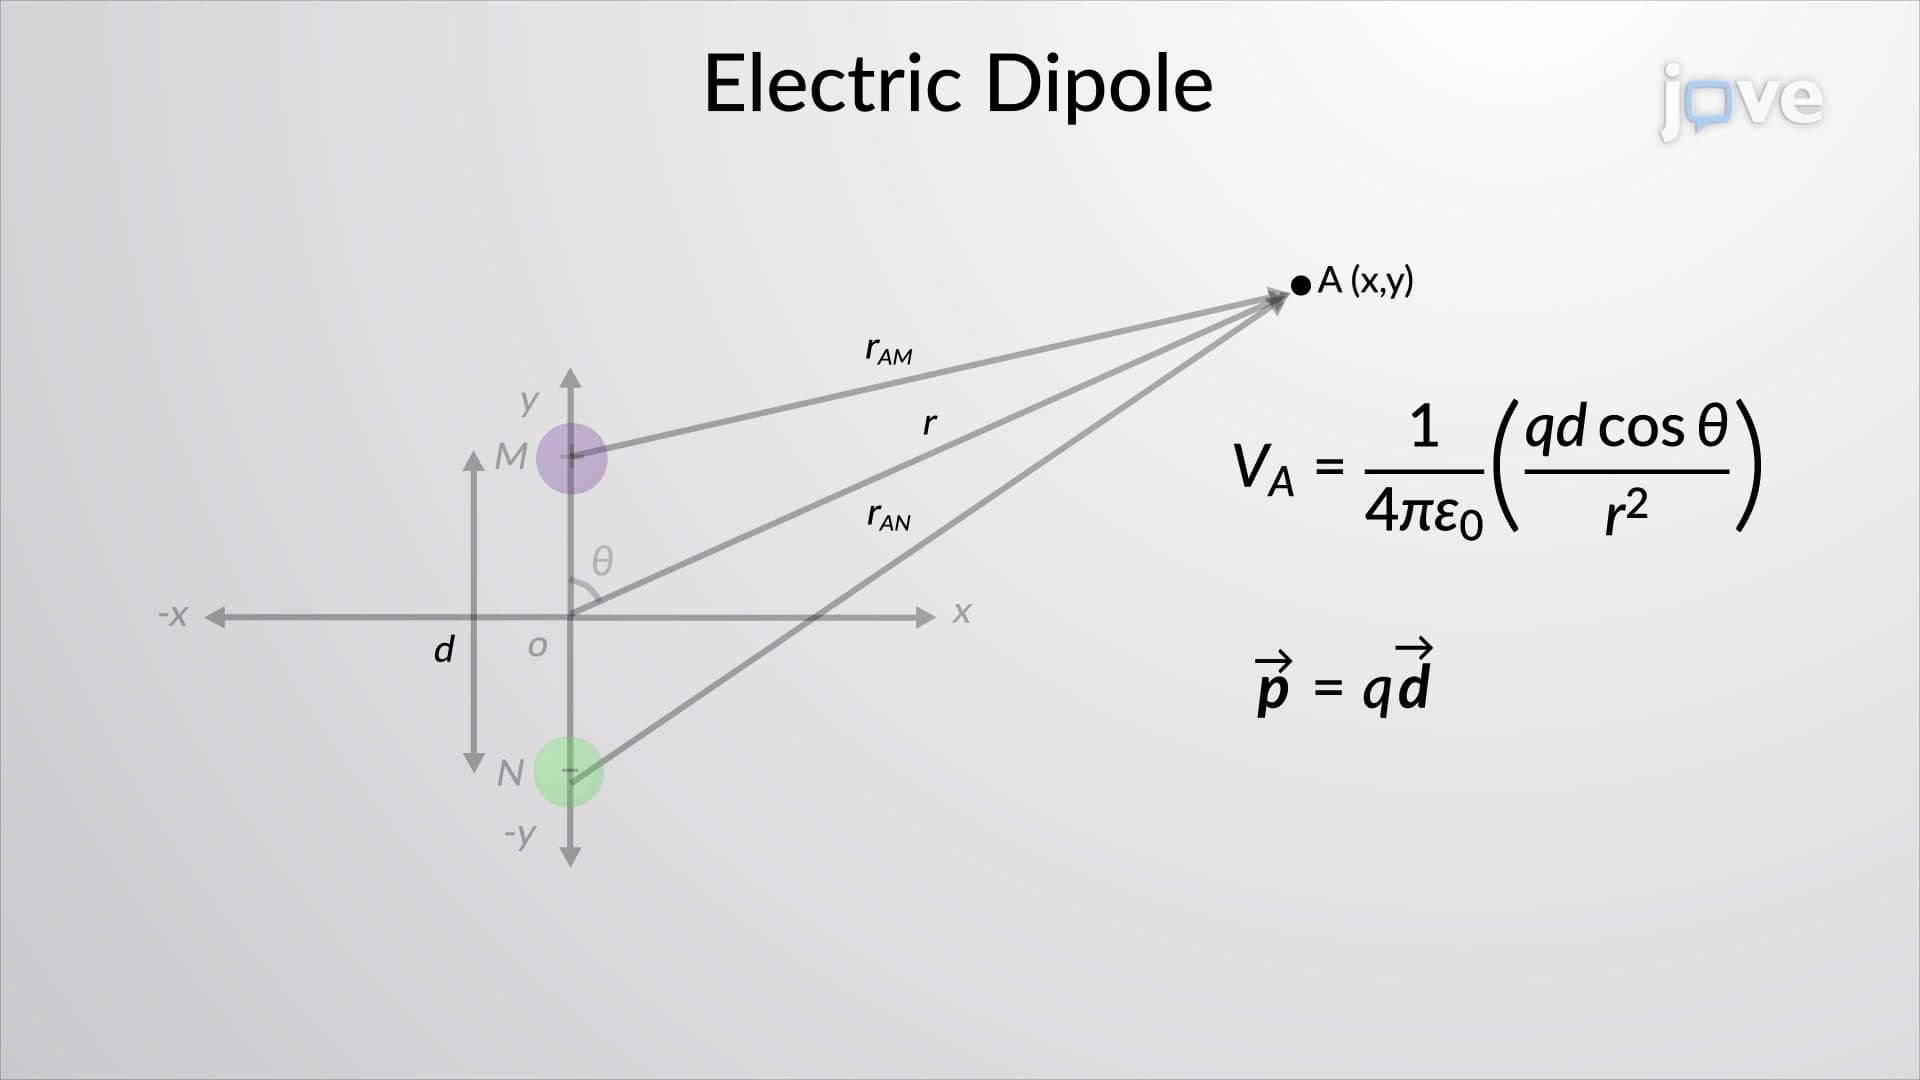
Calculations of Electric Potential II
Consider a...
 01:27
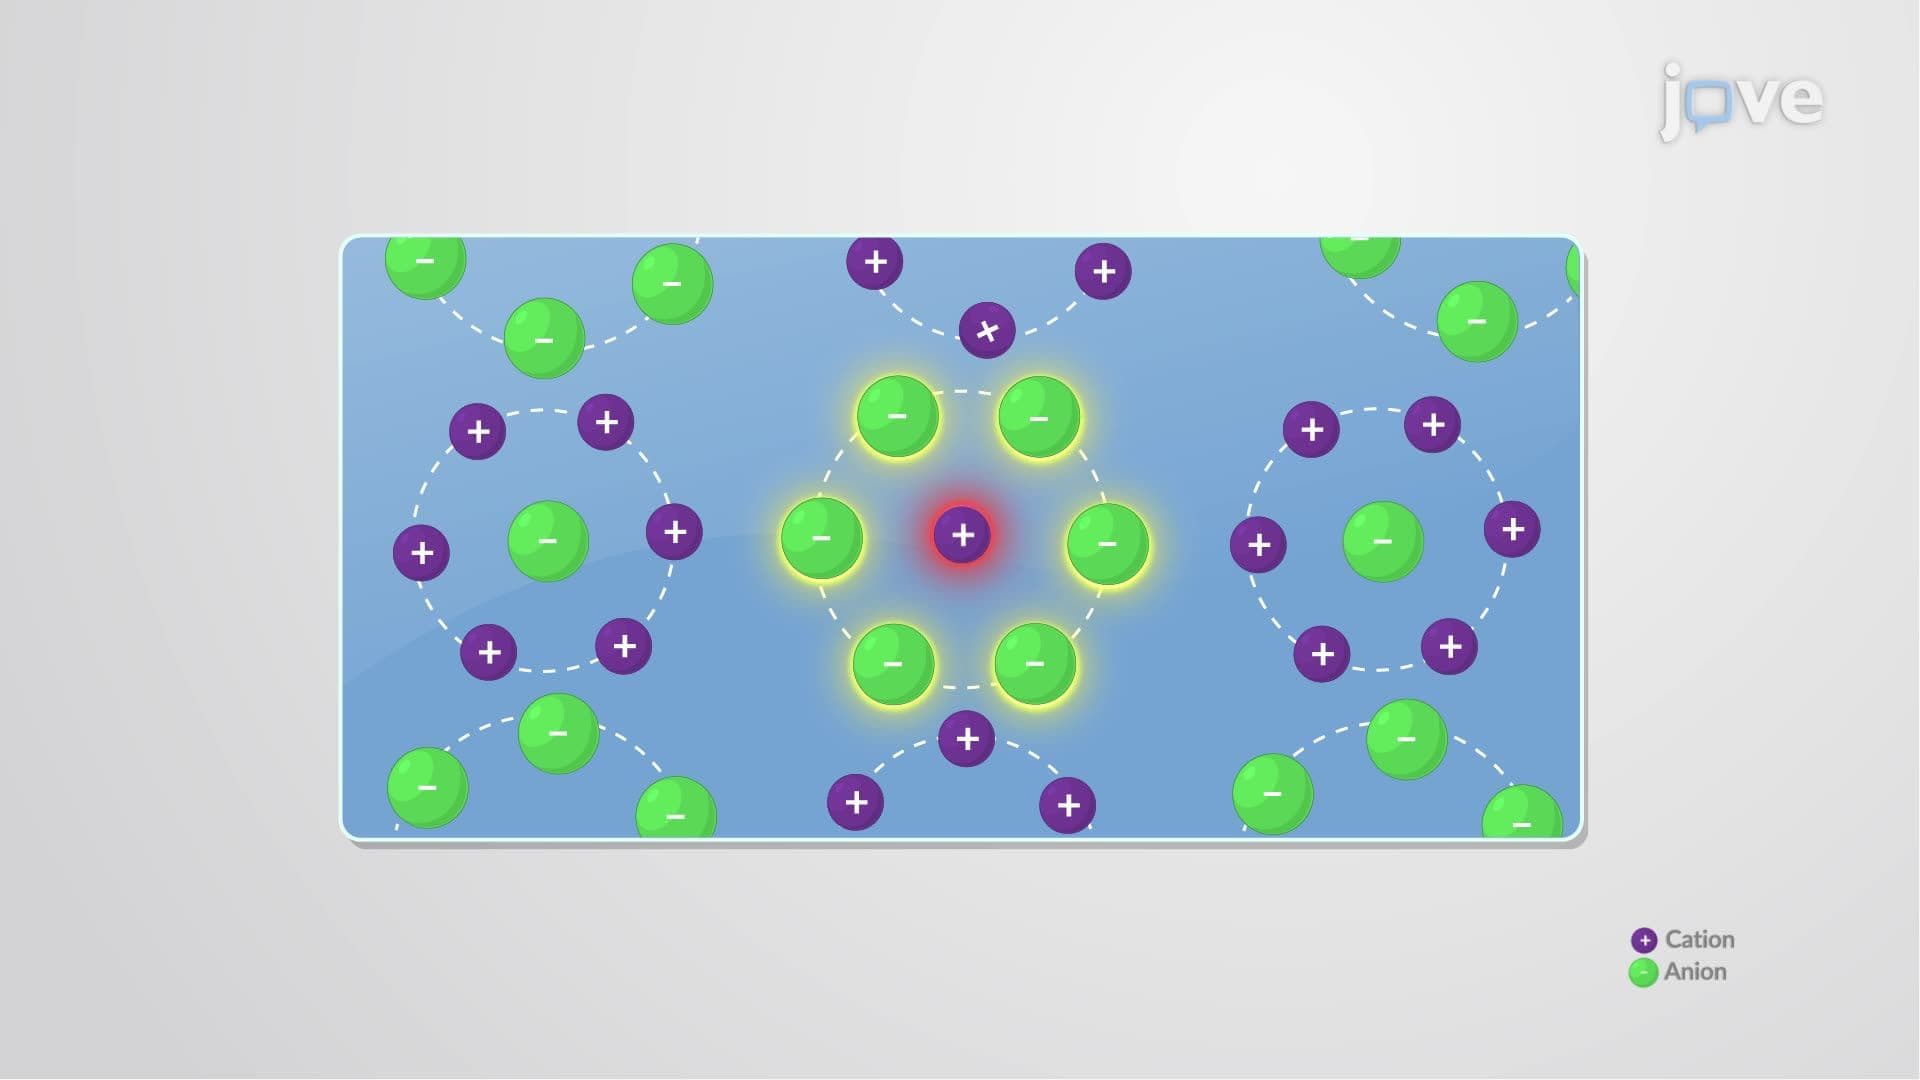
The Debye–Hückel Theory of Electrolyte Solutions
 01:30
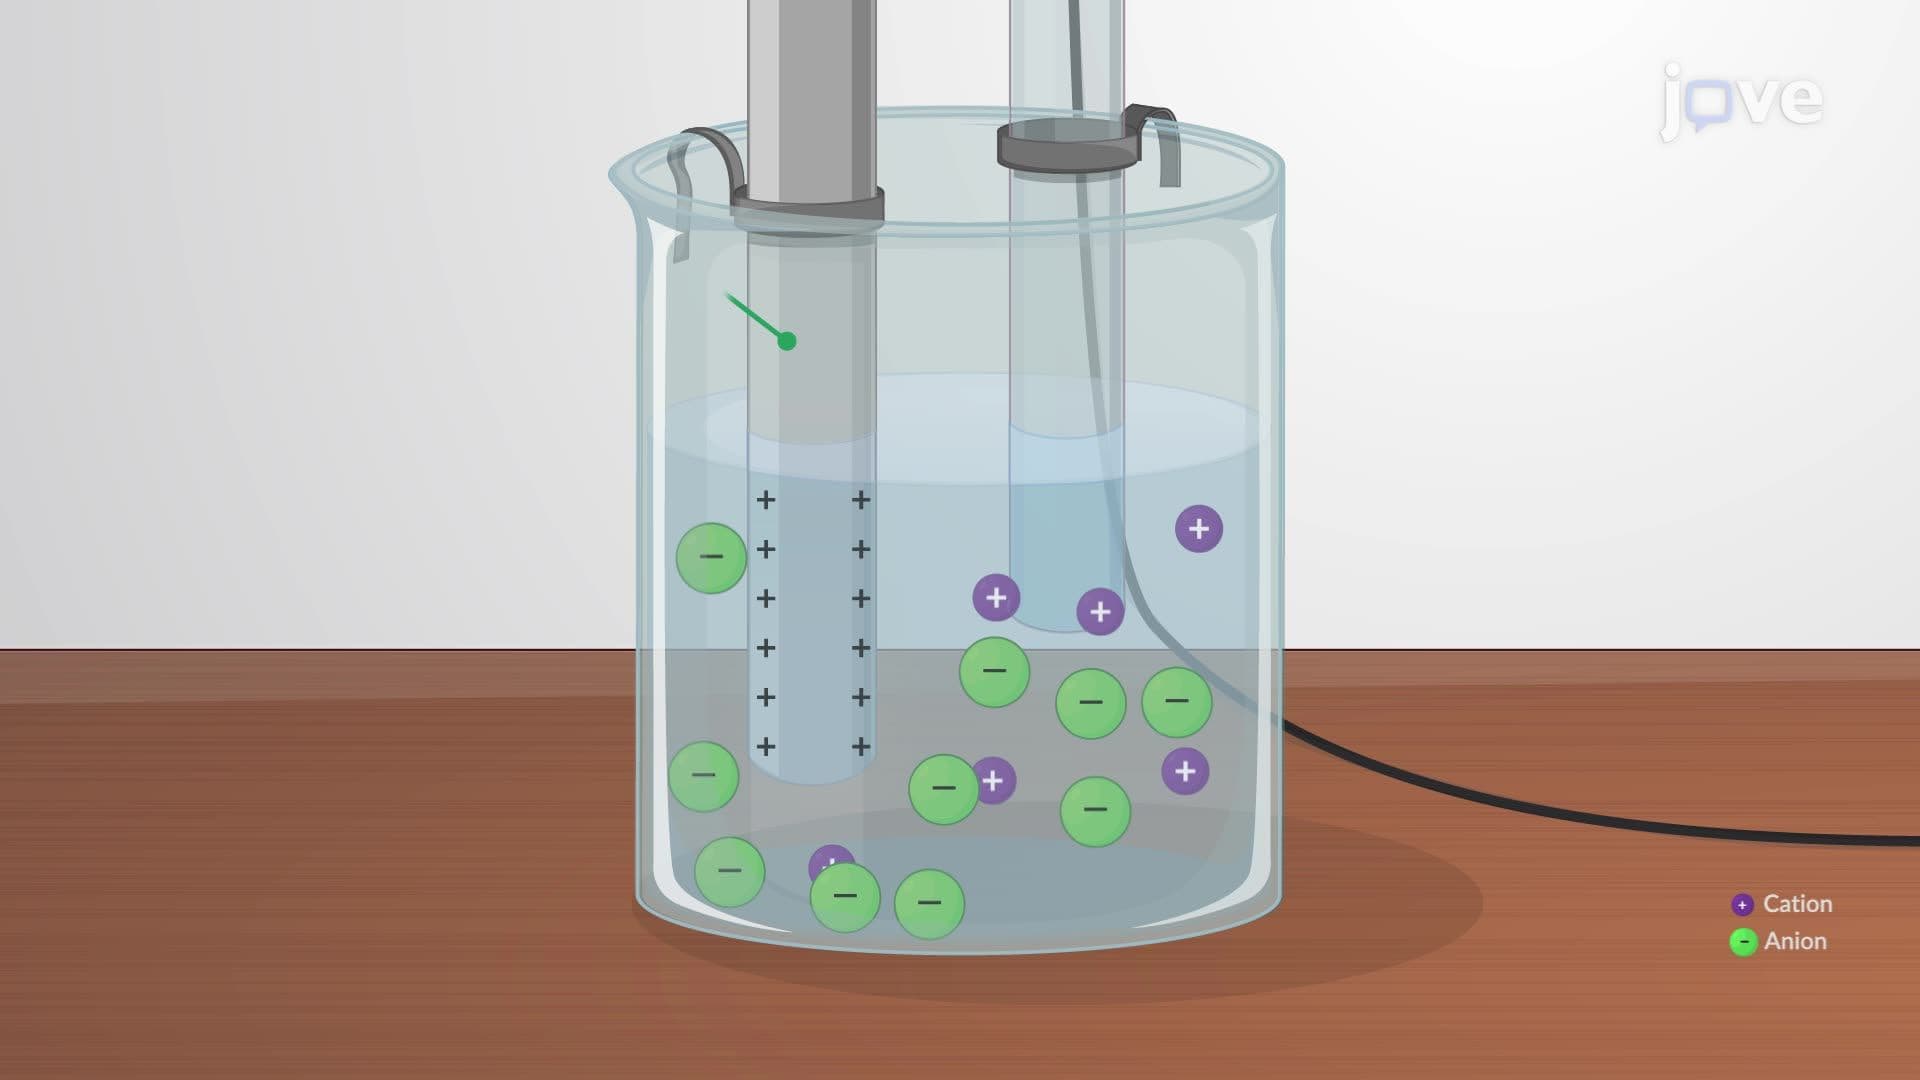
The Electrical Double Layer
 03:13
Intermolecular Forces
 01:26
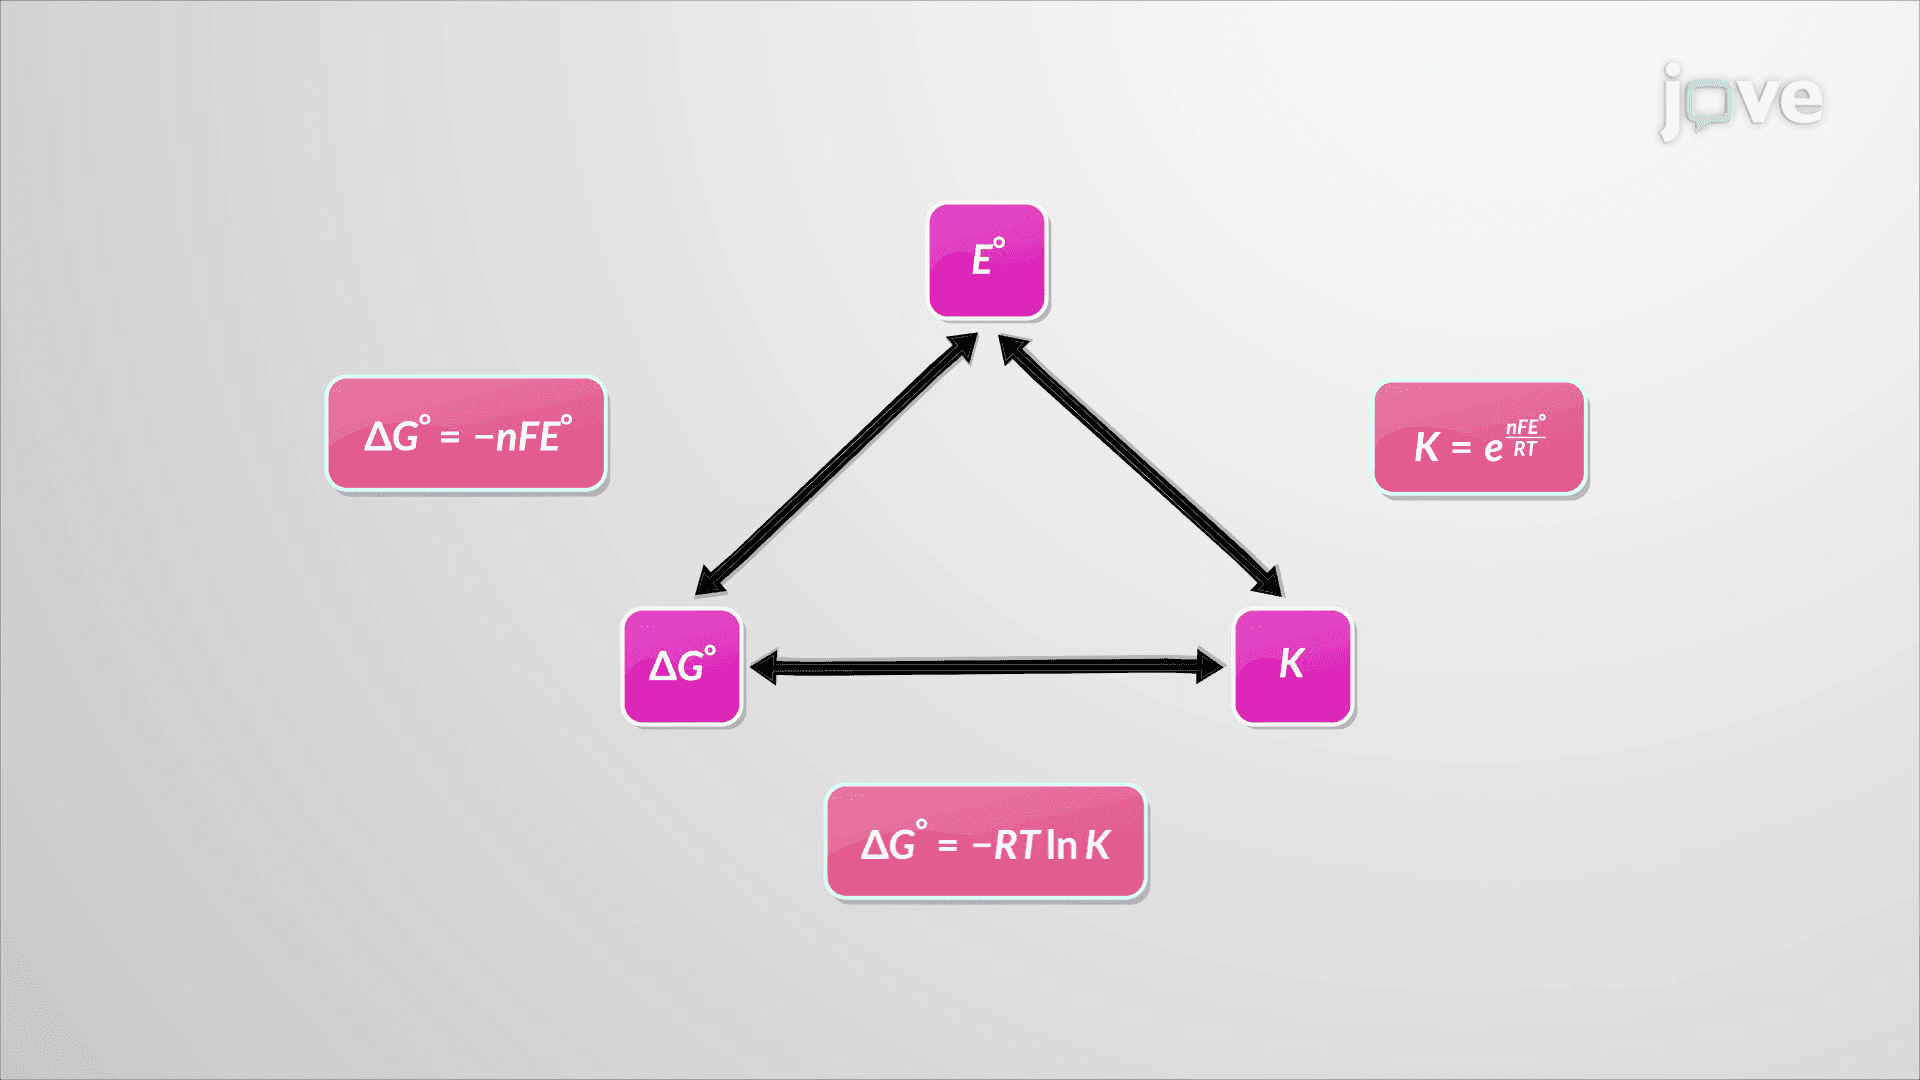
Applications of EMF Measurements