First principles simulations of transition metals for diodes: challenges and approaches to overcome inaccuracies in calculations
Related Experiment Video
Updated: May 8, 2026
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Marina Petroselli1, Elaheh Mohebbi1, Eleonora Pavoni1
1Department of Science and Engineering of Matter, Environment and Urban Planning, Polytechnic University of Marche Ancona Italy e.laudadio@staff.univpm.it.

Abstract:
The electronic, optical, and mechanical properties of transition metals belonging to groups 4B (Cr, Mo, and W), 8B (Ni, Pd, and Pt), and 9B (Cu, Ag, and Au) were investigated using Density Functional Theory (DFT). Three exchange-correlation functionals PBE, R2SCAN, and HSE06 were used and compared to evaluate their capability in prediction performance based on different physical properties. The results obtained from the density of states (DOS) calculations revealed a strong agreement between PBE and R2SCAN, while HSE06 introduced significant shifts in the valence and conduction regions due to the inclusion of the exact Hartree-Fock exchange. The optical properties of these metals highlighted that PBE and R2SCAN have reproduced the experimental curves with higher accuracy for metals with filled or nearly filled d bands, whereas HSE06 performs better for metals with more localized d states. The mechanical properties were considered in terms of Young's modulus; the outcomes revealed that metals which did not exhibit electronic d promotions possess higher values of the elastic modulus. This work highlights how electronic structure prediction, bond hybridization, and electronic promotion determine the properties of the considered metallic elements, highlighting the strengths and weaknesses of the three computational approaches and suggesting guidelines for obtaining reliable simulations for pure metal systems.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 11:33
All-electronic Nanosecond-resolved Scanning Tunneling Microscopy: Facilitating the Investigation of Single Dopant Charge Dynamics
Published on: January 19, 2018
Related Concept Videos
 01:14
Modeling of Diode Reverse Characteristics
When a reverse voltage applied to a Zener diode exceeds its breakdown voltage, the diode enters the breakdown region. At this point, the...
 01:19
Modeling of Diode Forward Characteristics
 01:22
Semiconductors
Metals such as copper (Cu), zinc (Zn), or lead (Pb) have low resistivity and feature conduction bands that are either not fully occupied or overlap with the valence band, making a bandgap non-existent. This allows electrons in the highest energy levels of the valence band to easily transition to the conduction band upon gaining...
 01:27
Biasing of Metal-Semiconductor Junctions
In Schottky junctions, where the semiconductor is n-type, applying a positive voltage to the metal relative to the semiconductor reduces its Fermi...
 01:24
Metal-Semiconductor Junctions
Schottky Barriers
Schottky barriers arise when a metal with a work function (Φm) contacts a semiconductor with a different work function (Φs). Initially, electrons transfer until the Fermi levels of the metal and semiconductor align at equilibrium. For instance, if Φm > Φs, the semiconductor Fermi level is higher than the metal's before contact. The semiconductor's...
 01:18
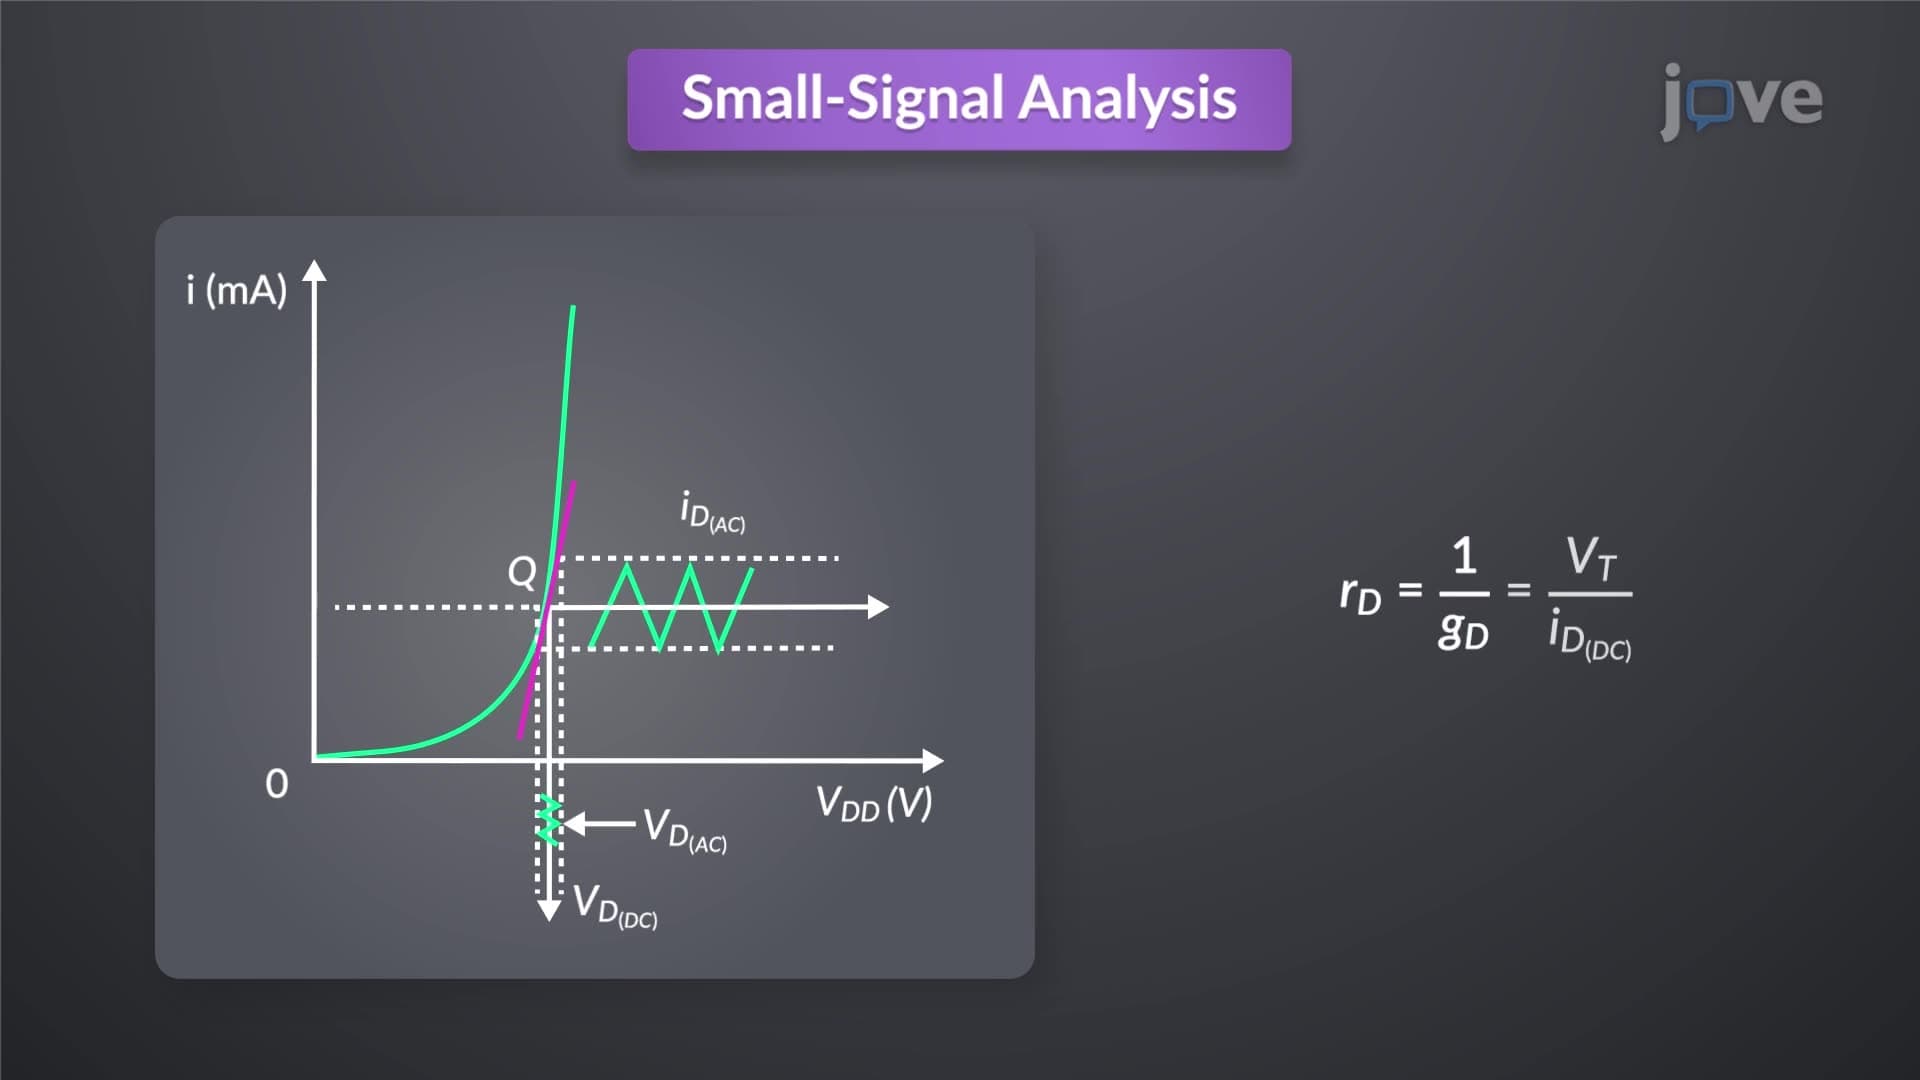
Small-signal Diode Model