Related Experiment Video
Updated: May 21, 2026
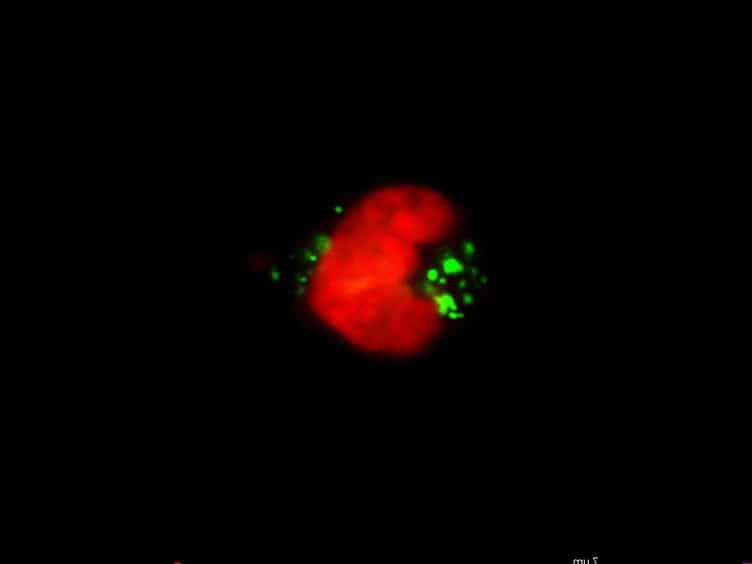

Cell Death Associated with Abnormal Mitosis Observed by Confocal Imaging in Live Cancer Cells
Published on: August 21, 2013
MAP4Ks drive cell death in response to Salmonella SpvB-induced actin depolymerization
Mitchell A Pallett1, Romina Tocci1, Andrea Majstorovic1
1Centre for Bacterial Resistance Biology, Department of Infectious Disease, Faculty of Medicine, Imperial College London, London, United Kingdom.

Abstract:
Many pathogens target the host actin cytoskeleton through the delivery of actin depolymerizing toxins, including mono-ADP-ribosyltransferases (mART), ultimately triggering host cell death. Despite the importance of mARTs in pathogen virulence, it remains unclear whether actin ribosylation is required for mART-dependent cell death, and how actin depolymerization leads to cell death. Using the non-typhoidal Salmonella enterica Typhimurium-encoded mART, SpvB, we report that cell death is induced exclusively through ribosylation of actin. We found cell death to be morphologically and mechanistically distinct from apoptosis as well as any previously reported mode of cell death. Instead, our data identify the Hippo signaling MAP4Ks as the essential host cell sensors of actin depolymerization signaling through JNK to facilitate vacuolization and host cell death. Cell death following treatment of cells with the actin depolymerizing agent latrunculin A followed the same pathway, identifying a conserved mechanism of cell death. Therefore, we identify MAP4K family members as key regulators of an atypical caspase-independent cell death induced by actin depolymerization, building on our understanding of host-cell death signaling and mechanisms of bacterial virulence.
Importance:
Host cell death plays a critical role as an intrinsic defense mechanism against infection and disease. However, many pathogens subvert cell death signaling to enhance their replication and survival. Here, we show that the mono-ADP ribosyl transferase family of toxins encoded by pathogens of global importance, including Salmonella spp., Neisseria spp. and C. difficile induces actin depolymerization leading to MAP4K activation and JNK-dependent cell death. Through mechanistically characterizing this atypical cell death pathway, our study identifies and positions key components of a previously undescribed cell death pathway and broadens our understanding of bacterial pathogenesis and virulence.
Insights
Pathogenic toxins that depolymerize actin trigger a novel cell death pathway. This atypical, caspase-independent cell death is regulated by MAP4K signaling, offering insights into bacterial virulence.
Area of Science:
- Microbiology
- Cell Biology
- Immunology
Background:
- Pathogens utilize toxins to disrupt host actin cytoskeleton, often inducing cell death.
- Mono-ADP-ribosyltransferases (mARTs) are key virulence factors, but their precise role in cell death remains unclear.
- Host cell death is a defense mechanism, yet pathogens can manipulate it for survival.
Purpose of the Study:
- To investigate the role of actin ribosylation in mART-induced cell death.
- To elucidate the mechanism by which actin depolymerization leads to host cell death.
- To identify host factors involved in sensing actin depolymerization and mediating cell death.
Main Methods:
- Utilized *Salmonella enterica* Typhimurium and its mART SpvB.
- Investigated cell death pathways using biochemical and cell imaging techniques.
- Administered actin depolymerizing agent latrunculin A to compare pathways.
Main Results:
- Actin ribosylation by SpvB is essential for inducing cell death.
- Identified a novel cell death pathway involving MAP4K activation and JNK signaling.
- Demonstrated that this pathway is distinct from apoptosis and conserved with latrunculin A treatment.
- Observed vacuolization as a feature of this atypical cell death.
Conclusions:
- MAP4K family members are critical sensors of actin depolymerization.
- This identifies a conserved, atypical, caspase-independent cell death pathway.
- Findings enhance understanding of bacterial pathogenesis and host cell death signaling.
More Related Videos
 08:51
Applying an Inducible Expression System to Study Interference of Bacterial Virulence Factors with Intracellular Signaling
Published on: June 25, 2015
 11:38
Application of Fluorescent Nanoparticles to Study Remodeling of the Endo-lysosomal System by Intracellular Bacteria
Published on: January 2, 2015
