Related Experiment Video
Updated: May 25, 2026

In Vitro Model for Studying Differentiation and Changes of Multi-Omics on Murine Airway Epithelial Cells Stimulated with Cigarette Smoke Extract
Published on: July 12, 2024
Rab26 mitigates ferroptosis in airway epithelium induced by cigarette smoke through suppression of VDAC1
Xin Tian1, Wen Zhang2, Zaichun You1
1Department of General Practice, Xinqiao Hospital, Third Military Medical University (Army Medical University), Chongqing, 400037, China.

Background:
Cigarette smoke (CS)-induced epithelial ferroptosis is a pivotal driver of airway inflammation and remodeling in chronic obstructive pulmonary disease (COPD). Although Rab26 is known to protect against inflammatory injury, the precise molecular mechanism linking Rab26 to mitochondrial integrity and ferroptotic cell death remains largely unexplored. This study aims to elucidate a novel regulatory axis involving Rab26, mitochondrial voltage-dependent anion channel 1 (VDAC1), and the cGAS-STING pathway in the pathogenesis of COPD.
Methods:
A COPD mouse model was established using Rab26-deficient (Rab26-/-) mice exposed to cigarette smoke, alongside wild-type mice treated with a pharmacological STING inhibitor to validate pathway involvement. Ferroptosis and inflammation were assessed in vitro and in vivo. Crucially, we employed co-immunoprecipitation (Co-IP), chemical cross-linking, and transmission electron microscopy (TEM) to investigate the molecular interaction between Rab26 and VDAC1, and to monitor VDAC1 oligomerization and mitochondrial DNA (mtDNA) release.
Results:
We observed significantly downregulated Rab26 expression in lung tissues from COPD patients and CS-exposed mice. In vivo, Rab26 deficiency markedly exacerbated airway inflammation, emphysema, and epithelial ferroptosis. Mechanistically, we identified a novel interaction wherein Rab26 physically binds to VDAC1. Under CS exposure, loss of Rab26 promotes VDAC1 oligomerization, leading to mitochondrial membrane permeabilization and the leakage of mtDNA into the cytosol. This cytosolic mtDNA subsequently triggers the aberrant hyperactivation of the cGAS-STING pathway, which represses the antioxidant SLC7A11/GPX4 axis and drives lethal ferroptosis. Conversely, restoring Rab26 or blocking STING signaling effectively inhibited VDAC1-mediated mtDNA release and alleviated ferroptotic injury.
Conclusions:
This study provides that Rab26 acts as a guardian of mitochondrial integrity by preventing VDAC1 oligomerization. We define a previously unrecognized Rab26-VDAC1-cGAS-STING signaling axis that regulates airway epithelial ferroptosis. Targeting this axis offers a compelling therapeutic strategy to arrest inflammation and disease progression in COPD.
Related Concept Videos
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
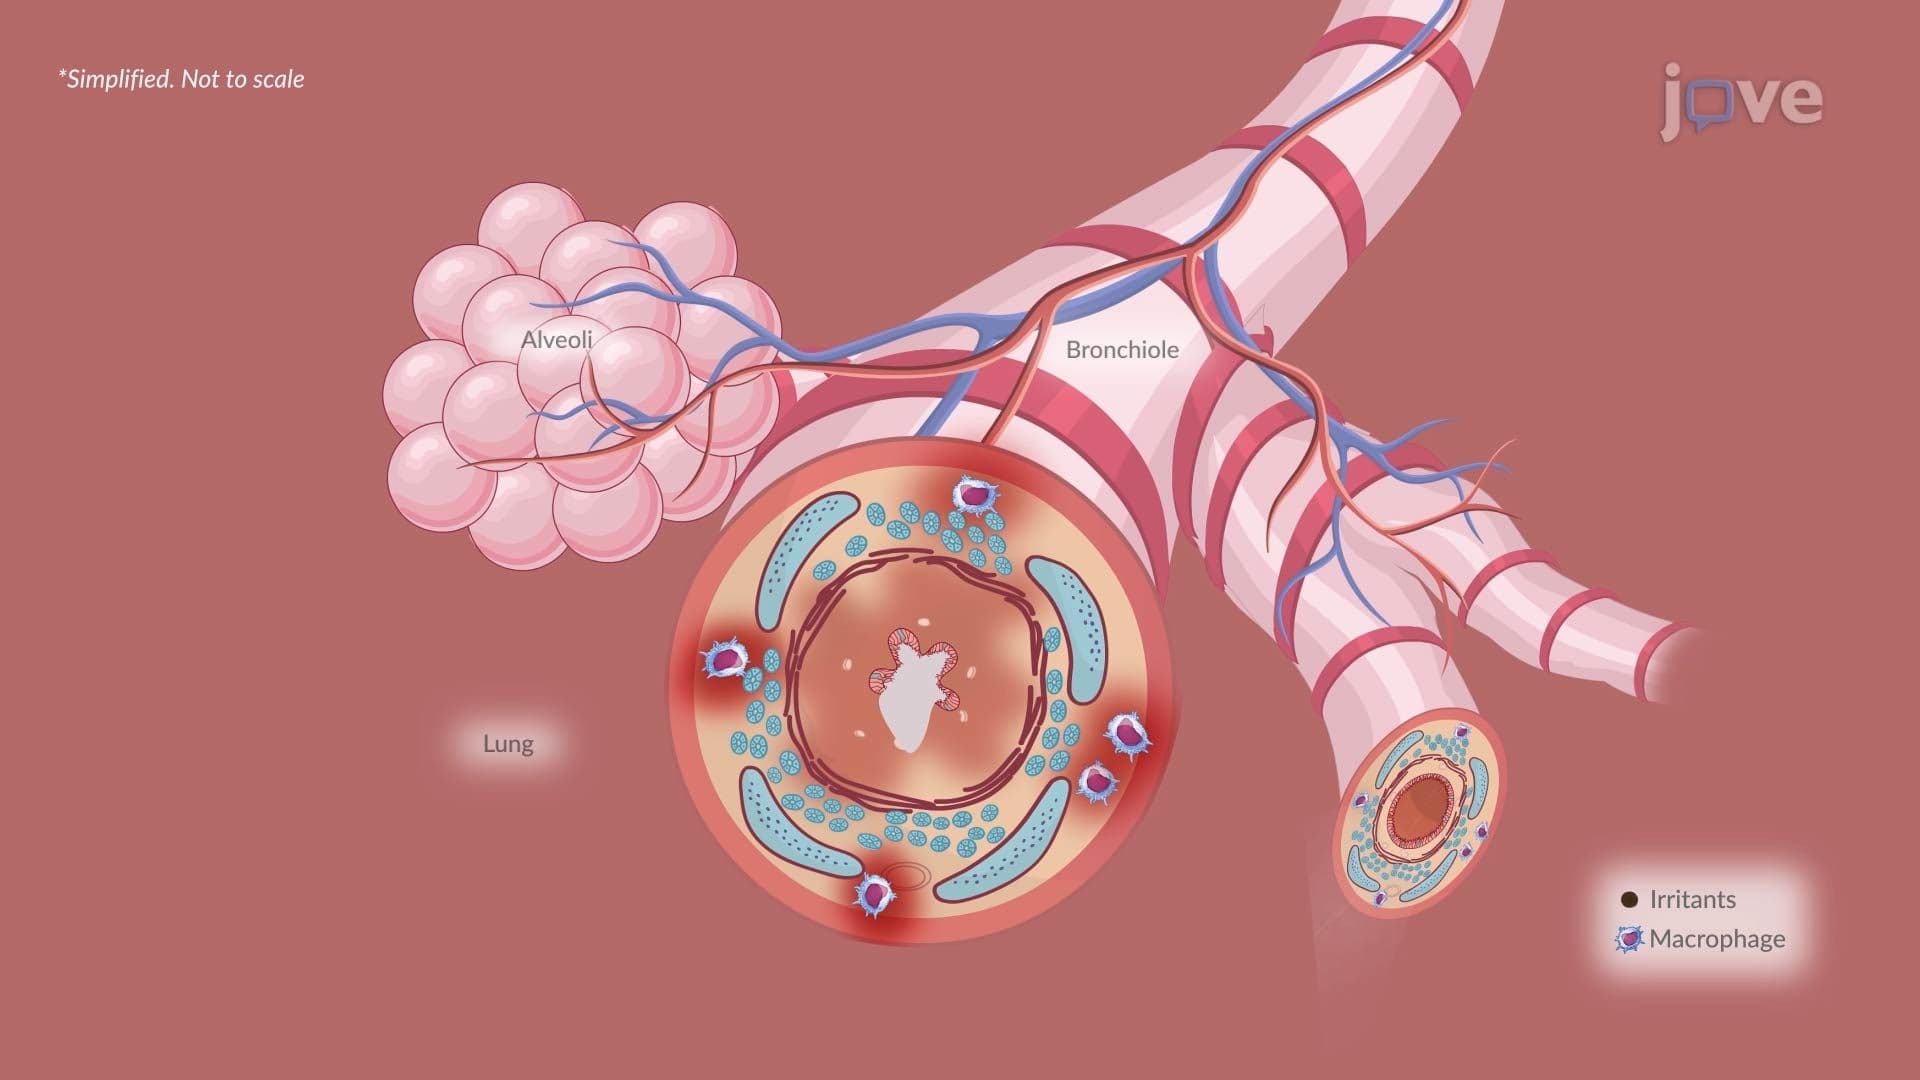
Chronic Inflammation
 01:24
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features