Related Experiment Video
Updated: Jun 12, 2026

Efficient Construction of Drug-like Bispirocyclic Scaffolds Via Organocatalytic Cycloadditions of α-Imino γ-Lactones and Alkylidene Pyrazolones
Published on: February 7, 2019
Predicting enantiomer migration order of levobunolol via sequential computational modeling
Pollyanna Pinto Maia1, Rafaela Maia Della-Sávia Freitas1, Luciana Guimarães1
1GPQCA: Grupo de Pesquisa em Química Computacional Aplicada, Departamento de Ciências Naturais (DCNAT), Universidade Federal de São João Del-Rei (UFSJ), Campus Dom Bosco, São João Del Rei, 36301-160, Minas Gerais, Brazil.

Context:
A detailed molecular-level rationale for the enantioseparation of the β-blocker levobunolol (BUN) by substituted β-cyclodextrins (β-CDs) is presented through high-level computational modeling. The calculations consistently reveal that the therapeutically active (+)-[S]-BUN enantiomer forms more stable inclusion complexes than the (-)-[R]-enantiomer with both carboxymethyl-β-CD (CM-β-CD) and sulfated-β-CD (SF-β-CD) selectors. Non-covalent interaction (NCI) reveals that the superior stability of the S-enantiomer arises from a more continuous dispersive interaction envelope and a more compact network of electrostatic and hydrogen bonds. For the most effective selector, SF-β-CD, the complexation Gibbs free energy (ΔG) in an aqueous medium is -29.5 kcal/mol for (+)-[S]-BUN versus -17.1 kcal/mol for (-)-[R]-BUN. This results in a large free energy difference (ΔΔG) of 12.4 kcal/mol, indicating exceptional enantioselectivity. The NCI isosurfaces also confirm that the S-isomer achieves optimal stereoelectronic complementarity with the host's sulfate and hydroxyl groups, showing lower steric penalties than the R-isomer. These findings provide a robust prediction of a longer electrophoretic migration time for the (+)-[S]-enantiomer and validate SF-β-CD as a highly efficient chiral selector for BUN, underscoring the power of in silico methods to elucidate complex chiral recognition mechanisms.
Methods:
Semiempirical geometry, frequency, non-covalent, 2nd version, eXtended Tight Binding (GFN2-xTB) and Density Functional Theory (DFT) (ωB97X-D3/6-31G(d,p) and ωB97X-D3/6-311 + G(d,p)) methods were employed in both gas and aqueous phases. NCI analysis was also performed. All DFT and semiempirical calculations were carried out using the ORCA 5.0 software package. The NCI analysis was carried out using the Multiwfn program.
More Related Videos
 11:04
Preparation of Enantiopure Non-Activated Aziridines and Synthesis of Biemamide B, D, and epiallo-Isomuscarine
Published on: June 13, 2022
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 02:24
Properties of Enantiomers and Optical Activity
 02:05
Prochirality
 02:30
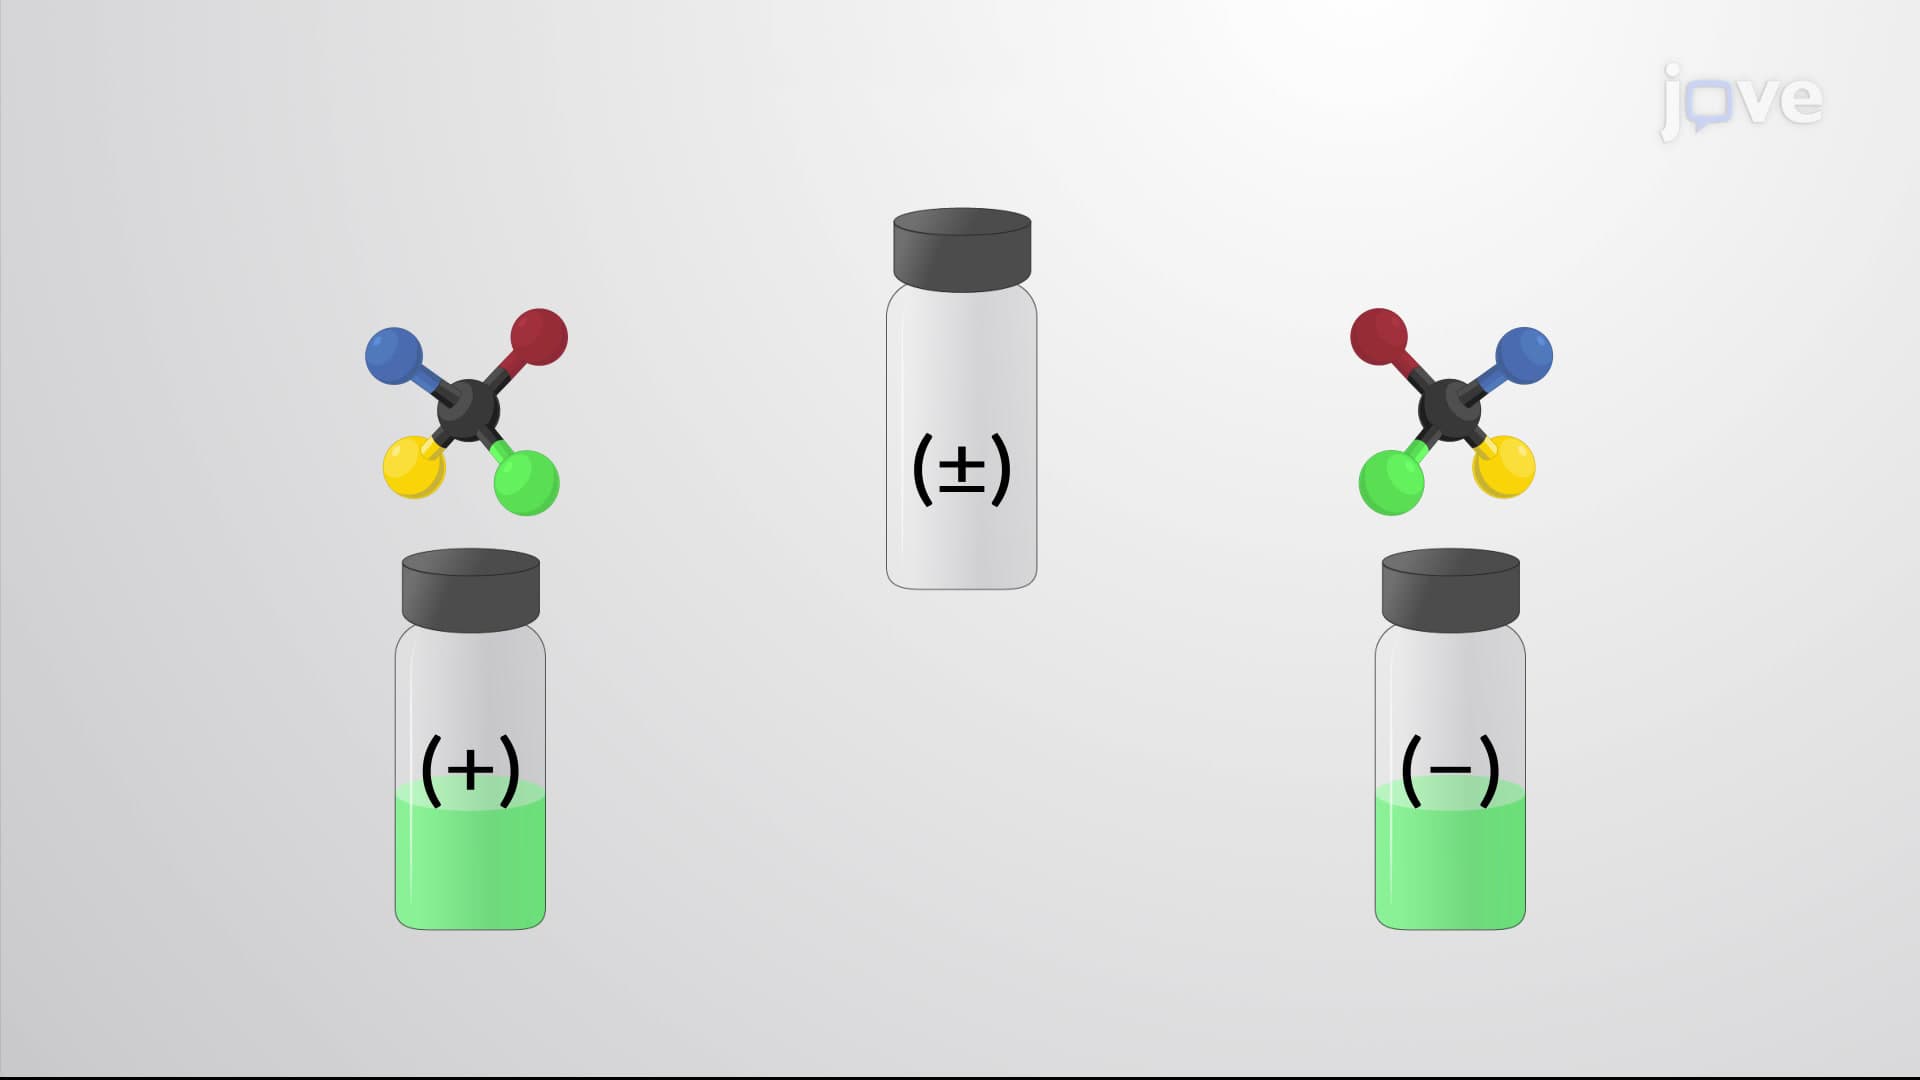
Racemic Mixtures and the Resolution of Enantiomers
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 02:21
Naming Enantiomers
 02:15
SN1 Reaction: Stereochemistry
In the first step of an SN1 reaction, the bond between the electrophilic carbon and the leaving group ionizes to generate the carbocation intermediate. The second step of the mechanism is the nucleophilic attack.
In the formed carbocation, the positively charged carbon is sp2 hybridized with a trigonal planar geometry. As all the three substituents lie on the same plane, a plane of symmetry for the...