Related Experiment Video
Updated: Jun 12, 2026

A Technical Guide for Performing Spectroscopic Measurements on Metal-Organic Frameworks
Published on: April 28, 2023
Pathological Pd-phenanthroline complex under standard DFT protocols
Artur Brotons-Rufes1, Simona Russo1, Doaa R Ramadan2,3
1Dipartimento di Chimica e Biologia "A. Zambelli", Università di Salerno, V. Giovanni Paolo II, Fisciano, SA, Italy.

Context:
The reliability of density functional theory (DFT) calculations for planar π-extended palladacycles can be strongly affected by the modelling of the surrounding chemical environment. In the present study, standard gas-phase calculations lead to an artificial folding of the complex, initially attributed to an overestimation of non-covalent interactions with aromatic additive species. Although improved dispersion treatments partially mitigate the distortion, the introduction of strongly donating ligands exacerbates the curvature even when modern D4 dispersion corrections are employed. The results indicate that the structural deformation does not originate solely from dispersion effects but rather from an artefact associated with the unrealistic empty space opposite the coordinating ligands in the gas-phase model. Importantly, the explicit inclusion of a single solvent molecule substantially reduces the curvature of the palladacycle while maintaining the same dispersion treatment. These findings highlight the importance of minimal yet chemically meaningful environmental models for obtaining physically consistent structures in computational studies of palladium complexes.
Methods:
Geometry optimisations were carried out with the Gaussian 16 package, using the BP86 functional with the Def2SVP basis set for the light atoms and SDD for the metal centre with the Stuttgart/Dresden ECPs. Furthermore, dispersion corrections were introduced with Grimme D3 and D4; ORCA 6 software was used for the latter. The energies were refined via single-point calculations on optimized structures using the Def2TZVPP with diffuse basis set for the electronegative atoms, testing dispersion effects on M06 and MN15 functionals, with the double hybrid B2PLYPD3 as reference. Solvent effects were introduced via implicit solvent model (PCM, toluene).
Related Concept Videos
 01:20
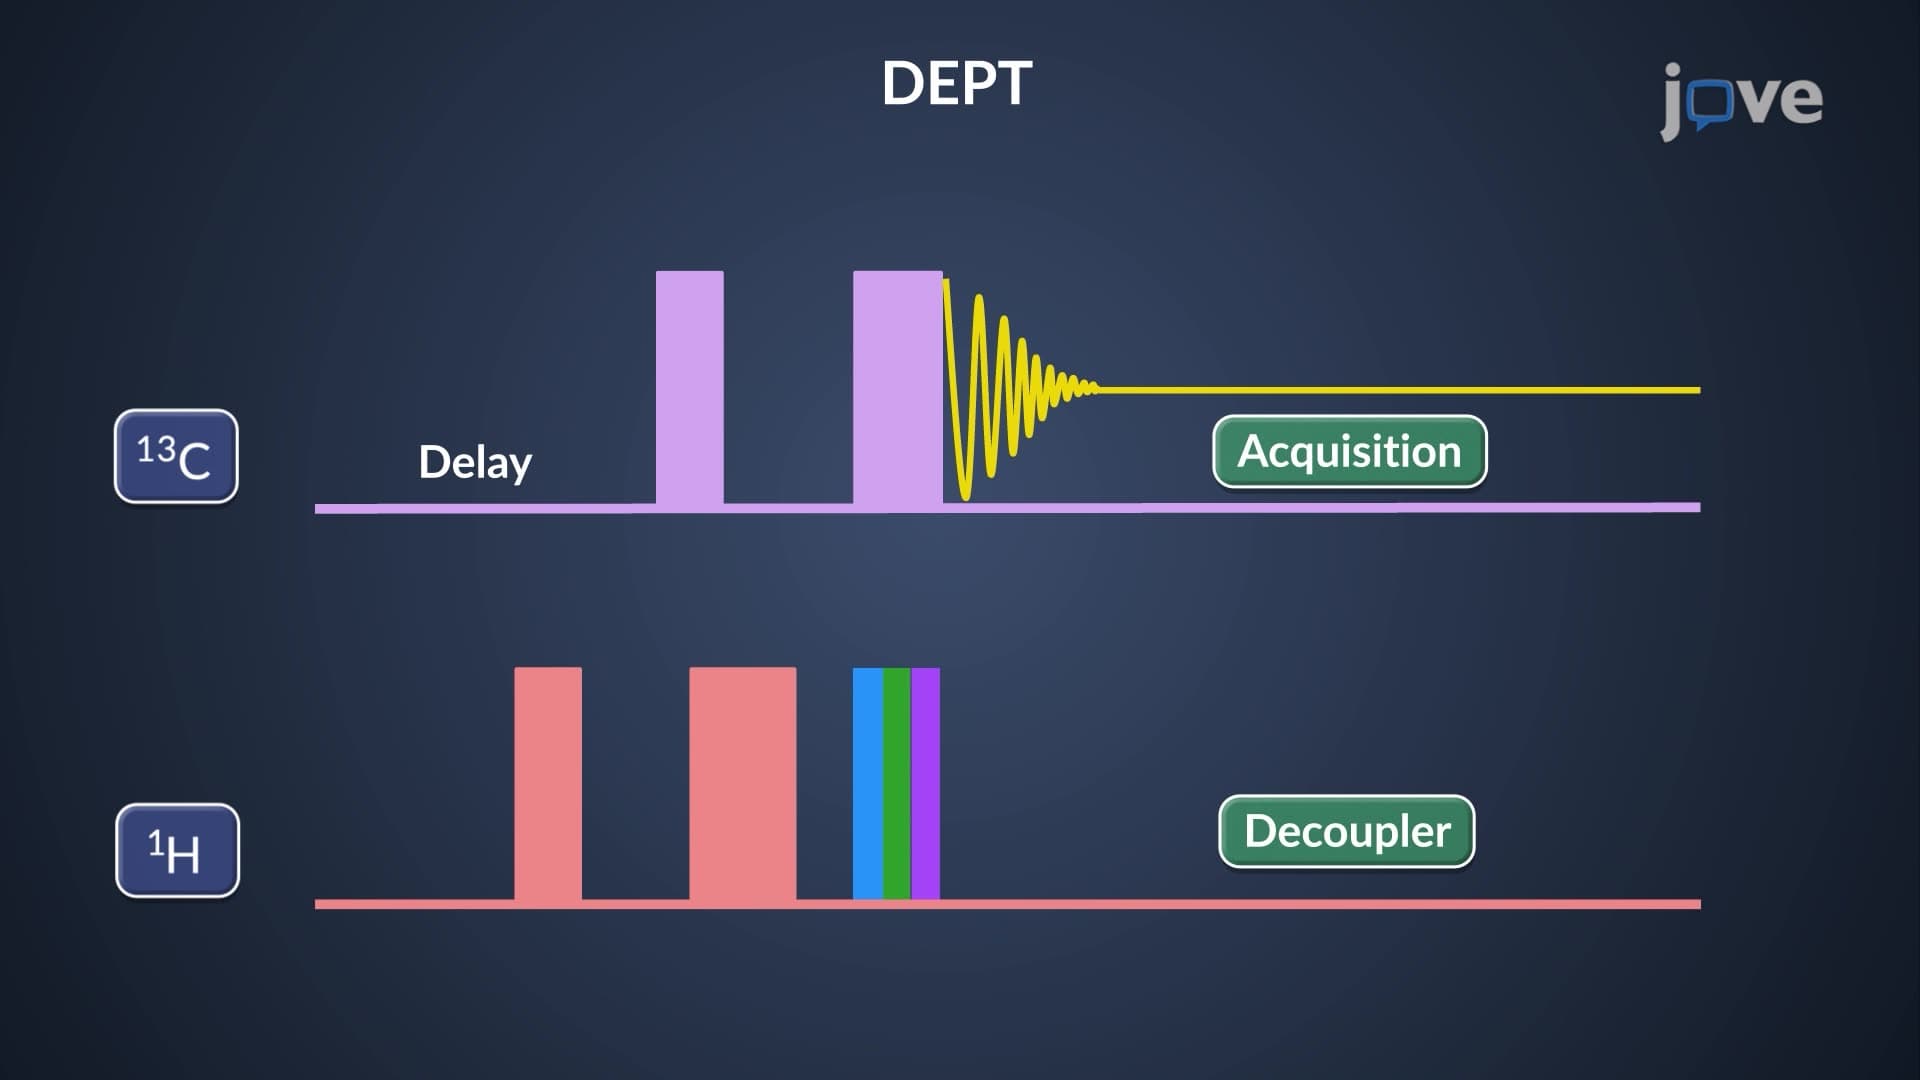
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory