Related Experiment Video
Updated: Jul 15, 2026

Identification of Antibacterial Immunity Proteins in Escherichia coli using MALDI-TOF-TOF-MS/MS and Top-Down Proteomic Analysis
Published on: May 23, 2021
Comparative genomic analysis of Bacteroides fragilis from intestinal and extra-intestinal sites
Renee E Oles1,2, Marvic Carrillo Terrrazas1, Luke R Loomis1
1Department of Pathology, University of California, San Diego, California, USA.

Abstract:
Bacteroides fragilis, a key member of the human gut microbiota, contributes to host health by maintaining intestinal homeostasis. Yet, it is also the most frequently isolated anaerobe in clinical infections. These contrasting roles raise questions about the genetic and ecological factors that explain why this common symbiont is disproportionately linked to infection. We analyzed 813 Division I B. fragilis genomes, including 147 new isolates from intestinal and extra-intestinal sites. Infection-associated isolates spanned all phylogroups, indicating no pathogenic lineage. We identified 16 phylogroups, distinguished by genes associated with capsule biosynthesis and interbacterial competition. Additionally, differential metabolomic analysis identified 12 metabolites associated with isolation source, while a microbial genome-wide association study uncovered 44 genes enriched in isolates from extra-intestinal sites, providing the first population-scale markers tied to clinical recovery sites. These results do not implicate a pathogenic lineage; instead, they point to associational links between accessory modules and recovery from extra-intestinal sites under permissive host conditions. This work underscores how genomic diversity and ecological context may jointly shape the clinical impact of gut commensals.IMPORTANCEBacteroides fragilis, a human gut resident, is paradoxically one of the most frequent anaerobes recovered from bloodstream and abscess infections. The genetic features that enable frequent recovery from extra-intestinal sites remain poorly defined. Using comparative genomic and metabolomic analyses of strains from intestinal and extra-intestinal sources, we show that strains isolated from infections are phylogenetically dispersed rather than restricted to a single lineage. We observe lineage-linked differences in capsule loci and competition systems, which suggests constrained gene flow and lineage-specific adaptation within the gut. Additionally, a subset of genes and metabolites is enriched among extra-intestinal isolates. Together, these findings suggest that extra-intestinal survival among B. fragilis strains reflects the interplay between species-wide genomic diversity and permissive host conditions, rather than the emergence of a single pathogenic lineage.
More Related Videos
 08:33
Quantitative Polymerase Chain Reaction-based Analyses of Murine Intestinal Microbiota After Oral Antibiotic Treatment
Published on: November 17, 2018
 09:44
Characterization of a Pathogenic Escherichia coli Strain Derived from Oreochromis spp. Farms Using Whole-Genome Sequencing
Published on: December 23, 2022
Related Concept Videos
 01:29
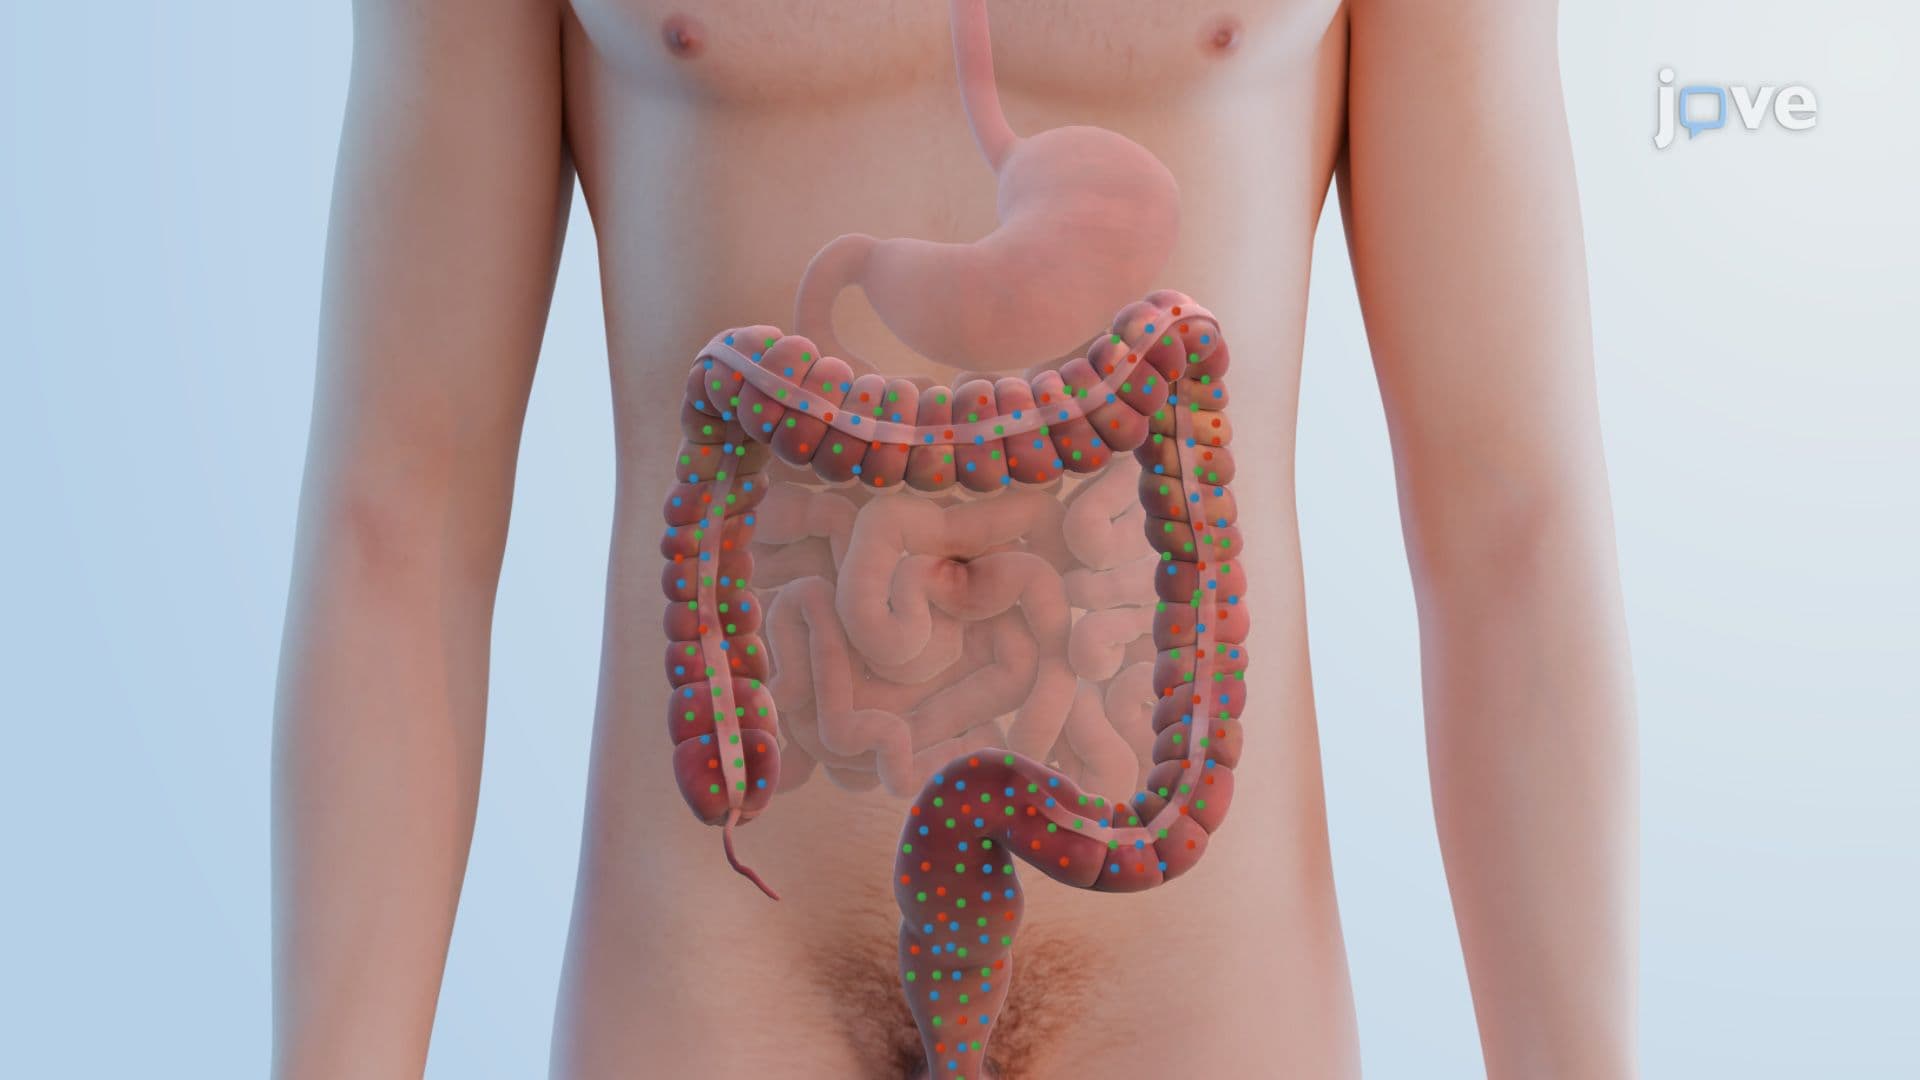
Bacterial Flora of the Large Intestine
The normal gut flora of the colon plays a critical role in generating essential vitamins such as vitamins K, B5, and B7.
 01:27
Microbiota of the Large Intestine
 01:26
Bacterial Phylum Bacteroidota
 01:29
Modern Molecular Taxonomy
 01:27
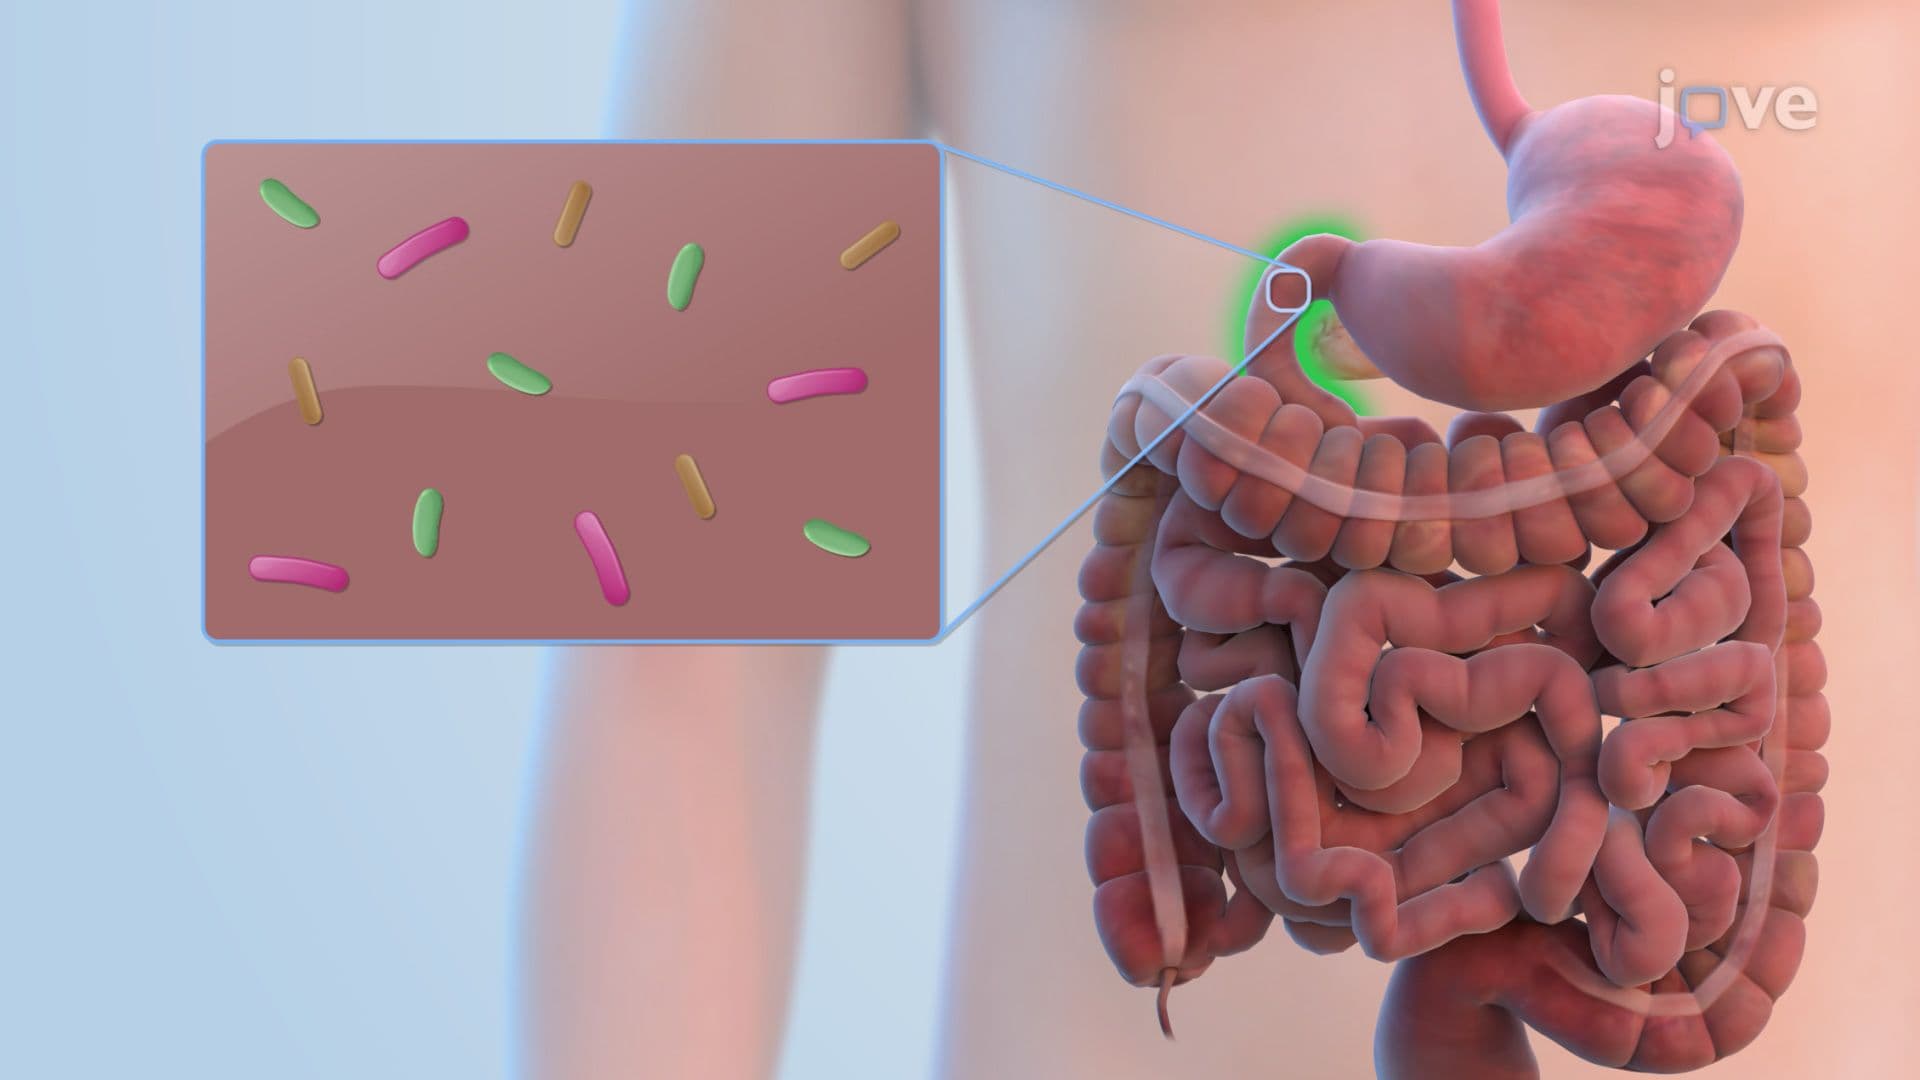
Microbiota of the Stomach and Small Intestine