Related Experiment Video
Updated: Aug 5, 2026

Genetically-encoded Molecular Probes to Study G Protein-coupled Receptors
Published on: September 13, 2013
A Ligand-Triggered Receptor Conformation Enables the Design of Selective Agonists for the Dopamine 3 Receptor (D3R)
Sandra Arroyo-Urea1, Antonina L Nazarova2,3, Alexander Knieb4
1Institute for Biocomputation and Physics of Complex Systems (BIFI) and Laboratory of Advanced Microscopy (LMA), University of Zaragoza, Zaragoza 50018, Spain.

Abstract:
While G protein-coupled receptors (GPCRs) represent the largest drug target family, designing subtype-selective molecules is still a challenge, especially to distinguish among closely related subtypes. One of the most challenging cases is the distinction between dopamine D2R and D3R, pivotal receptors in motor functions and cognition, and targets of Parkinson's disease treatments, schizophrenia, or substance use disorders. Attempts to design D3R-selective molecules with ligands binding toward the first transmembrane helix (the most sequence-diverse and conformationally flexible segment in GPCRs but rarely participating in ligand binding) allowed us to discover a ligand-induced ordering of TM1 unique to D3R, yielding an unexploited selectivity site for drug development. Using rational bitopic drug design and the ligand-triggered conformation of the D3R we designed, synthesized, and characterized the most selective D3R agonists to date, >100,000-fold more selective than available ligands. More specifically, we report D3R partial agonists AB12-82 (6d) and AB13-73A (11), with >575,000- and >750,000-fold subtype selectivity, picomolar potency, and 85% and 49% efficacy, respectively. We also present the most selective full agonists reported to date, AB13-08 (4b) and AB13-46A (9), presenting low and subnanomolar potencies with >2,800- and 6,300-fold selectivity for D3R. Overall, we introduce a first-in-class pharmacological toolbox to dissect the (patho)-physiology of D3R, open new avenues for the design of improved neurotherapeutics, and show that using ligand-induced TM1 reorganizations might represent a promising strategy for the design of subtype-selective molecules in other GPCRs.
More Related Videos
Related Concept Videos
 01:25
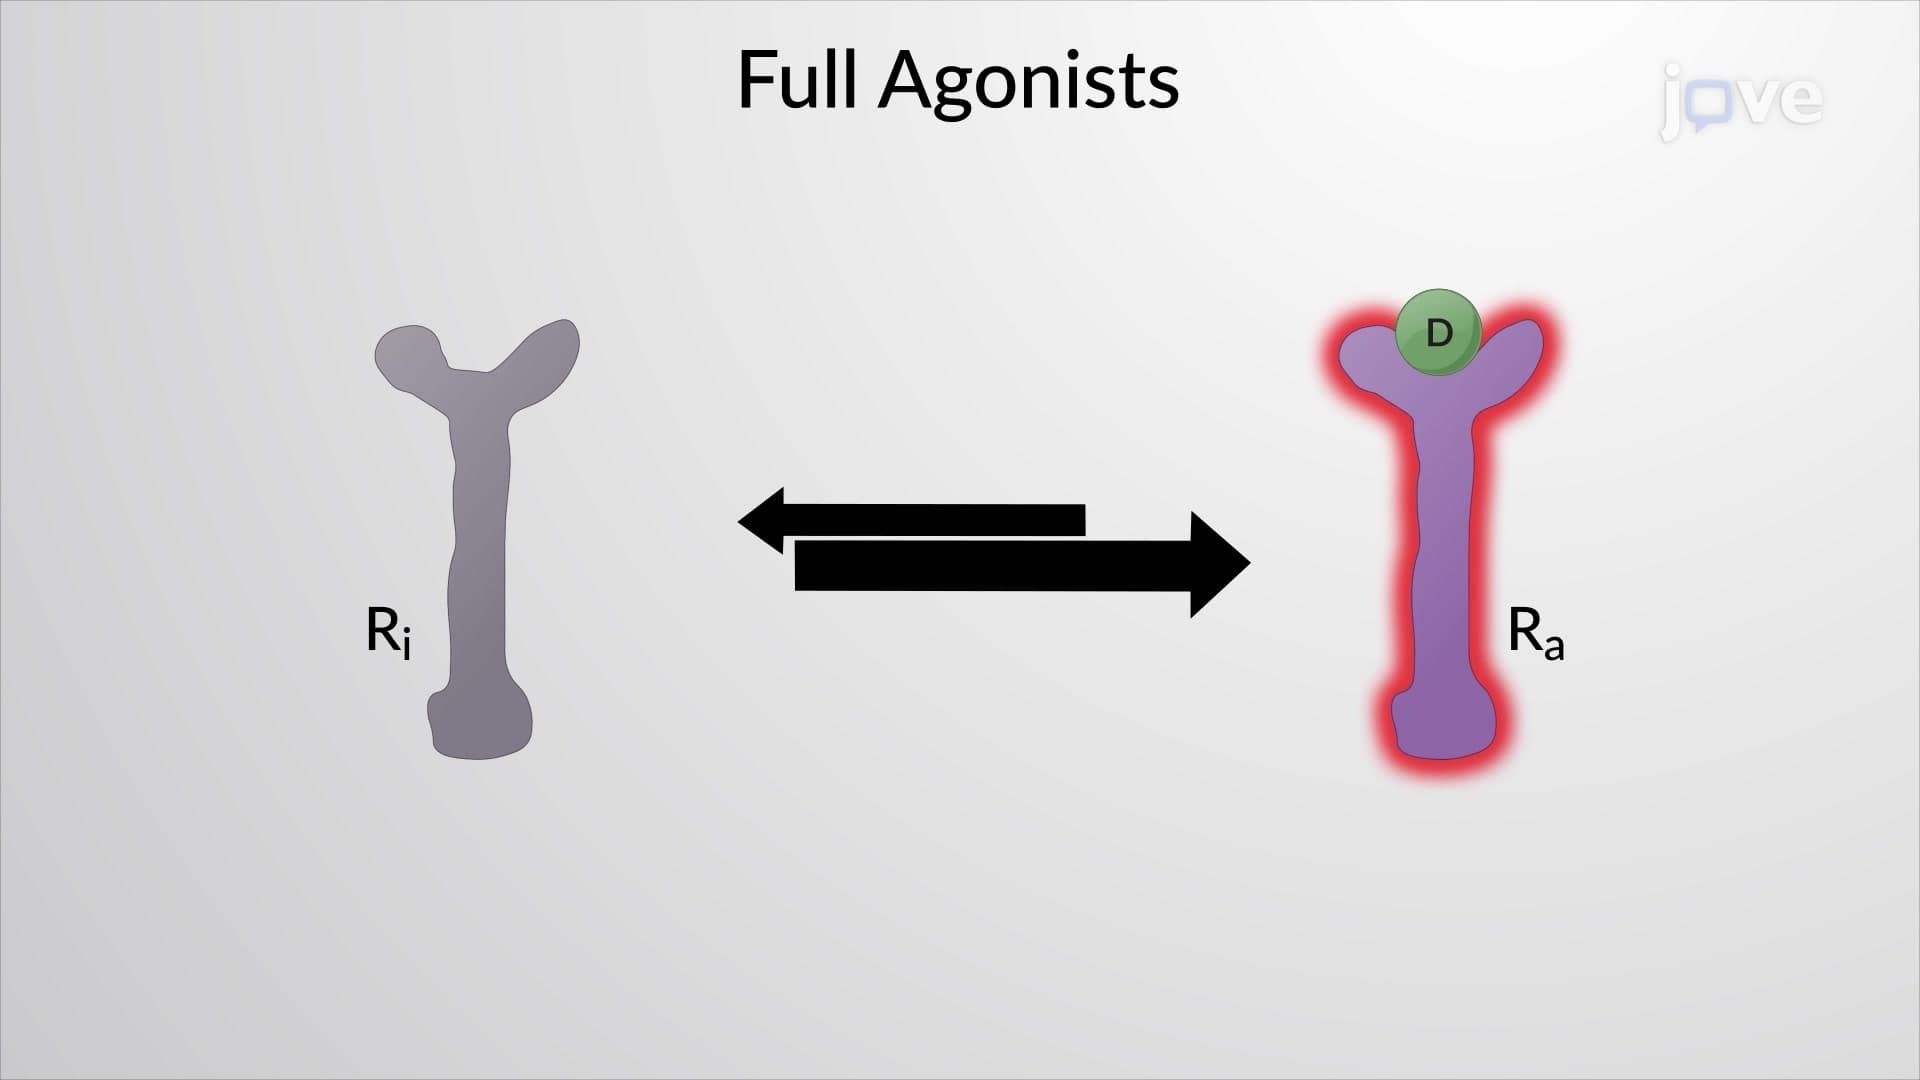
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:29
Drug-Receptor Interactions
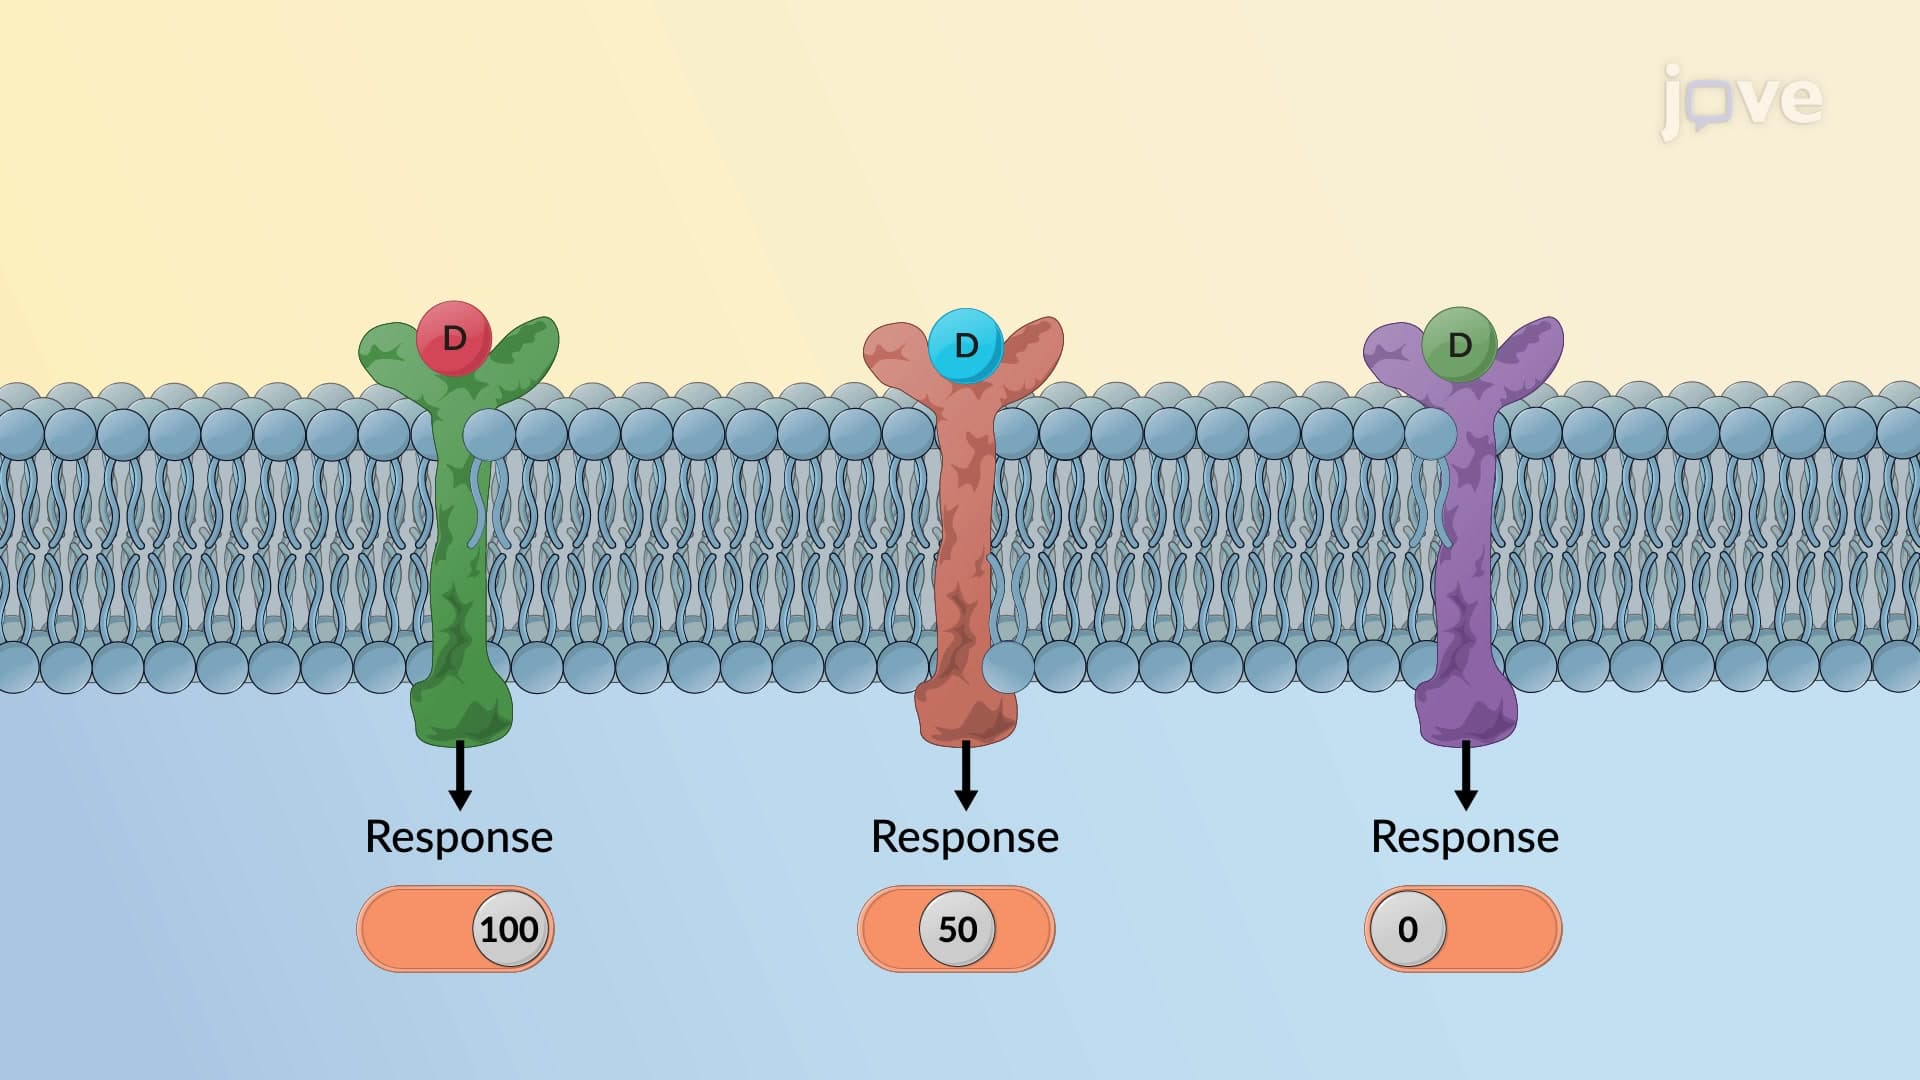
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:22
Opioid Receptors: Overview
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...