Related Experiment Video
Updated: Sep 16, 2026

In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
First-Principles Study on the Adsorption Mechanism of Oxygen on UN(100), (110), and (111) Surfaces
Tianyu Zhang1, Min Zhu1, Huang Huang1
1College of Nuclear Science and Technology, Naval University of Engineering, Wuhan 430030, China.

Abstract:
Uranium nitride (UN) is considered a promising candidate material for advanced nuclear fuels in Generation IV reactors due to its high thermal conductivity and excellent fission product retention capability. However, the surface corrosion behavior of UN in oxygen-containing environments severely limits its practical application. In this work, first-principles calculations based on density functional theory (DFT) were employed to systematically investigate the adsorption and dissociation behaviors of O2 molecules and O atoms on the UN(100), (110), and (111) surfaces. The electronic structure mechanisms underlying the adsorption were elucidated through analysis of surface work function, projected density of states (PDOS), Bader charge, and charge density difference. The calculated results show that O2 molecules undergo thermodynamically highly favorable dissociative chemisorption on all three UN surfaces at 0 K, with the O-O bond length stretched to 1.44-1.50 Å, characteristic of a peroxo-like (O22-) species, which is intermediate between the superoxide (O2-, ~1.33 Å) and complete dissociation. The most stable adsorption configurations on each surface are: (100)-H site (-4.05 eV), (110)-B(2) site (-4.81 eV), and (111)-H site (-5.03 eV), with the adsorption strength governed by the surface coordination environment. The adsorption energies of O atoms (-4.5 to -8.2 eV) are significantly higher than those of O2 molecules. The order of the most stable O atom adsorption sites is (110)-B(2) site (-8.22 eV) > (100)-H site (-6.97 eV) > (111)-T(U) site (-5.09 eV), which differs from that of O2, revealing the efficient O atom trapping effect of the groove structure on the (110) surface. Electronic structure analysis indicates that upon O atom adsorption, the work functions of the (100) and (110) surfaces increase by 0.62 eV and 0.39 eV, respectively, while that of the (111) surface decreases by 0.57 eV, suggesting that the (111) surface is more susceptible to further oxidation. In terms of bonding mechanisms, the interaction between O and U is primarily dominated by p-d hybridization between O-2p and U-6d orbitals. The U-5f orbital participates indirectly through f-d coupling with U-6d, while direct p-f hybridization is enhanced in the case of isolated O atom adsorption. This study provides an atomic-scale theoretical basis for understanding the initial oxidation mechanism of UN surfaces and offers important guidance for the surface protection design of UN-based nuclear fuels.

Related Concept Videos
 01:28
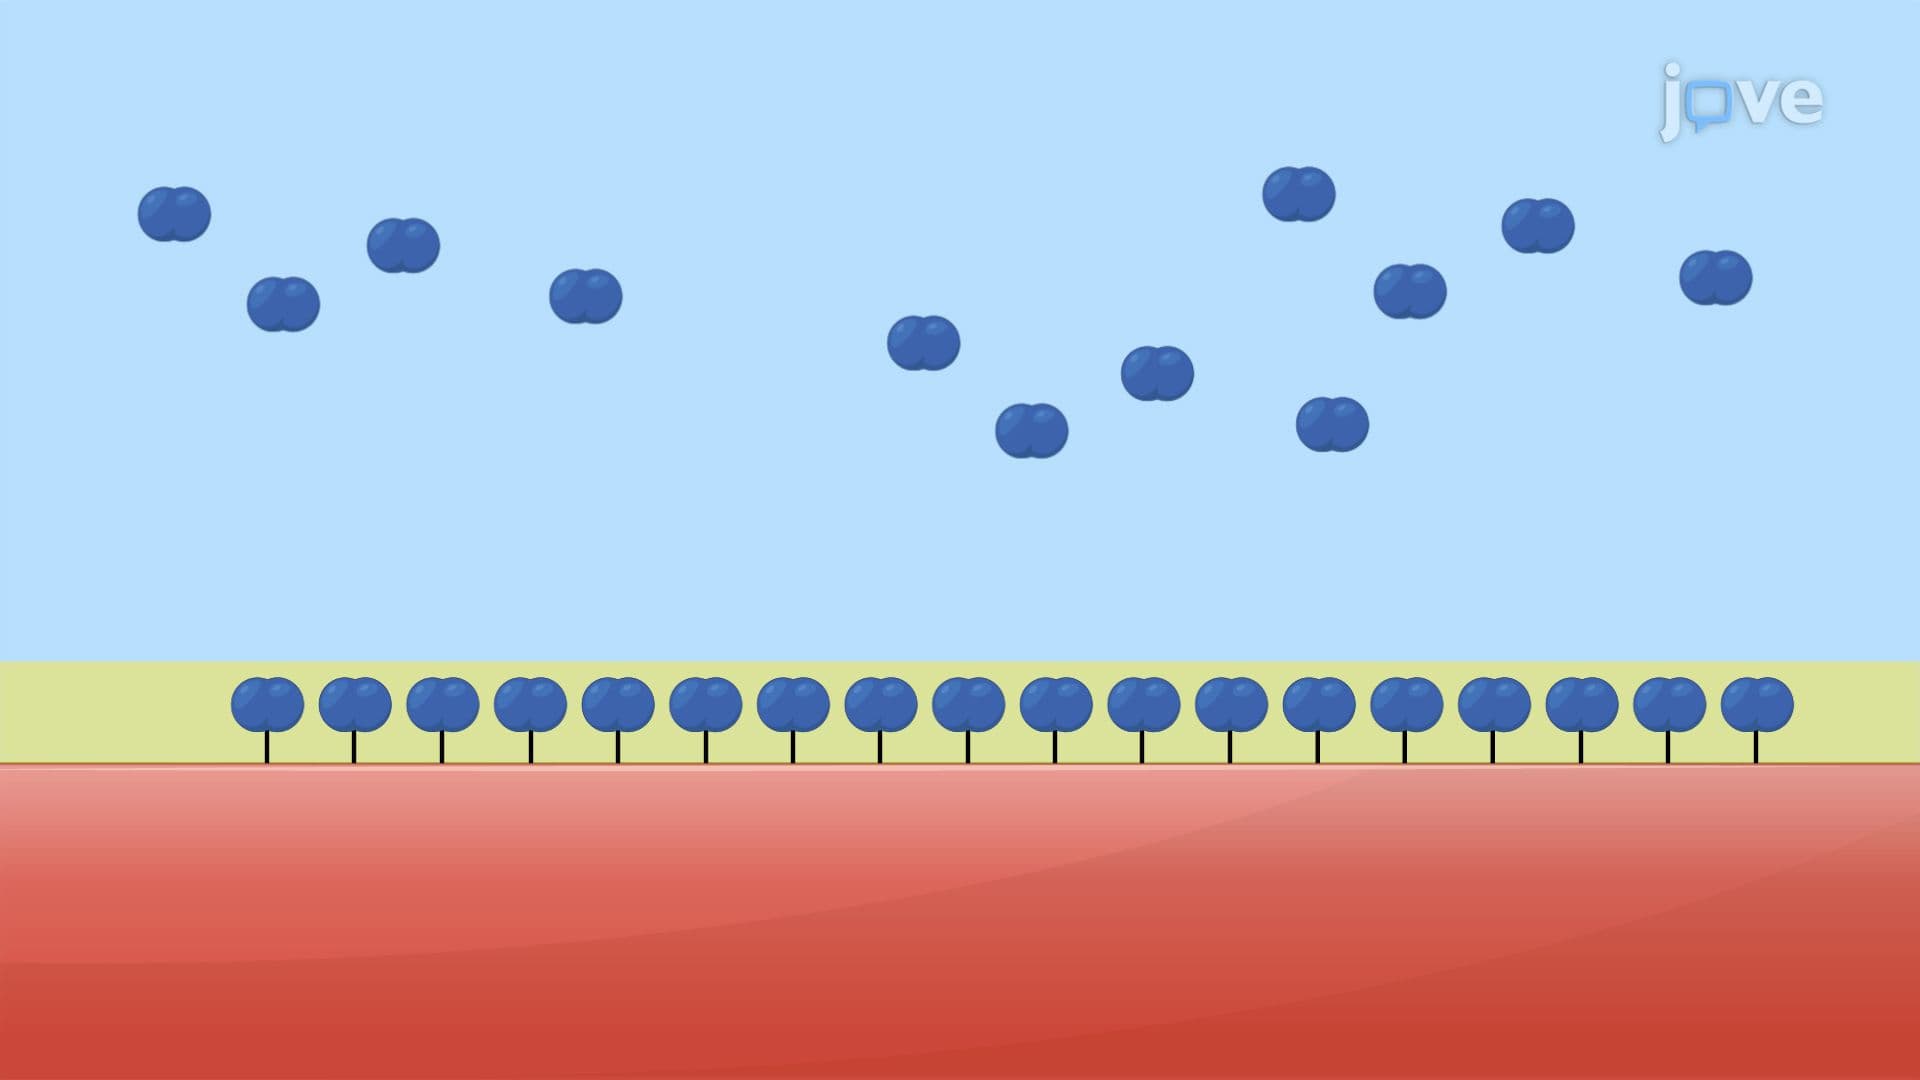
Adsorption of Gases on Solids
 01:29
Adsorption Isotherms I
 01:25
Adsorption Isotherms II
 01:22
Heterogeneous Catalysis
 01:16
Physical Principles Governing Gas Exchange
Gas Laws Governing Respiration
The behavior of gases is guided by Dalton's Law of partial pressures and Henry's Law.
Dalton's Law asserts that the total pressure exerted by...