Related Experiment Video
Updated: Sep 19, 2026
A Tandem Liquid Chromatography–Mass Spectrometry-based Approach for Metabolite Analysis of Staphylococcus aureus
Published on: March 28, 2017
Gut microbiota and fecal metabolomic alterations in hospitalized patients with Staphylococcus aureus infection: an
Yan Zhao1, Wei Chen1, Peiruo Chen1
1Department of Pulmonary and Critical Care Medicine, the Sixth Medical Center of Chinese PLA General Hospital, Beijing, China.

Introduction:
This study characterized gut microbial composition and fecal metabolomic profiles in hospitalized patients with Staphylococcus aureus infection (SAI) and explored their associations with inflammatory and organ-function indicators.
Methods:
This single-center exploratory observational study included 16 hospitalized patients with microbiologically confirmed and clinically adjudicated SAI and 10 healthy controls (HCs). The first qualified fecal sample was collected within 7 days after microbiological confirmation. Fifteen patients had received antimicrobial treatment before sampling, whereas one had not; none of the HCs had received antibiotics before fecal collection. Gut microbiota were profiled by 16S rRNA sequencing, and fecal metabolites were analyzed by LC-MS/MS-based untargeted metabolomics. Genus-level differential abundance was assessed using ANCOM-BC2 with Benjamini-Hochberg correction and pseudo-count sensitivity analysis.
Results:
Alpha-diversity indices were lower in the SAI group. Bray-Curtis PERMANOVA showed a statistically significant but modest group-associated difference in community composition (F = 2.8654, R² = 0.1067, P = 0.0001), whereas PERMDISP was not significant (P = 0.0885). ANCOM-BC2 identified four genera with robustly higher bias-corrected abundance in the SAI group: Corynebacterium, the [Clostridium] innocuum group, Enterococcus, and Eggerthella. Their effect directions remained positive in culture-confirmed and respiratory-infection-only analyses, although not all retained pseudo-count-robust significance. Untargeted metabolomics detected 2,956 features, including 2,491 with putative MS/MS-based annotations. Using VIP > 1 and raw P < 0.05, 381 candidate features were identified. Exploratory enrichment signals involved tryptophan, glutathione, sulfur, bile acid, amino acid, and lipid metabolism, but were not regarded as FDR-confirmed. No clinical-omics association remained significant after FDR correction.
Discussion:
Hospitalized patients with SAI showed group-associated microbial and metabolic differences compared with HCs. However, because nearly all patients were sampled after antimicrobial exposure, these patterns cannot be separated from treatment, hospitalization, disease severity, and other clinical interventions. The findings are therefore preliminary and hypothesis-generating rather than S. aureus-specific signatures.
More Related Videos
 09:49
Investigation of Microbial Cooperation via Imaging Mass Spectrometry Analysis of Bacterial Colonies Grown on Agar and in Tissue During Infection
Published on: November 18, 2022
 07:15
An In Vitro Batch-culture Model to Estimate the Effects of Interventional Regimens on Human Fecal Microbiota
Published on: July 31, 2019
Related Concept Videos
 01:22
Introduction to the Human Microbiota
 01:21
Microbiota Modulation by Antibiotics
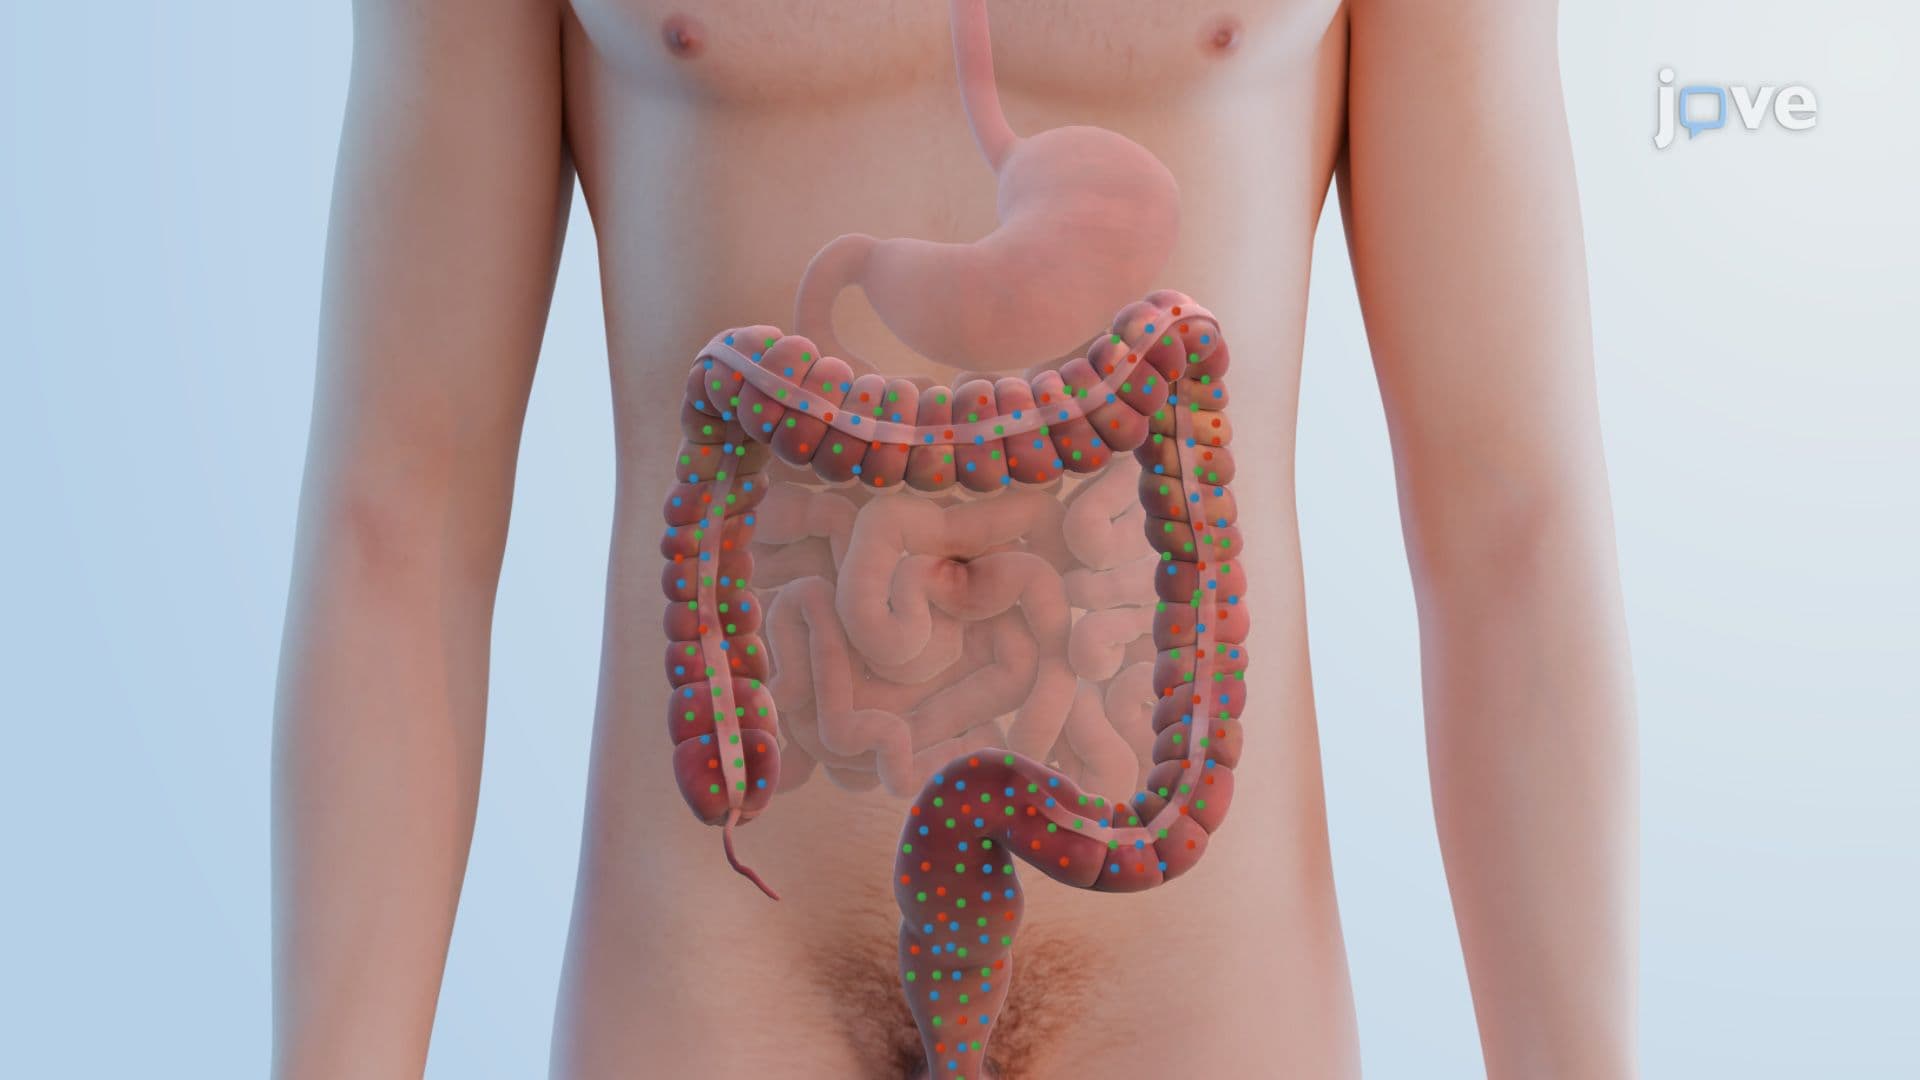 01:27
Microbiota of the Large Intestine
 01:30
Development of Human Microbiota
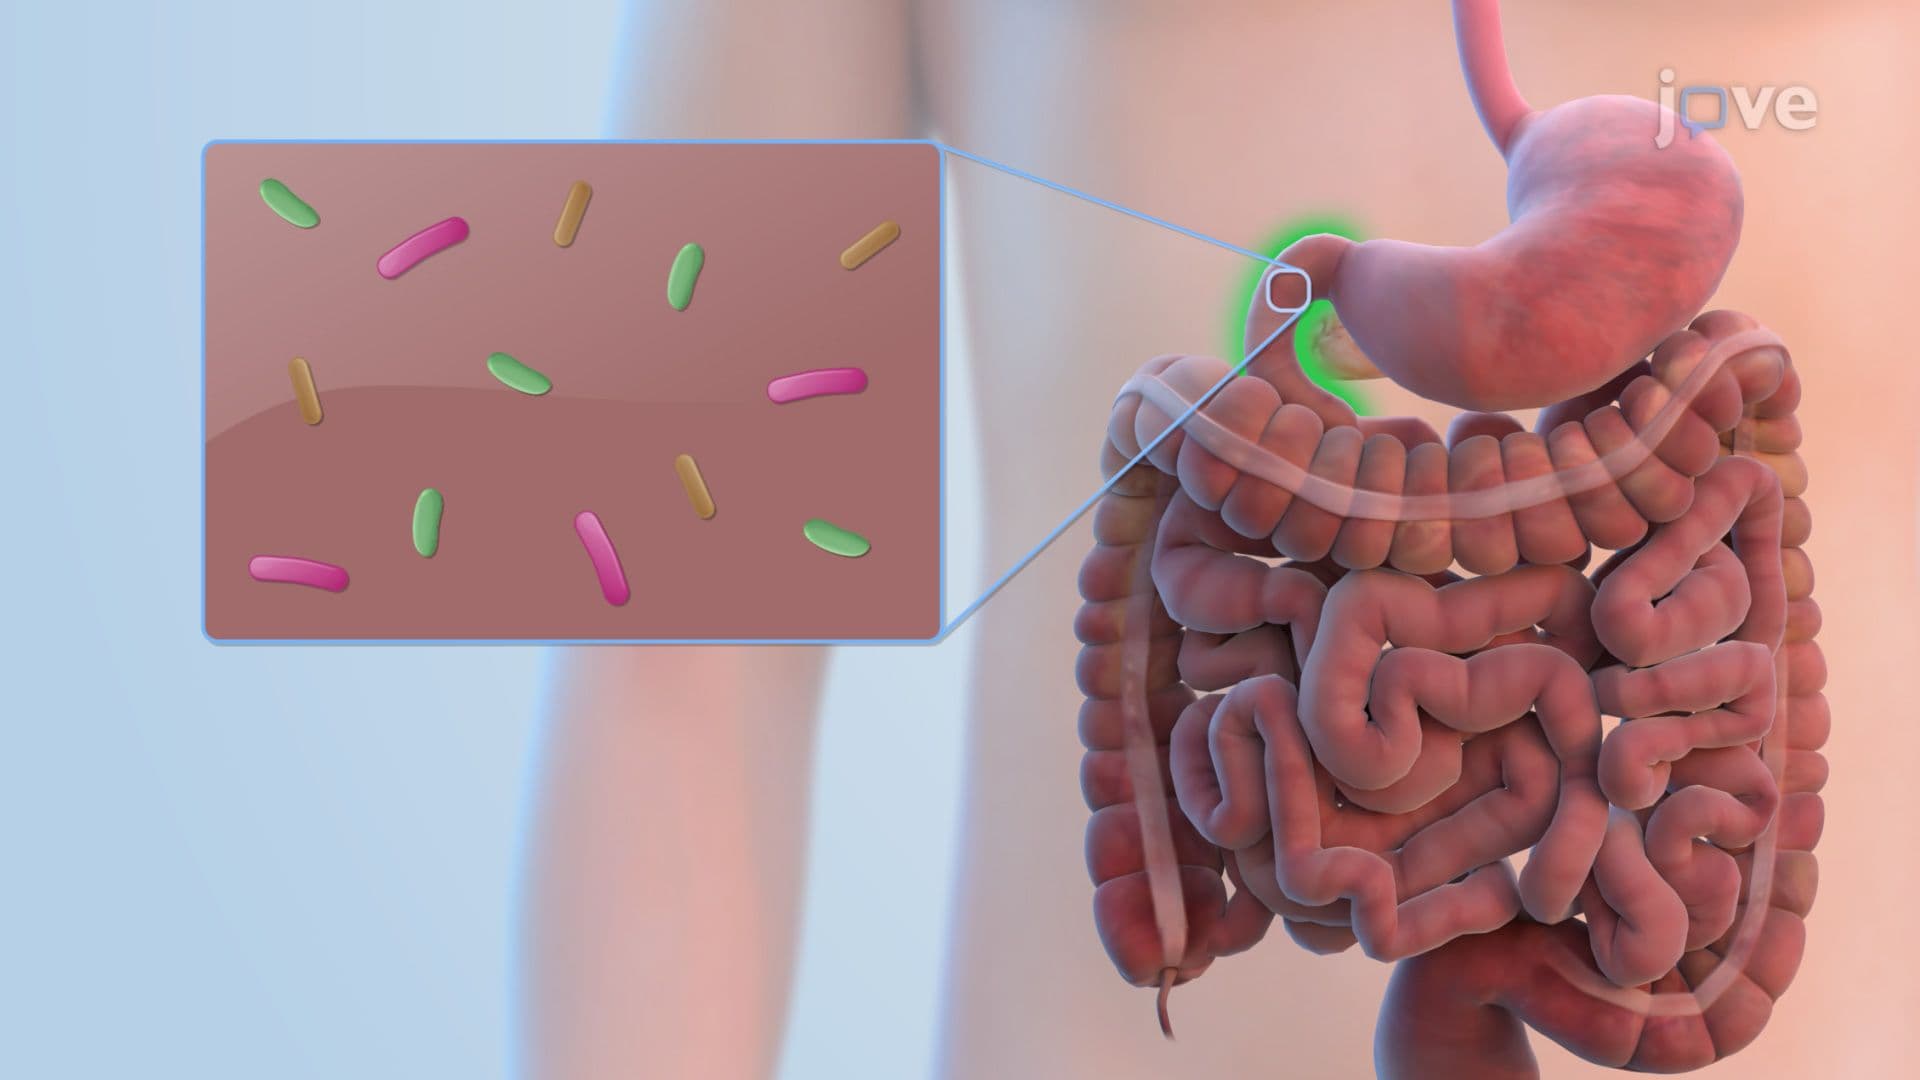 01:27
Microbiota of the Stomach and Small Intestine
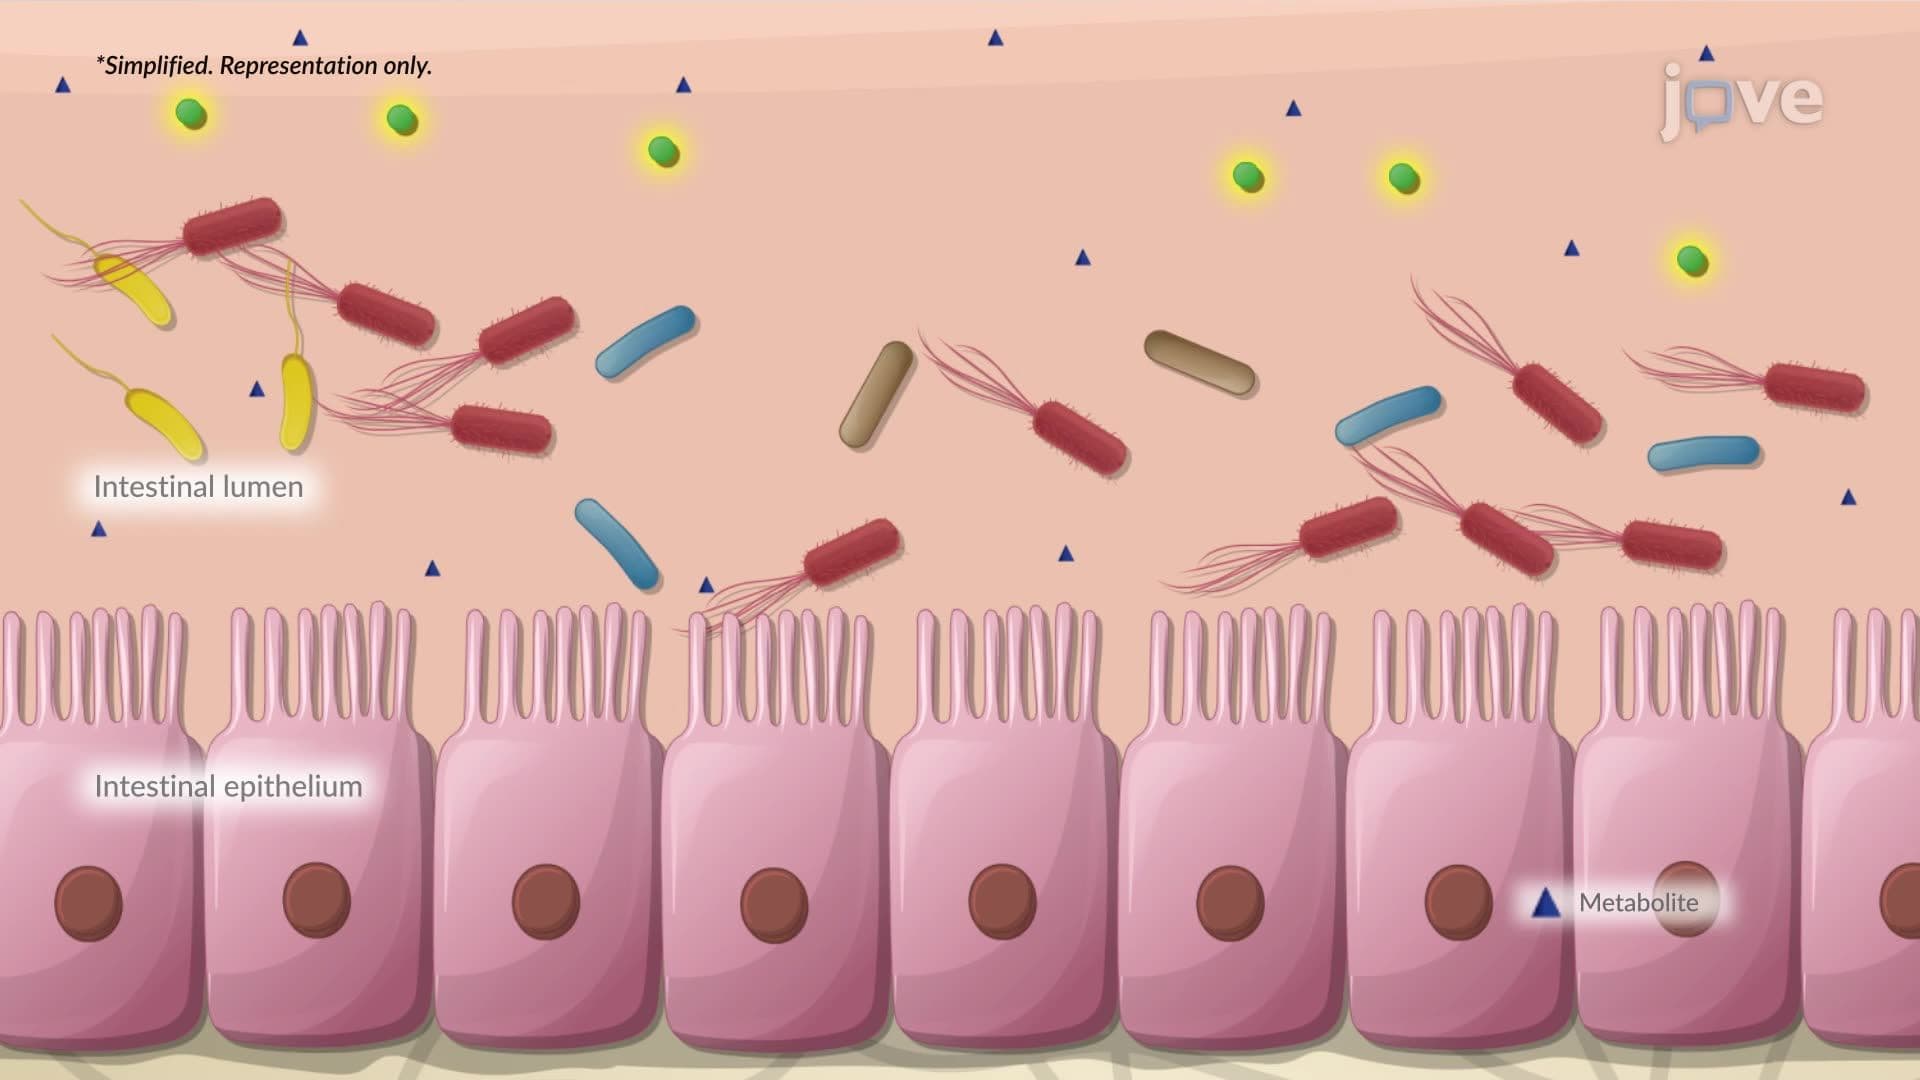 01:18
Dysbiosis of the Gut Microbiota