Related Experiment Video
Updated: Sep 27, 2026
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Assessment of the variational quantum eigensolver (VQE) for bond dissociation of H 2 , H 3 + , and CH 5 +
Amanda Marques de Lima1, Eivson Darlivam Rodrigues de Aguiar Silva2, Erico Souza Teixeira3
1Departamento de Química Fundamental, Universidade Federal de Pernambuco, Recife, PE, 50670-901, Brazil. amanda.aml002@gmail.com.

Context:
The variational quantum eigensolver (VQE) is a quantum-classical algorithm for estimating molecular ground-state energies. However, the accuracy depends on the choice of ansatz, active space, and initial variational parameters. Hence, it is relevant to perform assessments of VQE. The dissociation profiles of H , H , and CH were chosen for these assessments. For H and H , the UCCSD, GateFabric, and k-UpCCGSD ansätze reproduced the dissociation profiles with high accuracy. For CH , GateFabric and k-UpCCGSD provided the best compromise between accuracy and scalability. Hardware-efficient ansätze (HEA), particularly HEA-A and HEA-B, produced significant deviations from the reference energies. The results also demonstrate that larger numbers of parameters or logic gates do not necessarily improve VQE performance. Furthermore, some ansätze, especially GRSD and EHA, showed sensitivity to the initialization of the parameters.
Methods:
Reference geometries and energies were obtained at the CISD/STO-3 G level using Gaussian 09. VQE simulations were performed with the PennyLane library employing the Jordan-Wigner mapping. The ansätze evaluated were UCCSD, k-UpCCGSD, GateFabric, GRSD, HEA-A, HEA-B, HEA-C, and Entanglement-variational Hardware-efficient Ansatz (EHA). Dissociation profiles were generated by varying interatomic distances and angular coordinates for H and H , as well as the C-H dissociation coordinate for CH . Active-space ranging from 2/2 to 8/8 electrons/orbitals was employed for CH . Variational parameters were optimized using stochastic gradient descent with a convergence threshold of E . The VQE energies were obtained from averages over multiple runs using both zeros and random parameter initializations.
More Related Videos
 16:11
Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:36
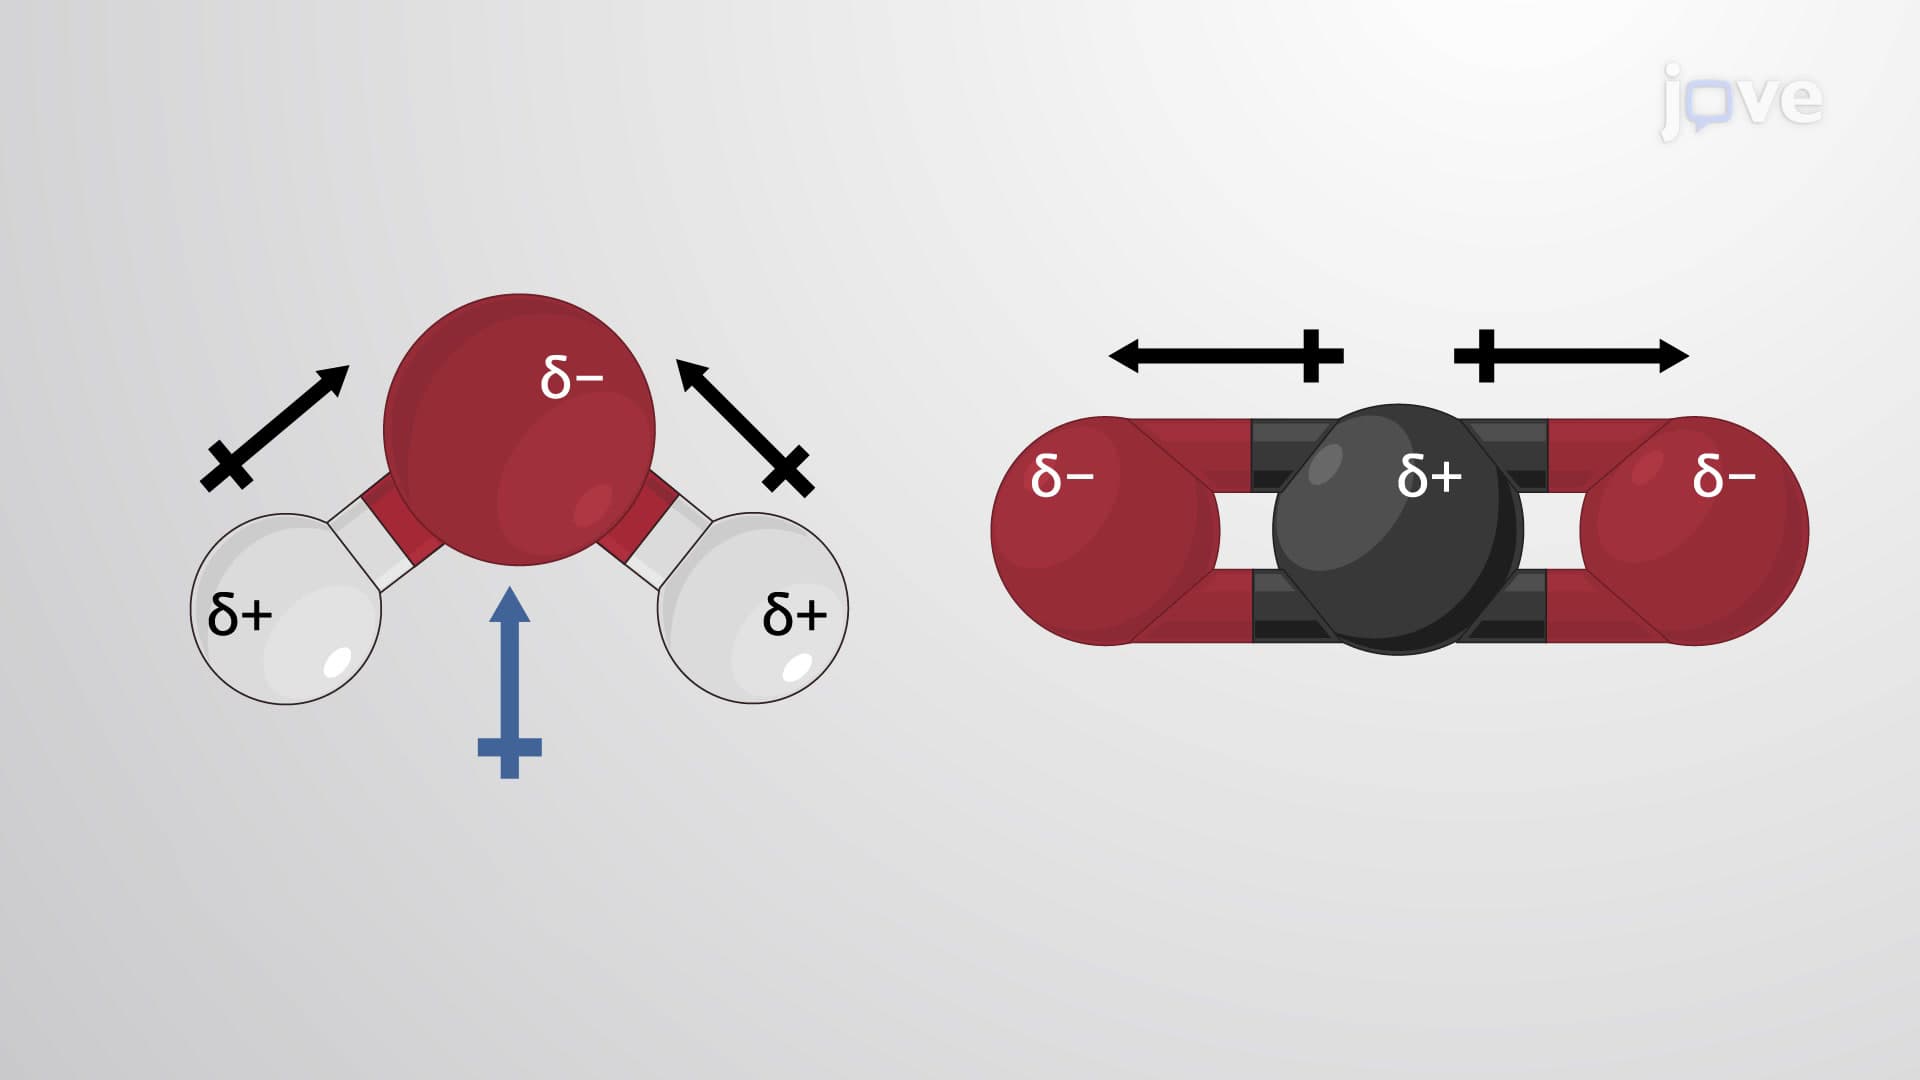
Molecular Geometry and Dipole Moments
 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 02:37
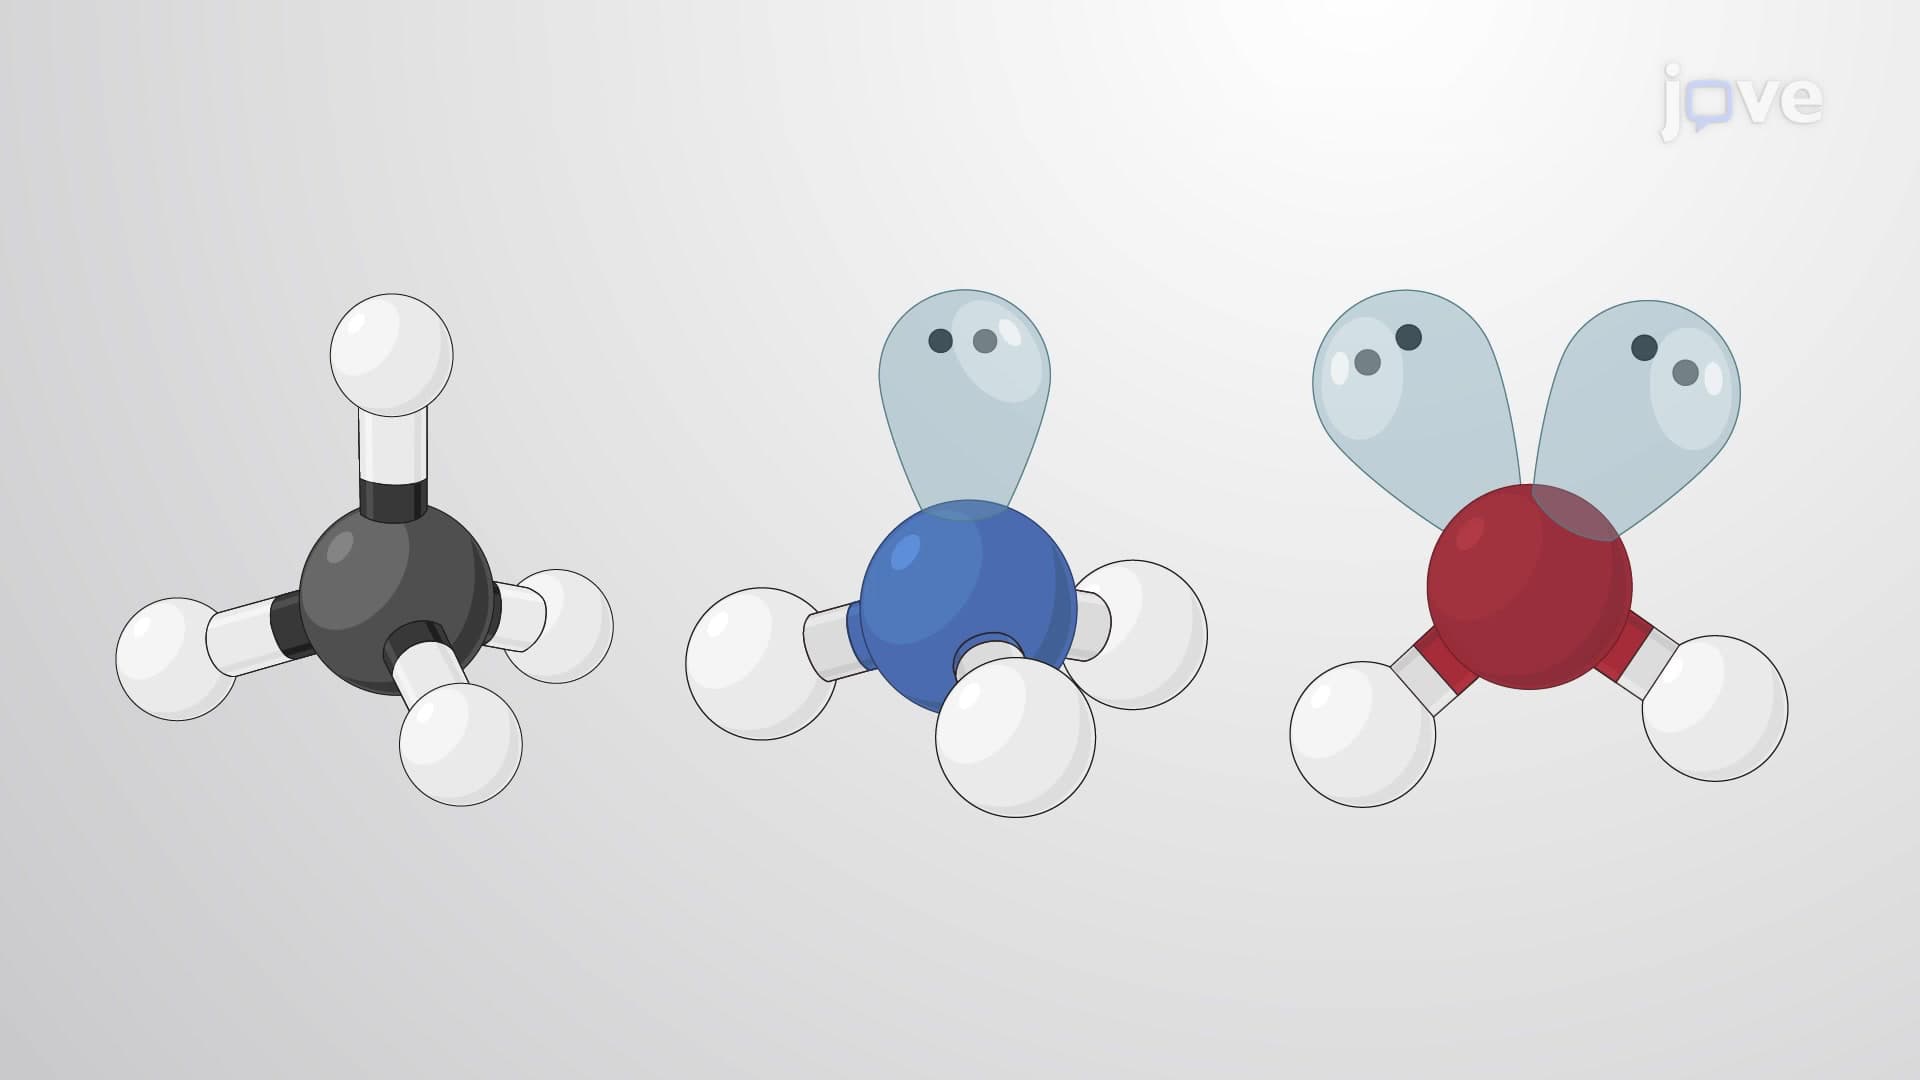
VSEPR Theory
 02:35
Molecular Orbital Theory I
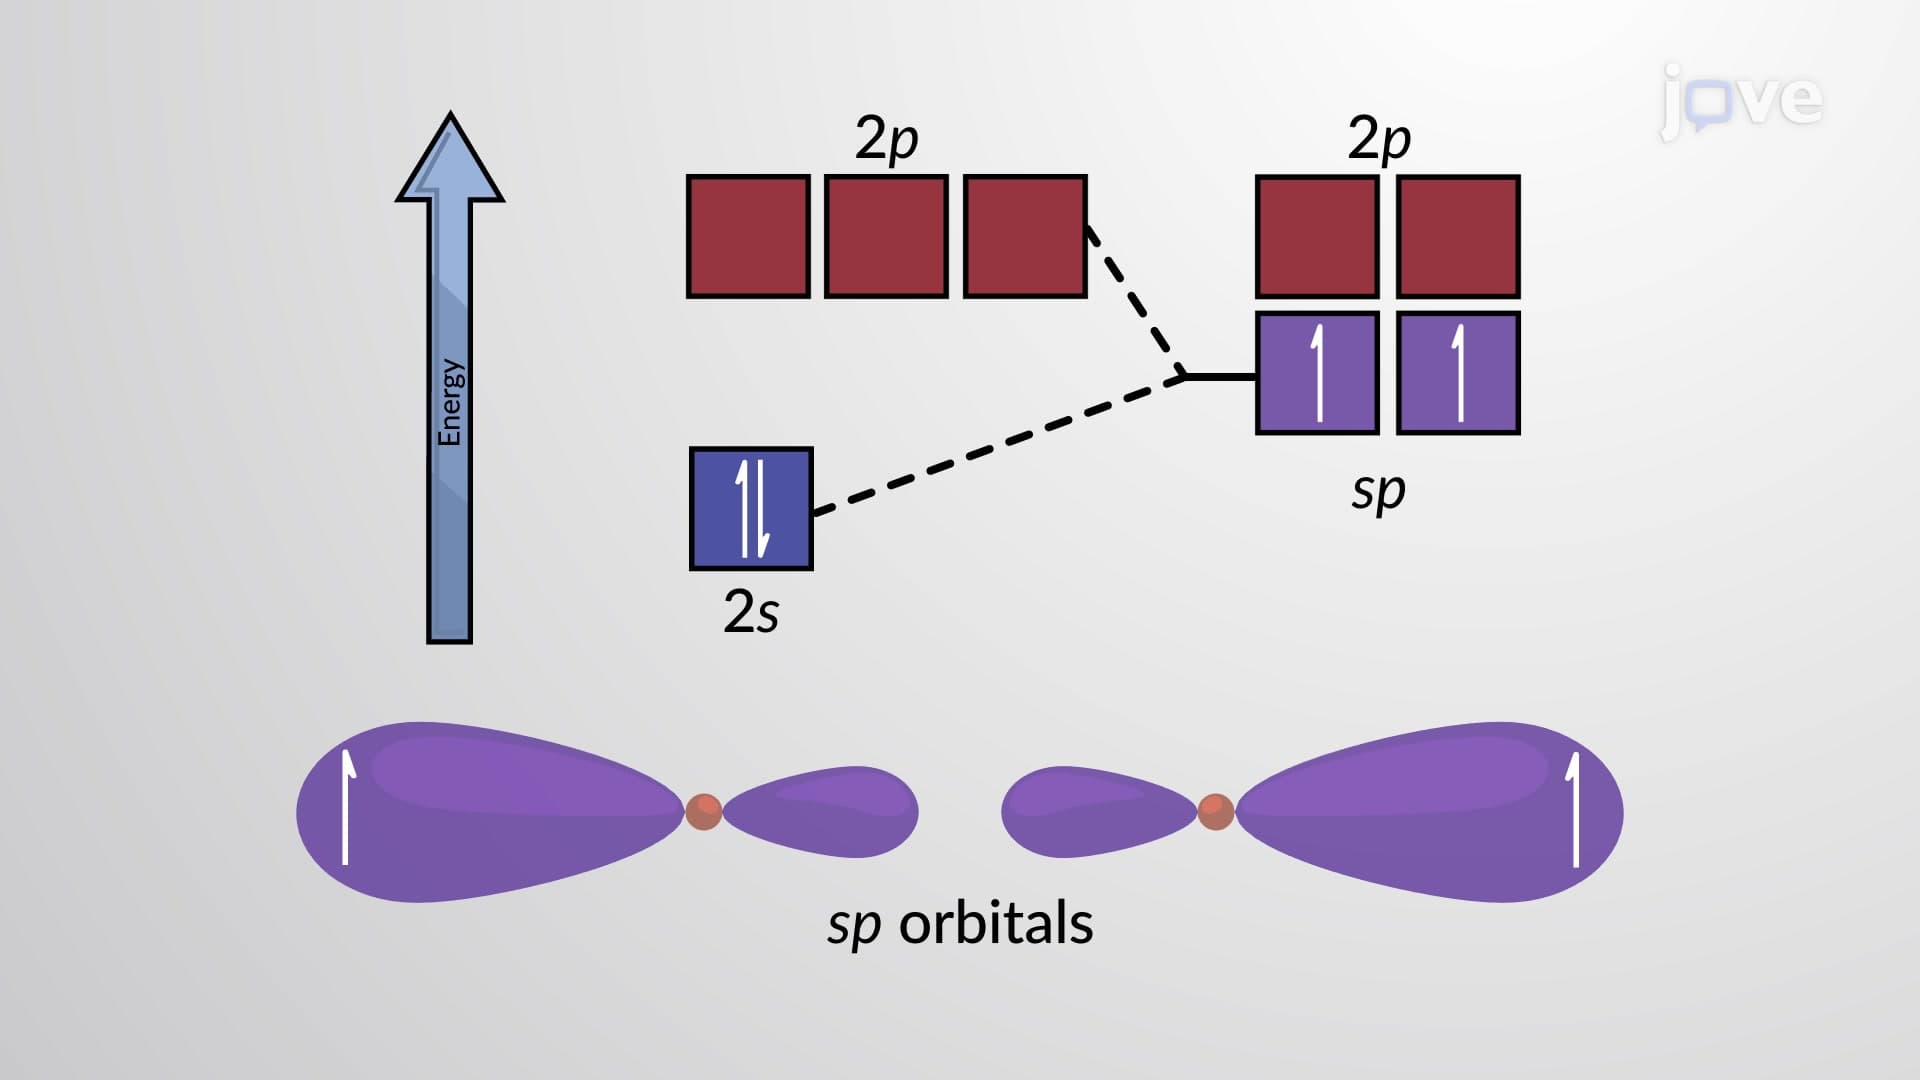 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...