Related Experiment Video
Updated: Oct 9, 2026
Preparation of Carbon Nanosheets at Room Temperature
Published on: March 8, 2016
Molecular dynamics study on surface adsorption of hexyl trimethylammonium bromide, octyl trimethylammonium bromide,
Wenyan Zhang1, Hao Yan1, Shuyan Liu1
1Department of Fundamental Physical Chemistry, School of Chemistry and Biological Engineering, University of Science and Technology Beijing, Beijing, 100083, China.

Context:
Molecular dynamics simulations were performed on four surfactants (hexyl trimethylammonium bromide, octyl trimethylammonium bromide, sodium hexyl sulfate, and sodium heptyl sulfate) to investigate their surface adsorption behavior in aqueous solution. In this work of simulations, surfactant molecules were initially distributed in both the surface and bulk regions of the solution, enabling them to form micelles spontaneously within the aqueous solution. The results show that the solute surface concentration and bulk concentration for both cationic surfactants (hexyl trimethylammonium bromide and octyl trimethylammonium bromide) and anionic surfactants (sodium hexyl sulfate and sodium heptyl sulfate) can be accurately described by a Padé approximant, which is consistent with the results of our liquid surface adsorption theory. The type of hydrophilic head groups and the length of hydrophobic tail groups affect the speed of surface adsorption. Molecular dynamics simulation demonstrated that the thickness of the surface layer in surfactant solution increases as the temperature and concentration increase. This is due to the weak intermolecular interaction between solvent molecules.
Method:
Molecular dynamics simulations were carried out in the NVT ensemble using the GROMACS software package. The interaction potential was chosen to be the Lennard-Jones 12-6 model, with long-range electrostatic interactions treated using the Particle Mesh Ewald (PME) method. A cutoff distance of 1.0 nm was taken for both non-bonded and electrostatic interactions. The V-rescale thermostat was used to realize a thermal equilibrium state at 298.15 K. The time steps were all taken as 2 fs. A typical total simulation time was 35 ns, of which the initial period was used to bring the system to equilibrium, and the remaining steps were used to dump the trajectories for subsequent analysis via VMD.
More Related Videos
 10:28
Probing the Structure and Dynamics of Interfacial Water with Scanning Tunneling Microscopy and Spectroscopy
Published on: May 27, 2018
 06:35
Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
Related Concept Videos
 01:29
Adsorption Isotherms I
 01:27
Surface Active Agents
 01:09
Analyte Adsorption and Distribution
 01:30
Micelles
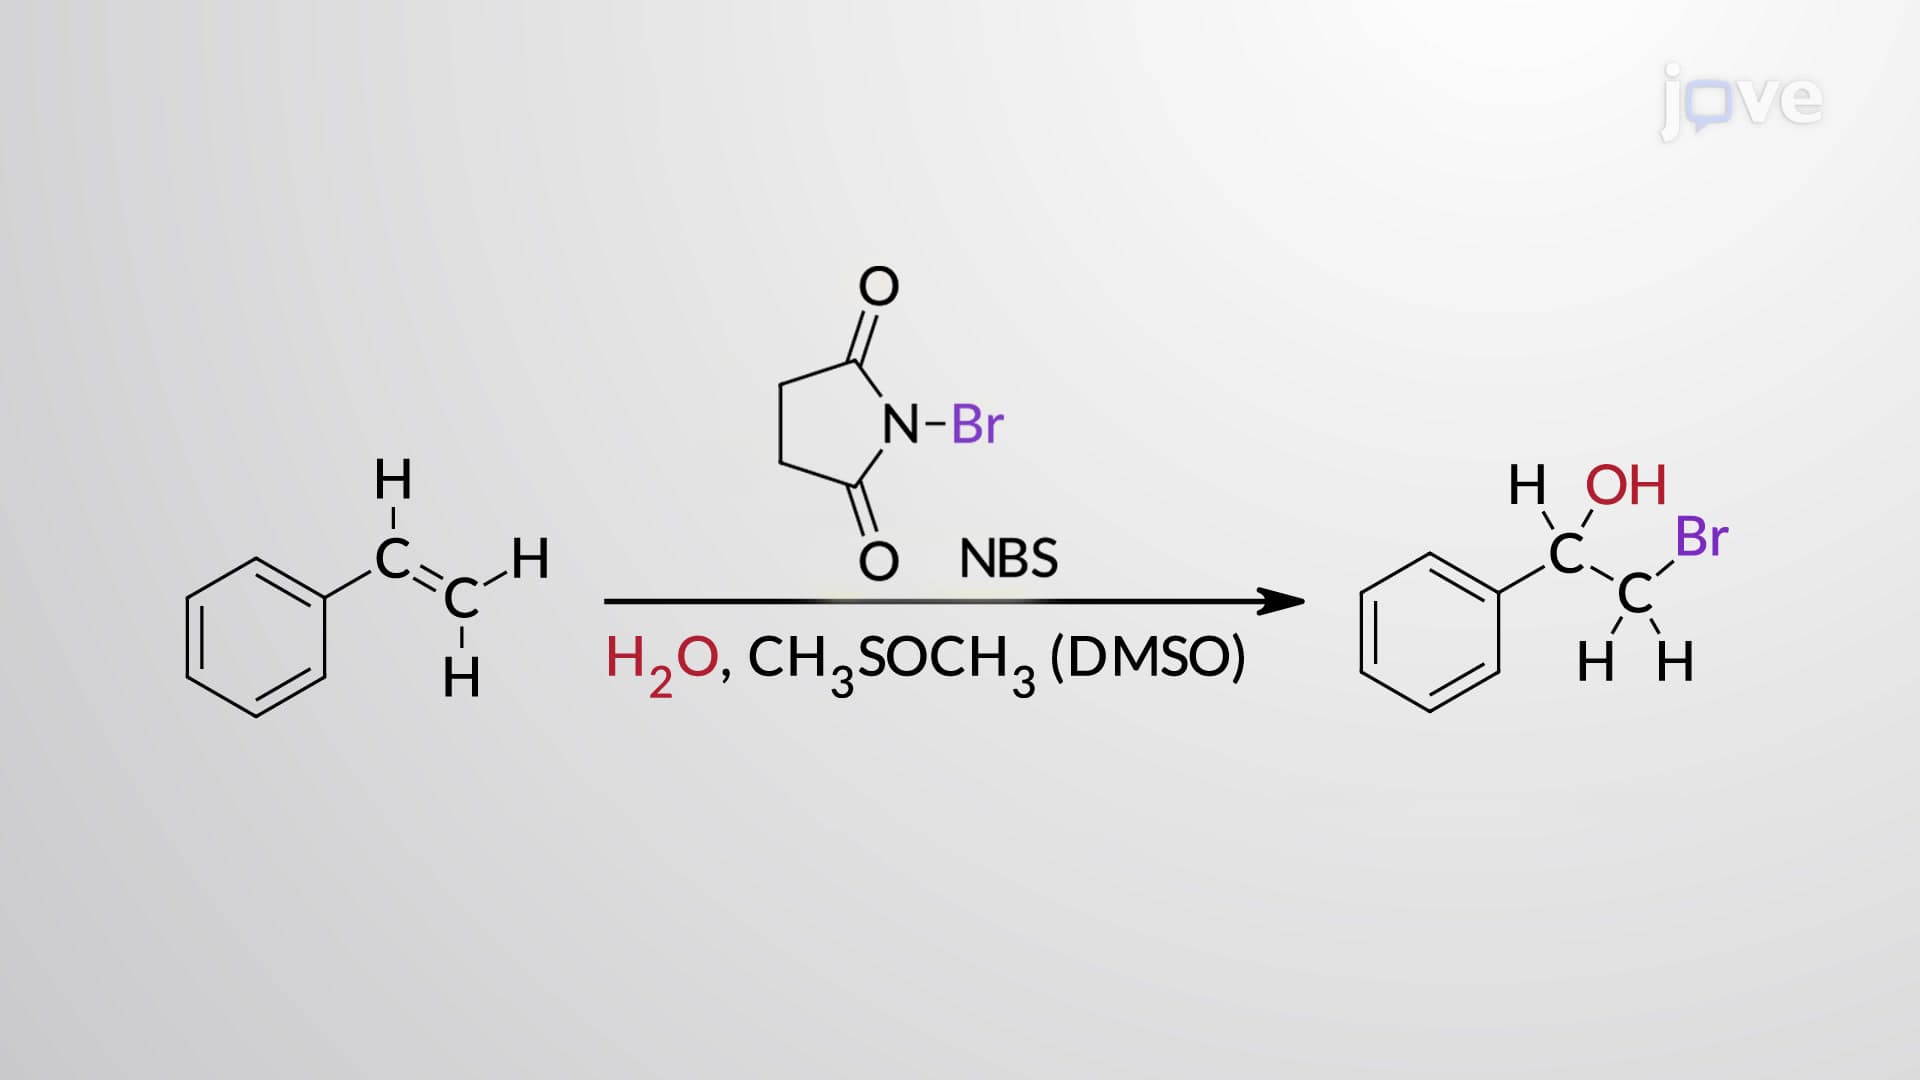 02:41
Formation of Halohydrin from Alkenes
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration