Related Experiment Video
Updated: Oct 13, 2026
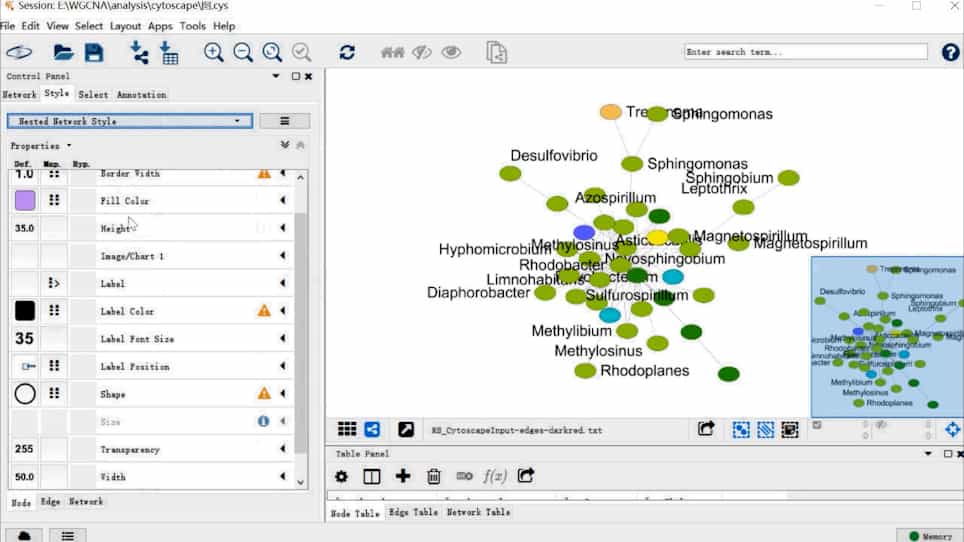

Divergence of Root Microbiota in Different Habitats based on Weighted Correlation Networks
Published on: September 25, 2021
Comparative transcriptional network analysis reveals ecological specialization in infant-associated species of
Felipe Melis-Arcos1, Masanori Arita2, Daniel Garrido1
1Department of Chemical and Bioprocess Engineering, School of Engineering, Pontificia Universidad Católica de Chile, Santiago, Chile.

Abstract:
Bifidobacterium species are important microorganisms in the infant gut microbiota displaying substantial genomic diversity, yet how this variability is reflected in transcriptional regulatory architecture remains an open question. Here, we integrated pangenome analysis with transcription factor binding site inference and network-based approaches to compare regulatory architectures across Bifidobacterium longum subsp. infantis, Bifidobacterium breve, and B. longum subsp. longum using over 300 high-quality genomes. All three species exhibited open pangenomes, with regulatory interactions frequently involving accessory genes. Inferred network metrics were compared with score-randomized controls across motif-score thresholds. Increasing score stringency reduced network size while increasing the number of detected communities, indicating greater fragmentation of the retained networks rather than a measured increase in modularity. NrdR showed an intermediate overlap in the Clusters of Orthologous Groups (COG) category sets represented among its predicted targets and species-dependent target prevalence. HMO-related regulators (FucR, NanR, and NagR) differed descriptively between Bifidobacterium infantis and B. breve, the two species included in the complete three-regulator comparison, in predicted target counts, represented COG category sets, and genomic-prevalence patterns; only FucR showed a significant difference in accessory-target fraction. These findings identify species-dependent differences in predicted regulon composition and genomic prevalence and provide testable hypotheses about how regulatory architecture may relate to ecological specialization in the infant gut.
Importance:
Bifidobacterium species are among the most abundant microorganisms in the breast-fed infant gut microbiota, where they display distinct patterns of genomic diversification and host association. However, the transcriptional regulatory mechanisms underlying these species-specific differences remain incompletely characterized. Here, we reconstructed and compared genome-scale transcriptional regulatory networks across three infant-associated Bifidobacterium species, integrating pangenome structure with transcription factor binding site inference across more than 300 genomes. Our analyses identified predicted regulatory interactions involving accessory genes and species-specific differences in network structure, regulon composition, and target prevalence. These associations provide hypotheses for experimental studies of how gene content and regulatory architecture may contribute to niche adaptation. This pangenome-scale framework can support the prioritization of regulatory hypotheses for future functional studies of infant-associated Bifidobacterium.
Related Concept Videos
 01:30
Development of Human Microbiota
 01:29
Coordination of Gene Expression Processes in Bacteria
 01:21
Microbiota Modulation by Antibiotics
 00:43
Bacterial RNA Polymerase
In most genes, the transcription site is a single base present upstream of the coding sequence. Though RNAP is a catalytically efficient enzyme, it does not recognize...
 01:18
Deep Sea Microbial Ecology
 01:28
Development of the Oral Microbiota