Related Experiment Video
Updated: Oct 14, 2026
A Syngeneic Orthotopic Osteosarcoma Sprague Dawley Rat Model with Amputation to Control Metastasis Rate
Published on: May 3, 2021
Rational Design and Evaluation of Novel ATR Inhibitors for Osteosarcoma Therapy: An Integrated 3D-QSAR, Molecular
Panchen Zhao1, Zheng Lian1, Xuexiao Ma1
1The Affiliated Hospital of Qingdao University, Qingdao, Shandong, China.

Background:
Ataxia Telangiectasia and Rad3-related protein kinase (ATR) is a central regulator of the DNA damage response and represents an attractive therapeutic target in tumors with high replication stress, including osteosarcoma. This study aimed to characterize the structure- activity relationships of a distinct ATR-targeting scaffold and to guide the rational design of new inhibitors using integrated computational approaches.
Methods:
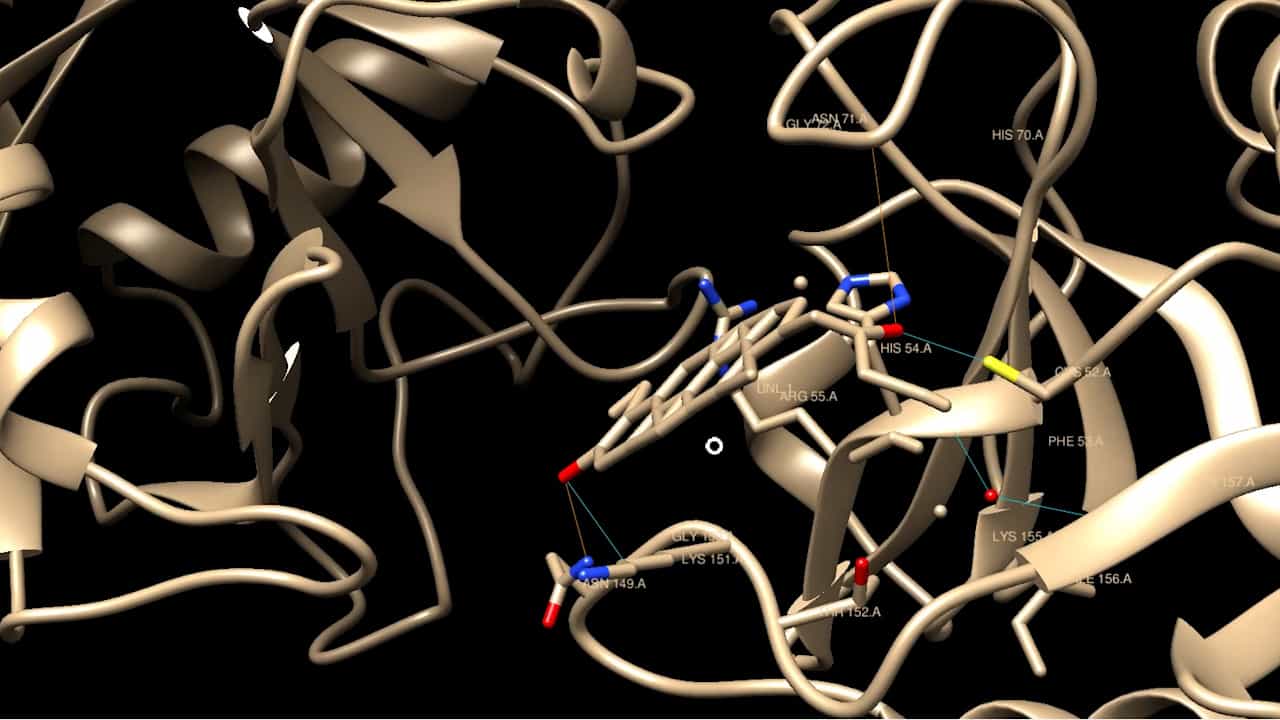
A dataset of 23 previously reported derivatives was used to construct a Comparative Molecular Similarity Indices Analysis (CoMSIA) 3D-QSAR model, with 18 compounds assigned to the training set and five to an external test set. Steric, electrostatic, hydrophobic, hydrogen-bond donor, and hydrogen-bond acceptor fields were evaluated. Model performance was assessed by leave-oneout cross-validation, Y-randomization, bootstrap resampling, external validation, Golbraikh-Tropsha criteria, rm 2-based metrics, and applicability-domain analysis. CoMSIA contour maps were then used to guide analogue design. Molecular docking was independently performed against both the engineered ATR-pocket surrogate 5UK8 and the recently resolved native human ATR structure 9L40 to assess the robustness of the predicted binding mode across distinct receptor models. The resulting 5UK8-19h-5 and 9L40-19h-5 complexes were subsequently subjected to molecular dynamics simulations to evaluate the dynamic stability and persistence of the predicted ligand-protein interactions.
Results:
The optimal CoMSIA model showed good statistical performance, with a cross-validated q² of 0.521 and a non-cross-validated r² of 0.996. External validation yielded an R² of 0.691 and a concordance correlation coefficient of 0.831, while randomization and bootstrap analyses further supported model stability. Electrostatic and hydrophobic interactions were the major contributors to activity, followed by hydrogen-bonding effects. Guided by the resulting structure-activity patterns, five new analogues were designed, of which 19h-5 showed the highest predicted activity (pIC₅₀ = 8.521). Cross-structure docking revealed a broadly conserved binding mode of 19h-5 in the 5UK8 and 9L40 binding pockets, and subsequent molecular dynamics simulations supported the stability of both predicted complexes.
Discussion:
The findings indicate that ATR inhibitory activity within this scaffold is governed by coordinated electrostatic, hydrophobic, and hydrogen-bonding features. Compared with previously studied aminopyrazine- and morpholinopyrimidine-based ATR inhibitors, the present scaffold provides a distinct structural framework and expands the chemical space available for ATR-targeted drug design. The concordance between the surrogate and native ATR models, together with the dynamic stability observed in molecular dynamics simulations, reduces the structural uncertainty associated with relying solely on the engineered 5UK8 receptor.
Conclusion:
Integrating CoMSIA-based 3D-QSAR modeling with cross-structure molecular docking and molecular dynamics simulations identified a scaffold-specific SAR framework and prioritized 19h-5 for further evaluation. The consistent structural behavior observed in both the ATR-mimetic and native ATR receptor models provides additional computational support for the proposed binding hypothesis, while experimental validation remains necessary.