Related Experiment Video
Updated: Oct 14, 2026
Protein Target Prediction and Validation of Small Molecule Compound
Published on: February 23, 2024
In silico Prediction of Protein Targets for Benzofuranone Ring-containing Agents and Molecular Docking Analysis
Fatma K Ata1,2, Zerrin Çalışkan1, Nehir Ö Özgentürk1
1Department of Molecular Biology and Genetics, Yıldız Technical University, Istanbul, Turkey.

Background:
Benzofuranone and its derivatives, which serve as drug precursors and intermediates due to their biological activity, are of great importance in the pharmaceutical industry. These heterocyclic molecules, composed of benzene and furan chains, possess antifungal, antiinflammatory, antidepressant, and anticancer properties. Given their pharmaceutical importance, mapping benzofuranone targets is a crucial step toward elucidating their molecular mechanisms of action and predicting potential side effects or cross-reactivity.
Methods:
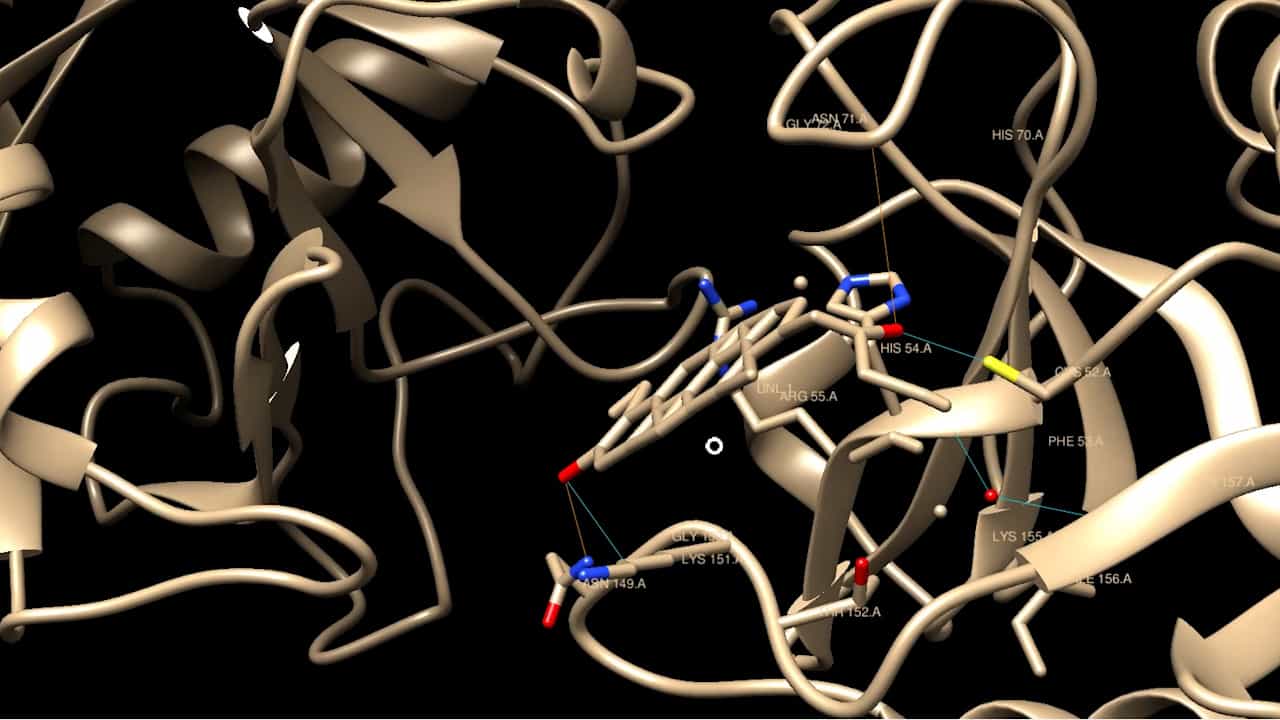
In this study, we evaluated the in silico pharmacokinetic properties and potential protein targets of 15 derivatives of 2(3H)-Benzofuranone. Chemical structures were constructed and optimized using Spartan software. Next, SwissADME and ProTox servers were used to analyze pharmacokinetic features, while SwissTargetPrediction and STRING/Reactome were used to predict protein targets. In the final stage, the binding affinities of benzofuranone-hybrid molecules for Homo sapiens CDK1 and PLK1 proteins were calculated using molecular docking.
Results:
Ligands M and N were classified as class IV toxicity and met all Lipinski, Ghose, Veber, Egan, and Muegge filter conditions. The estimated LD50 values (rat acute oral toxicity) of these ligands were found to be 2000 mg/kg. Protein target prediction showed that the M ligand interacts with CDK1 and PLK1 proteins responsible for microtubule stability and spindle fiber organization. Molecular docking results predicted favorable binding of ligand M within the CDK1 and PLK1 binding sites, with mean binding energies of -8.2 kcal/mol and -7.7 kcal/mol, respectively.
Discussion:
These results were comparable to those obtained for the reference CDK1 inhibitor Abemaciclib (-10 kcal/mol) and the PLK1 inhibitor Volasertib (-8.2 kcal/mol). Furthermore, structural comparison with the reference inhibitor Abemaciclib predicted a partially overlapping binding pattern in the CDK1 binding domain (Lys33, Ile10, and Leu135) of ligand M.
Conclusion:
In our study, we identify ligand M as a promising starting scaffold for derivatizing benzofuranone- based compounds. In silico information provided in this study may serve as a basis for future researchers to design, synthesize, and evaluate new derivatives of this ligand with improved biological properties.
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...