Related Experiment Video
Updated: Oct 25, 2026
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Accelerating Chemical Potential Calculations with Minimal Normalizing Flows
Philippe B Baron1, Athanassios Z Panagiotopoulos1
1Department of Chemical and Biological Engineering, Princeton University, Princeton, New Jersey08540, United States.

Abstract:
Chemical potentials are among the most important properties that can be obtained from a molecular simulation since they define many technologically relevant collective properties such as solubilities and activity coefficients. The chemical potential of a species in solution is typically obtained by computing the free energy change of adding that species into a bulk system, a calculation typically very expensive for systems of technological interest such as electrolytes, due to the lack of phase space overlap between "not-inserted″ and "inserted″ states. Recently, normalizing flows have been introduced as a way to accelerate free energy computations by learning a bijective function to map the configuration space of one Boltzmann distribution onto another. Currently, these trainable mappings are constructed to be as expressive as possible, theoretically having the ability to create a perfect mapping between states. This expressivity makes them difficult to train, limiting their ability to be generated "on-the-fly″ for any new free energy estimation challenge, and in practice these mappings have shown only modest performance improvements for liquid systems. We address these issues by introducing the concept of a "minimal″ normalizing flow (MNF). This is a trainable bijective mapping that is intentionally limited in expressivity and instead applies low-dimensional, physically informed transformations. Useful MNFs can be trained in approximately 1 min of GPU time due to their simplicity and our introduction of a novel loss function based on the Bhattacharyya distance as an alternative to the Kullback-Leibler divergence. We show how calculations of chemical potentials of pure and binary Lennard-Jones particle systems can be accelerated by at least an order of magnitude with a simple radial mapping. For a more complex illustration of the approach, we apply a two-dimensional (radial and orientational) mapping to the solvation of sodium and chloride ions in water, showing that MNFs can increase the effective sample size by as much as 3 times for charging free energy calculations and as much as 8 times for calculating free energy changes due to force field perturbations. This provides the foundation for the development of physically informed normalizing flows that can accelerate complex free energy calculations while retaining low training costs.

Related Concept Videos
 01:10
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
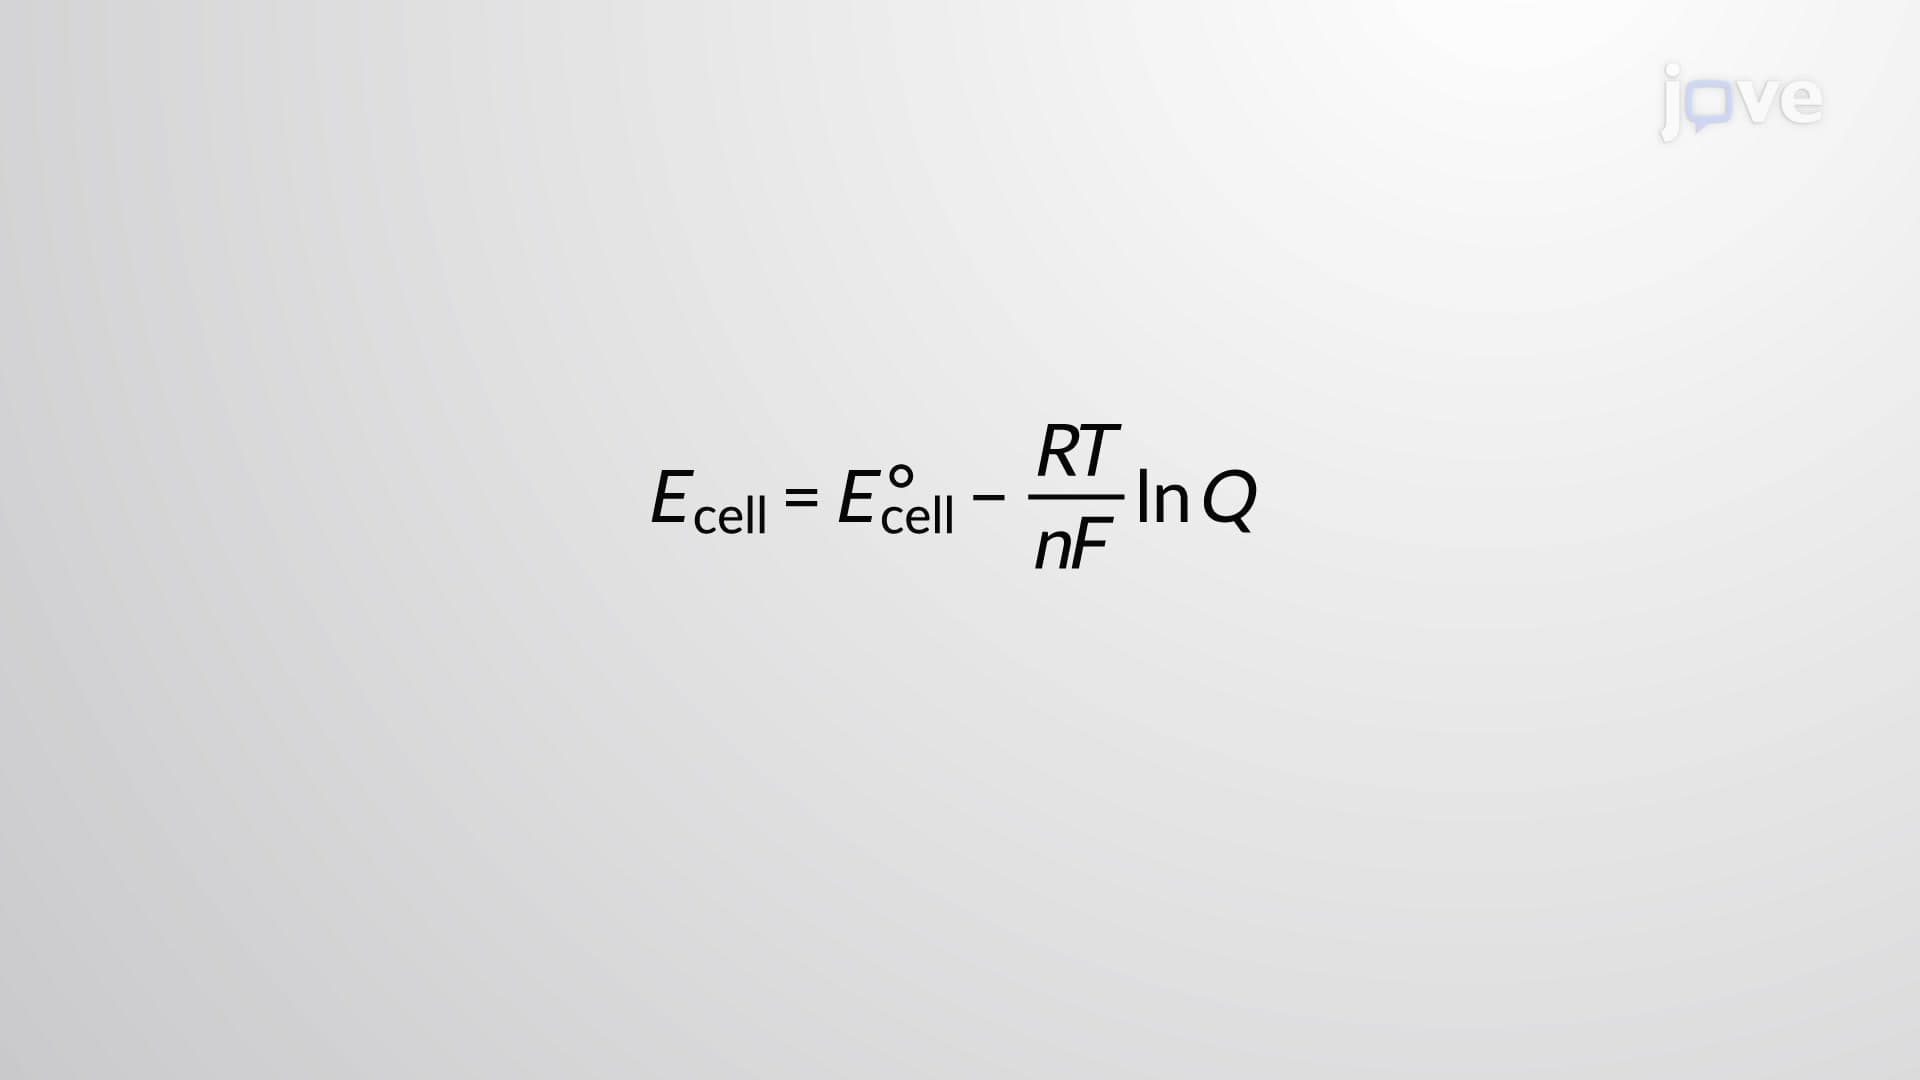 02:59
The Nernst Equation
The interconnection between standard cell potentials and various thermodynamic parameters such as the standard free energy change ΔG° and equilibrium constant K has been previously explored. For example, a redox reaction involving zinc(II) and tin(II) ions at 1 M concentration with Eºcell = +0.291 V and ΔG° = −56.2 kJ is spontaneous.
 01:26
Thermodynamic Potentials
 02:20
The Small x Assumption
 01:21
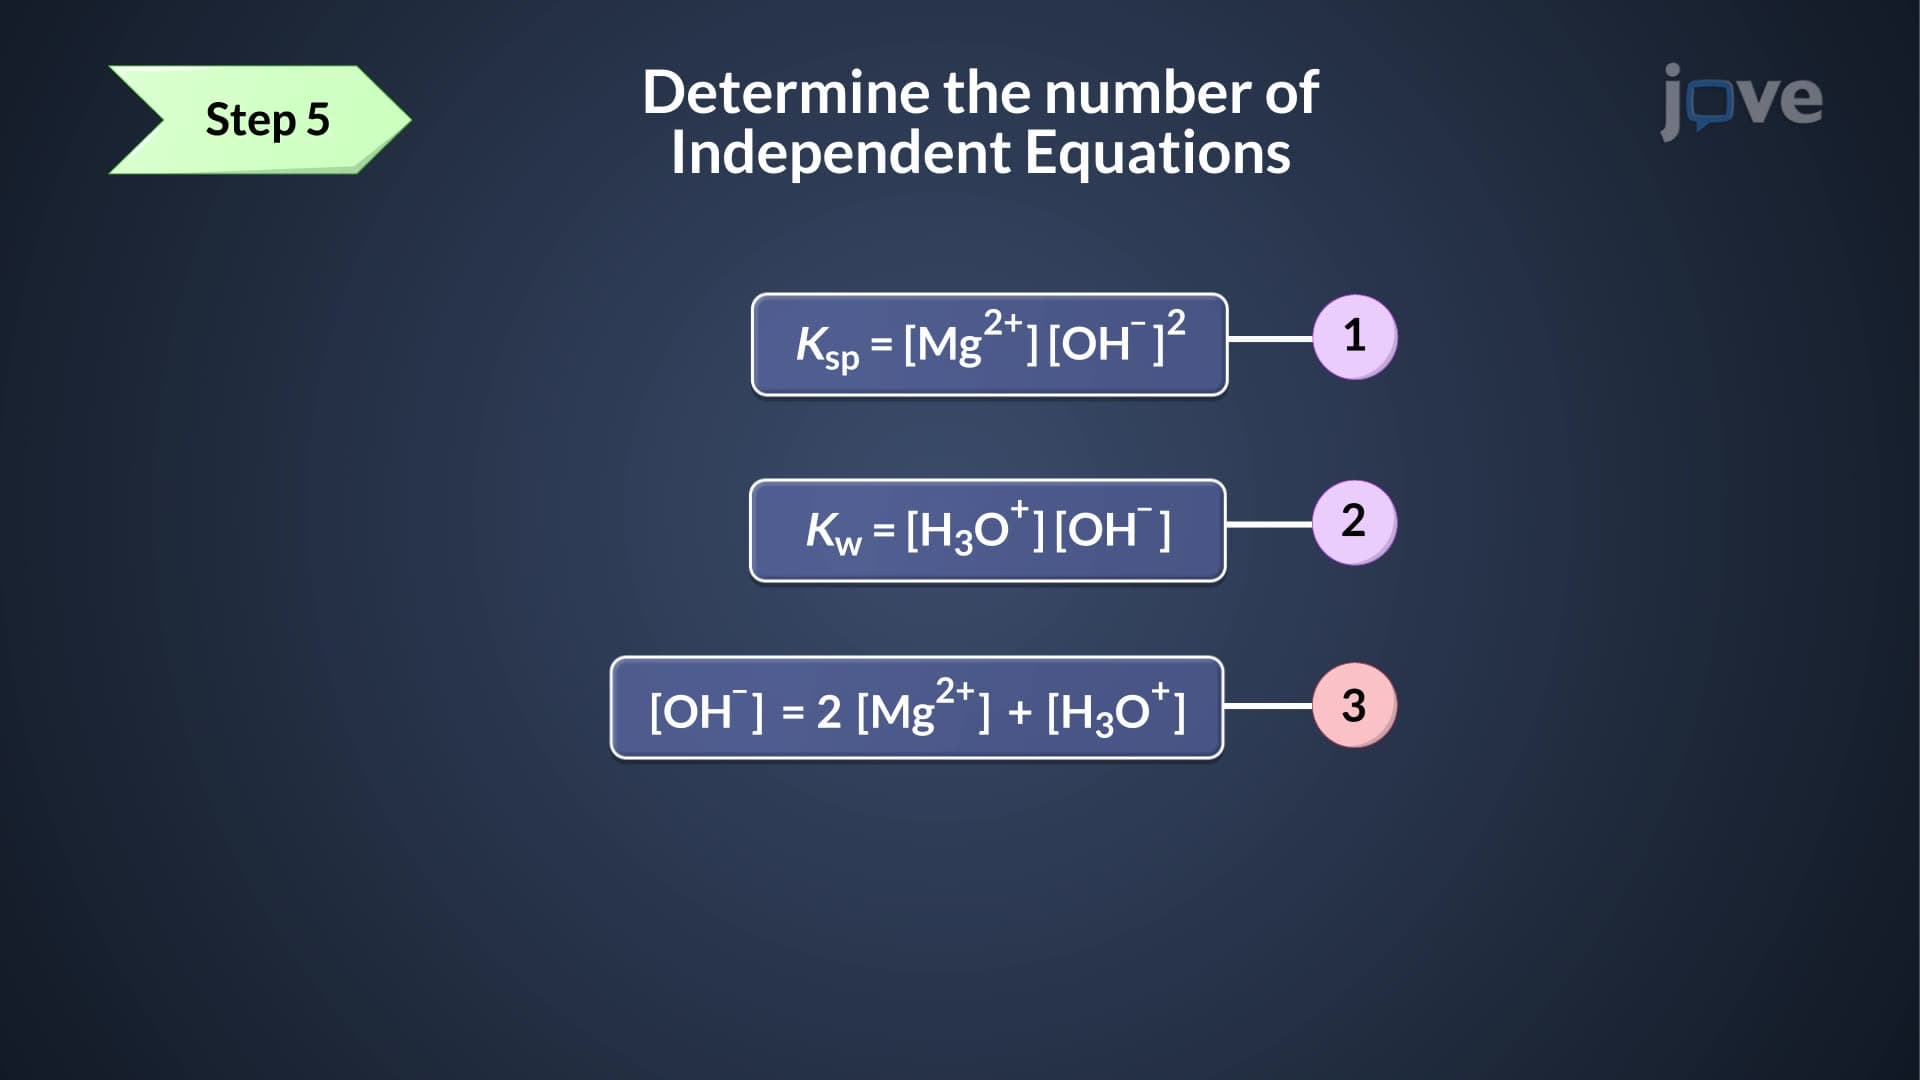
Chemical Equilibria: Systematic Approach to Equilibrium Calculations
The first step is to identify all the chemical reactions involved, The...
 01:17
Accelerating Fluids
The motion of the liquid within this infinitesimal cylinder is considered to obtain the pressure difference. Three vertical forces act on this liquid: